Related Experiment Video
Updated: Jul 23, 2026
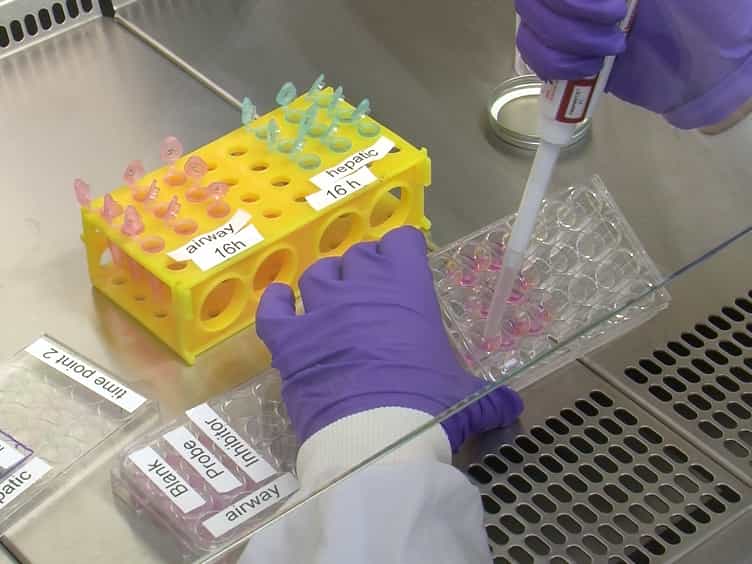

Mass Spectrometry and Luminogenic-based Approaches to Characterize Phase I Metabolic Competency of In Vitro Cell Cultures
Published on: March 28, 2017
Metabolism of carbamazepine by CYP3A6: a model for in vitro drug interactions studies
E Mesdjian1, E Sérée, B Charvet
1Centre Saint Paul et Laboratoire de Biologie Moléculaire appliquée au médicament, Marseille, France.

Abstract:
Carbamazepine (CBZ) is widely used in the treatment of epilepsy. The drug is principally metabolized by CYPs to 10, 11-epoxy carbamazepine (CBZ-E) but this metabolite more toxic than the parent drug, does possess anticonvulsant properties. In humans, CYP3A4, CYP2C8 and CYP1A2 have been shown to be implicated in CBZ biotransformation. Our purpose was to establish an experimental model to determine the interaction of CBZ with other antiepileptic drugs. We first identified the CYP isoforms that metabolized CBZ in rabbit. We used liver microsomes from rabbit treated with various compounds known to induce principally some CYPs subfamilies. Having tested all the compounds we demonstrated that only the animals treated with CYP3A inducers were able to metabolize CBZ strongly. The CBZ biotransformation was inhibited by anti CYP3A antibodies. All the CYP3A subfamily substrates specifically decrease CBZ-E formation. In our experiment we did not observe any inhibition with CYP2C substrate. These data provide evidence that in rabbit the CYP3A subfamily is primarily involved in CBZ metabolism. Using this model we investigated the interaction of CBZ with phenobarbital, phenytoin, ethosuccimide, primidone, progabide, vigabatrin and lamotrigine.
Related Concept Videos
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion, mediated...
 01:25
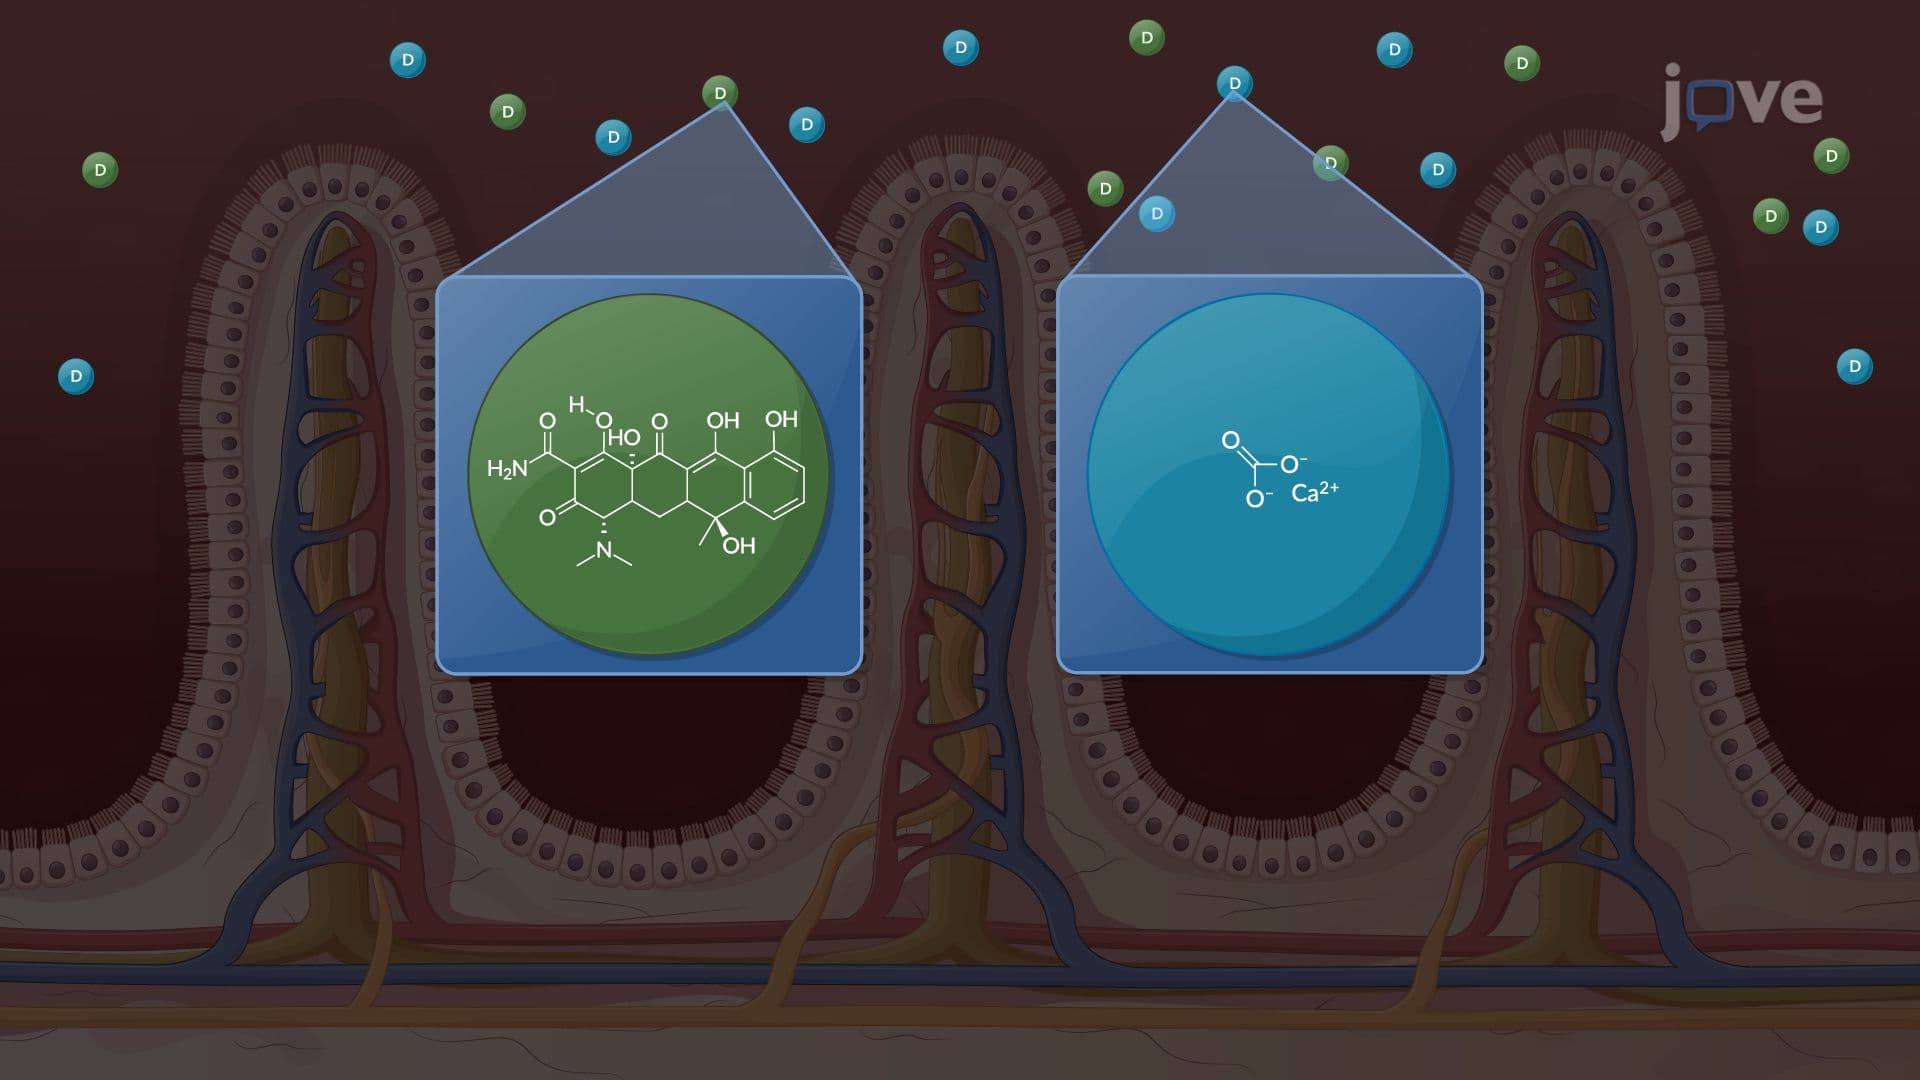
Pharmacokinetics: Drug–Drug Interactions
 01:26
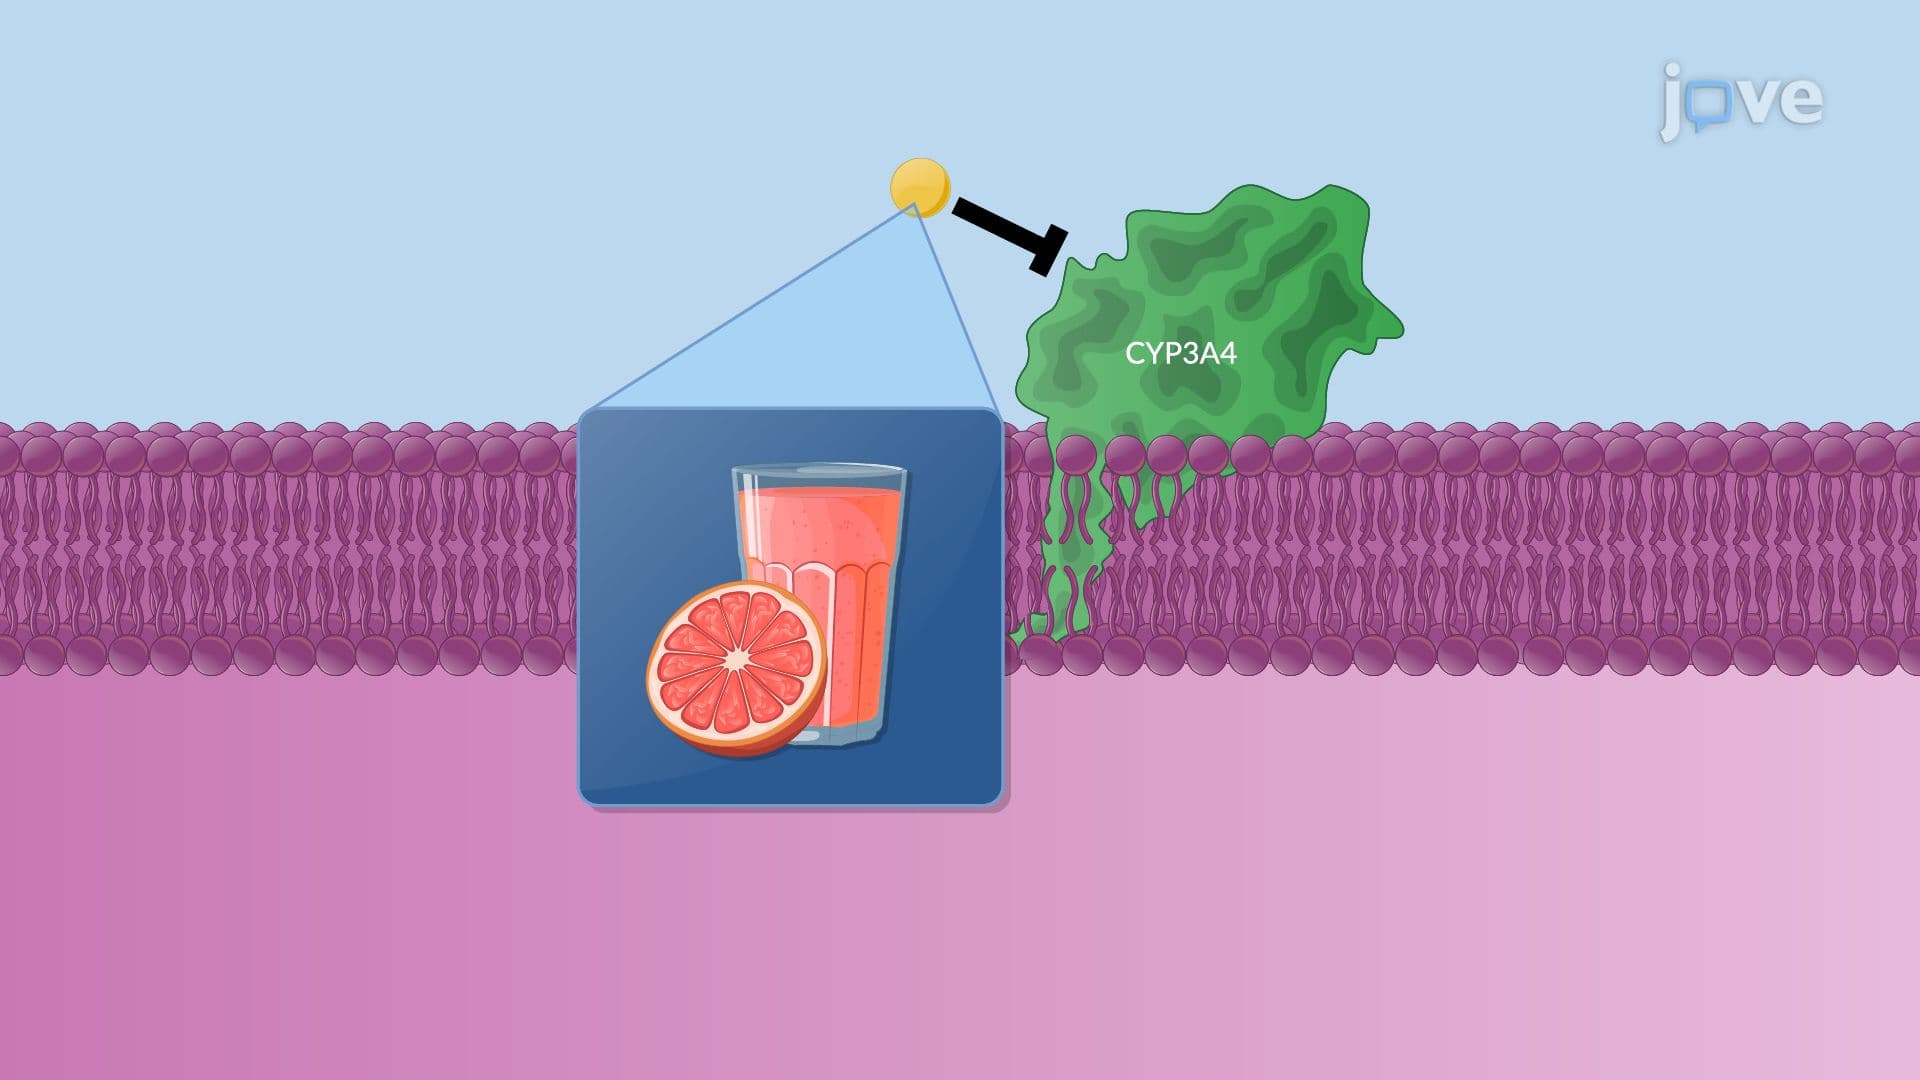
Pharmacokinetics: Drug–Food and Drug–Viral Interactions
 01:20
Pharmacokinetics in Obese Patients: Drug Metabolism and Excretion
 01:27
Pharmacogenetics of Drug Metabolism: Overview
 01:28
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes