5'-N-substituted carboxamidoadenosines as agonists for adenosine receptors
Related Experiment Video
Updated: Oct 21, 2026
Drug-induced Sensitization of Adenylyl Cyclase: Assay Streamlining and Miniaturization for Small Molecule and siRNA Screening Applications
Published on: January 27, 2014
M de Zwart1, A Kourounakis, H Kooijman
1Division of Medicinal Chemistry, Leiden/Amsterdam Center for Drug Research, Leiden University, P.O. Box 9502, 2300 RA Leiden, The Netherlands.

Abstract:
Novel as well as known 5'-N-substituted carboxamidoadenosines were prepared via new routes that provided shorter reaction times and good yields. Binding affinities were determined for rat A1 and A2A receptors and human A3 receptors. EC50 values were determined for cyclic AMP production in CHO cells expressing human A2B receptors. On all receptor subtypes relatively small substituents on the carboxamido moiety were optimal. Selectivity for the A3 receptor was found for several analogues (1a, 1d, 1h, and 1k). On A1 receptors a number of compounds, but not 5'-N-ethylcarboxamidoadenosine (NECA, 1b), showed small GTP shifts, which could be indicative of lower intrinsic activities at the A1 receptor. At the A2B receptor, derivatives 1i-k with modified ethyl substituents had reduced activities compared to the A2B reference agonist NECA (1b). Thiocarboxamido derivatives (8b and 8c) displayed considerable although decreased A2B receptor activity. The X-ray structure determination of compound 8b was carried out. Due to intramolecular hydrogen bonding between the carboxamido NH and the purine N3 in the crystal structure, the ribose moiety of this compound is in a syn conformation. However, theoretical calculations support that NECA (1b), and less so 8b, can readily adopt both the syn and the anti conformation, therefore not excluding the proposed anti mode of binding to the receptor.
Related Concept Videos
 01:25
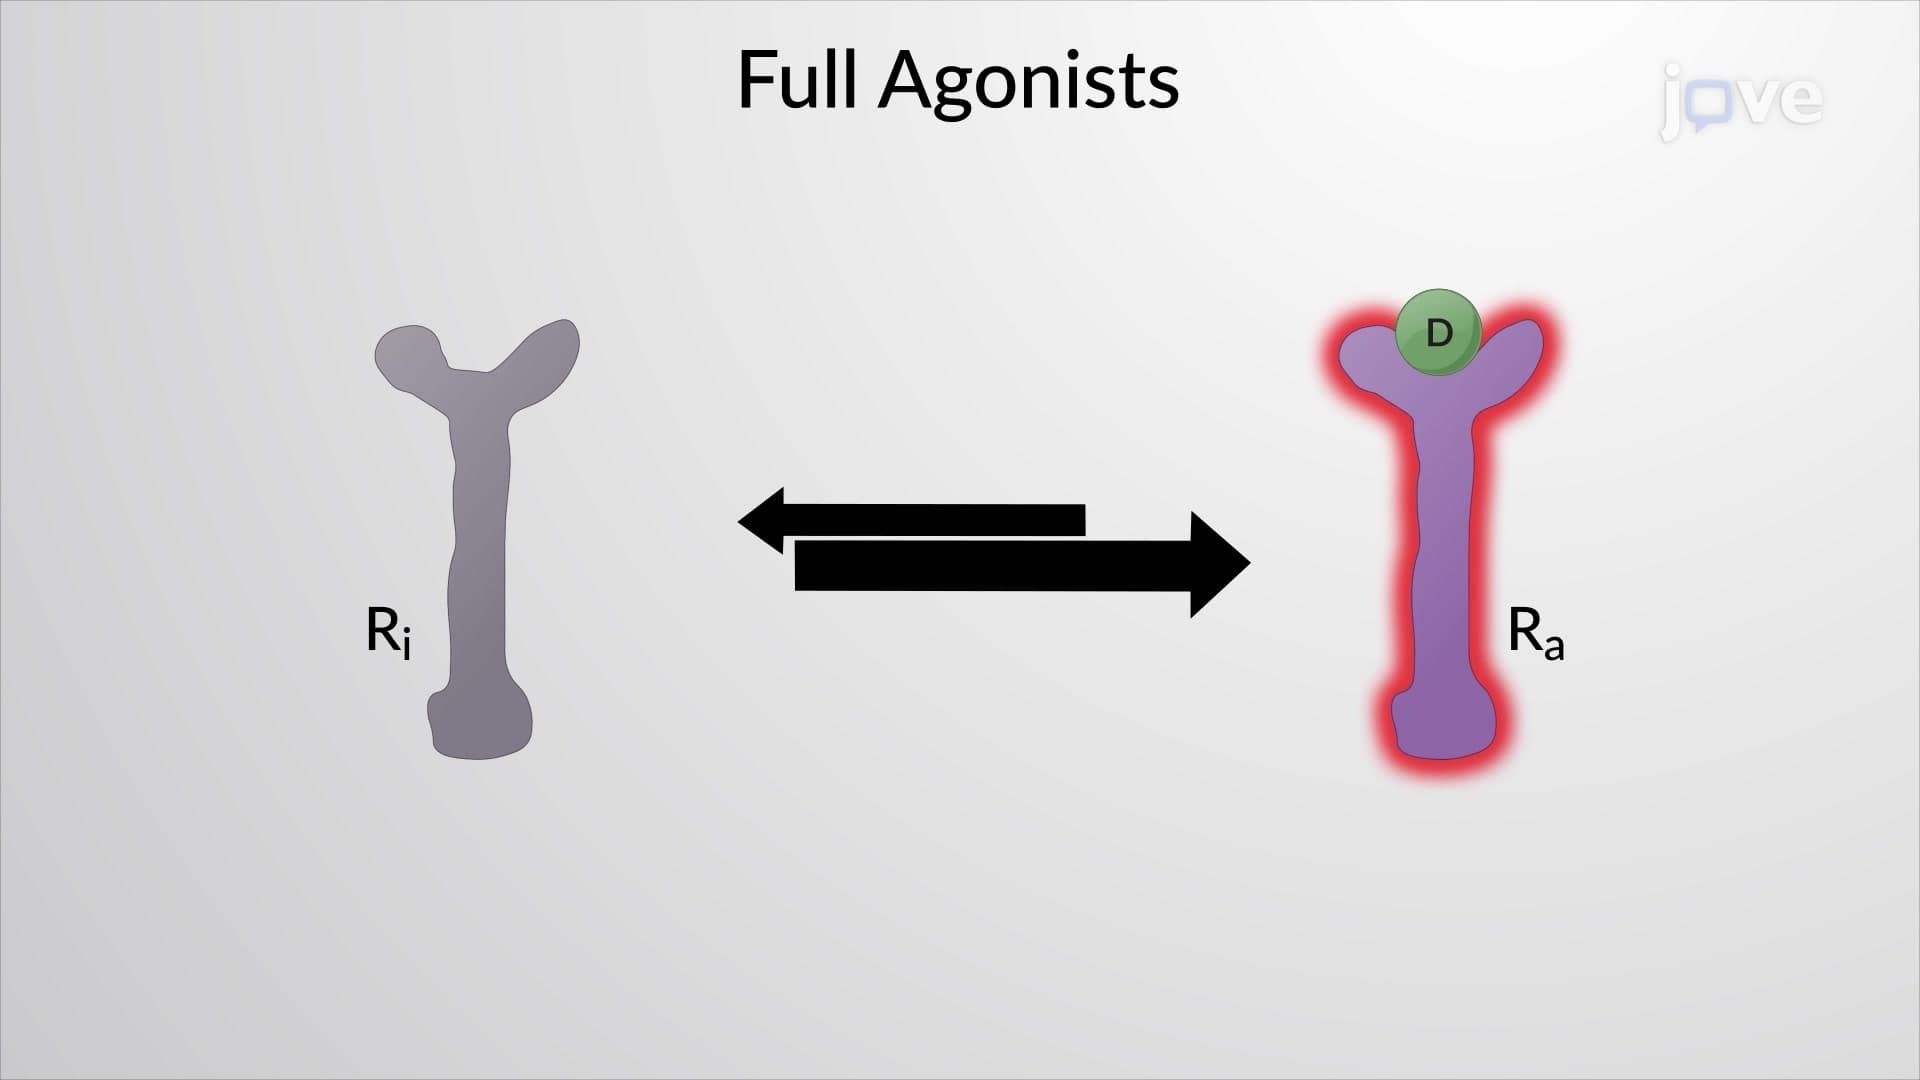
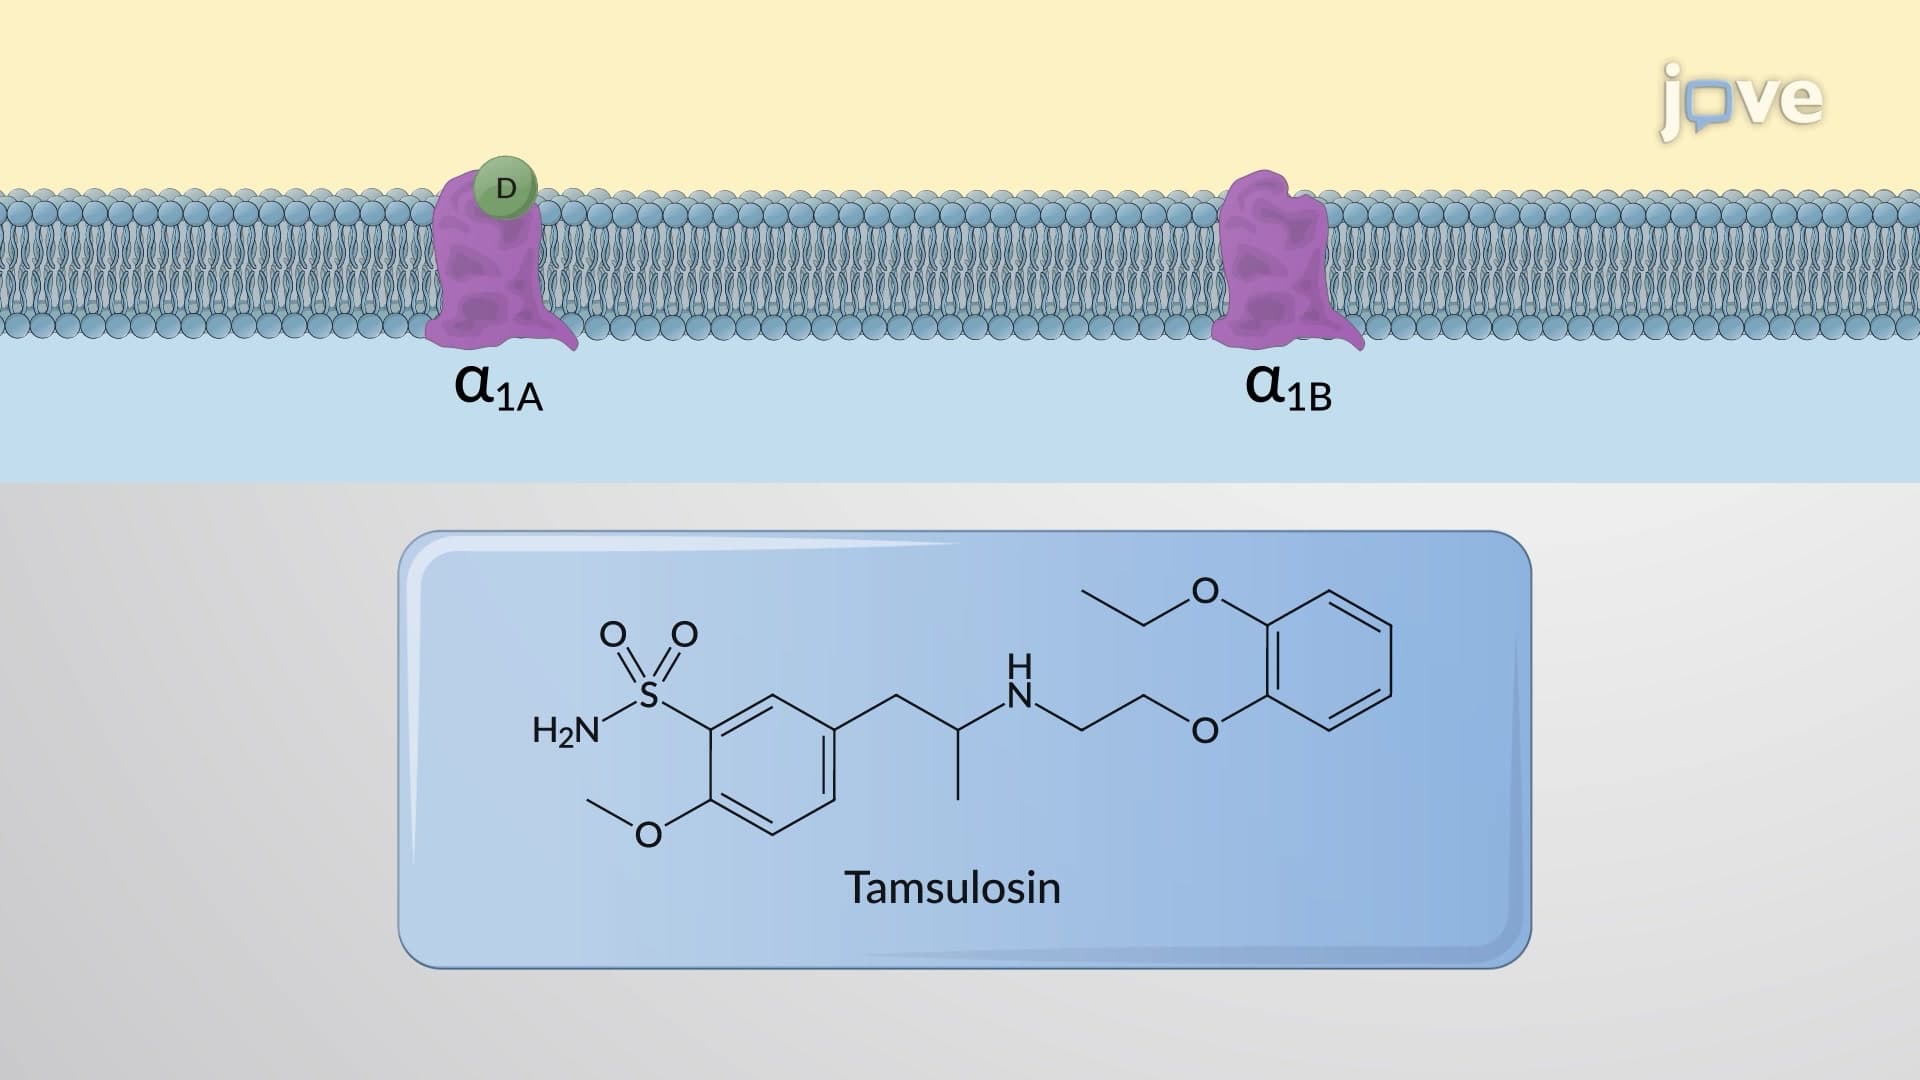
Drug-Receptor Interaction: Agonist
Agonists can bind to receptors in different ways. Some agonists bind directly to the receptor's active site, mimicking the endogenous ligand's action.
 01:31
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase C—inositol-1,4,5-trisphosphate...
 01:30
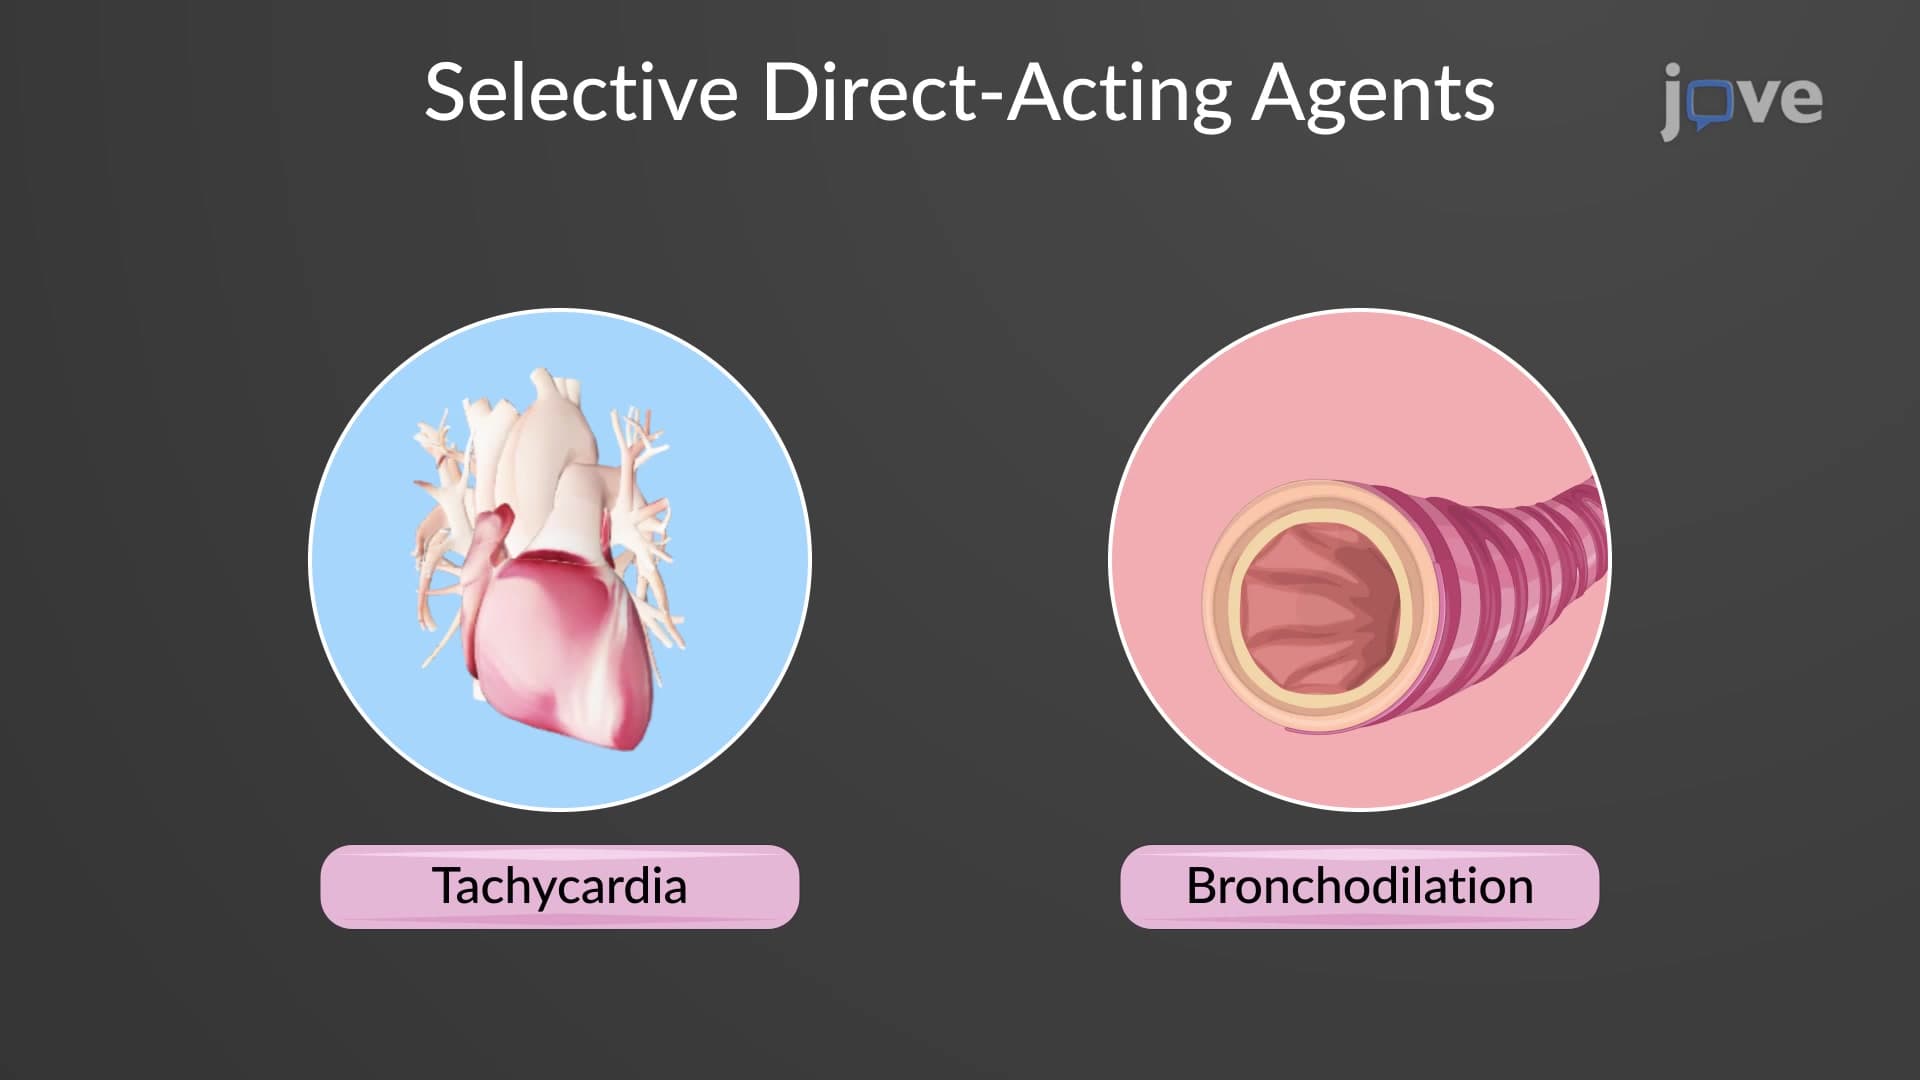
Adrenergic Agonists: Direct-Acting Agents
These agents can be classified...
 01:25
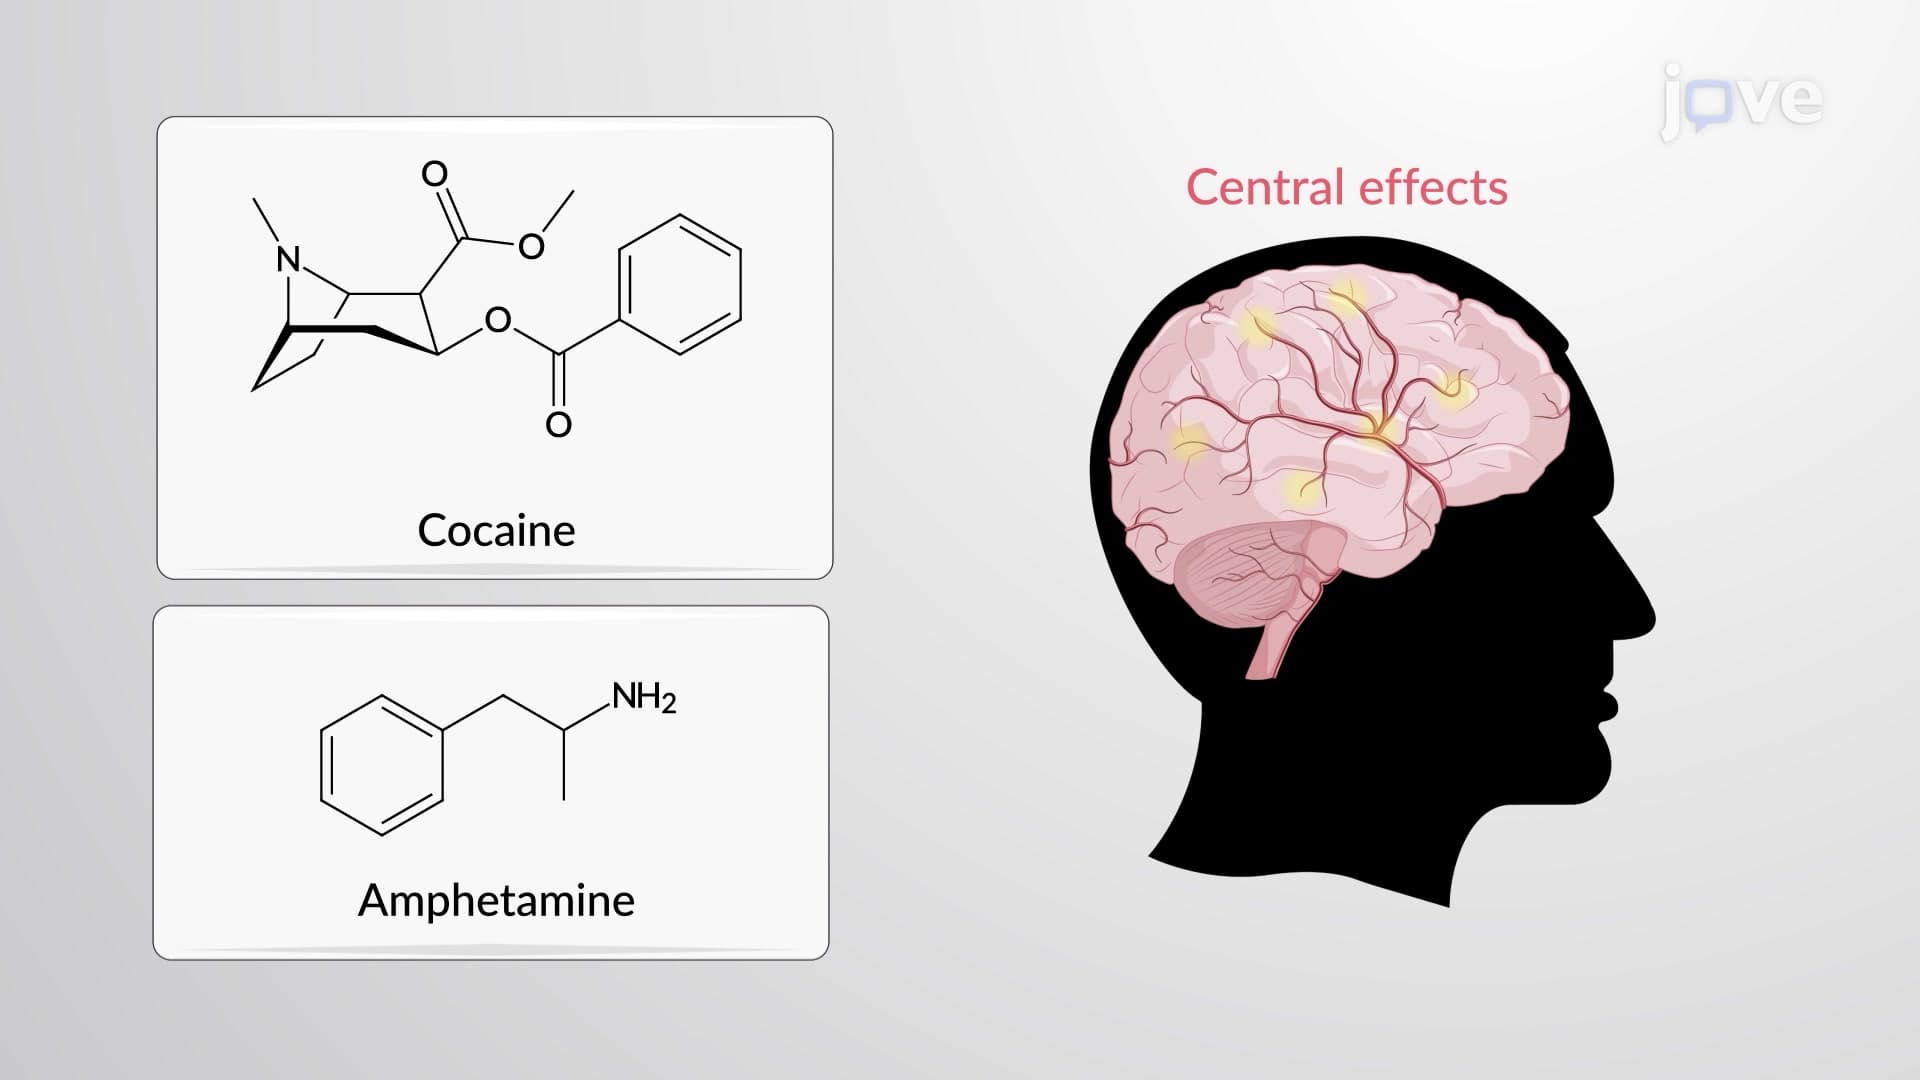
Adrenergic Agonists: Indirect-Acting Agents
One mechanism involves depleting stored catecholamines by displacing them from synaptic vesicles. These agents, known as "displacers," are transported into vesicles at the expense of noradrenaline. Examples include amphetamine and tyramine, which lack a catechol moiety, resulting in prolonged action, improved oral bioavailability, and...
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:17
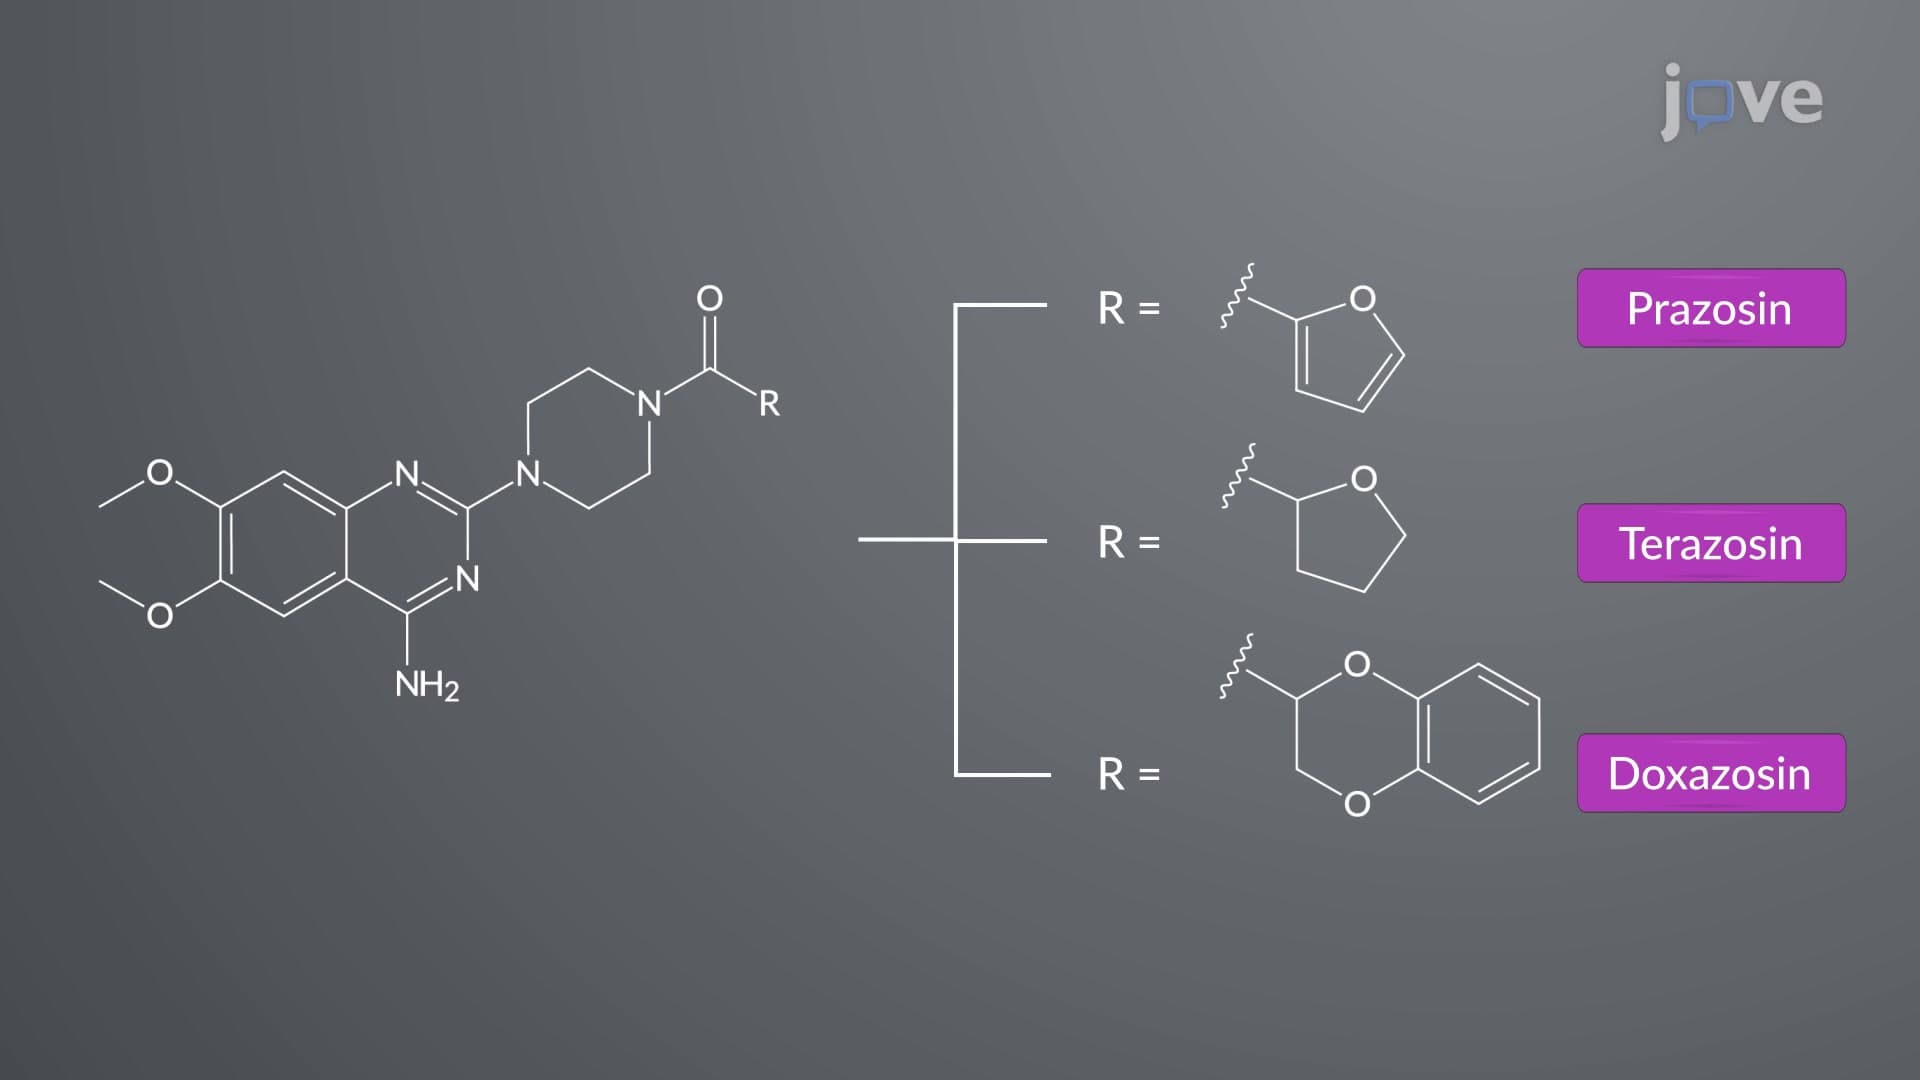
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...