N6,5'-Disubstituted adenosine derivatives as partial agonists for the human adenosine A3 receptor
Related Experiment Video
Updated: Oct 23, 2026
HPLC-based Assay to Monitor Extracellular Nucleotide/Nucleoside Metabolism in Human Chronic Lymphocytic Leukemia Cells
Published on: July 20, 2016
E W van Tilburg1, J von Frijtag Drabbe Künzel, M de Groote
1Division of Medicinal Chemistry, Leiden/Amsterdam Center for Drug Research, P.O. Box 9502, 2300 RA Leiden, The Netherlands. vtilburg@chem.leidenuniv.nl

Abstract:
5'-(Alkylthio)-substituted analogues of N6-benzyl- and N6-(3-iodobenzyl)adenosine were synthesized in 37-61% overall yields. The affinities of these compounds for the adenosine A1, A2A, and A3 receptors were determined using rat brain cortex, rat brain striata, and stably transfected human A3 receptors in HEK 293 cells, respectively. The compounds proved to be selective for the adenosine A3 receptor and displayed affinities in the nanomolar range. Compounds 8, 10, and 11 had the highest affinities for the A3 receptor with Ki values ranging from 8.8 to 27.7 nM. In the N6-benzyl series, compound 4 (LUF 5403), with a 5'-methylthio group, maintained a reasonable affinity and had the highest selectivity for the A3 receptor. Compound 12 (LUF 5411), with an N6-(3-iodobenzyl) group and a 5'-(n-propylthio) substituent, had the highest A3 selectivity of all of the compounds and also displayed high affinity for this receptor (Ki = 44.3 nM). The compounds were also evaluated for their ability to stimulate [35S]GTPgamma[S] binding in cell membranes expressing the human adenosine A3 receptor. It appeared that the N6,5'-disubstituted adenosine derivatives behaved as partial agonists. Compounds 2, 4, 8, and 10 had the highest intrinsic activities. Additionally, when tested in a cAMP assay, these compounds also behaved as partial agonists.
Related Concept Videos
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:31
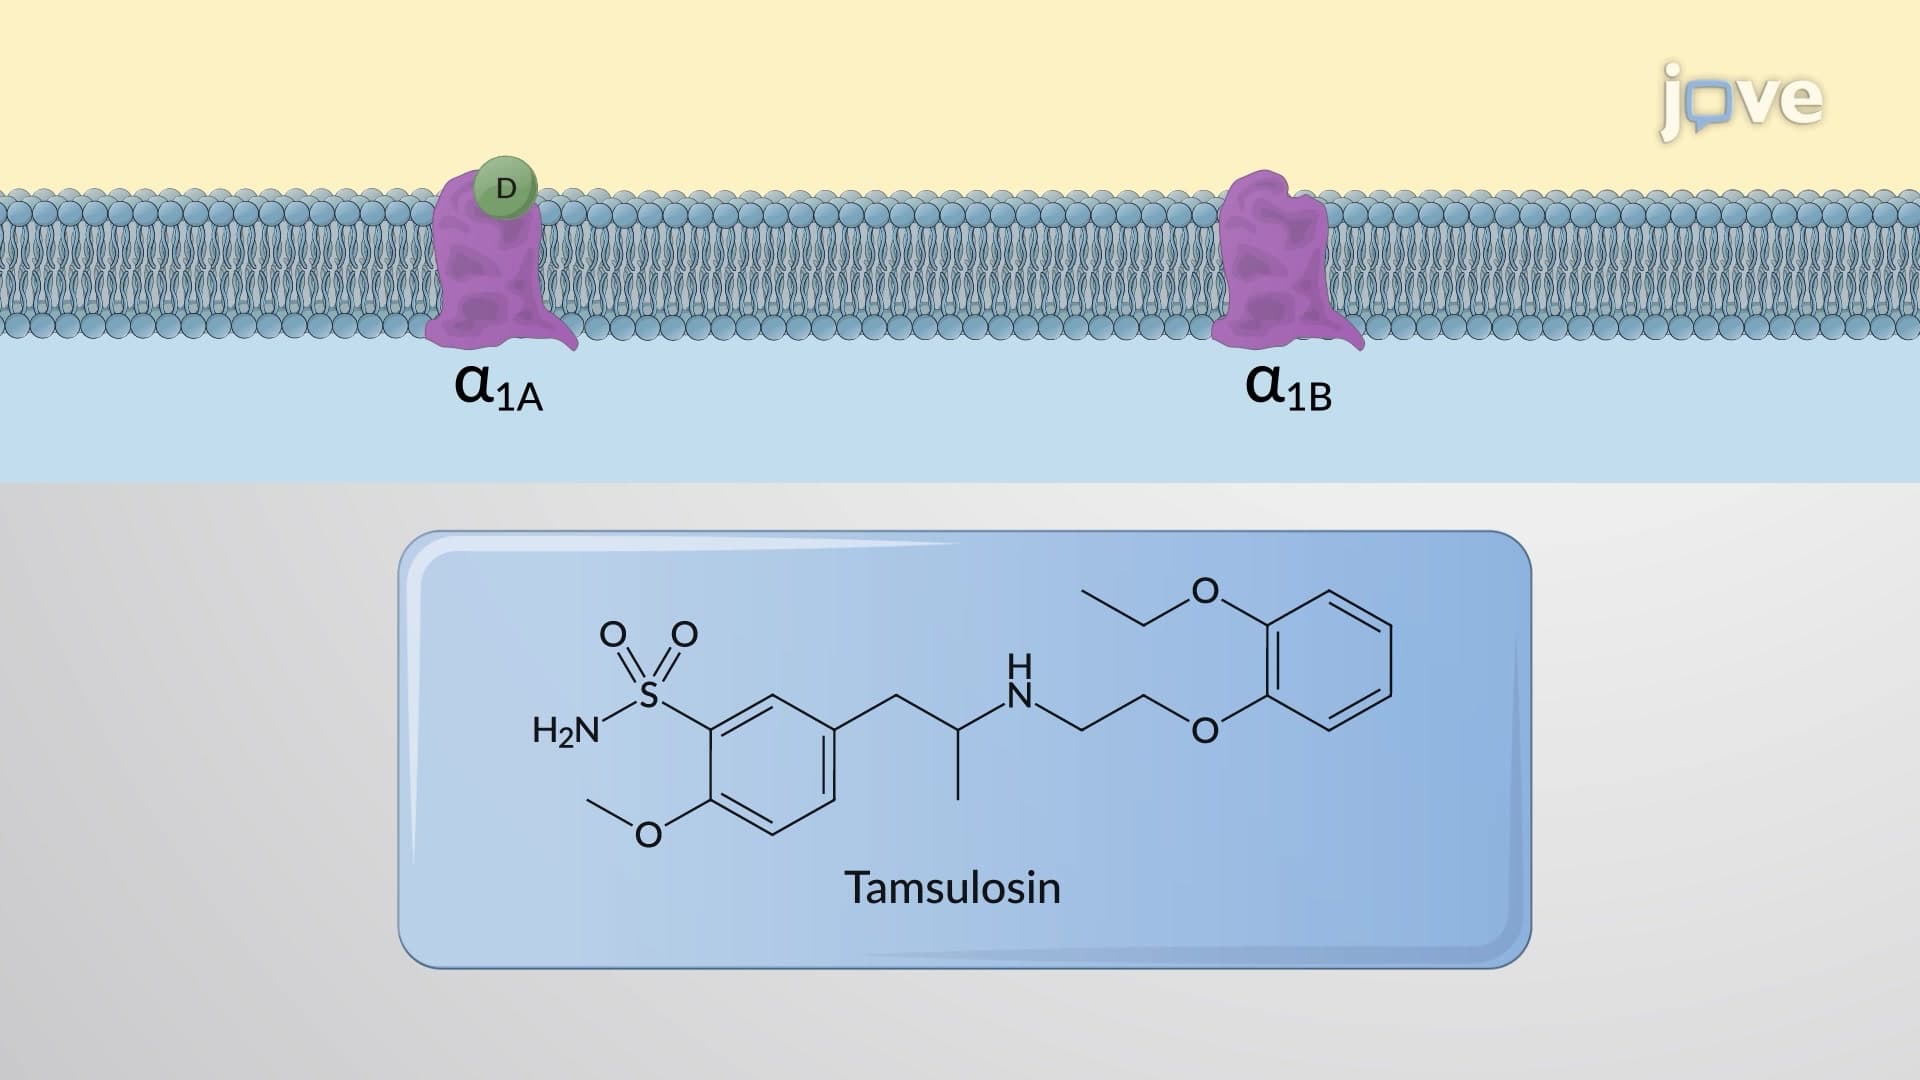
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase C—inositol-1,4,5-trisphosphate...
 01:17
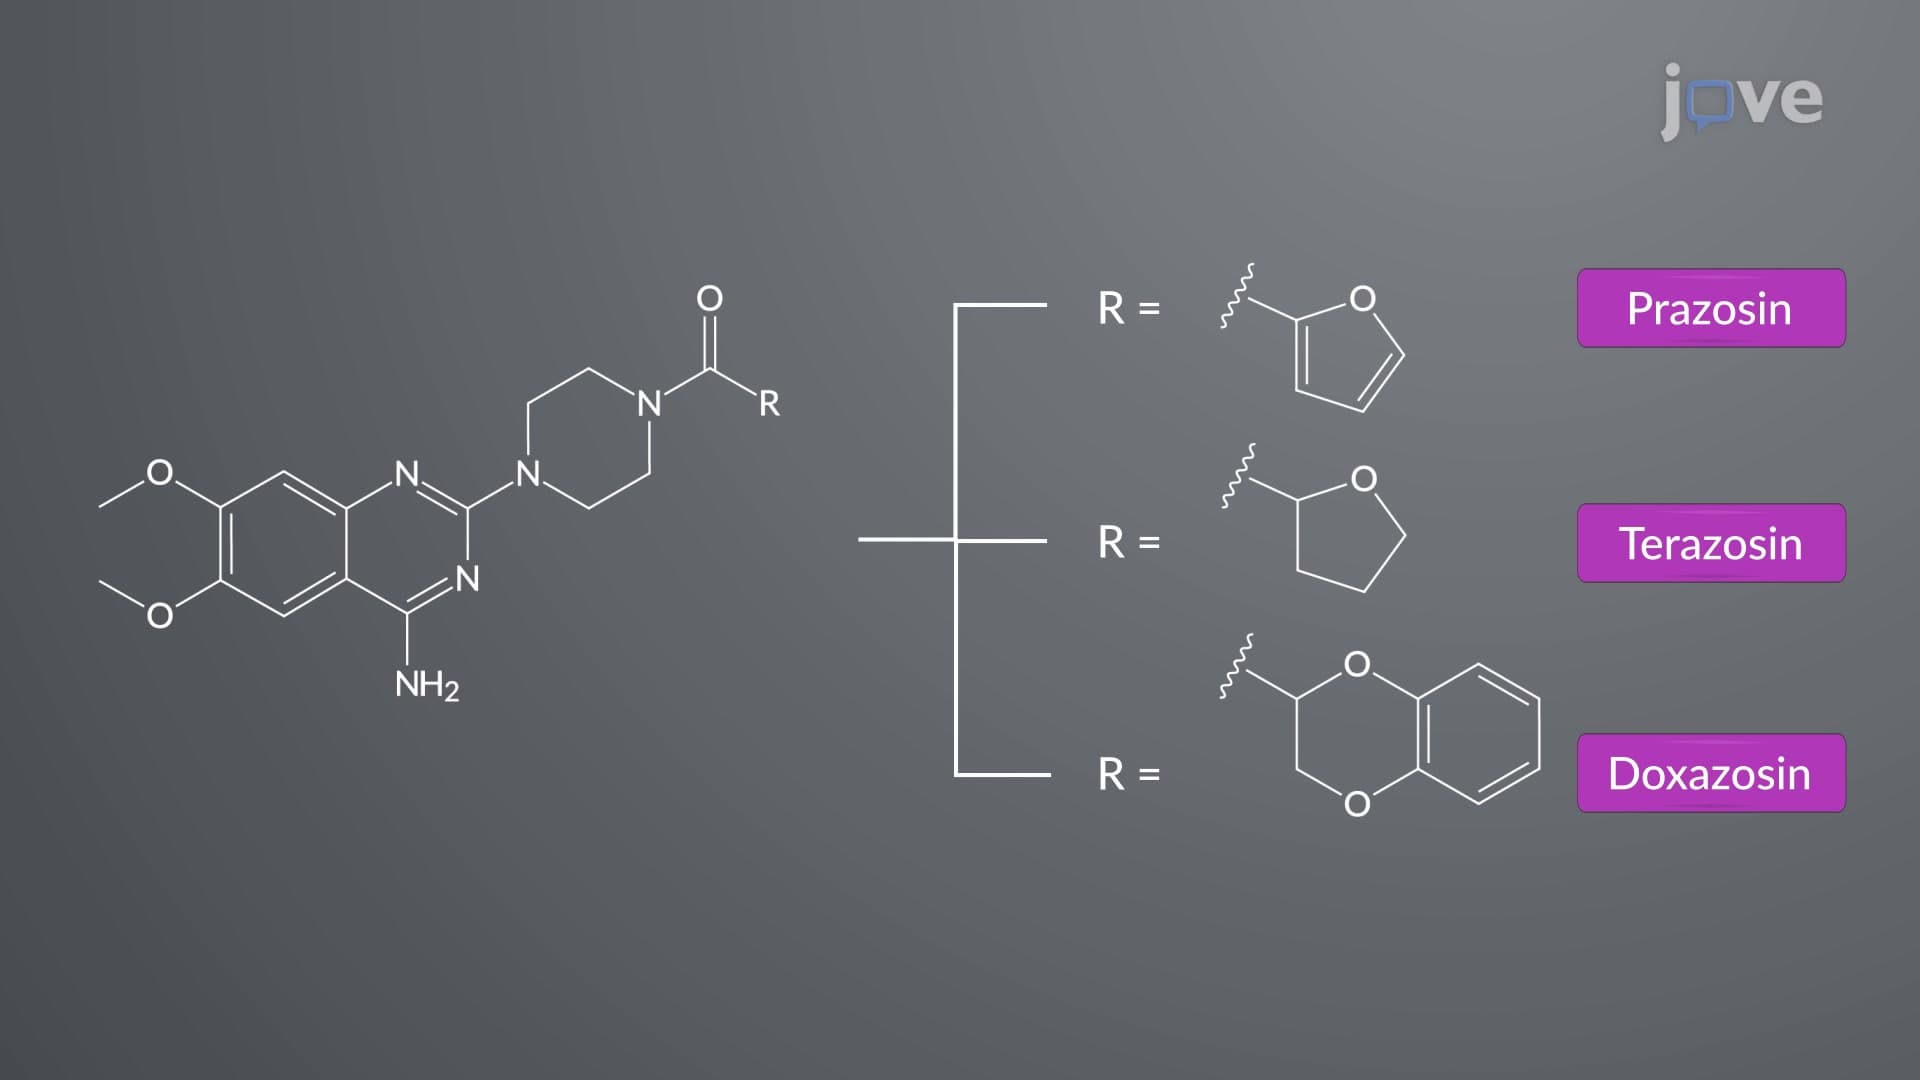
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...
 01:30
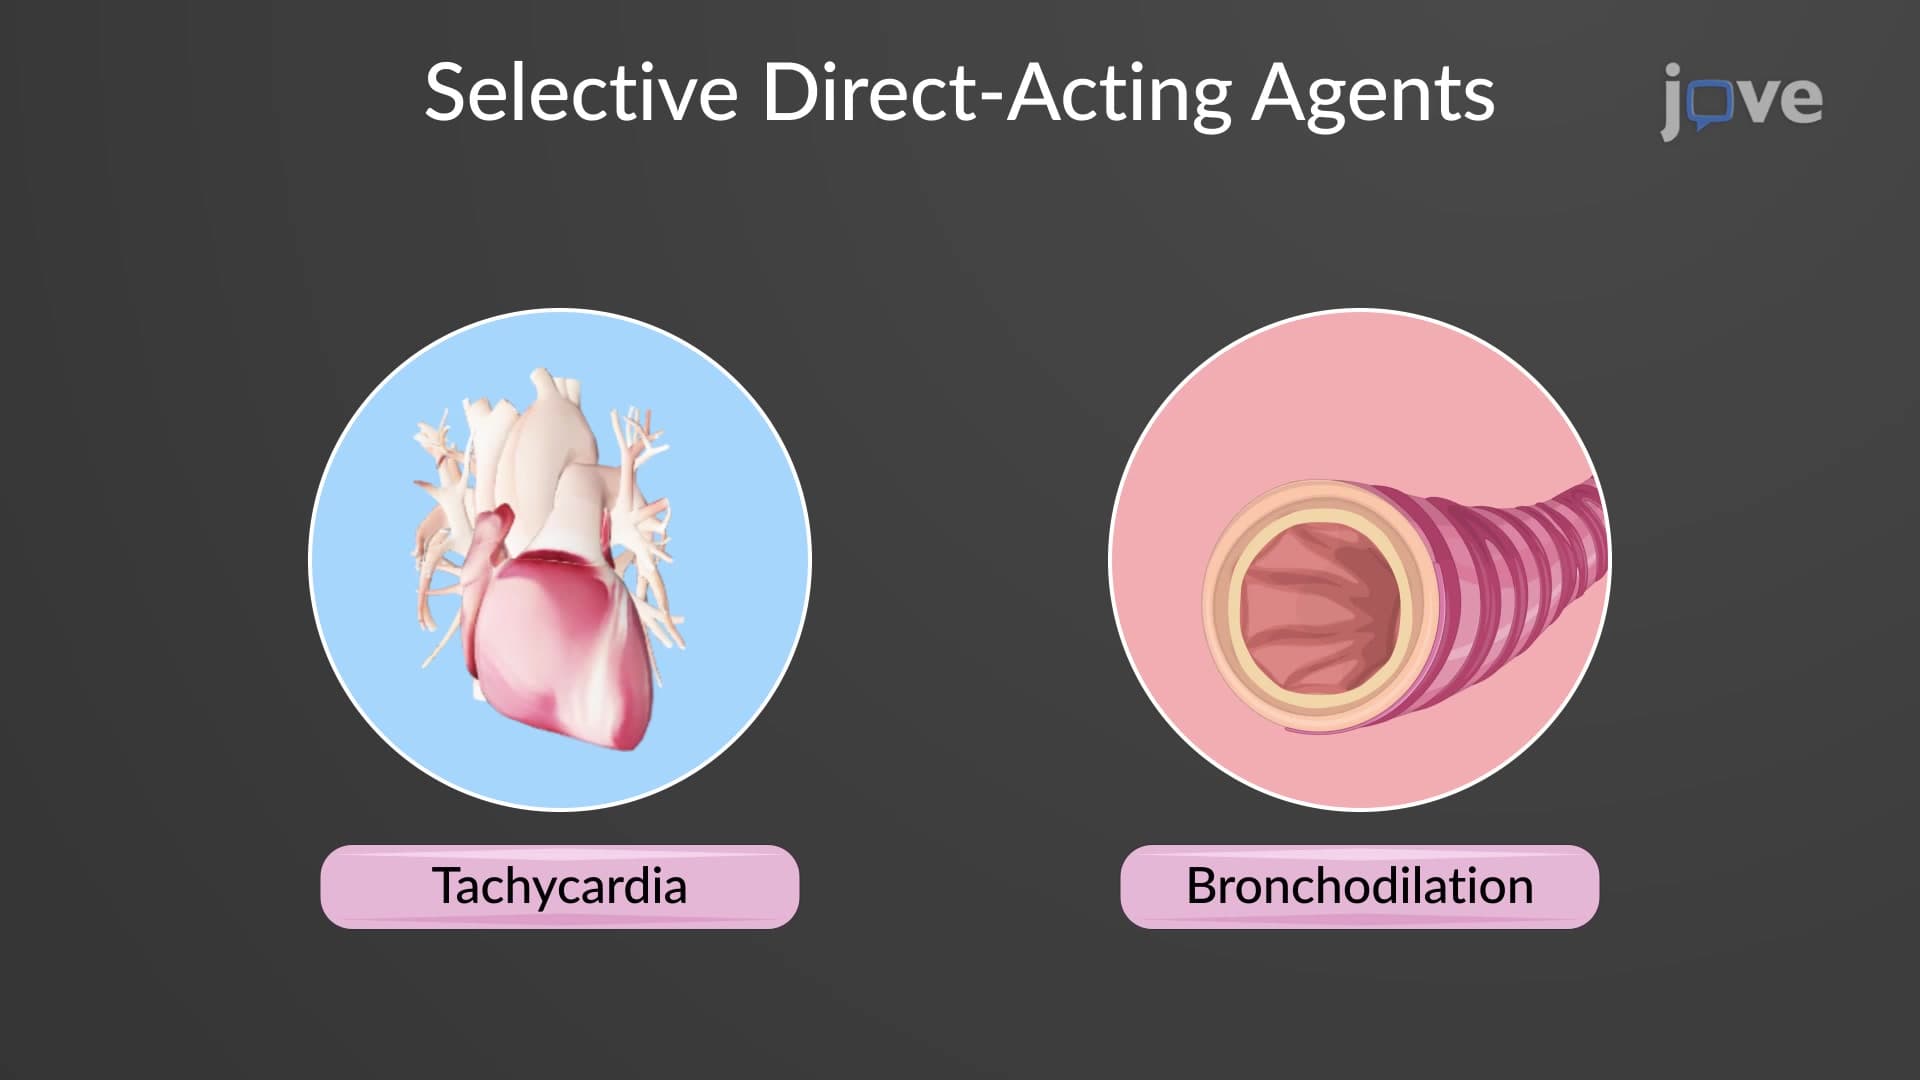
Adrenergic Agonists: Direct-Acting Agents
These agents can be classified...
 01:22
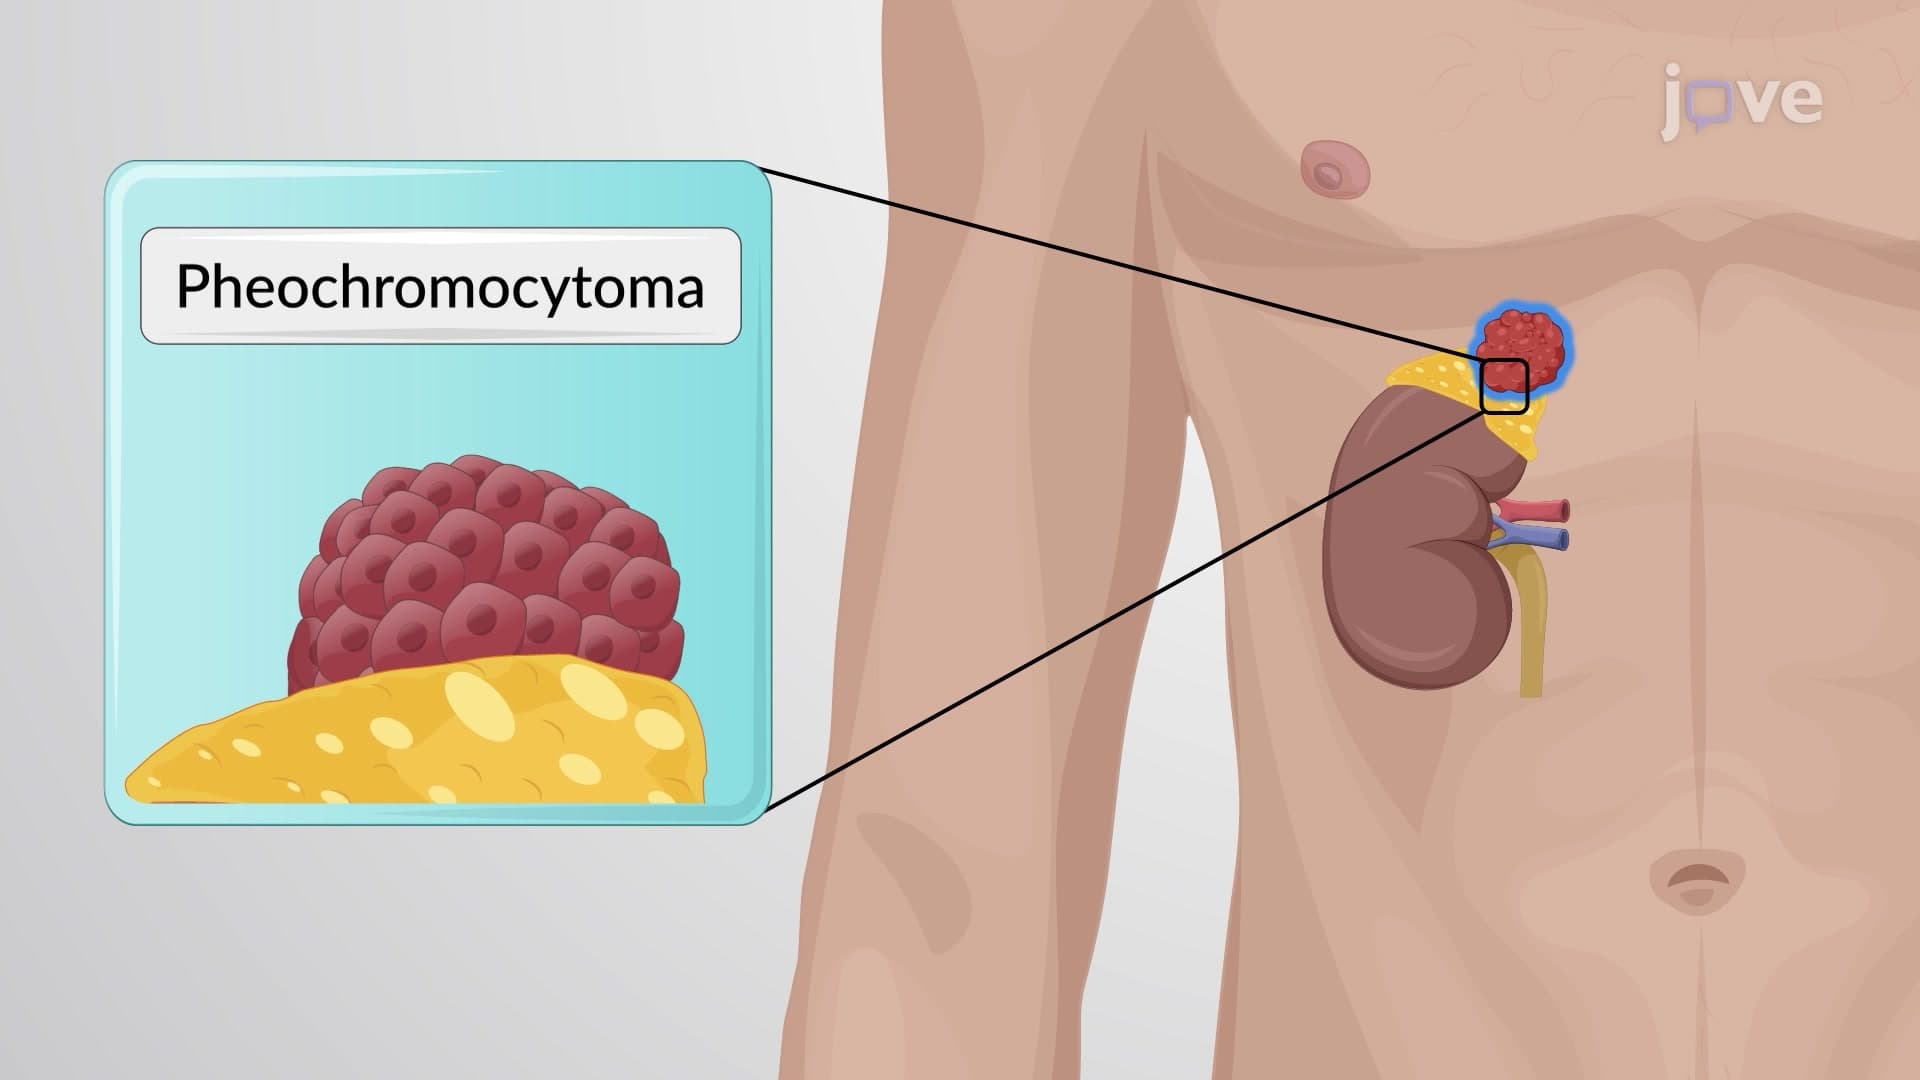
Adrenergic Antagonists: Pharmacological Actions of ɑ-Receptor Blockers
α1-blockers: These drugs inhibit α1-adrenoceptors on smooth muscle cells, resulting in vasodilation. This vasodilation lowers blood pressure, making α1-blockers valuable in treating hypertension. Additionally, α1-blockers effectively address urinary obstruction...
 01:25
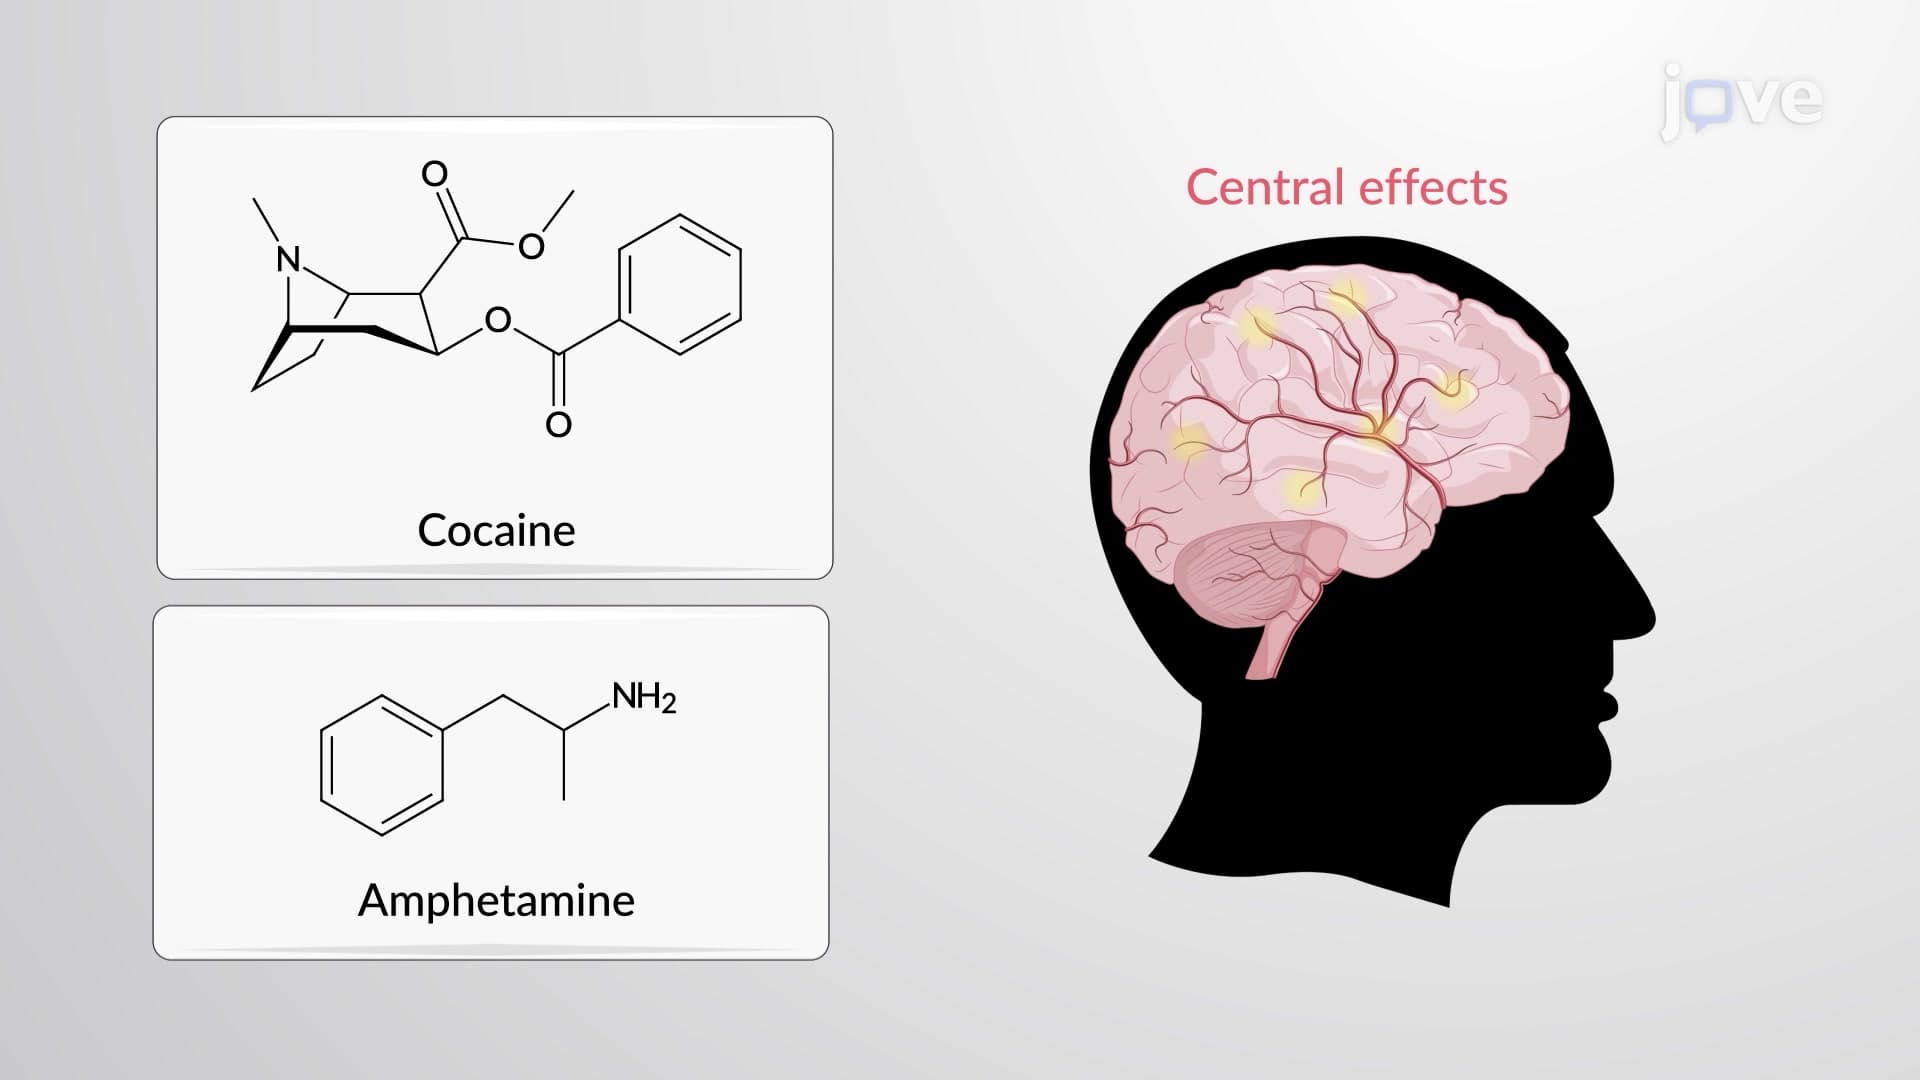
Adrenergic Agonists: Indirect-Acting Agents
One mechanism involves depleting stored catecholamines by displacing them from synaptic vesicles. These agents, known as "displacers," are transported into vesicles at the expense of noradrenaline. Examples include amphetamine and tyramine, which lack a catechol moiety, resulting in prolonged action, improved oral bioavailability, and...