Hyperornithinaemia-hyperammonaemia-homocitrullinuria syndrome is caused by mutations in a gene encoding a

J A Camacho1, C Obie, B Biery

  • 1Institute for Genetic Medicine, Johns Hopkins University School of Medicine, Baltimore, Maryland, USA.

Nature Genetics
|June 16, 1999
PubMed

Related Concept Videos

Urea Cycle01:23

Urea Cycle

The urea cycle describes how liver cells convert ammonia to urea. Ammonia is a toxic waste product of protein catabolism. Land animals must convert ammonia into the less toxic urea which can be safely eliminated by the kidneys through urine. Marine animals excrete ammonia directly, and the surrounding water dilutes the ammonia to safe levels.
Translation01:37

Translation

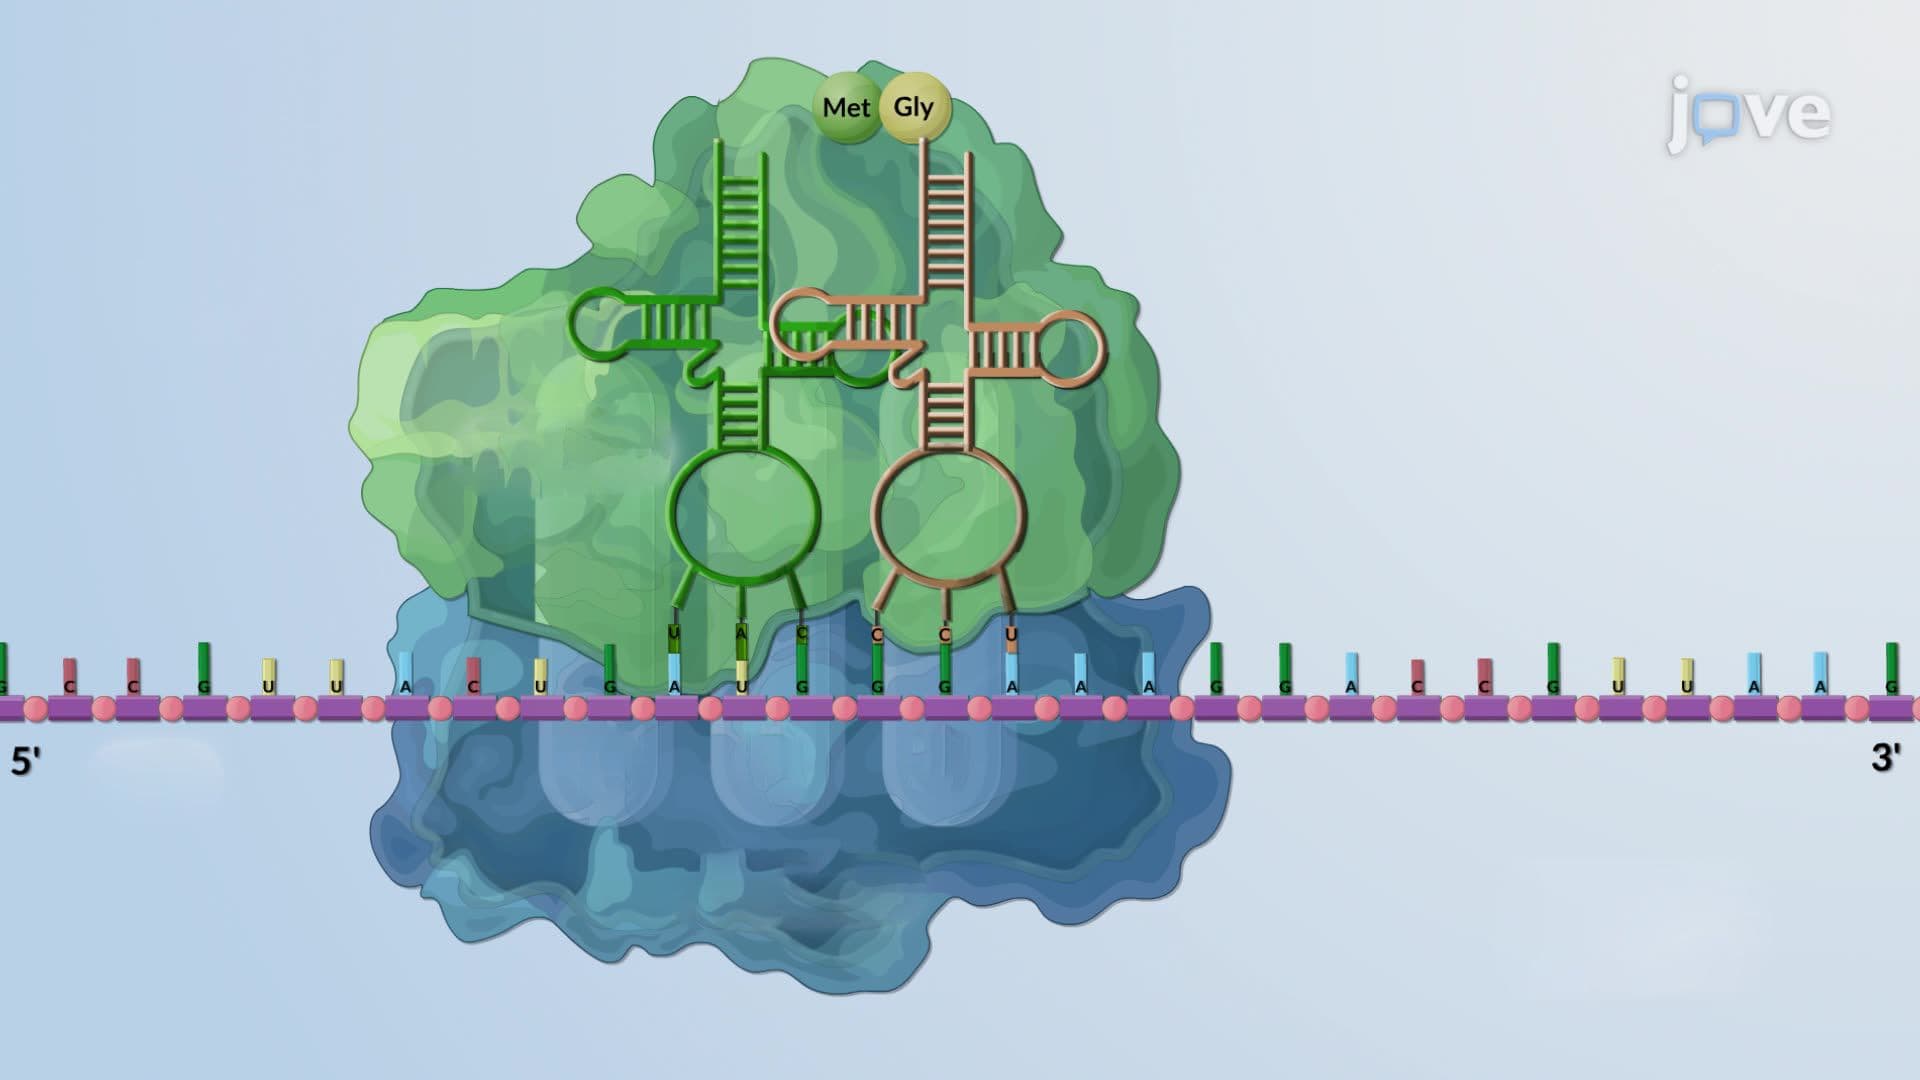
Translation is the process of synthesizing proteins from the genetic information carried by messenger RNA (mRNA). Following transcription, it constitutes the final step in the expression of genes. This process is carried out by ribosomes, complexes of protein and specialized RNA molecules. Ribosomes, transfer RNA (tRNA), and other proteins produce a chain of amino acids—the polypeptide—as the end product of translation.Translation Produces the Building Blocks of LifeProteins are called the...
Inborn Errors of Metabolism01:20

Inborn Errors of Metabolism

Phenylketonuria (PKU) is a protein metabolism disorder characterized by high blood levels of the amino acid phenylalanine. This results from a mutation in the gene responsible for phenylalanine hydroxylase, an enzyme that converts phenylalanine into tyrosine. When this enzyme is deficient, phenylalanine builds up in the blood, leading to symptoms such as vomiting, rashes, seizures, growth deficiency, and severe mental retardation. An early diagnosis and a diet restricting phenylalanine intake...
ATP Synthase: Mechanism01:48

ATP Synthase: Mechanism

In animals, the mitochondrial F1F0 ATP synthase is the key protein that synthesizes ATP molecules through a complex catalytic mechanism. While the nuclear genome encodes the majority of ATP synthase subunits, the mitochondrial genome encodes some of the enzyme's most critical components. The formation of this multi-subunit enzyme is a complex multi-step process regulated at the level of transcription, translation, and assembly. Defects in one or more of these steps can result in decreased ATP...
Lysosomal Hydrolases01:22

Lysosomal Hydrolases

Lysosomes are the site for the degradation of macromolecules and biological polymers released during membrane trafficking events such as secretory, endocytic, autophagic, and phagocytic pathways. The membrane-enclosed area of the lysosome, called the lumen, contains hydrolytic enzymes active in an acidic environment. These acid hydrolases are functional at a pH between 4.5 and 5 and are involved in cellular processes such as cell signaling, energy metabolism, restoration of the plasma membrane,...