
A calcium signaling defect in the pathogenesis of a mitochondrial DNA inherited oxidative phosphorylation deficiency

Abstract:
In recent years, genetic defects of the mitochondrial genome (mtDNA) were shown to be associated with a heterogeneous group of disorders, known as mitochondrial diseases, but the cellular events deriving from the molecular lesions and the mechanistic basis of the specificity of the syndromes are still incompletely understood. Mitochondrial calcium (Ca2+) homeostasis depends on close contacts with the endoplasmic reticulum and is essential in modulating organelle function. Given the strong dependence of mitochondrial Ca2+ uptake on the membrane potential and the intracellular distribution of the organelle, both of which may be altered in mitochondrial diseases, we investigated the occurrence of defects in mitochondrial Ca2+ handling in living cells with either the tRNALys mutation of MERRF (myoclonic epilepsy with ragged-red fibers) or the ATPase mutation of NARP (neurogenic muscle weakness, ataxia and retinitis pigmentosa). There was a derangement of mitochondrial Ca2+ homeostasis in MERRF, but not in NARP cells, whereas cytosolic Ca2+ responses were normal in both cell types. Treatment of MERRF cells with drugs affecting organellar Ca2+ transport mostly restored both the agonist-dependent mitochondrial Ca2+ uptake and the ensuing stimulation of ATP production. These results emphasize the differences in the cellular pathogenesis of the various mtDNA defects and indicate specific pharmacological approaches to the treatment of some mitochondrial diseases.
Insights
Genetic defects in mitochondrial DNA (mtDNA) cause mitochondrial diseases. Researchers found impaired calcium handling in MERRF cells, but not NARP cells, suggesting distinct cellular pathologies and potential drug targets.
Area of Science:
- Cellular Biology
- Genetics
- Neuroscience
Background:
- Mitochondrial diseases arise from genetic defects in mitochondrial DNA (mtDNA).
- Mitochondrial calcium (Ca2+) homeostasis is crucial for organelle function and is linked to the endoplasmic reticulum.
- The cellular basis and disease specificity of mtDNA defects remain unclear.
Purpose of the Study:
- Investigate mitochondrial Ca2+ handling defects in living cells with specific mtDNA mutations.
- Compare cellular pathogenesis in MERRF (tRNALys mutation) and NARP (ATPase mutation) mitochondrial diseases.
- Explore potential therapeutic strategies targeting mitochondrial Ca2+ transport.
Main Methods:
- Utilized living cell models harboring MERRF and NARP mutations.
- Assessed mitochondrial and cytosolic Ca2+ responses.
- Evaluated the effects of pharmacological agents on Ca2+ transport and ATP production.
Main Results:
- Mitochondrial Ca2+ homeostasis was disrupted in MERRF cells, but not NARP cells.
- Cytosolic Ca2+ signaling remained normal in both cell types.
- Drug treatment targeting organellar Ca2+ transport partially restored mitochondrial Ca2+ uptake and ATP production in MERRF cells.
Conclusions:
- Mitochondrial Ca2+ handling defects contribute to the pathogenesis of specific mtDNA disorders like MERRF.
- Cellular mechanisms differ between various mtDNA defects, impacting disease presentation.
- Targeting mitochondrial Ca2+ transport offers a potential therapeutic avenue for certain mitochondrial diseases.
More Related Videos
 09:40
Phosphorus-31 Magnetic Resonance Spectroscopy: A Tool for Measuring In Vivo Mitochondrial Oxidative Phosphorylation Capacity in Human Skeletal Muscle
Published on: January 19, 2017
 07:35
Assessment of Open Probability of the Mitochondrial Permeability Transition Pore in the Setting of Coenzyme Q Excess
Published on: June 1, 2022
Related Concept Videos
 02:59
Animal Mitochondrial Genetics
 01:45
Mitochondrial Membranes
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...
 01:48
ATP Synthase: Mechanism
 01:19
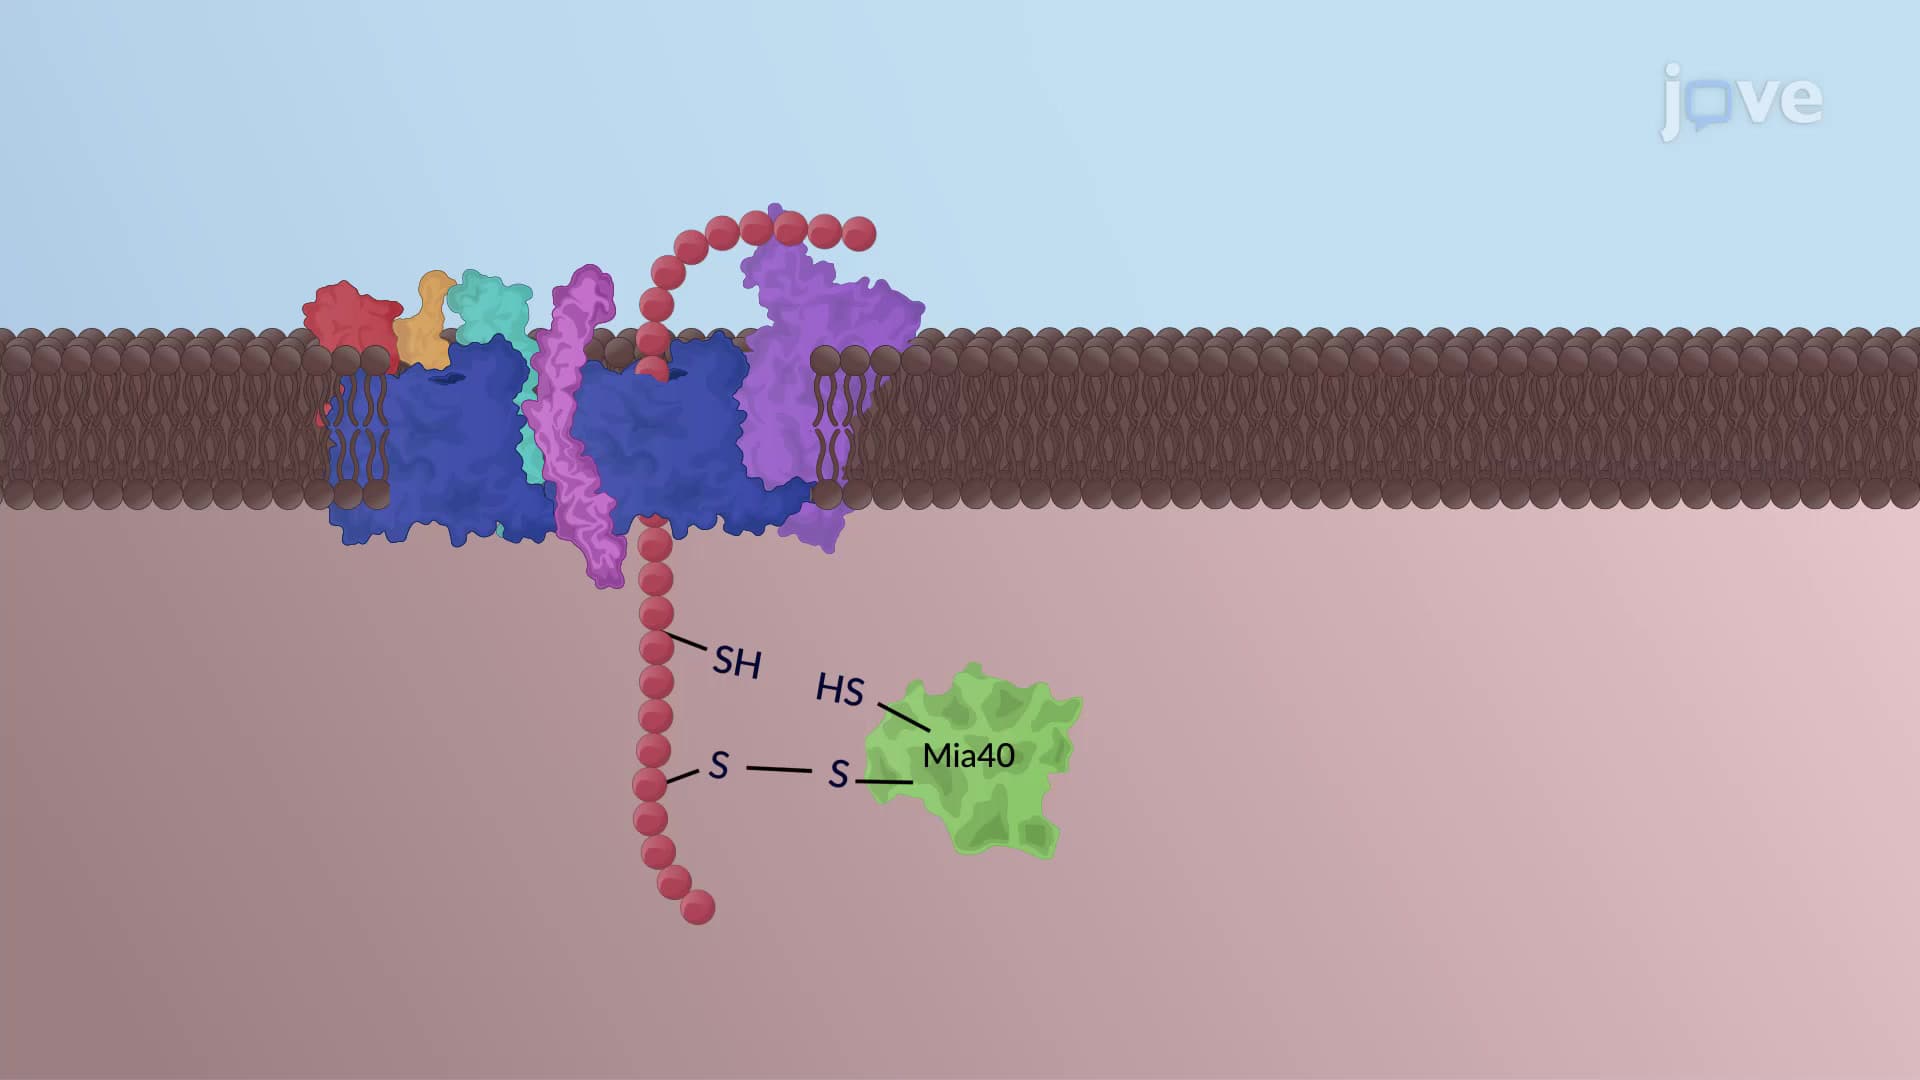
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
01:45Mitochondrial Membranes