Related Experiment Video
Updated: Jul 28, 2026

Measuring Biomolecular DSC Profiles with Thermolabile Ligands to Rapidly Characterize Folding and Binding Interactions
Published on: November 21, 2017
Stability, blood partition, and protein binding of NQ12, a new naphthoquinone derivative

Abstract:
The stability of a new phospholipase A2 inhibitor, NQ12, in various pH solutions, human plasma, urine, and gastric juices, and rat liver homogenate, the blood partition of NQ12 between plasma and blood cells of rat blood, and the factors influencing the binding of NQ12 to 4% human serum albumin (HSA) using an equilibrium dialysis technique were evaluated. NQ12 was unstable in various pH solutions, human plasma and urine, and rat liver homogenate when incubated in a water-bath shaker kept at 37 degrees C and at a rate of 50 oscillations per min. However, NQ12 was stable for up to 3-hr incubation in human gastric juice. The plasma-to-blood cell concentration ratios of NQ12 were independent of NQ12 rat blood concentrations when the whole blood was incubated for up to 2-hr; the mean (+/- standard deviation) values were 0.112 +/- 0.0650 and 0.172 +/- 0.105 at initial blood NQ12 concentrations of 10 and 20 microg/ml, respectively. The binding of NQ12 to 4% HSA was considerable (higher than 99%) at NQ12 concentrations ranging from 10 to 100 microg/ml in 4% HSA. The unbound fraction of NQ12 was dependent on NQ12 concentrations, pHs of buffer, and HSA concentrations, but was independent of the concentrations of sulfisoxazole or salicylic acid, incubation temperature, buffers containing various concentrations of chloride ion, and concentrations of heparin and alpha-1-acid glycoprotein.
Related Concept Videos
 02:18
The Equilibrium Binding Constant and Binding Strength
 01:21
Factors Affecting Dissolution: Drug pKa, Lipophilicity and GI pH
A drug's pKa and the pH of the gastrointestinal (GI) tract play crucial roles in drug...
 01:30
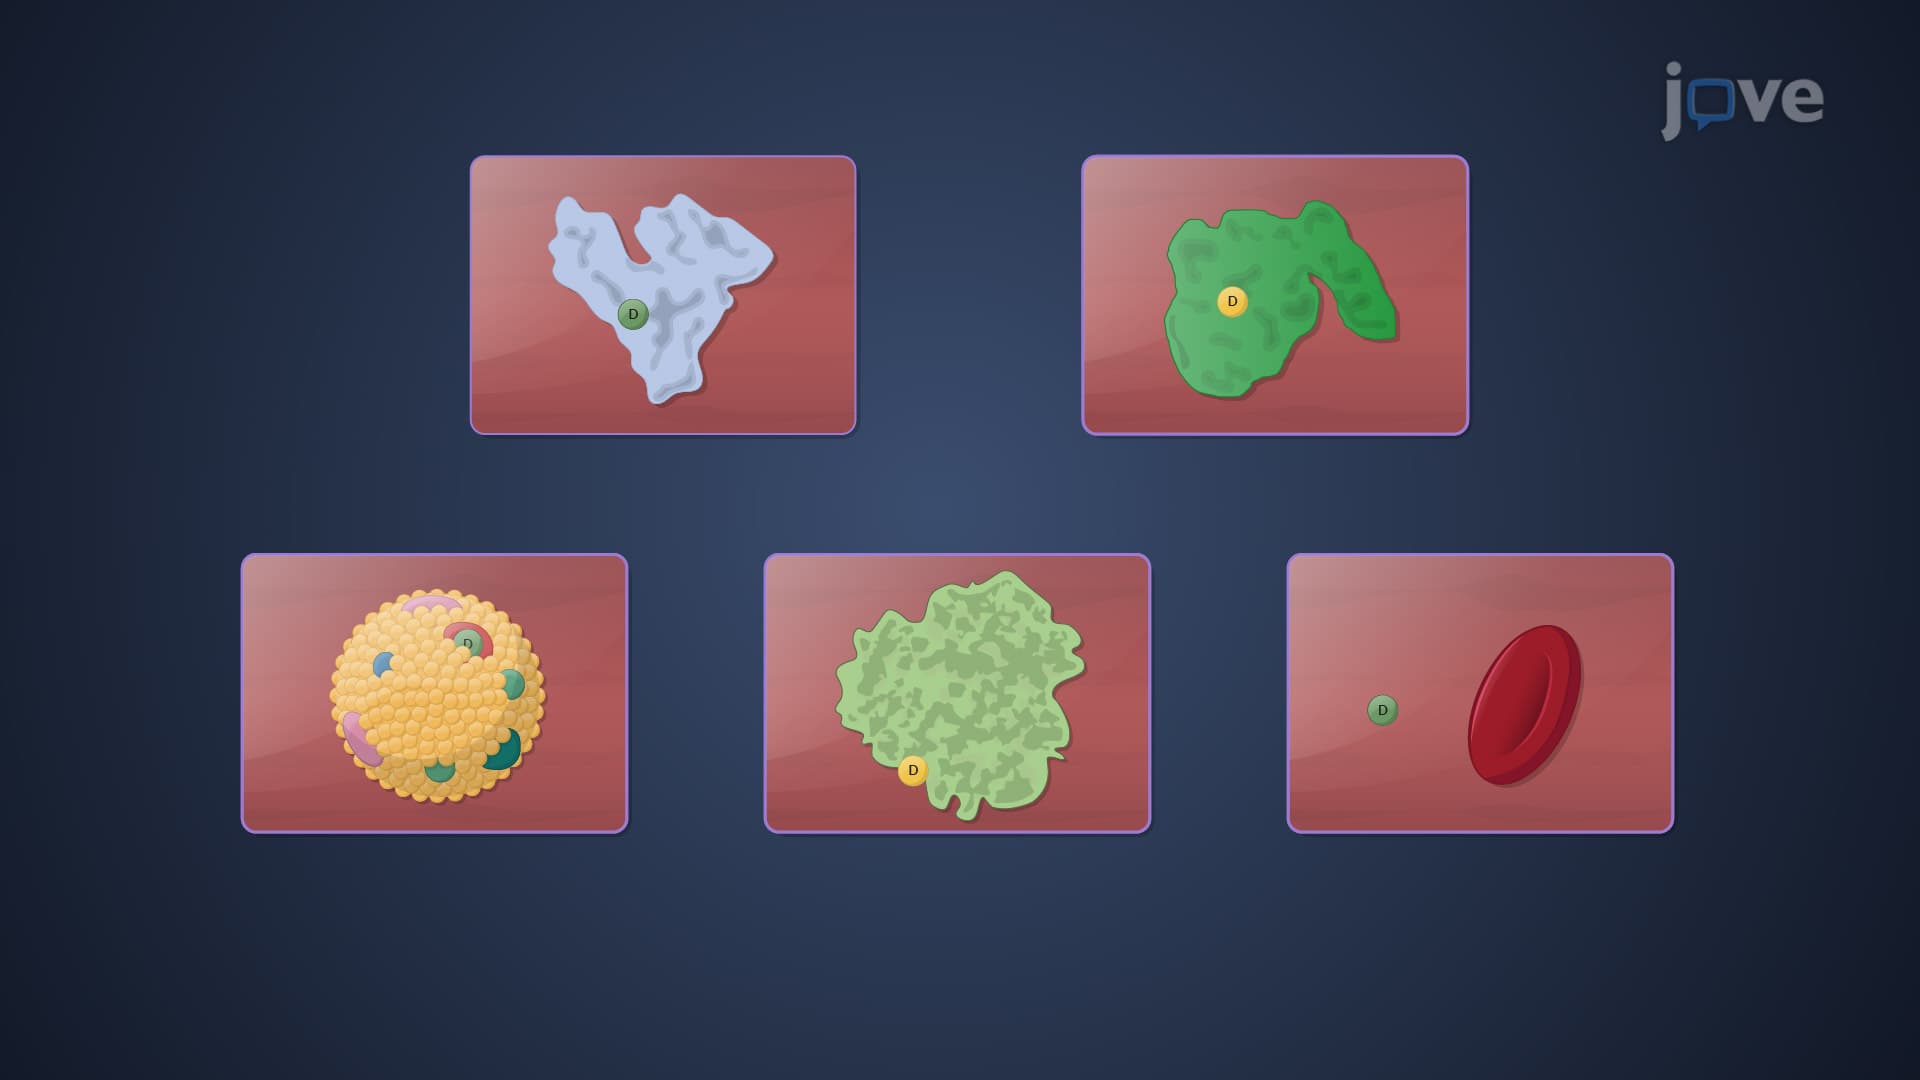
Drug Binding to Blood Components
HSA is the most abundant plasma protein and is vital in drug binding. It contains distinct drug-binding sites, with different drugs exhibiting affinity for specific sites. There are three main drug-binding domains for HSA: sites I, II, and III. These domains are further...
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 01:22
Nonlinear Pharmacokinetics: Bioavailability and Protein-Drug Binding
To quantify the extent of bioavailability, pharmacologists often use a parameter called .
 01:22
Pharmaceutical Alternatives: Stability-Related Therapeutic Nonequivalence