Related Experiment Video
Updated: Aug 19, 2026

Site Directed Spin Labeling and EPR Spectroscopic Studies of Pentameric Ligand-Gated Ion Channels
Published on: July 4, 2016
Correlation between drug release kinetics from proteineous matrix and matrix structure: EPR and NMR study
I Katzhendler1, K Mäder, M Friedman
1Department of Pharmaceutics, School of Pharmacy, The Hebrew University of Jerusalem, P.O.B. 12065, Jerusalem 91120, Israel.

Abstract:
The present study was conducted in order to probe the microstructure, microviscosity, and hydration properties of matrices containing two model drugs, naproxen sodium (NS) and naproxen (N), and egg albumin (EA) as matrix carrier. The results suggested that N release from EA matrix was controlled by a bulk erosion mechanism in combination with additional processes (crystal dissolution/crystallization rate) compared with NS matrix, which behaved as a non-erodible matrix and drug release occurred by diffusion through the gel. Using EPR technique it has been shown that incorporating NS into EA matrix strongly influences the microstructure of the protein gel, and hence the transport of the penetrant within the matrix, compared with matrices containing N. The presence of NS increased the protein chain mobility and hydration which supports our previous results showing that NS cause unfolding of EA. In contrast, N caused only marginal effect on EA chain mobility. The gel formed in EA/NS matrices was more porous compared with EA/N matrices as revealed by the lower rotational correlation time of PCA (lower microviscosity) in EA/NS matrices compared with EA/N. However, EA/N gelled matrices were more heterogeneous, i.e., containing a higher number of components having different mobility. The T(1) and T(2) relaxation studies by NMR provided an additional support for the higher chain hydration in EA/NS matrices compared with EA/N as indicated by the higher relaxation rates in the gelled matrices. Internal pH measurements by EPR revealed that the micro-pH inside 100% EA and 50/50 EA/N matrices were lower than 50/50 EA/NS matrices and in all cases lower than the penetrating buffer pH. The lower pH compartment formed in N matrices affected N solubility and crystal dissolution rate, which can explain its lower release rate compared with EA, from the same formulation. The EPR and NMR data supports our findings that NS caused unfolding of the protein, affected matrix structure, and converted it to a hydrophobic non-erodible matrix compared with EA/N matrix in which the native properties of EA were mainly retained.
Related Concept Videos
 01:17
Electron Paramagnetic Resonance (EPR) Spectroscopy: Organic Radicals
 01:16
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:22
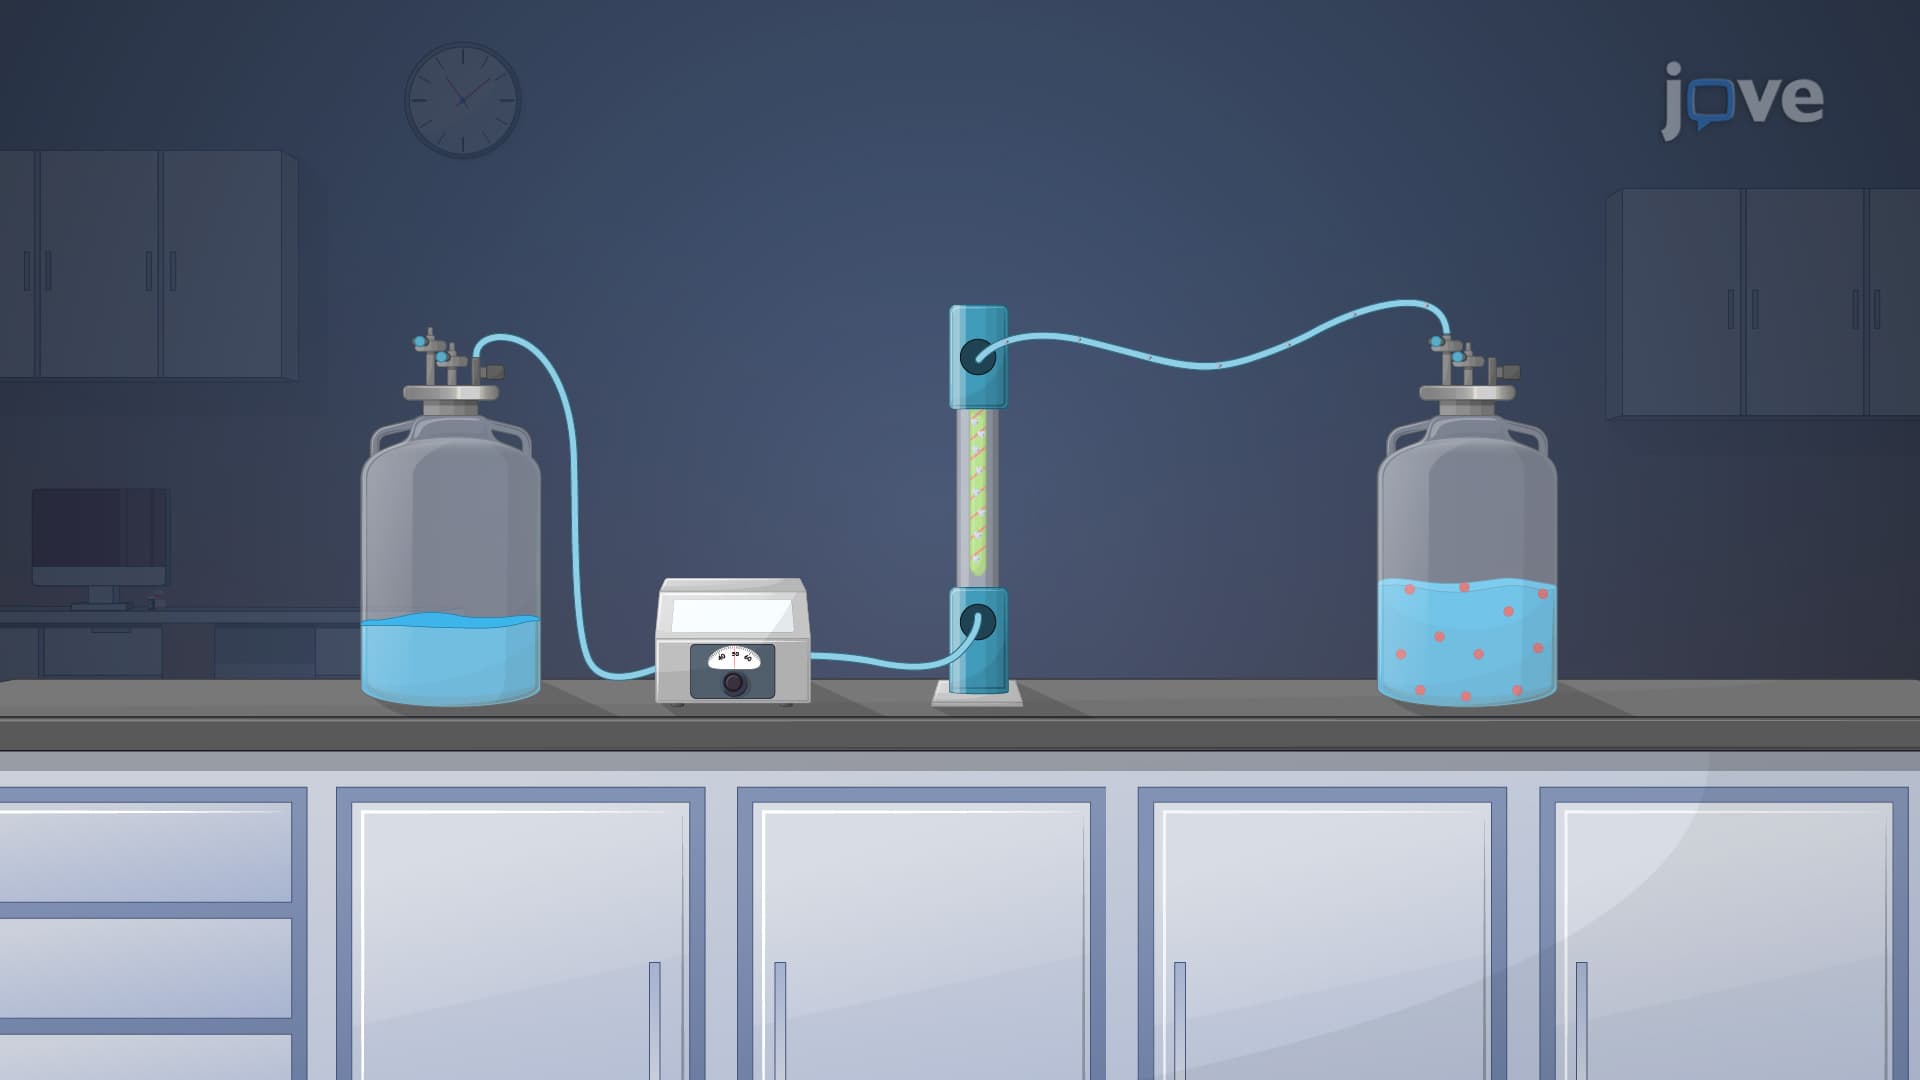
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:20
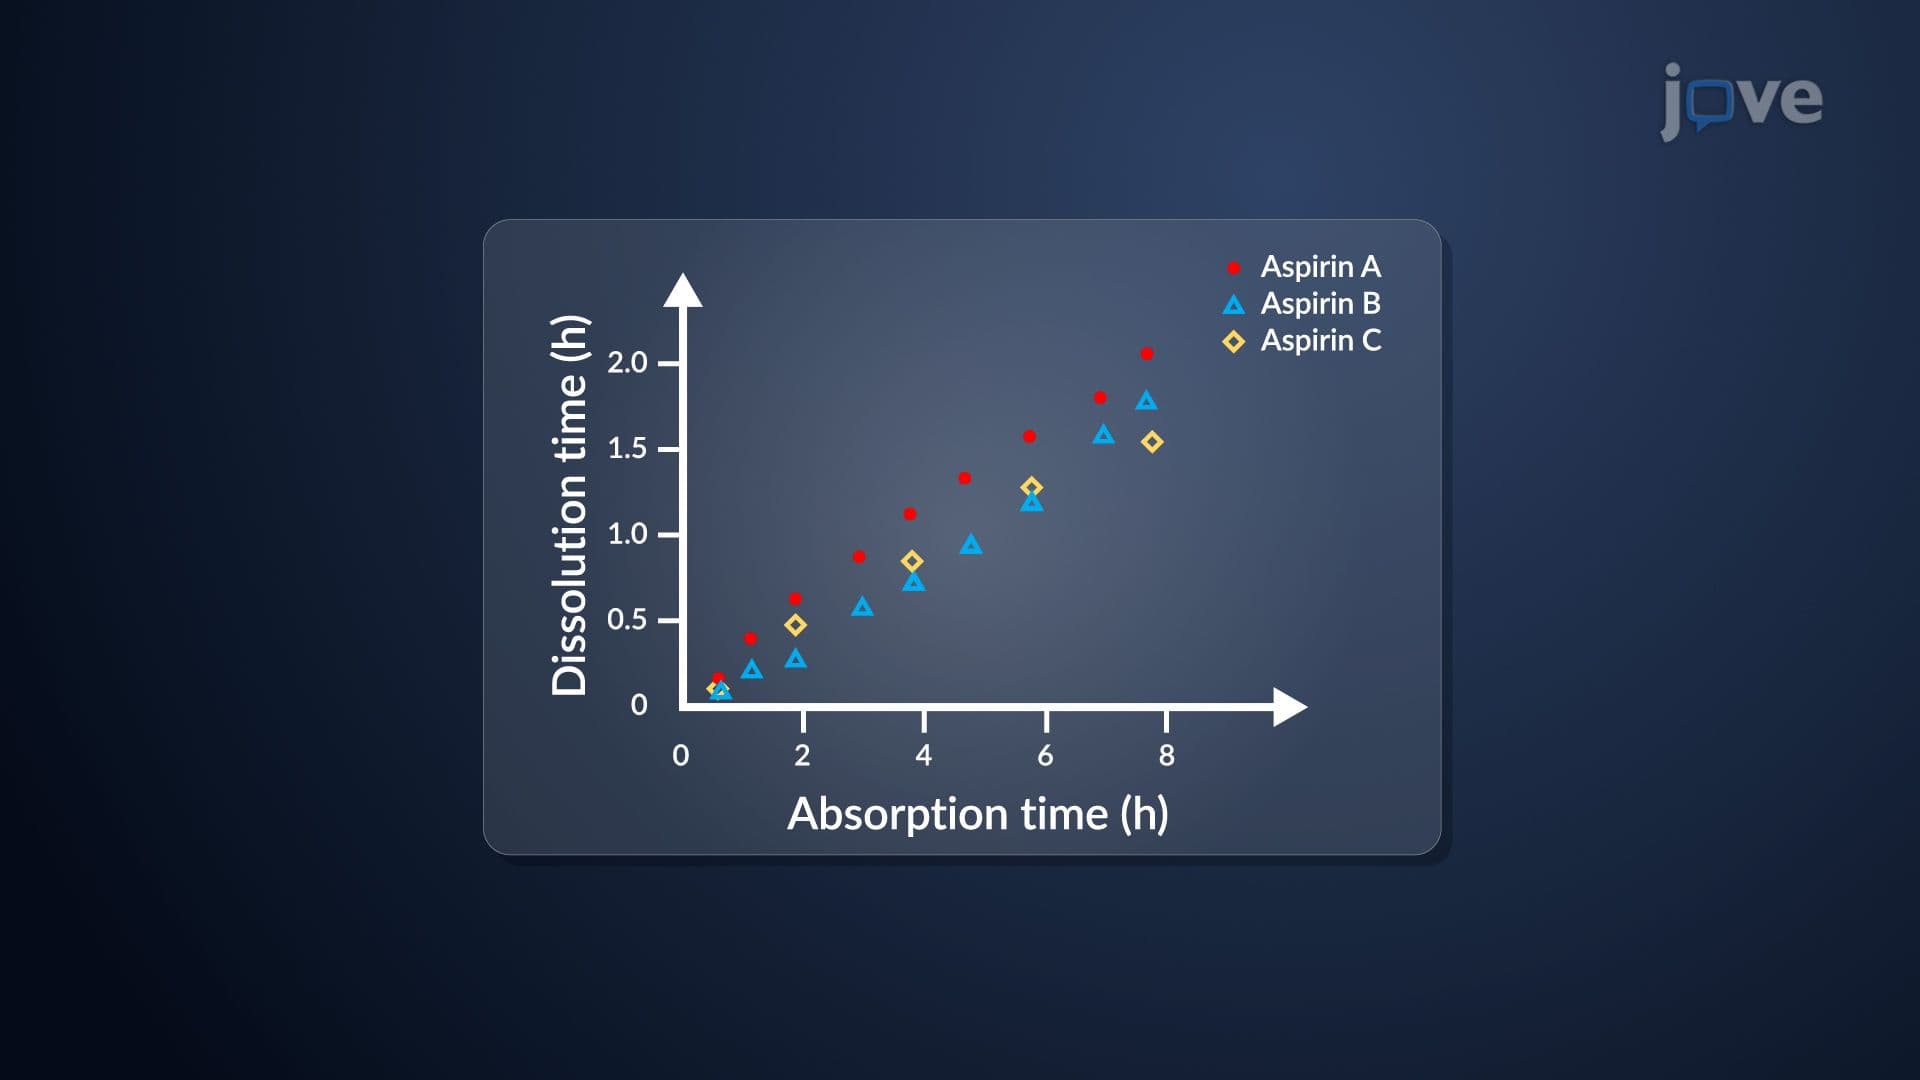
Drug Product Performance: In Vitro–In Vivo Correlation
 01:24
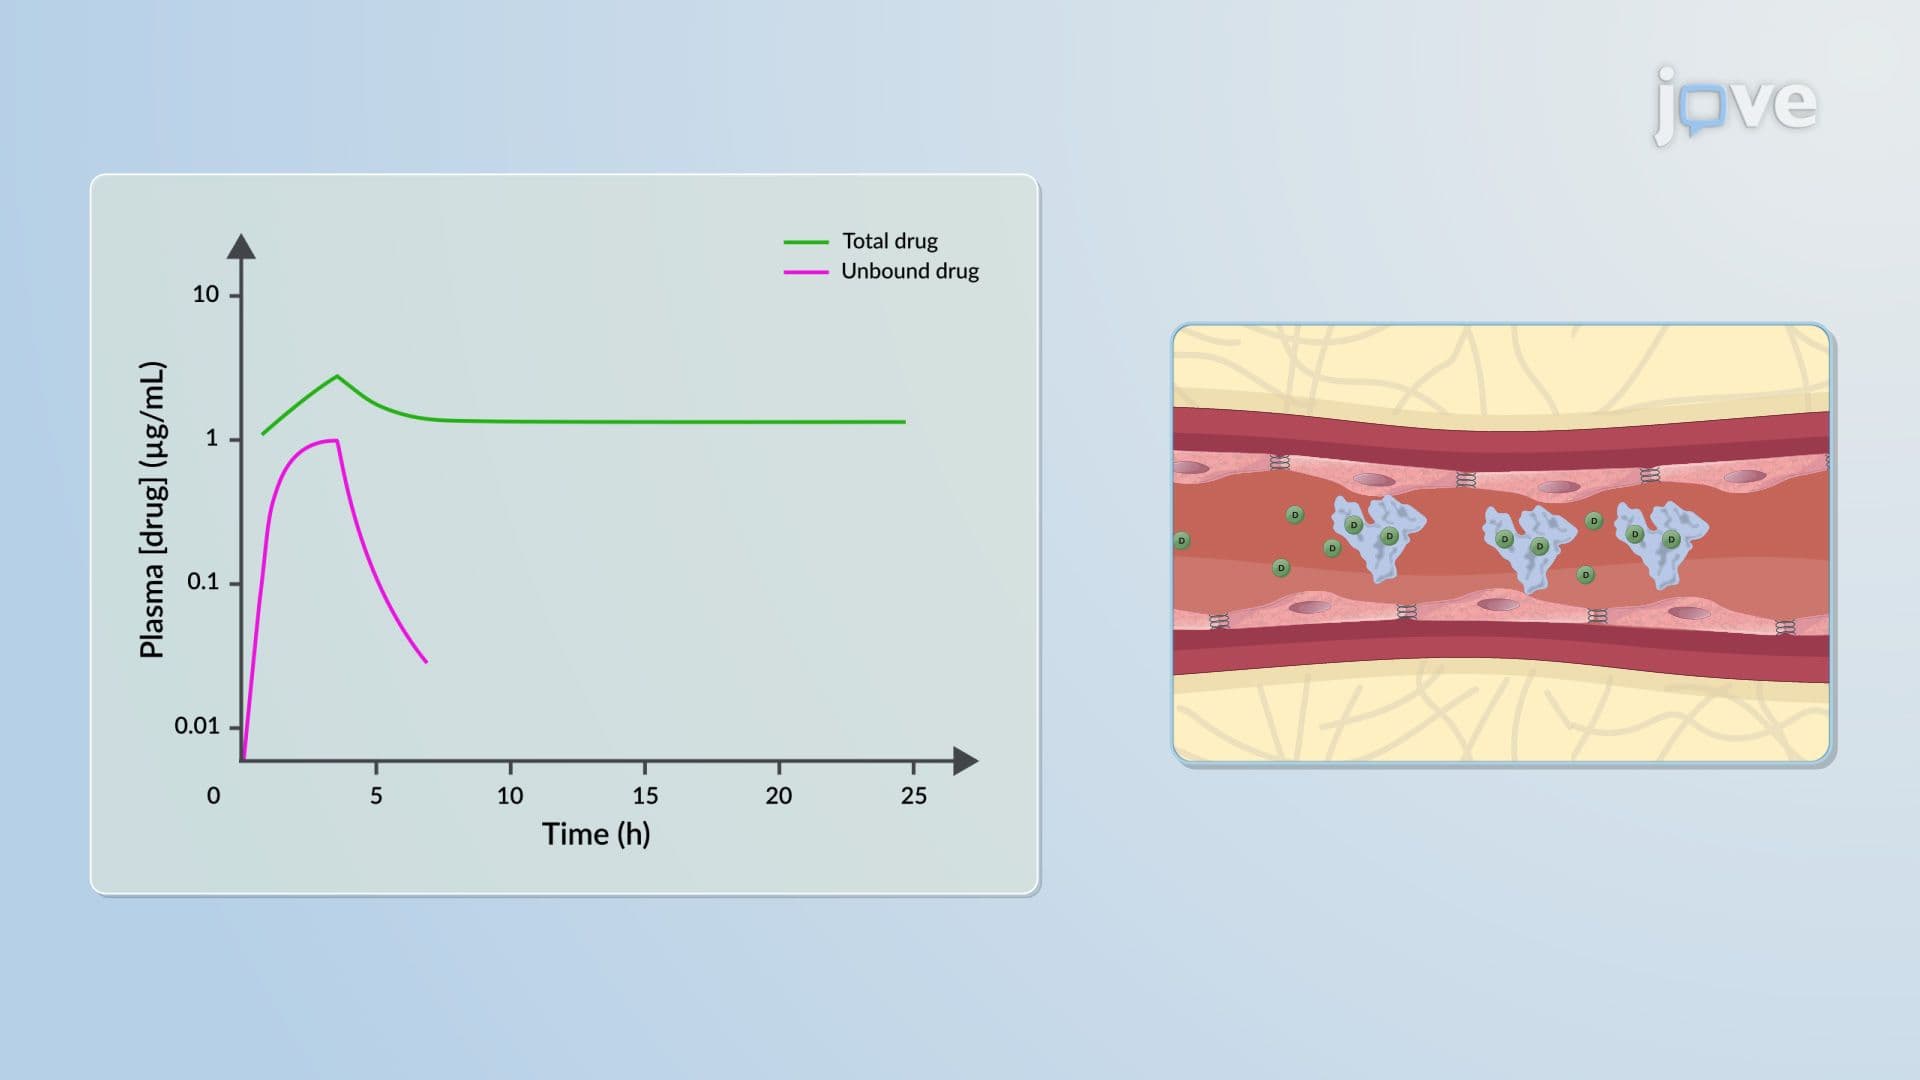
Pharmacokinetic–Pharmacodynamic Relationship: Problems
 01:18
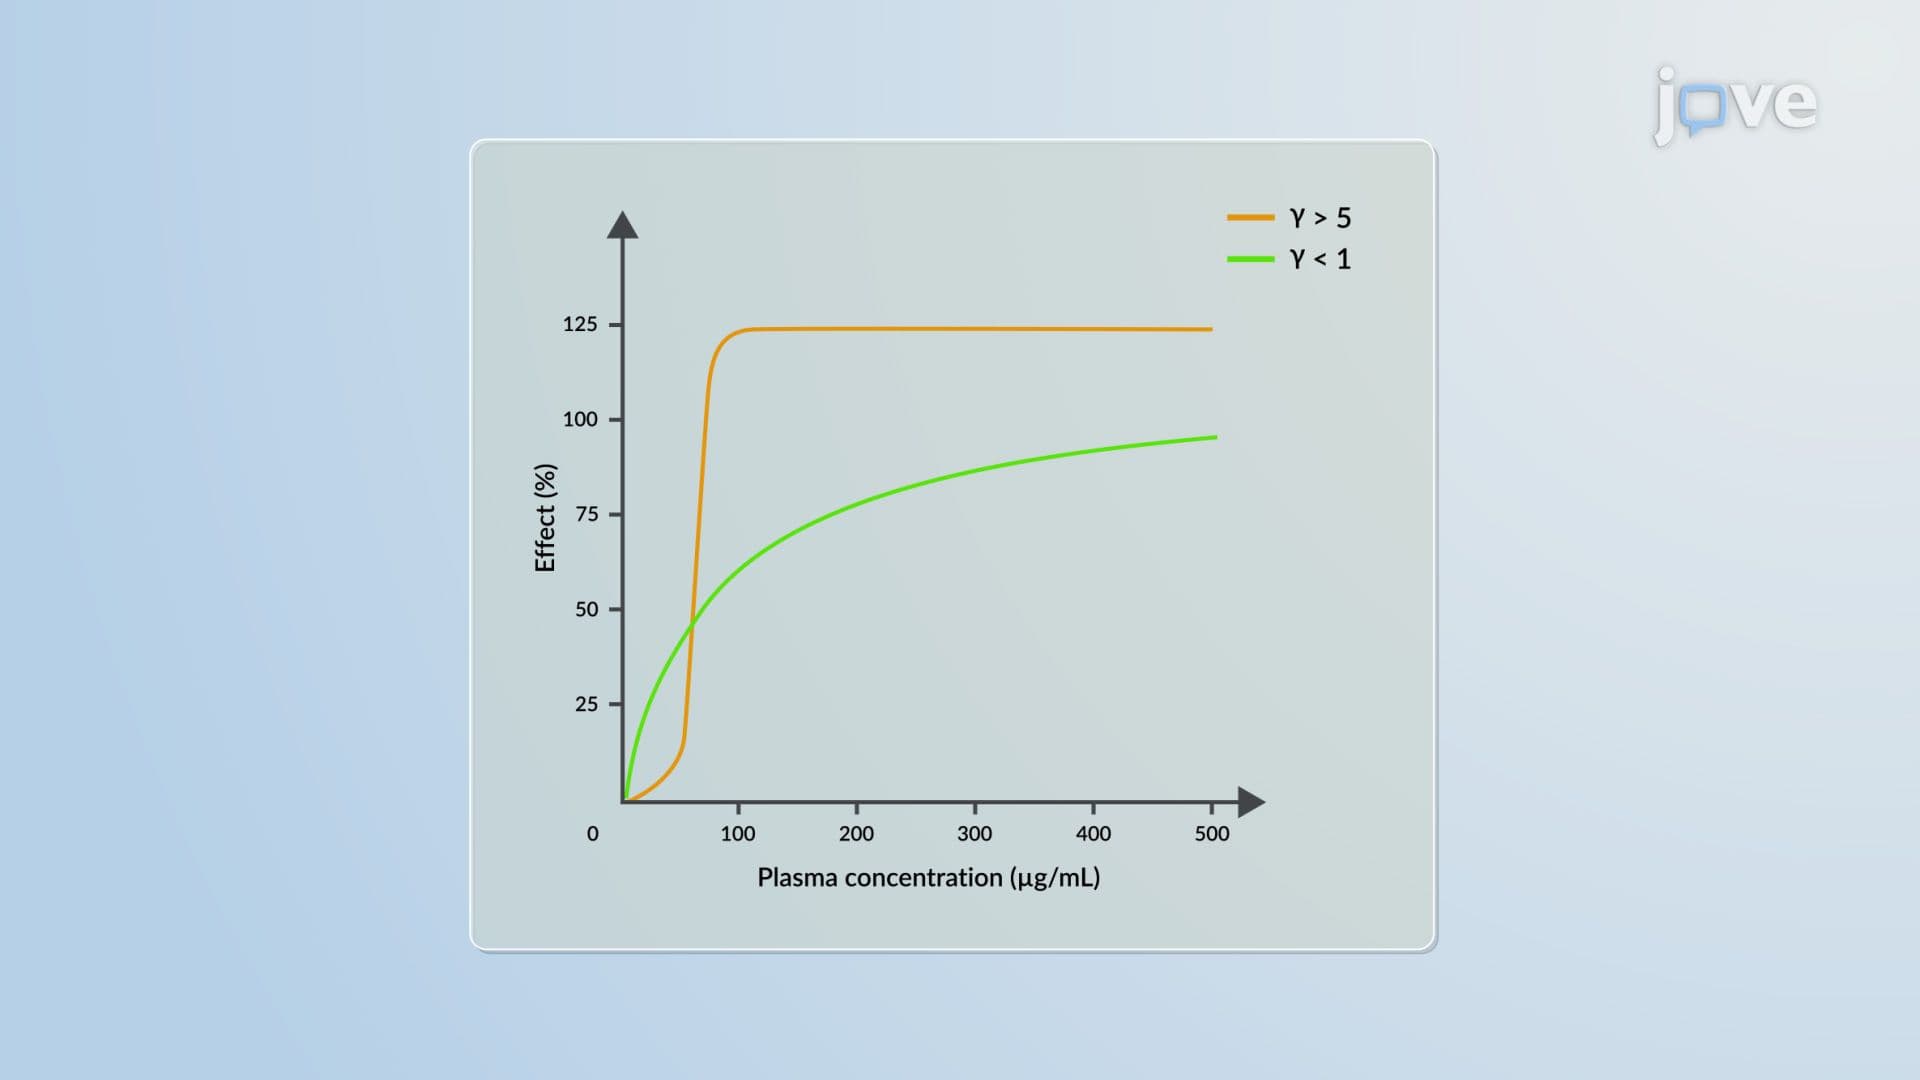
Pharmacodynamic Models: Emax Drug–Concentration Effect Model