Related Experiment Video
Updated: Aug 12, 2026

Atomic Scale Structural Studies of Macromolecular Assemblies by Solid-state Nuclear Magnetic Resonance Spectroscopy
Published on: September 17, 2017
A novel energy-based stochastic method for positioning polar protons in protein structures from X-rays
1Department of Pharmaceutical Chemistry, School of Pharmacy, Hebrew University of Jerusalem, Israel.

Abstract:
A novel automated method for the optimal placement of polar hydrogens in a protein structure is presented. The algorithm adds initially, to a protein data bank file of the protein, nonrotatable hydrogens such as peptide backbone hydrogens according to geometric considerations. Then, water protons and polar side chain protons of lysine, serine, threonine, tyrosine, aspartic acid, glutamic acid, and the C and N termini of a protein are added according to energy considerations. A unique stochastic approach has been developed to overcome a combinatorial explosion in the search for the lowest energy structure. First, the system is divided into ensembles. Each ensemble is treated separately: N conformations are sampled at random, their energies computed, whereas common components of high-energy combinations are gathered on one hand, and low-energy combinations on the other. Components that yield only high-energy conformations and do not contribute to any low energies are excluded. This is reiterated while the total amount of combinations is decreased along the iterative process. When the total number of combinations is lower than a user defined threshold, all remaining combinations are evaluated by exhaustive search. Energy evaluations use nonbonding energy expressions alone. The program was tested on five high-resolution crystal structures: bovine pancreatic trypsin inhibitor (Brookhaven Protein Data Bank file 5PTI), RNase-A (5RSA), trypsin (1NTP), and carbon monoxymyoglobin (2MB5), for which neutron diffraction structures are available, as well as phosphate binding protein (1IXH) for which very high resolution X-ray crystallography was used. The low RMS values prove the efficiency of this algorithm as a tool for positioning protons in proteins. It may be used for other biological structures.
More Related Videos
 10:10
Neutron Crystallography Data Collection and Processing for Modelling Hydrogen Atoms in Protein Structures
Published on: December 1, 2020
 07:24
Paramagnetic Relaxation Enhancement for Detecting and Characterizing Self-Associations of Intrinsically Disordered Proteins
Published on: September 23, 2021
Related Concept Videos
 02:06
Newman Projections
The organic molecules rotate across the single bonds leading to numerous temporary three-dimensional structures of varying energy known as conformers.
 01:20
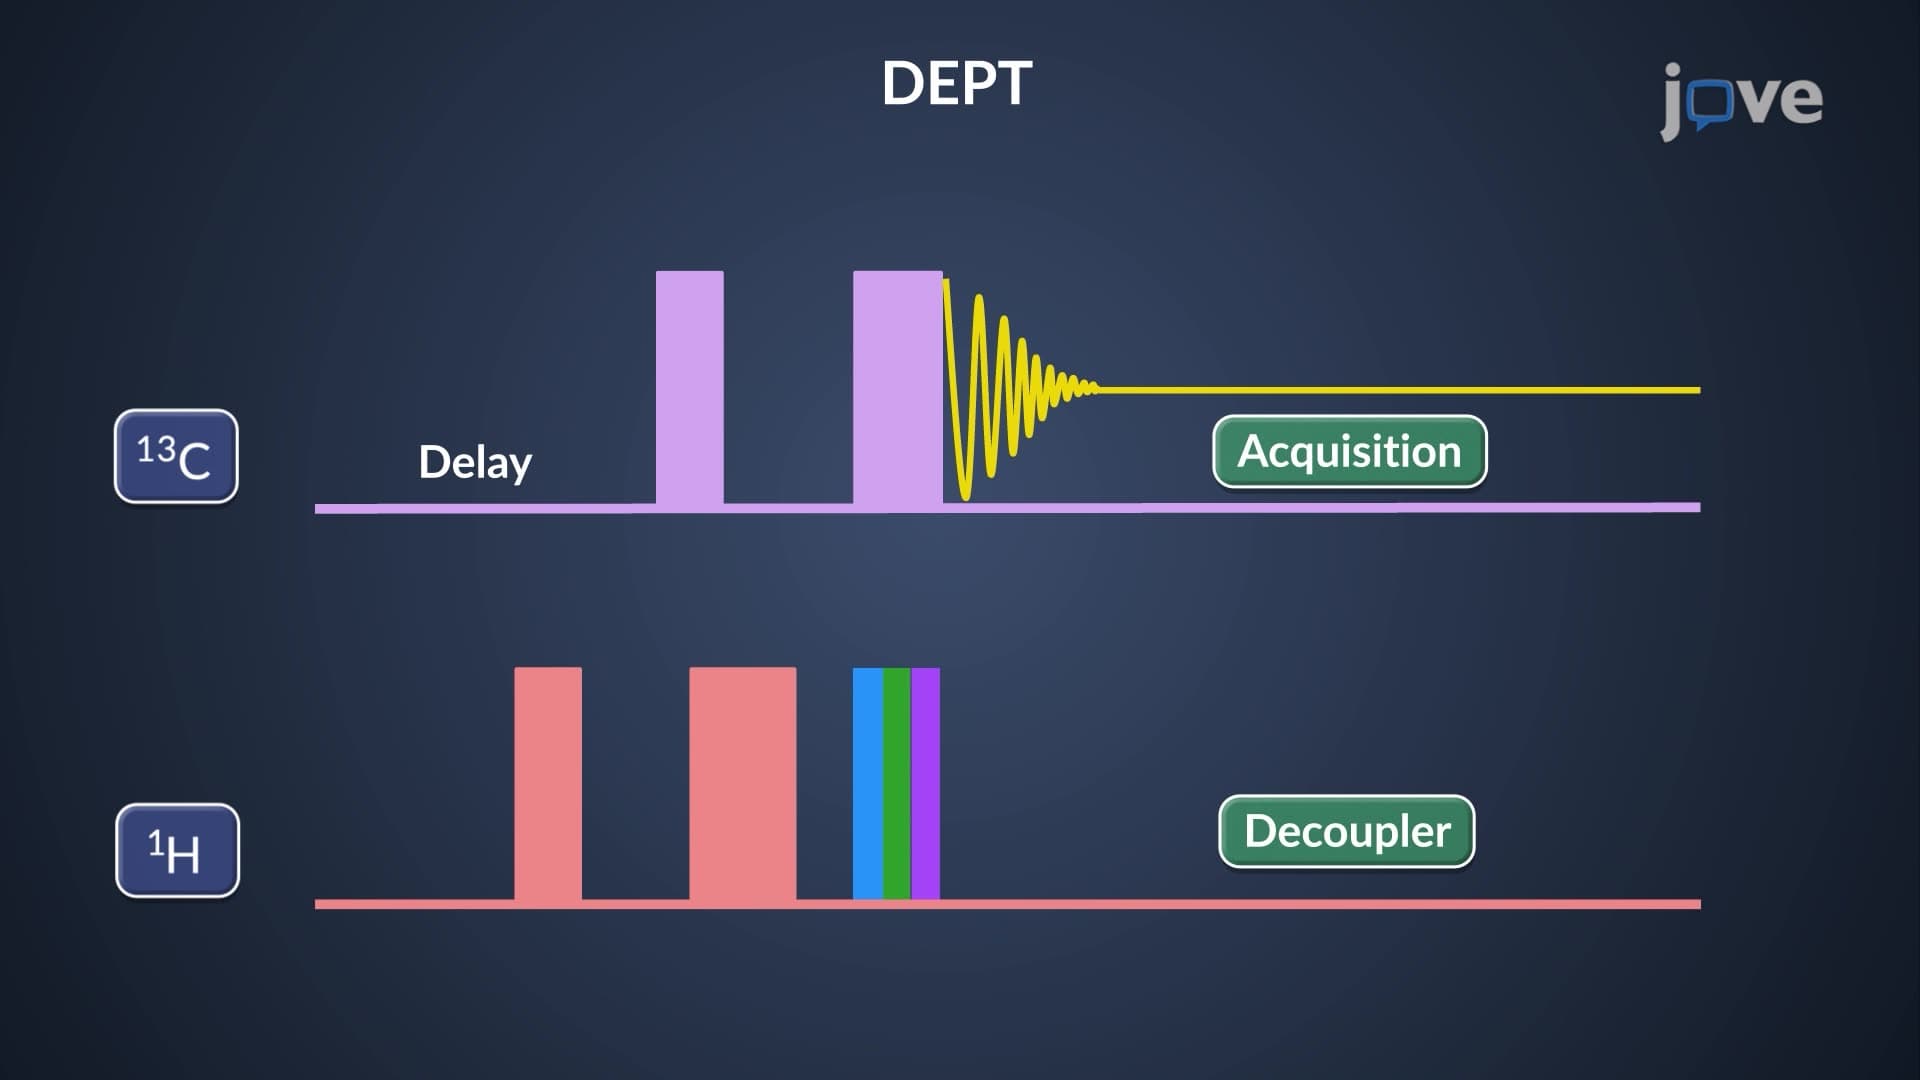
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 01:24
Protein Organization
The primary structure of a protein is its amino acid sequence.
 01:03
Atomic Nuclei: Nuclear Spin State Overview
 01:15
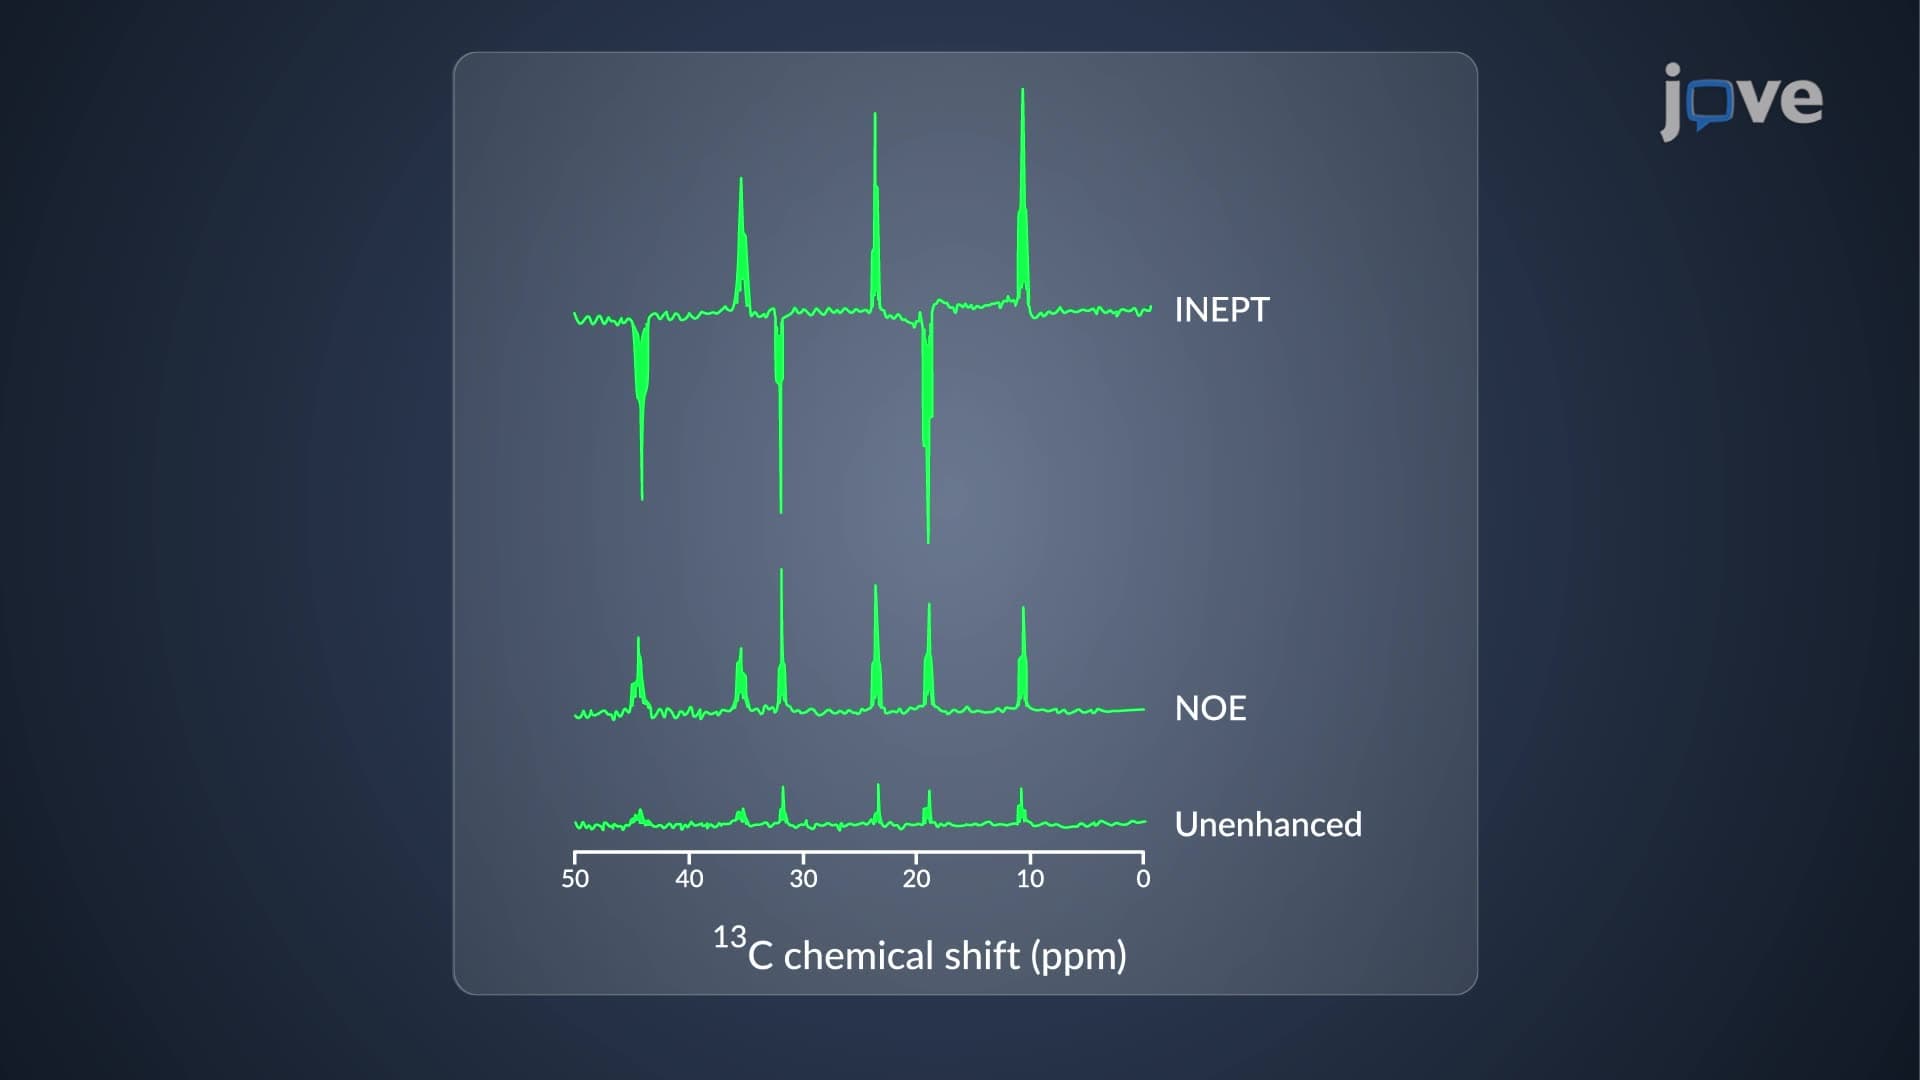
Insensitive Nuclei Enhanced by Polarization Transfer (INEPT)