Related Experiment Video
Updated: Aug 14, 2026
Preparation of N-(2-alkoxyvinyl)sulfonamides from N-tosyl-1,2,3-triazoles and Subsequent Conversion to Substituted Phthalans and Phenethylamines
Published on: January 3, 2018
Design, synthesis, and structure-activity relationships of phthalimide-phenylpiperazines: a novel series of potent
G H Kuo1, C Prouty, W V Murray
1Drug Discovery Division, Analytical Development, Global Chemical & Pharmaceutical Development Division, The R.W. Johnson Pharmaceutical Research Institute, Raritan, New Jersey 08869, USA. gkuo@prius.jnj.com

Abstract:
Beginning from the screening hit and literature alpha(1)-adrenergic compounds, a hybridized basic skeleton A was proposed as the pharmacophore for potent and selective alpha(1a)-AR antagonists. Introduction of a hydroxy group to increase the flexibility afforded B which served as the screening model and resulted in the identification of the second-generation lead 1. Using the Topliss approach, a number of potent and selective alpha(1a)-AR antagonists were discovered. In all cases, binding affinity and selectivity at the alpha(1a)-AR of S-hydroxy enantiomers were higher than those of the R-hydroxy enantiomers. As compared to the des-hydroxy analogues, the S-hydroxy enantiomers had slightly lower binding affinity at alpha(1a)-AR but gained more than 2-fold selectivity for alpha(1a)-AR over alpha(1b)-AR, and 2- to 6-fold selectivity for alpha(1a)-AR over alpha(1d)-AR. They also had less cross activities against a panel of 25-35 peripheral and CNS receptors. The S-hydroxy enantiomers 23 and 24 (K(i) = 0.29 nM, 0.33 nM; alpha(1b)/alpha(1a) >5690, >6060; alpha(1d)/alpha(1a) = 186, 158, respectively) were slightly less potent but much more selective at alpha(1a)-AR than tamsulosin (K(i) = 0.13 nM, alpha(1b)/alpha(1a) = 14.8, alpha(1d)/alpha(1a) = 1.4). In the functional assay, the S-hydroxy enantiomers 20, 23, and 24 were less potent than tamsulosin in inhibiting contractions of rat prostate tissue but more selective in the inhibition of tissue contractions of rat prostate versus rat aorta. Compound 24 was selected as the development candidate for the treatment of BPH.
More Related Videos
 07:16
Methods for the Discovery of Novel Compounds Modulating a Gamma-Aminobutyric Acid Receptor Type A Neurotransmission
Published on: August 16, 2018
 10:01
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Related Concept Videos
 01:22
Direct-Acting Cholinergic Agonists: Chemistry and Structure-Activity Relationship
The direct-acting...
 01:29
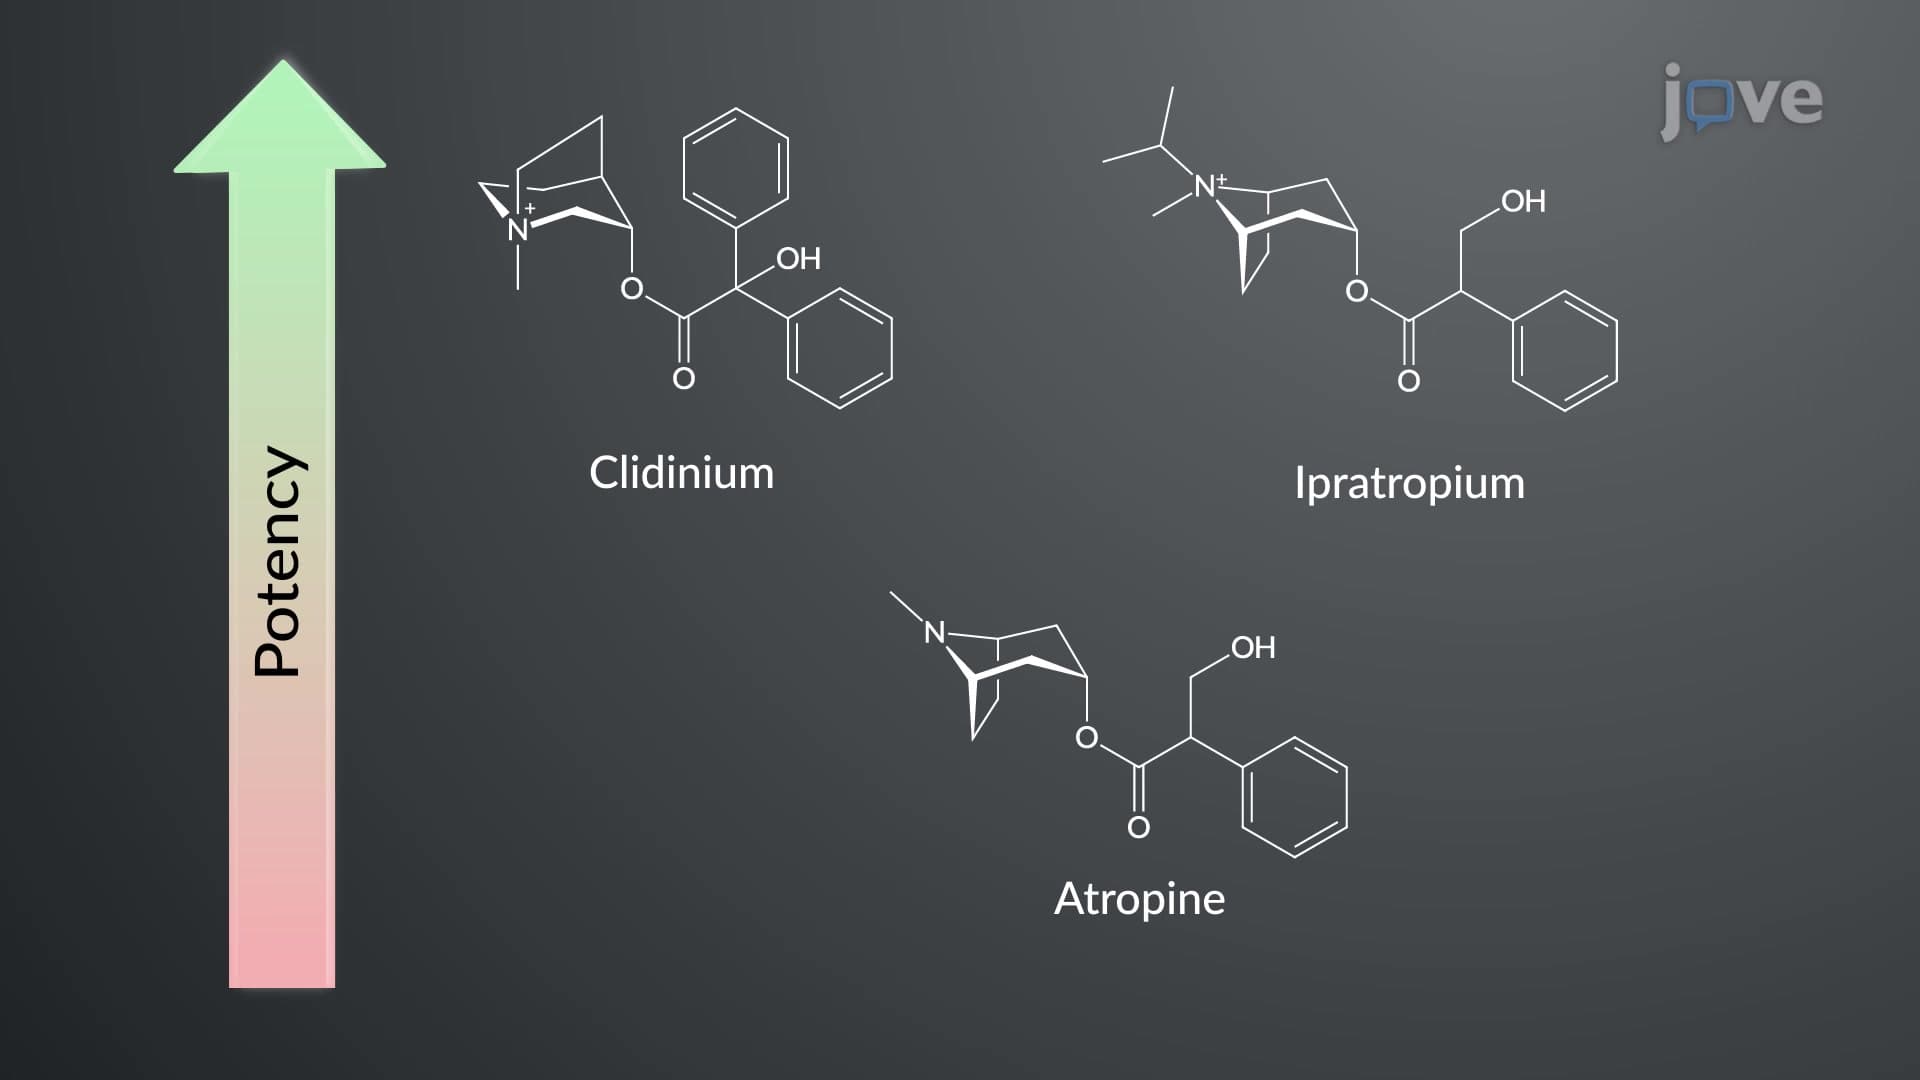
Cholinergic Antagonists: Chemistry and Structure-Activity Relationship
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:17
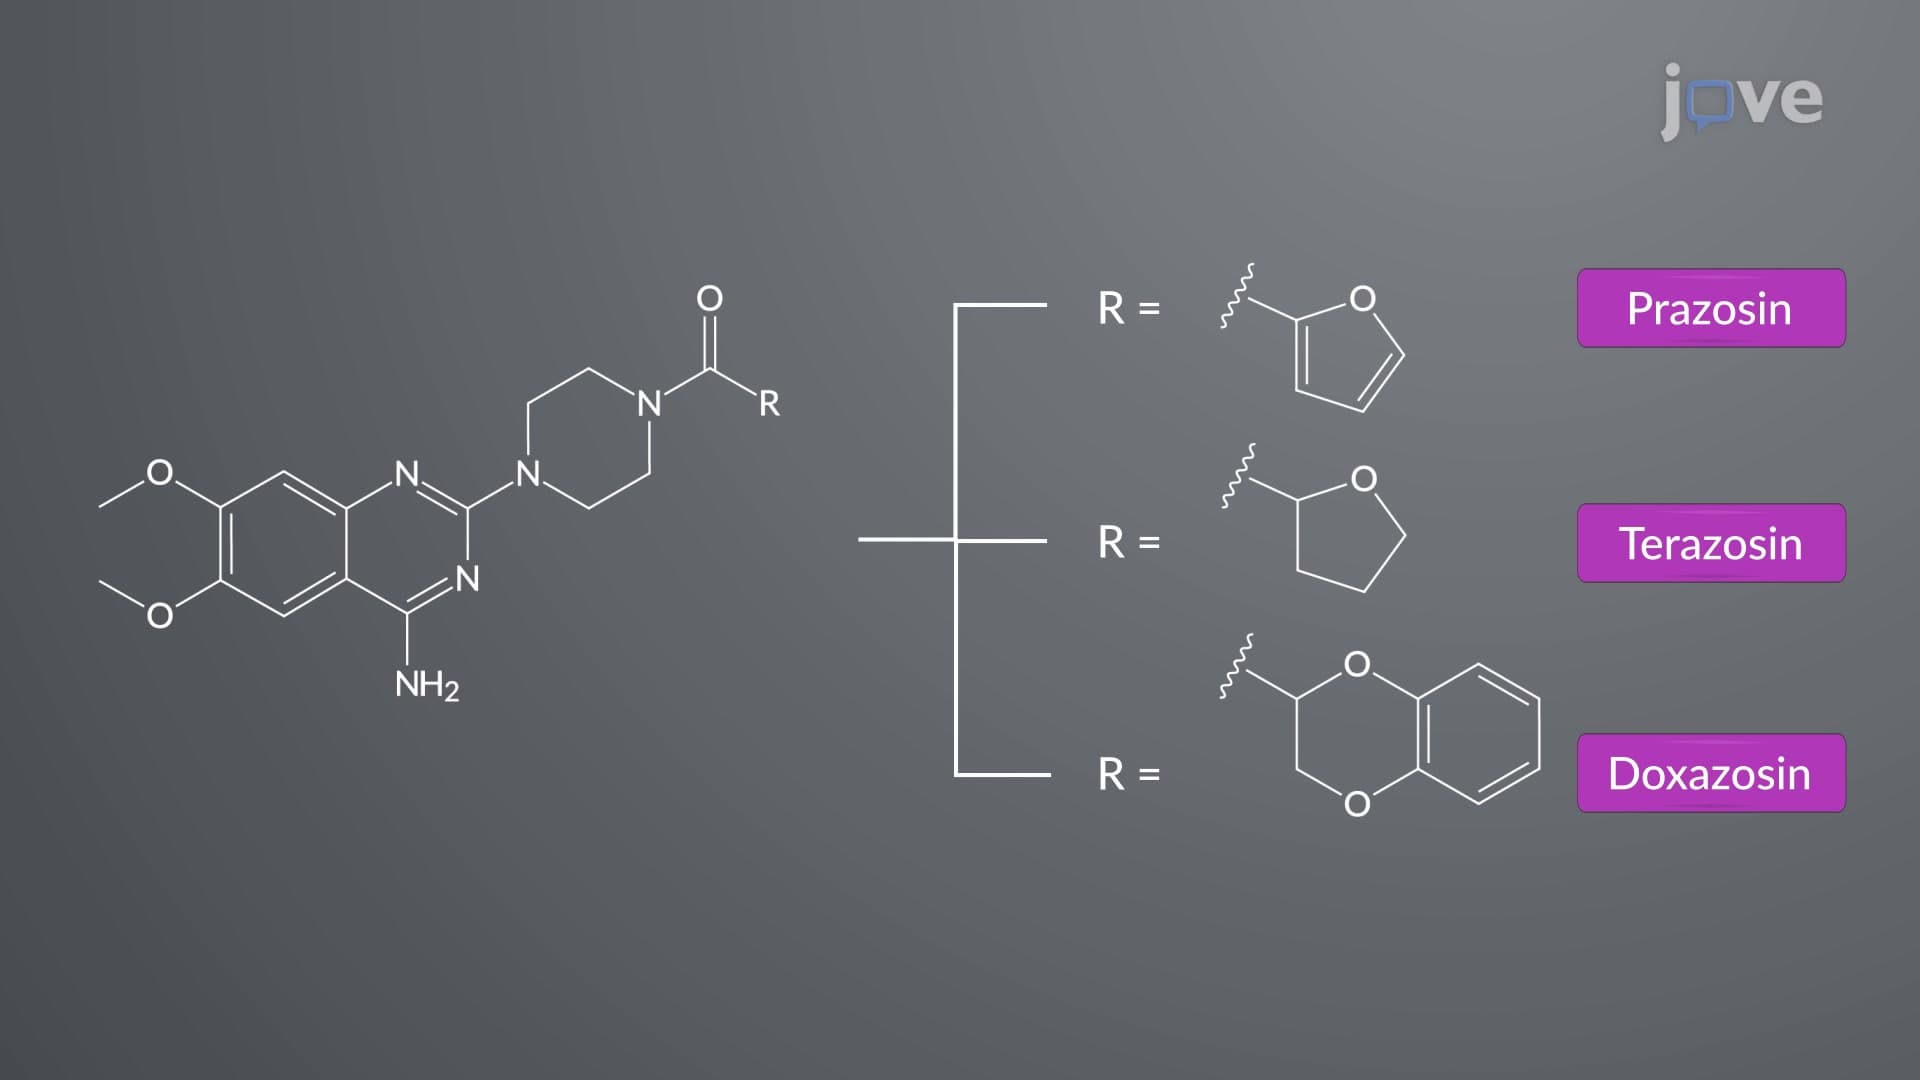
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...
 01:22
Adrenergic Antagonists: Pharmacological Actions of ɑ-Receptor Blockers
α1-blockers: These drugs inhibit α1-adrenoceptors on smooth muscle cells, resulting in vasodilation. This vasodilation lowers blood pressure, making α1-blockers valuable in treating hypertension. Additionally, α1-blockers effectively address urinary obstruction...
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...