An algorithm for predicting proton nuclear magnetic resonance deshielding over a carbon-carbon double bond
Related Experiment Video
Updated: Oct 14, 2026
Atomic Scale Structural Studies of Macromolecular Assemblies by Solid-state Nuclear Magnetic Resonance Spectroscopy
Published on: September 17, 2017
N H Martin1, N W Allen, J D Brown
1Department of Chemistry, University of North Carolina, Wilmington 28403-3297, USA. martinn@uncwil.edu

Abstract:
Hydrogen nuclei located over a carbon-carbon double bond in a strong magnetic field experience NMR shielding effects that result from the magnetic anisotropy of the nearby double bond and various other intramolecular shielding effects. We have used GIAO, a subroutine in Gaussian 98, to calculate isotropic shielding values and to predict the proton NMR shielding increment for a simple model system: methane held in various orientations, positions, and distances over ethene. The average proton NMR shielding increments of several orientations of methane have been plotted versus the Cartesian coordinates of the methane protons relative to the center of ethene. A single empirical equation for predicting the NMR shielding experienced by protons over a carbon-carbon double bond has been developed from these data. The predictive capability of this equation has been validated by comparing the shielding increments for several alkenes calculated using our equation to the experimentally observed shielding increments. This equation predicts the NMR shielding effects more accurately than previous models that were based on fewer geometries of methane over ethene. In fact, deshielding is predicted by this equation for protons over the center and within about 3 A of a carbon-carbon double bond. This result is in sharp contrast to predictions made by the long-held McConnell "shielding cone" model found in nearly every textbook on NMR, but is consistent with experimental observations. The algorithm for predicting the (de)shielding increment for a proton over an alkene can be used in a spreadsheet on a PC or incorporated into software that estimates chemical shifts using additive substituent constants or a database of structures. In either application its use can substantially improve the accuracy of the estimated chemical shift of a proton in the vicinity of a carbon-carbon double bond, and thus assist in spectral assignments and in correct structure determination.
More Related Videos
Related Concept Videos
 01:04
¹³C NMR: ¹H–¹³C Decoupling
A broadband decoupling technique is used to simplify these complex, sometimes overlapping, signals. Broadband decoupling relies on a...
 01:27
π Electron Effects on Chemical Shift: Overview
 01:20
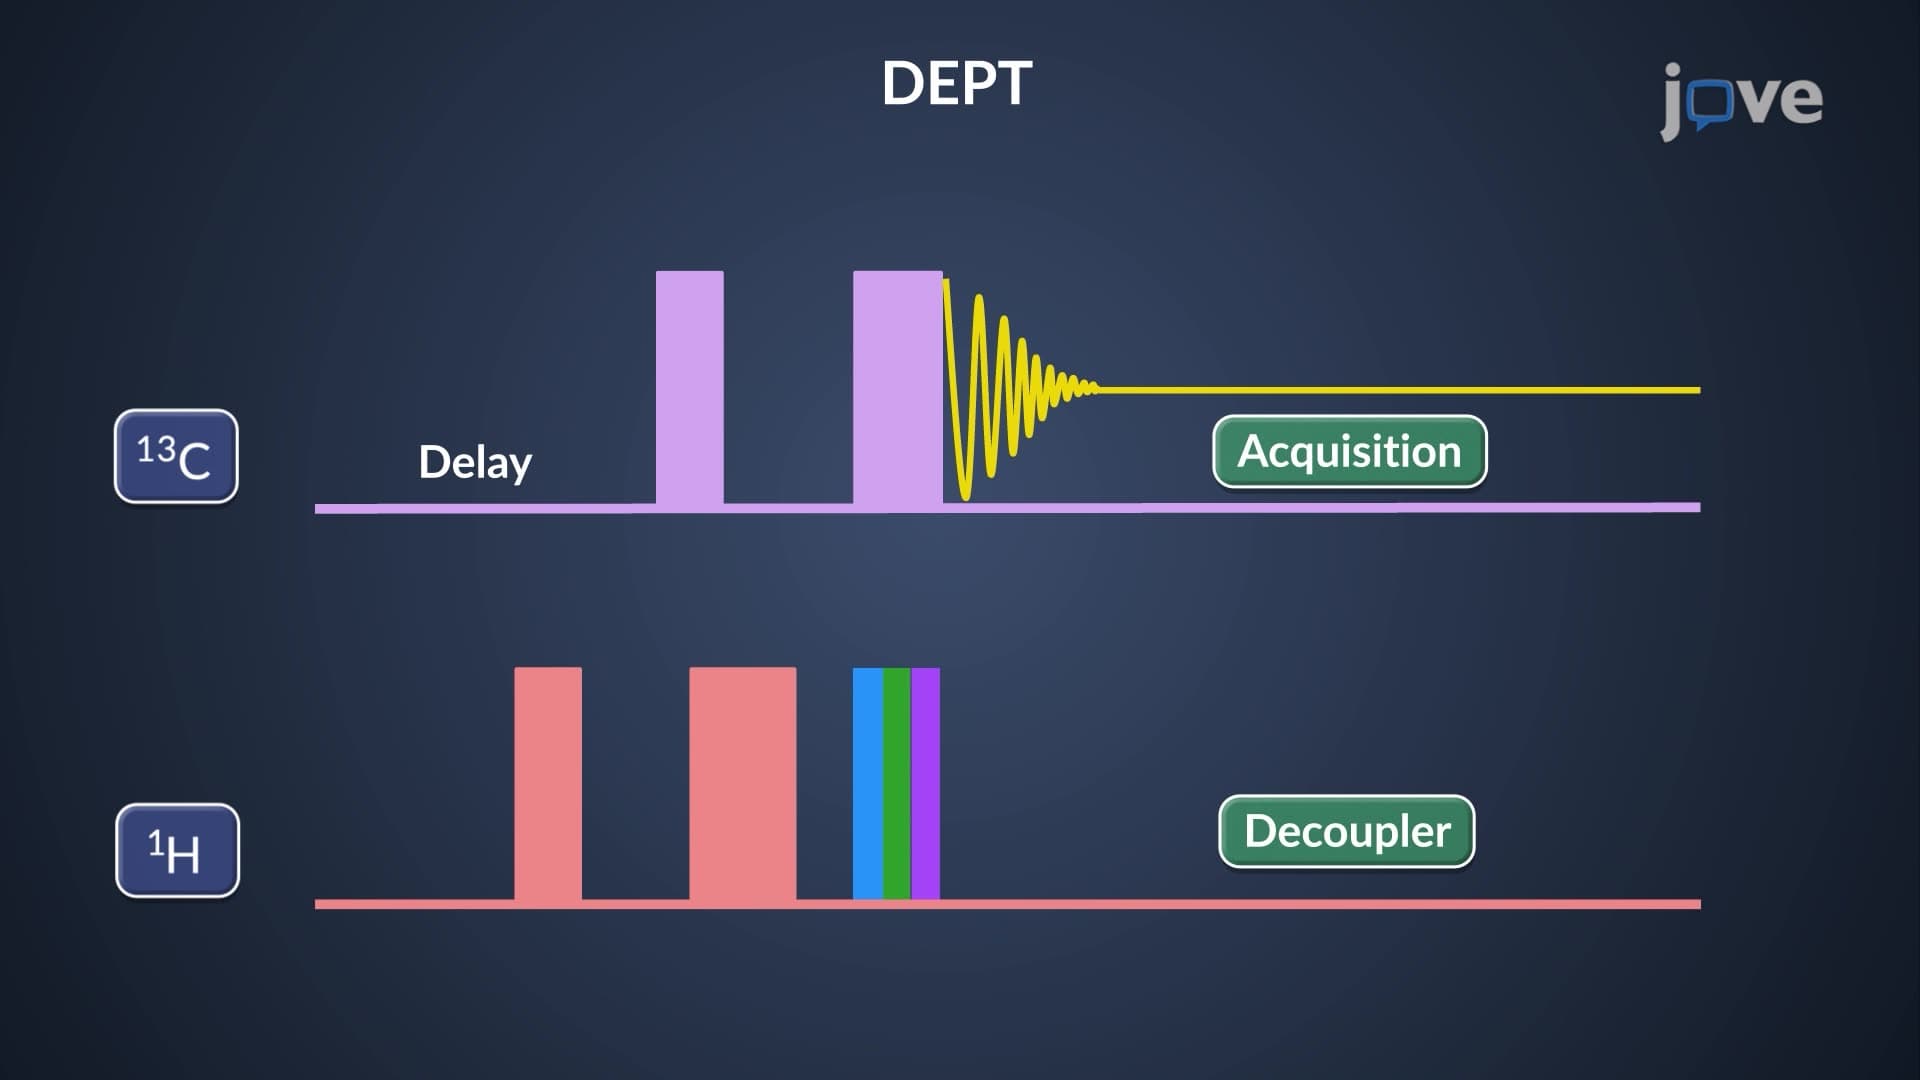
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 01:13
¹H NMR: Complex Splitting
Splitting diagrams or splitting tree diagrams are routinely used to depict such complex couplings. While drawing splitting diagrams, the splitting with the larger coupling constant is usually applied first.
 01:12
Double Resonance Techniques: Overview
Spin decoupling is usually achieved by...
 01:30
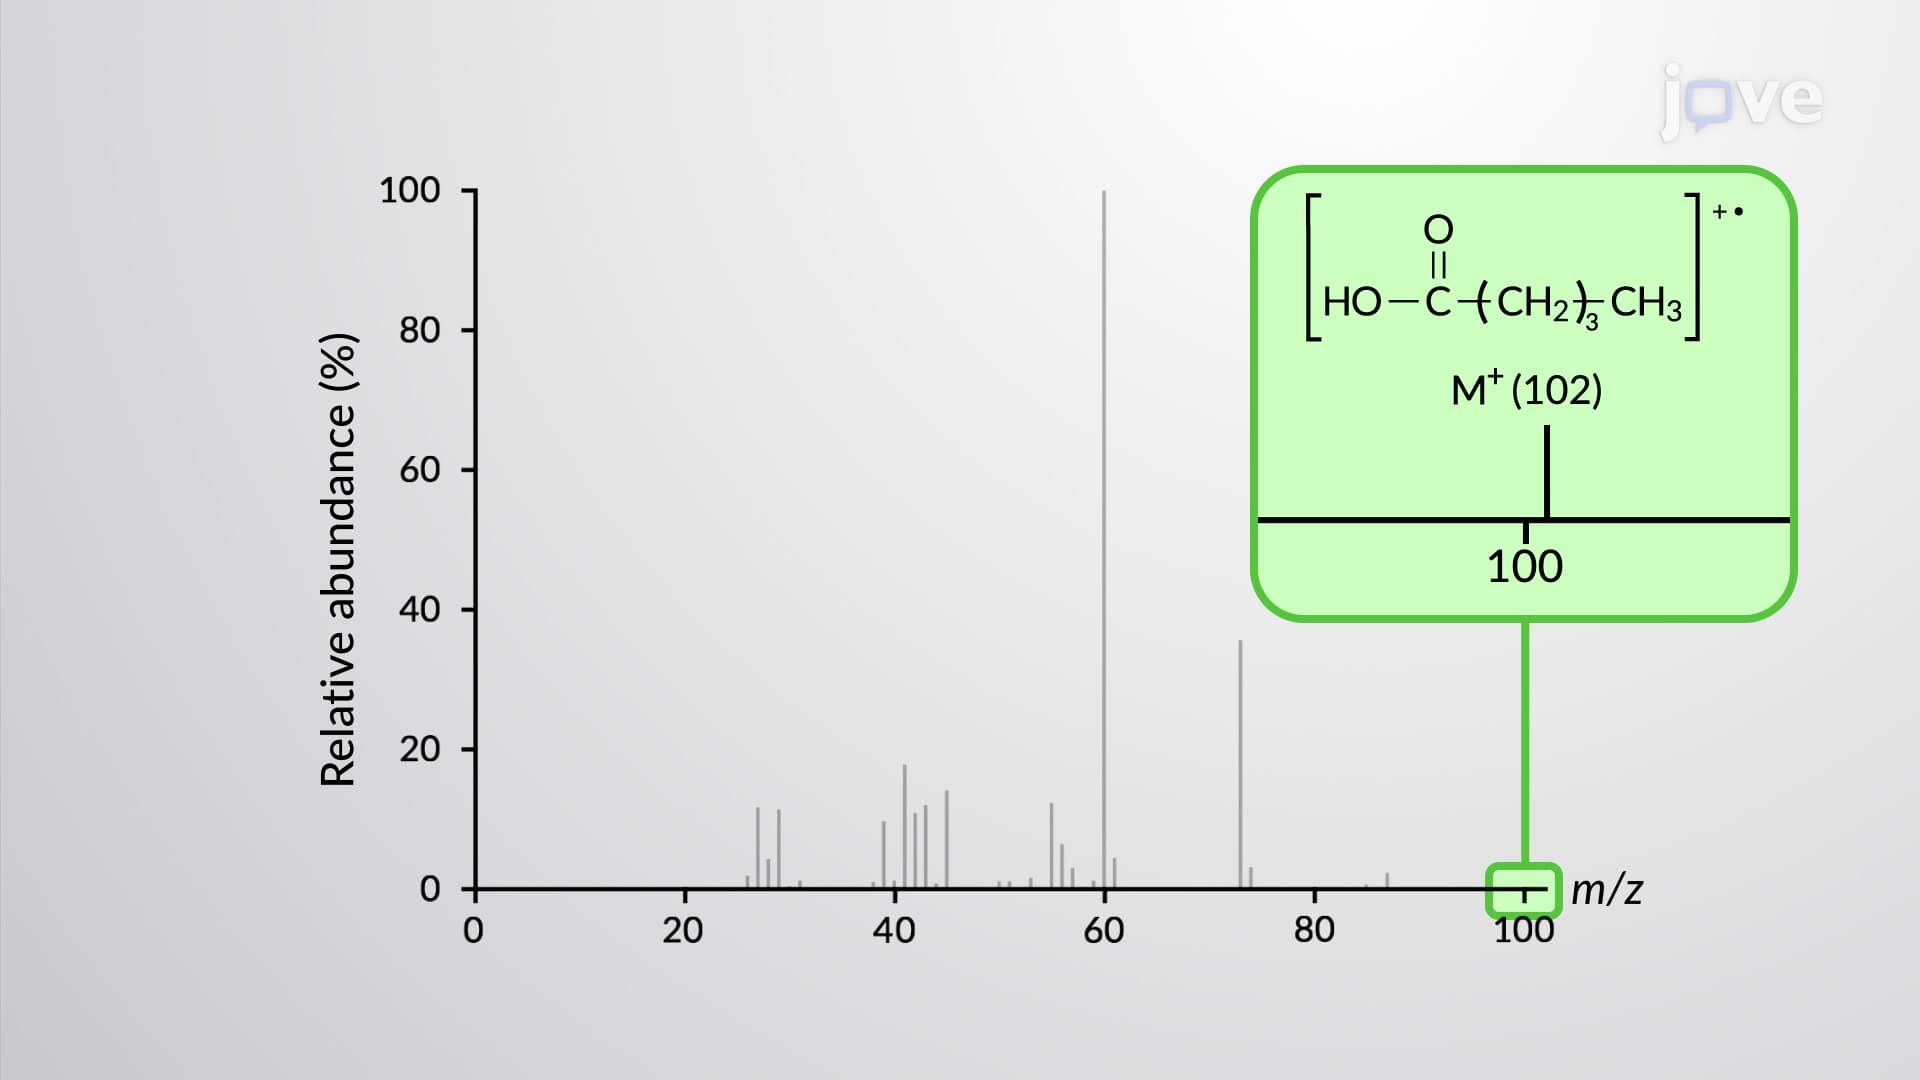
NMR and Mass Spectroscopy of Carboxylic Acids
While α protons of carboxylic acids absorb at 2–2.5 ppm, β protons absorb further upfield.
Carboxylic acids are easily identified by dissolving them in deuterium oxide, which results in a rapid exchange of the acidic protons with deuterium. This leads to the disappearance of the acidic...