Mitochondrial disease: beyond etiology unknown
Related Experiment Video
Updated: Oct 13, 2026
Using Live Cell STED Imaging to Visualize Mitochondrial Inner Membrane Ultrastructure in Neuronal Cell Models
Published on: June 30, 2023
1University of Massachusetts, Lowell, USA. cread@mediaone.net

Insights
Mitochondrial dysfunction, caused by genetic mutations affecting energy production, is a common cause of childhood degenerative diseases. This review covers its role in health, clinical features, and nursing care strategies for families.
Area of Science:
- Biochemistry
- Genetics
- Pediatrics
Background:
- Mitochondria are vital for cellular energy production.
- Mitochondrial dysfunction arises from genetic mutations impacting energy metabolism.
- This dysfunction is increasingly recognized as a cause of pediatric degenerative diseases.
Observation:
- Genetic defects in mitochondrial or nuclear genomes disrupt essential enzyme synthesis.
- This leads to a spectrum of pediatric health issues.
- Affected children may experience developmental delays, seizures, diabetes, and organ failure.
Findings:
- Mitochondrial dysfunction plays a significant role in various pediatric health conditions.
- Clinical manifestations are diverse, ranging from neurological to metabolic disorders.
- Case studies illustrate the challenges faced by affected children and their families.
Implications:
- Understanding mitochondrial disease is crucial for early diagnosis and intervention in children.
- Effective nursing strategies are essential for managing symptoms and supporting families.
- Further research into genetic causes and therapeutic targets is warranted.
Abstract:
Mitochondrial dysfunction is now recognized as a relatively common cause of degenerative disease in children. Mutations in either the mitochondrial or the nuclear genome that cause errors in the synthesis of enzymes essential for energy production and metabolism lead to a wide variety of pediatric problems, including developmental delays, sensorimotor impairment, seizures, diabetes, and organ failure. This article reviews the role of mitochondria in health and illness, discusses the clinical aspects of mitochondrial dysfunction, describes the experiences of three children with mitochondrial disease, and presents nursing strategies for affected families.
Related Concept Videos
 02:59
Animal Mitochondrial Genetics
 01:19
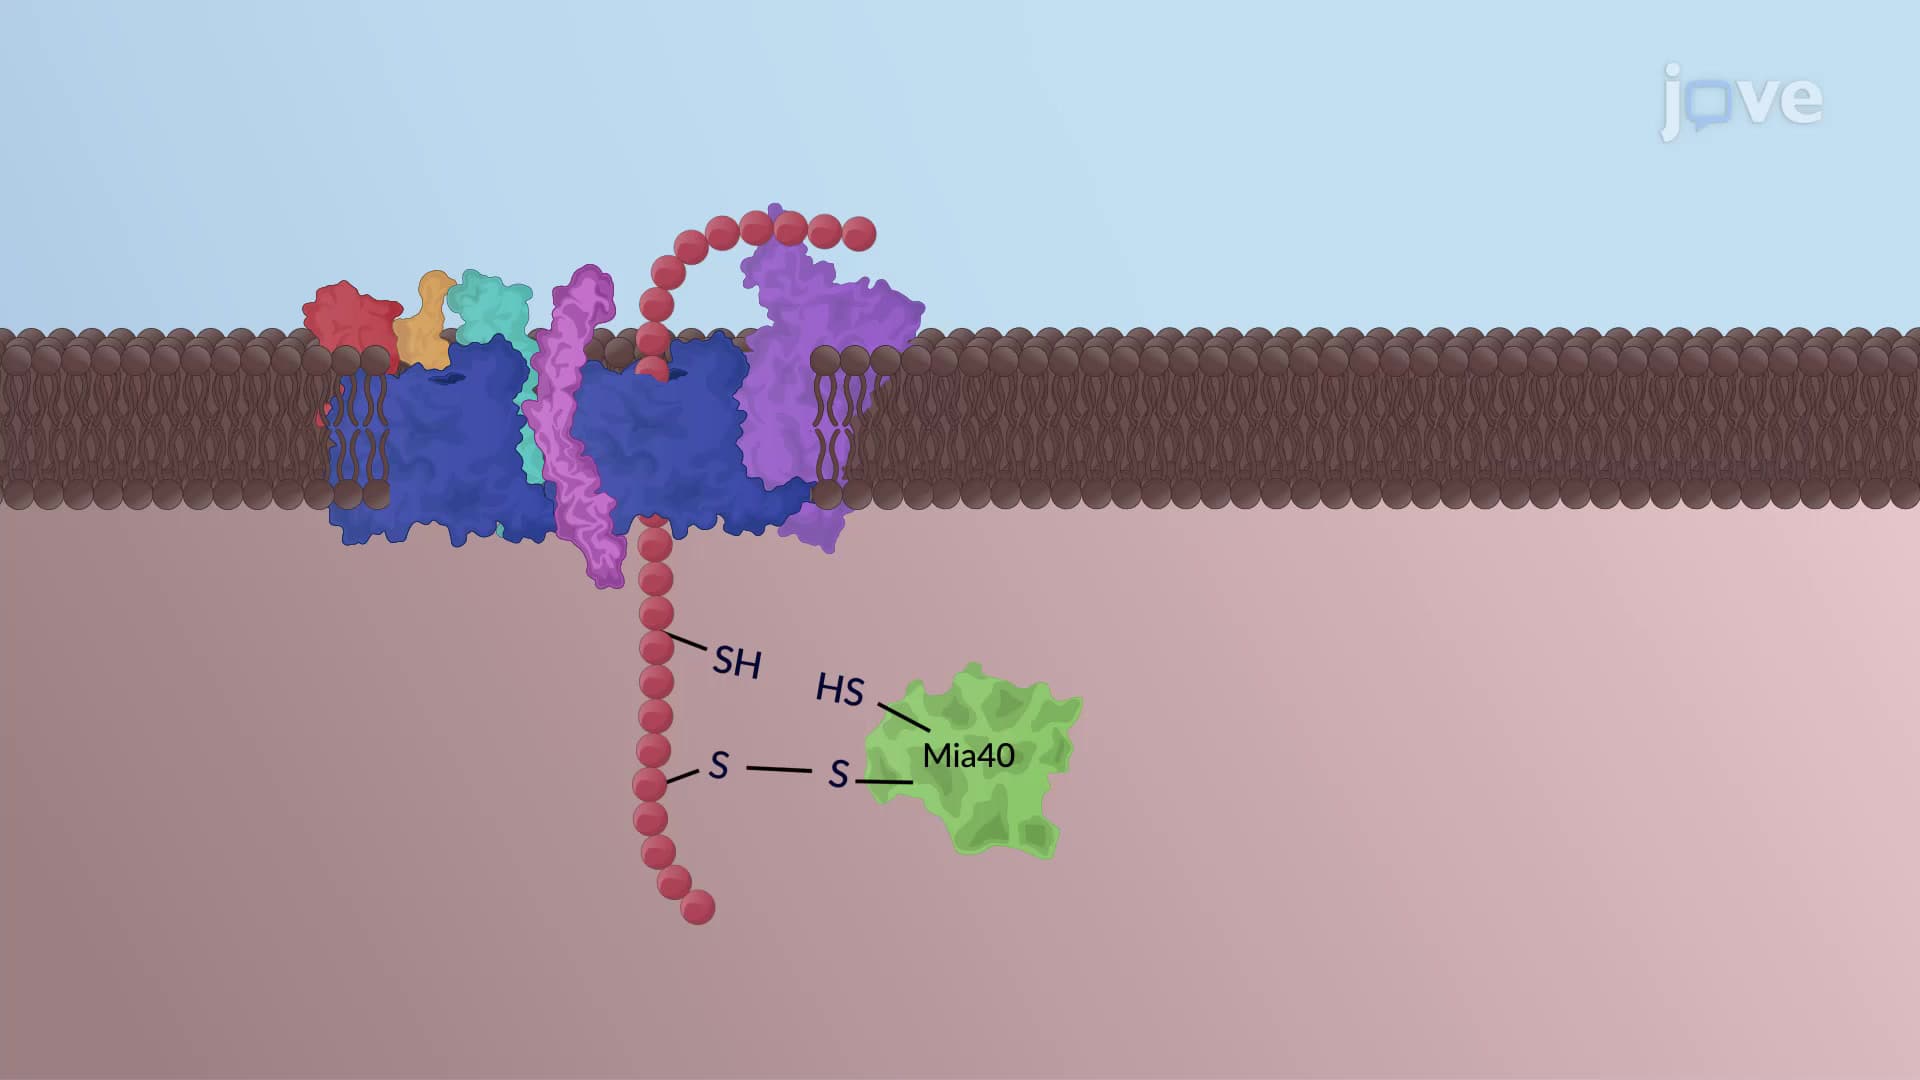
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:39
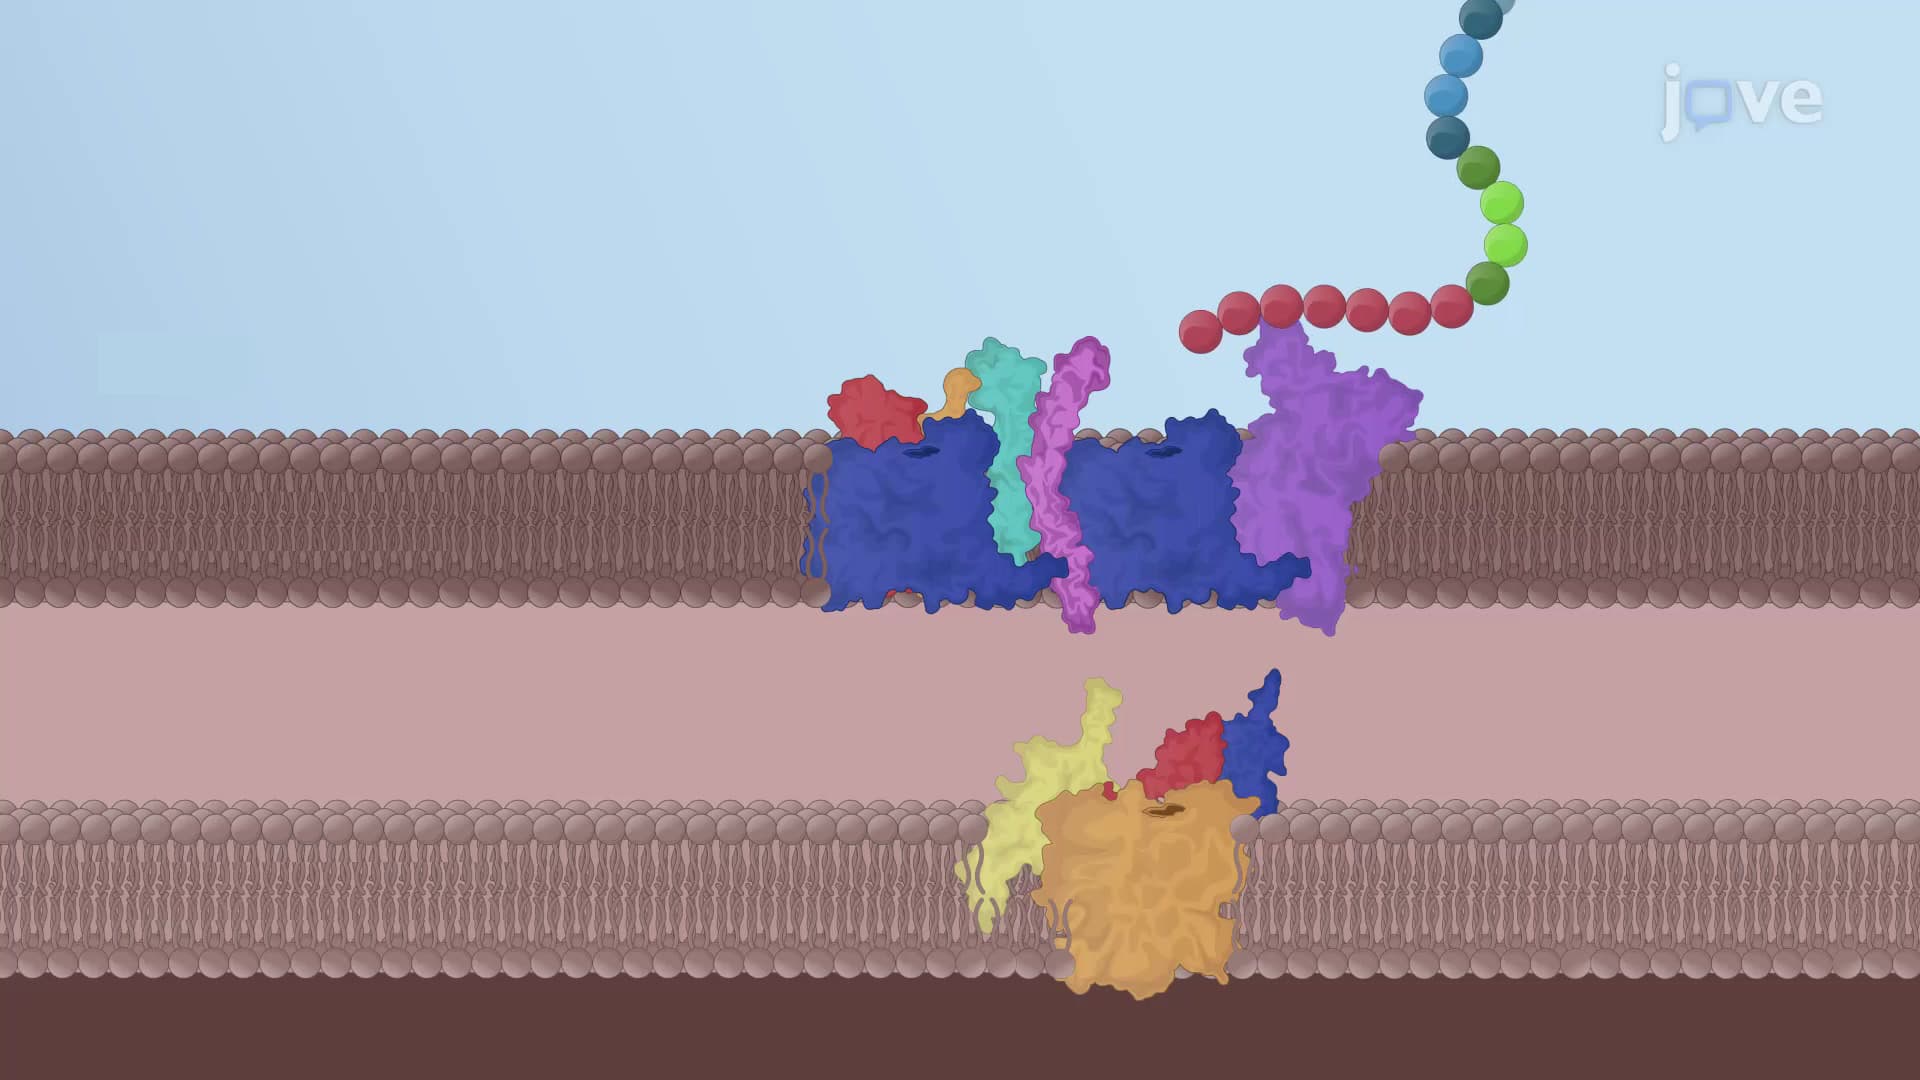
Mitochondrial Precursor Proteins
Most of the mitochondrial precursors...
 01:48
ATP Synthase: Mechanism
 01:24
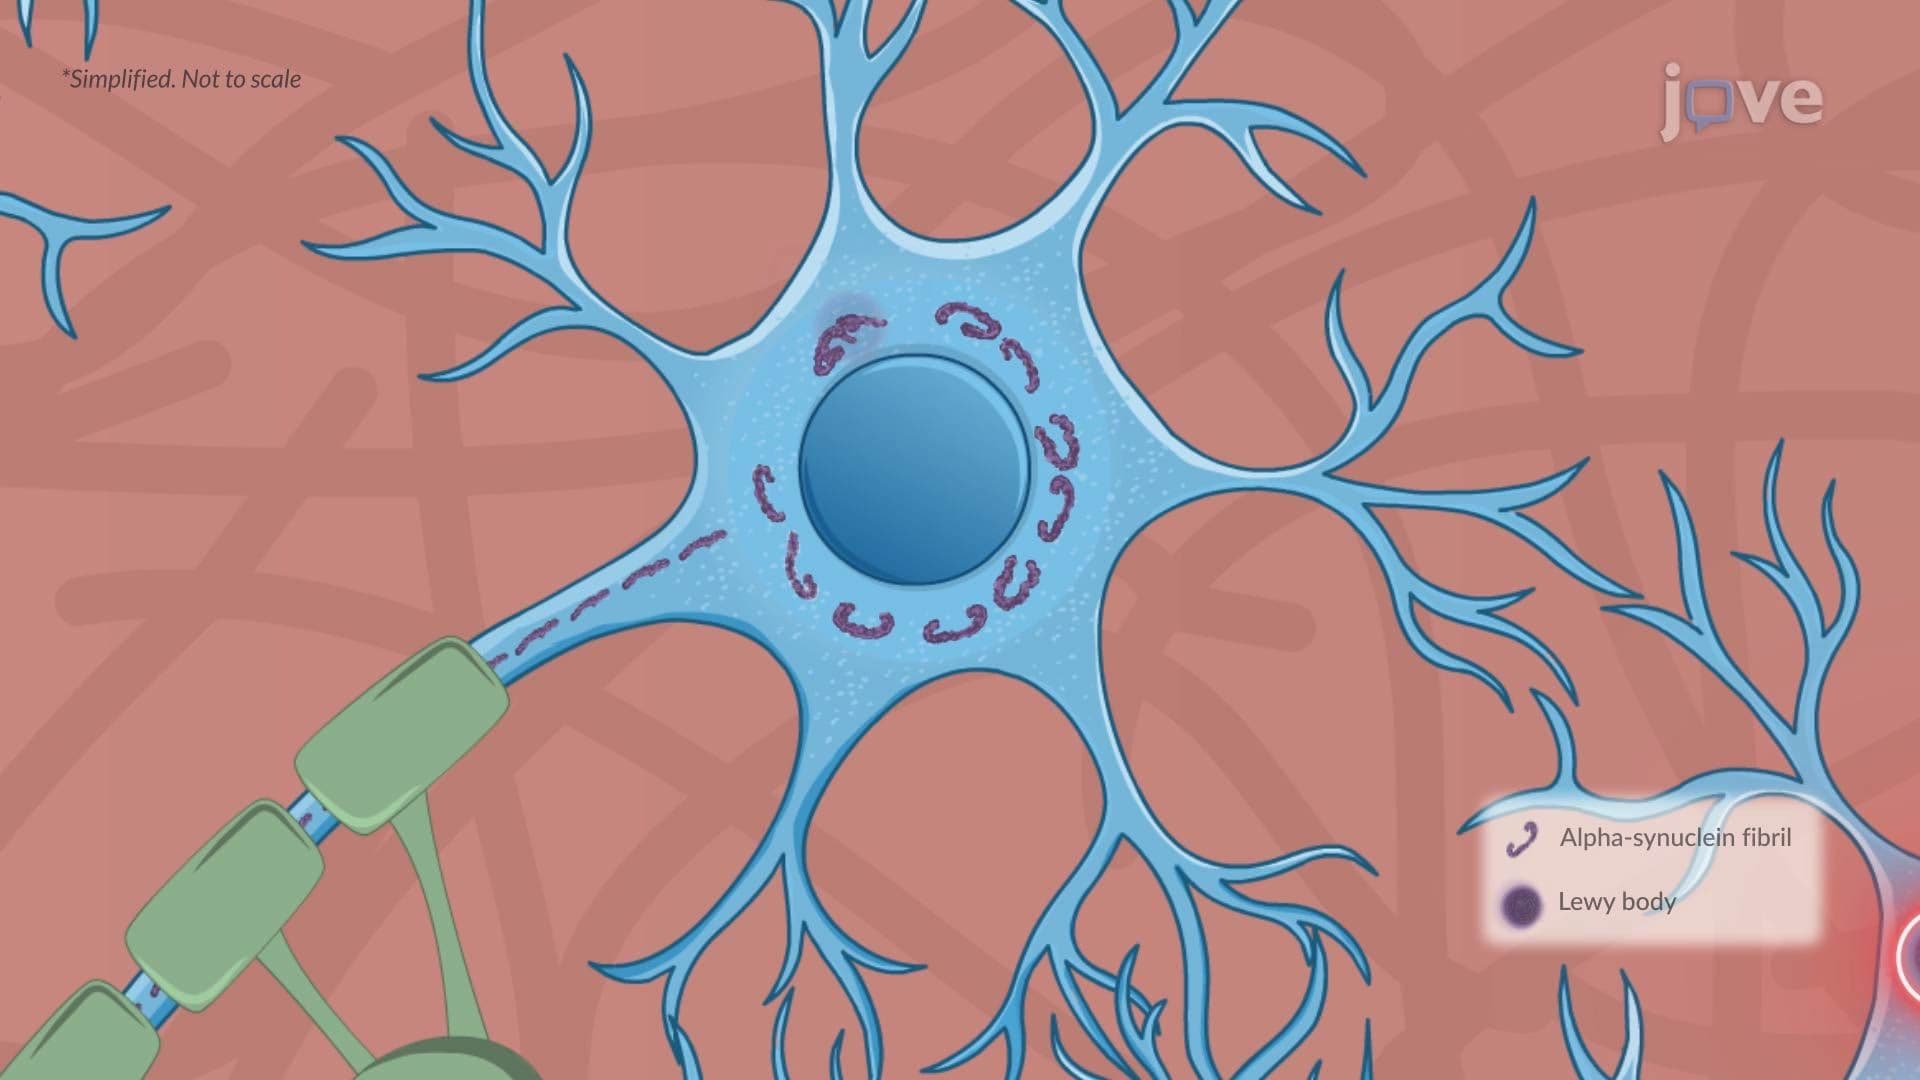
Parkinson Disease ll: Pathophysiology
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...