Using symmetry to monitor geared bond rotation in aromatic amides by dynamic NMR

Bragg1, Clayden

  • 1Department of Chemistry, University of Manchester, Oxford Road, Manchester M13 9PL, U.K.

Organic Letters
|October 13, 2000
PubMed

Related Concept Videos

¹H NMR of Conformationally Flexible Molecules: Temporal Resolution00:52

¹H NMR of Conformationally Flexible Molecules: Temporal Resolution

At room temperature, the chair conformer of cyclohexane undergoes rapid ring flipping between two equivalent chair conformers at a rate of approximately 105 times per second. These two chair conformers are in equilibrium. The rapid ring flipping results in the interconversion of the axial proton to an equatorial proton and an equatorial to the axial proton. Such interconversions are too rapid and cannot be detected on the NMR timescale. Hence, the NMR spectrometer cannot distinguish between the...
IR Spectrum Peak Splitting: Symmetric vs Asymmetric Vibrations01:08

IR Spectrum Peak Splitting: Symmetric vs Asymmetric Vibrations
Identical bonds within a polyatomic group can stretch symmetrically (in-phase) or asymmetrically (out-of-phase). Similar to hydrogen bonding, these vibrations also influence the shape of the IR peak. Generally, asymmetric stretching frequencies are higher than symmetric stretching frequencies. For example, primary amines exhibit two distinct IR peaks between 3300–3500 cm−1 corresponding to the symmetric and asymmetric N-H stretching, while secondary amines exhibit a single stretching vibration...
NMR Spectroscopy of Aromatic Compounds01:14

NMR Spectroscopy of Aromatic Compounds

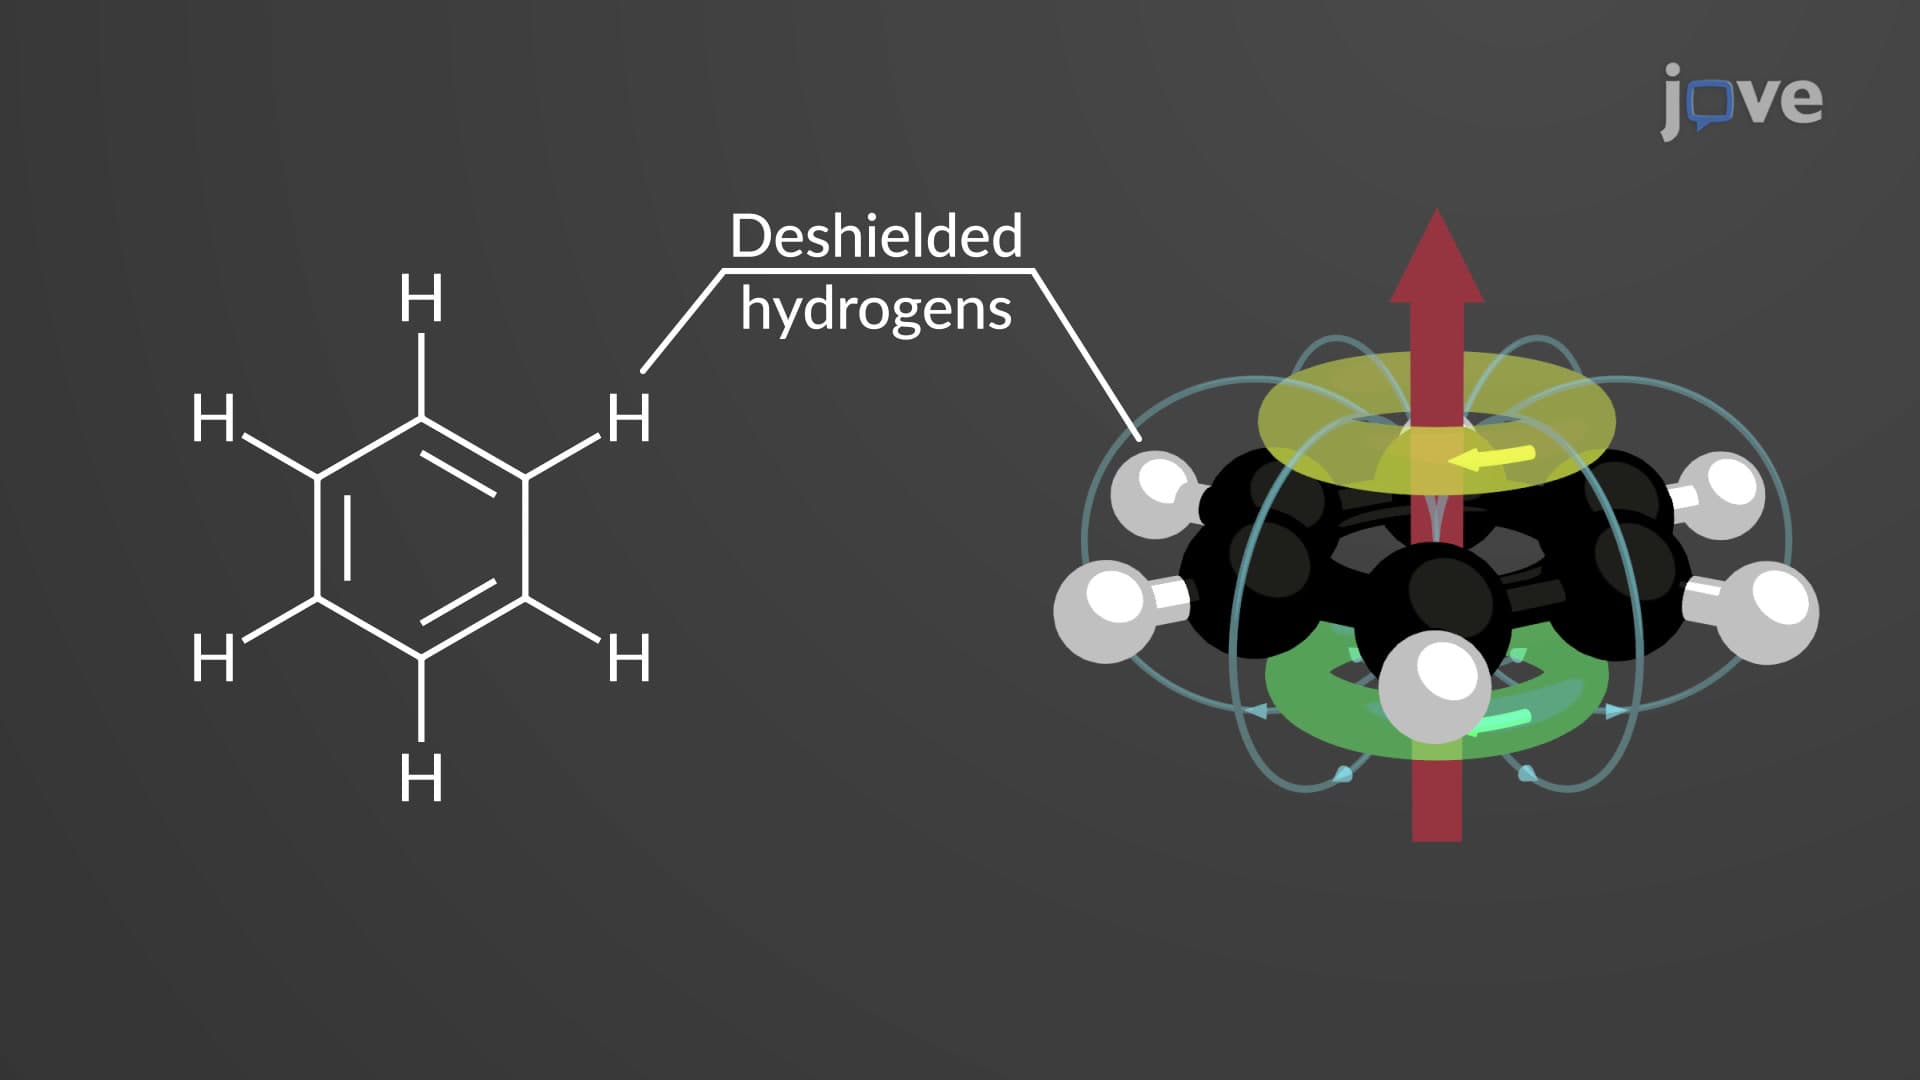
Aromatic compounds can be identified or analyzed using proton NMR and carbon‐13 NMR. Typically, aromatic hydrogens or hydrogens directly bonded to the aromatic rings are strongly deshielded by the aromatic ring current. Therefore, they absorb in the range of 6.5–8.0 ppm in proton NMR spectra. For instance, aromatic hydrogens directly bonded to the benzene ring absorb at 7.3 ppm. However, aromatic hydrogens of larger rings absorb farther upfield or downfield than the ideal range. Consider...
Spin–Spin Coupling: Two-Bond Coupling (Geminal Coupling)01:20

Spin–Spin Coupling: Two-Bond Coupling (Geminal Coupling)

Two NMR-active nuclei bonded to a central atom can be involved in geminal or two-bond coupling. Geminal coupling is commonly seen between diastereotopic protons in chiral molecules and unsymmetrical alkenes, among others.
The central atom need not be NMR-active because its electrons are affected by the electron polarization of the spin-active atoms. However, spin information is transmitted less effectively than in one-bond coupling, and 2J values are usually weaker than 1J values. The energy of...
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR01:15

¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
The axial and equatorial protons in cyclohexane can be distinguished by performing a variable-temperature NMR experiment. In this process, except for one proton, the remaining eleven protons are replaced by deuterium. The deuterium substitution avoids the possible peak splitting caused by the spin-spin coupling between the adjacent protons. The remaining proton flips between the axial and equatorial positions.
NMR Spectroscopy Of Amines01:19

NMR Spectroscopy Of Amines

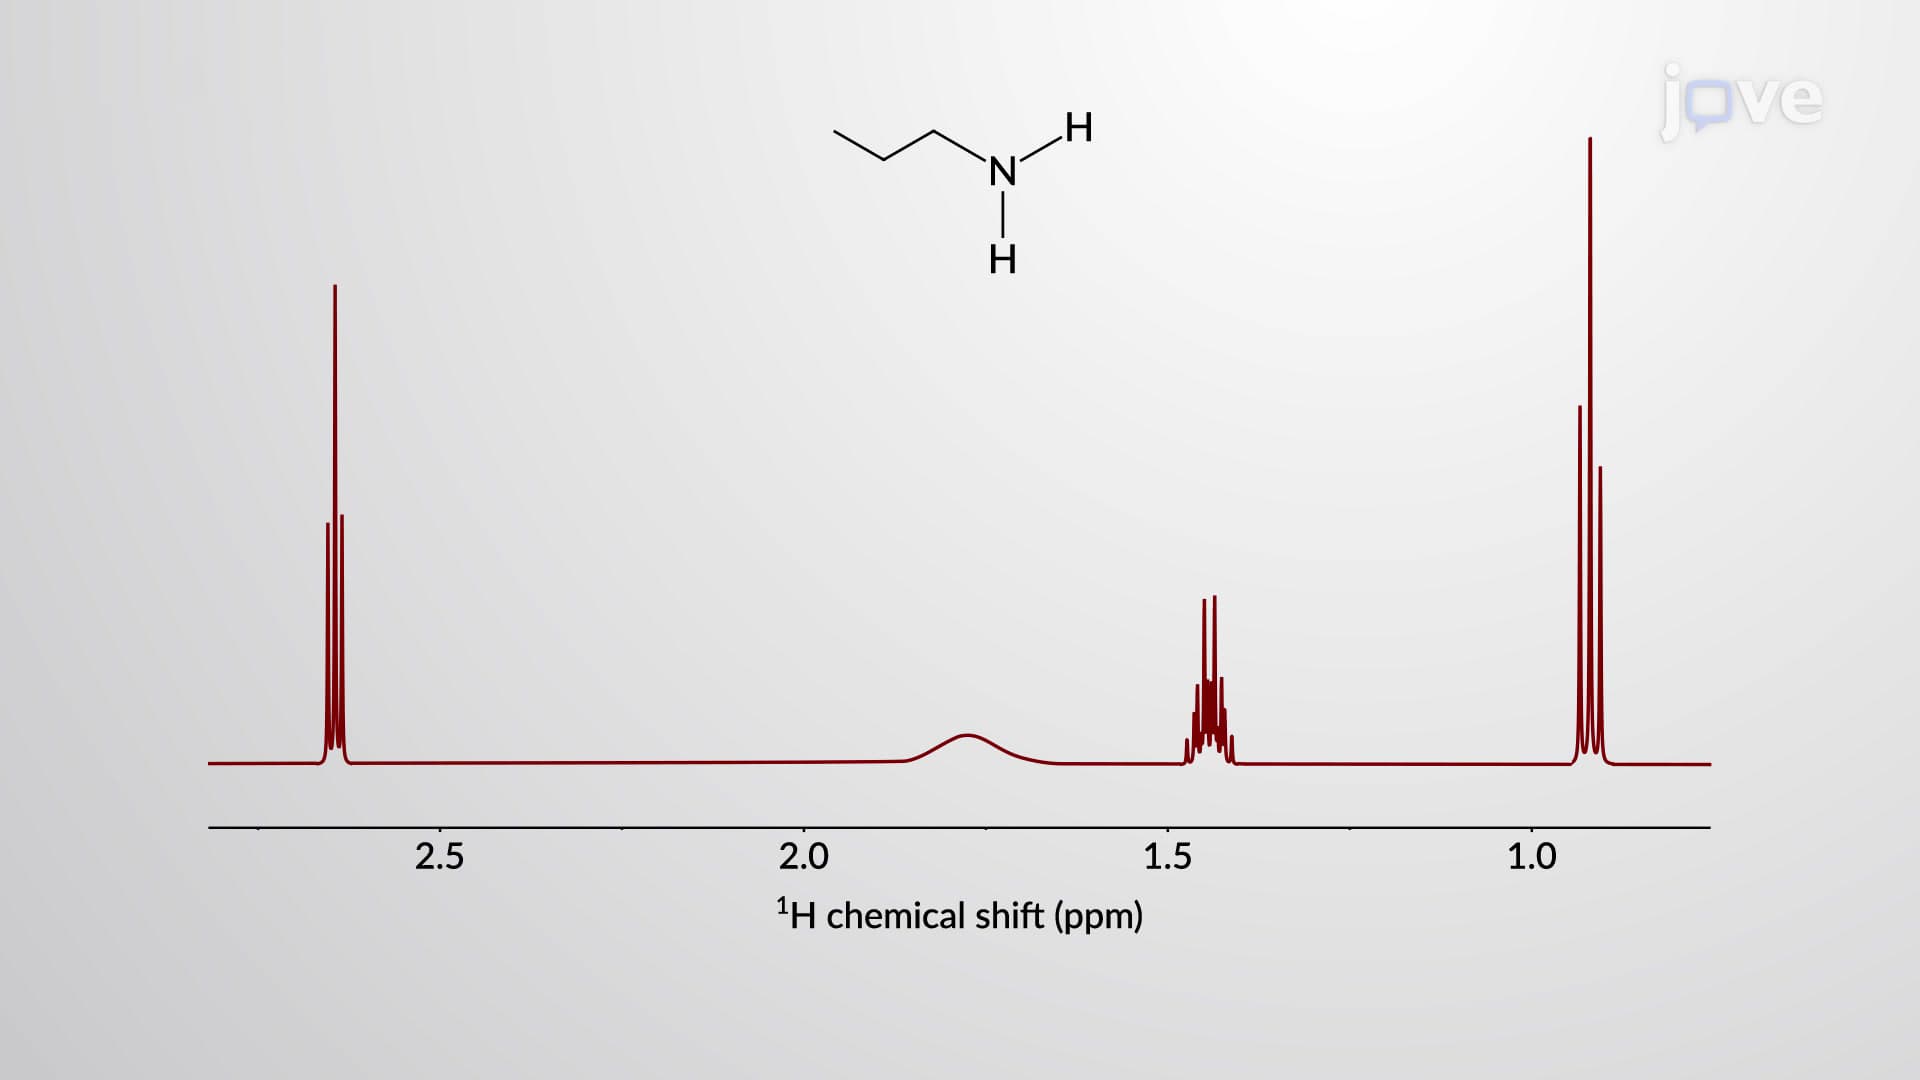
In proton NMR spectroscopy, primary amines and secondary amines showcase their N–H protons as a broad signal in the chemical shift range between δ 0.5 and 5 ppm. The exact position in this range depends on several factors, including sample concentration, hydrogen bonding, and the type of solvent used. Since amine protons undergo fast proton exchange in solution, the protons are labile and therefore do not participate in any splitting with adjacent protons. Thus, the observed peak is broad and...