Related Experiment Video
Updated: Aug 10, 2026
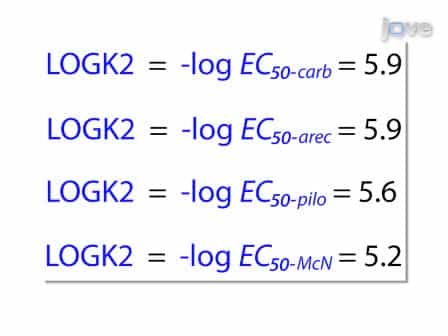

Quantifying Agonist Activity at G Protein-coupled Receptors
Published on: December 26, 2011
SCH-202676: An allosteric modulator of both agonist and antagonist binding to G protein-coupled receptors
A B Fawzi1, D Macdonald, L L Benbow
1Central Nervous System/Cardiovascular Research, Schering-Plough Research Institute, Kenilworth, New Jersey 07033-0539, USA. ahmad.fawzi@spcorp.com

Abstract:
A novel thiadiazole compound, SCH-202676 (N-(2,3-diphenyl-1,2, 4-thiadiazol-5-(2H)-ylidene)methanamine), has been identified as an inhibitor of both agonist and antagonist binding to G protein-coupled receptors (GPCRs). SCH-202676 inhibited radioligand binding to a number of structurally distinct, heterologously expressed GPCRs, including the human mu-, delta-, and kappa-opioid, alpha- and beta-adrenergic, muscarinic M1 and M2, and dopaminergic D1 and D2 receptors, but not to the tyrosine kinase epidermal growth factor receptor. SCH-202676 had no direct effect on G protein activity as assessed by [35S]guanosine-5'-O-(gamma-thio)triphosphate binding to purified recombinant G(oalpha)- or G(betagamma)-stimulated ADP-ribosylation of G(oalpha) by pertussis toxin. In addition, SCH-202676 inhibited antagonist binding to the beta2-adrenergic receptor expressed in Escherichia coli, a system devoid of classical heterotrimeric G proteins. SCH-202676 inhibited radiolabeled agonist and antagonist binding to the alpha2a-adrenergic receptor with an IC50 value of 0.5 microM, decreased the Bmax value of the binding sites with a slight increase in the KD value, and inhibited agonist-induced activation of the receptor. The effects of SCH-202676 were reversible. Incubation of plasma membranes with 10 microM SCH-202676 did not alter subsequent radioligand binding to the alpha2a-adrenergic receptor and the dopaminergic D1 receptor. Taken together, our data suggest that SCH-202676 has the unique ability to allosterically regulate agonist and antagonist binding to GPCRs in a manner that is both selective and reversible. The scope of the data presented suggests this occurs by direct interaction with a structural motif common to a large number of GPCRs or by activation/inhibition of an unidentified accessory protein that regulates GPCR function.
More Related Videos
 07:41
A Kinetic Fluorescence-based Ca2+ Mobilization Assay to Identify G Protein-coupled Receptor Agonists, Antagonists, and Allosteric Modulators
Published on: February 20, 2018
 07:16
Methods for the Discovery of Novel Compounds Modulating a Gamma-Aminobutyric Acid Receptor Type A Neurotransmission
Published on: August 16, 2018
Related Concept Videos
 01:09
GPCRs Regulate Adenylyl Cylase Activity
Two...
 01:30
Transducer Mechanism: G Protein–Coupled Receptors
GPCRs are also called heptahelical, 7TM, or...
 01:25
Cholinergic Receptors: Muscarinic
The subtypes M1, M3, and M5 couple with the Gq subunit and activate the phospholipase C (PLC) activity, mobilizing intracellular Ca2+. Activation...
 01:22
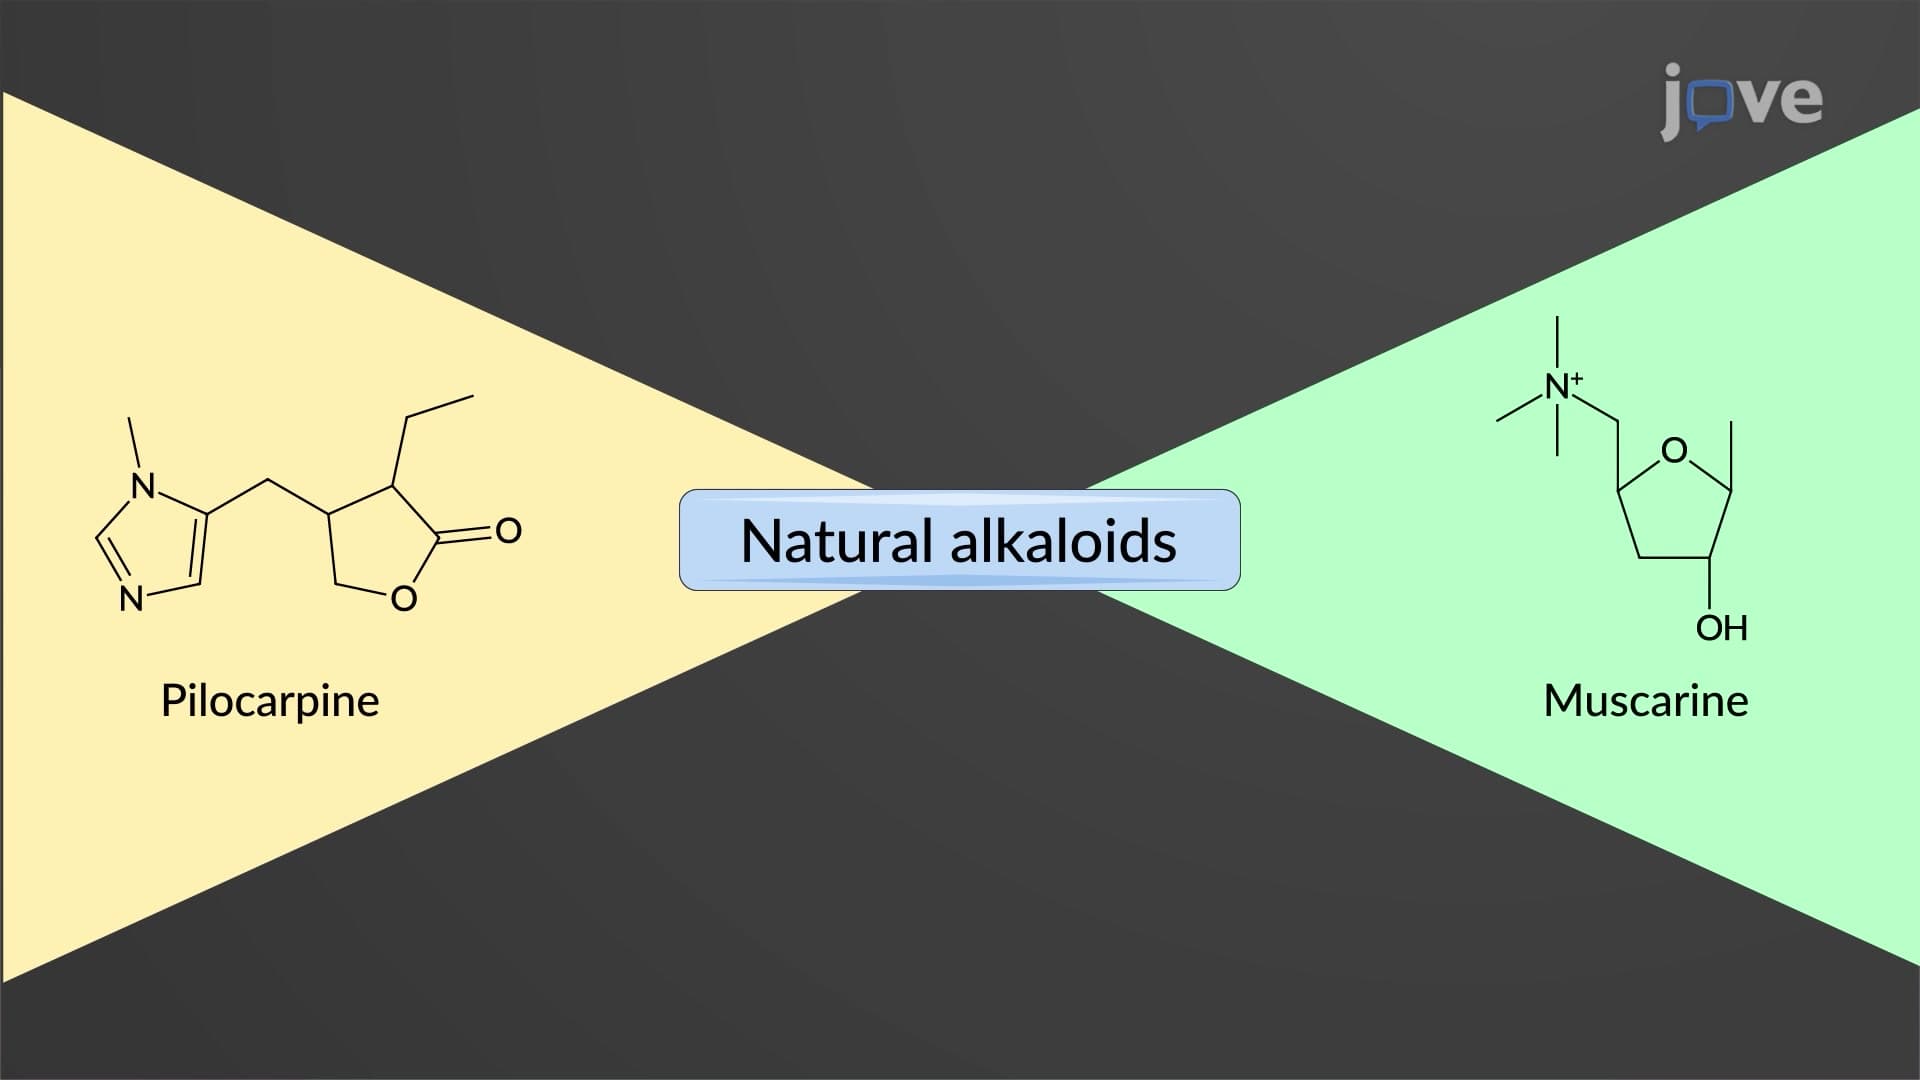
Direct-Acting Cholinergic Agonists: Chemistry and Structure-Activity Relationship
The direct-acting...
 01:29
Indirect-Acting Cholinergic Agonists: Chemistry and Structure-Activity Relationship
Reversible inhibitors display short to medium durations of action. Short-acting agents include simple alcohols with...
 01:29
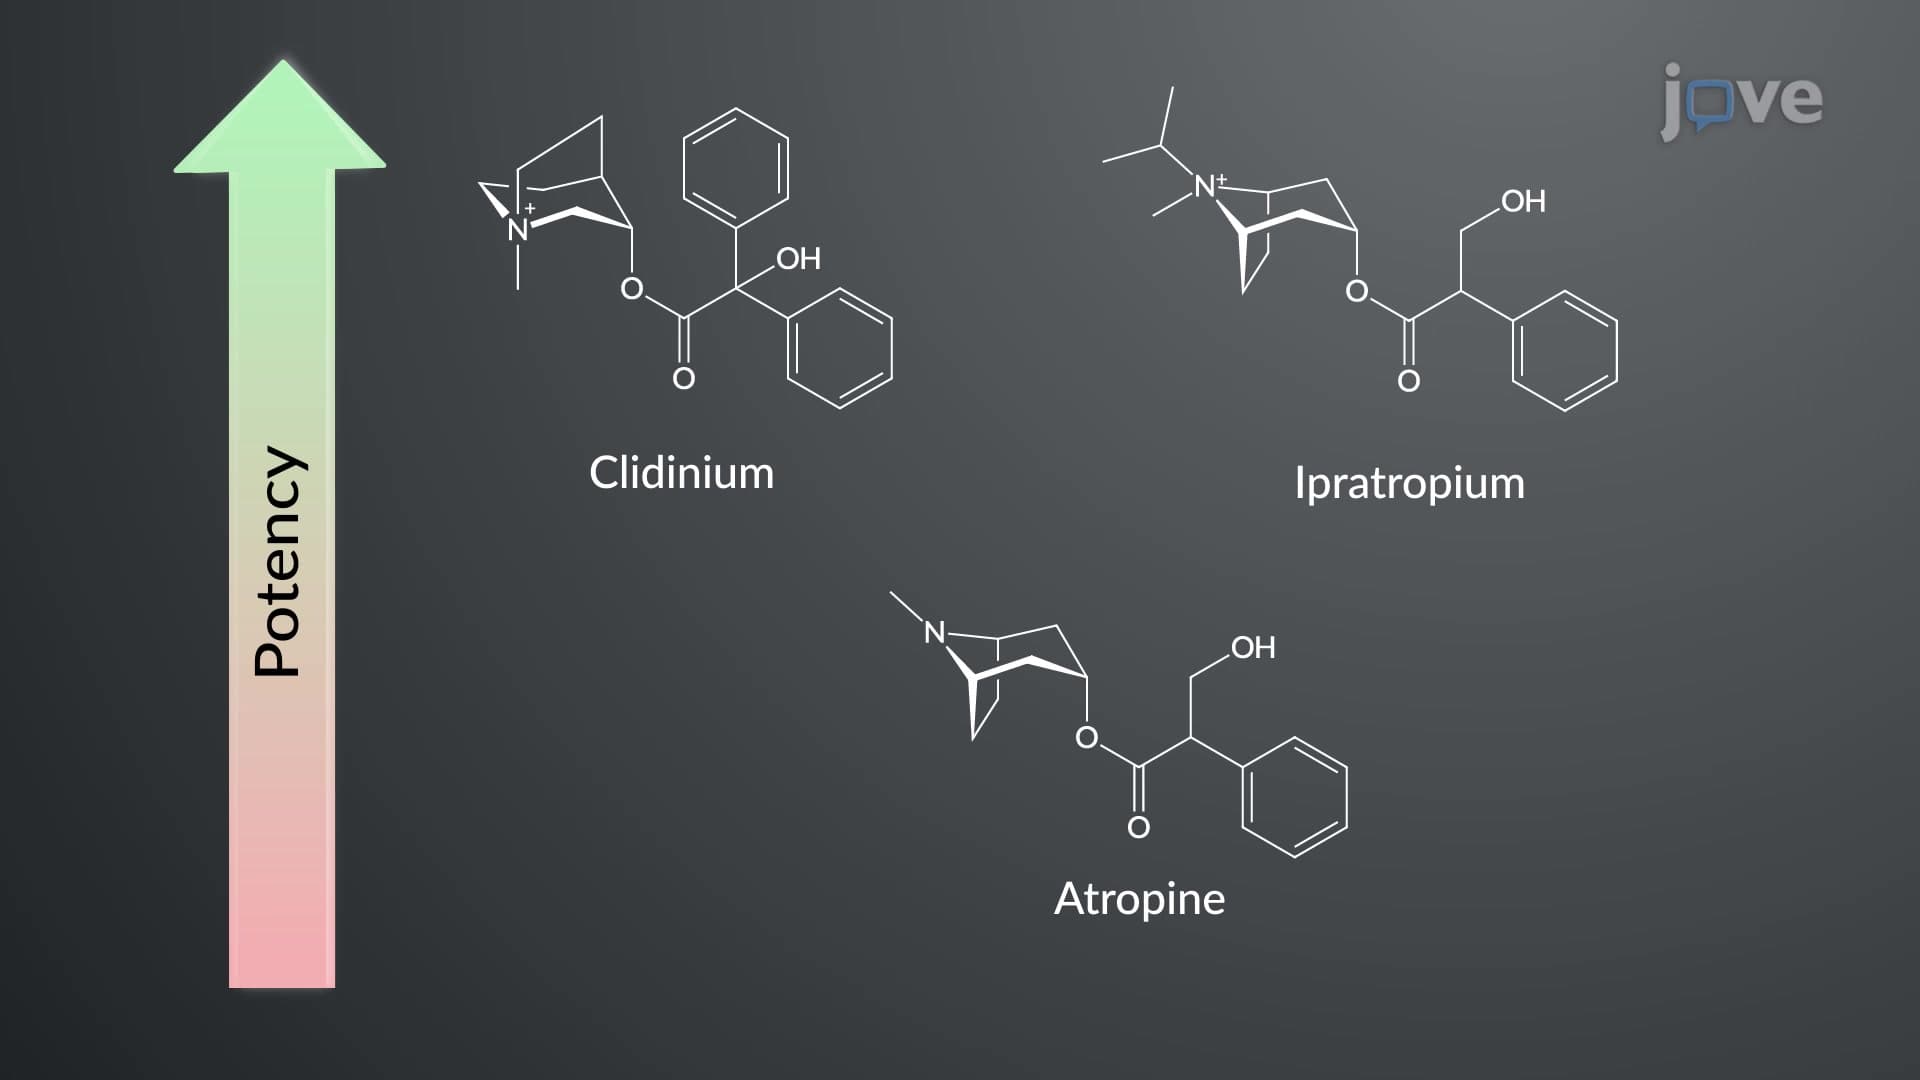
Cholinergic Antagonists: Chemistry and Structure-Activity Relationship