Related Experiment Video
Updated: Sep 20, 2026
Quantifying Agonist Activity at G Protein-coupled Receptors
Published on: December 26, 2011
A quantitative analysis of antagonism and inverse agonism at wild-type and constitutively active hamster
P Hein1, M Goepel, S Cotecchia
1Nephrol. Lab. IG 1, Klinikum Essen, Germany.

Abstract:
In order to characterize inverse agonism at alpha1B-adrenoceptors, we have compared the concentration-response relationships of several quinazoline and non-quinazoline alpha1-adrenoceptor antagonists at cloned hamster wild-type (WT) alpha1B-adrenoceptors and a constitutively active mutant (CAM) thereof upon stable expression in Rat-1 fibroblasts. Receptor activation or inhibition thereof was assessed as [3H]inositol phosphate (IP) accumulation. Quinazoline (alfuzosin, doxazosin, prazosin, terazosin) and non-quinazoline alpha1-adrenoceptor antagonists (BE 2254, SB 216,469, tamsulosin) concentration-dependently inhibited phenylephrine-stimulated IP formation at both WT and CAM with Ki values similar to those previously found in radioligand binding studies. At CAM in the absence of phenylephrine, the quinazolines produced concentration-dependent inhibition of basal IP formation; the maximum inhibition was approximately 55%, and the corresponding EC50 values were slightly smaller than the Ki values. In contrast, BE 2254 produced much less inhibition of basal IP formation, SB 216,469 was close to being a neutral antagonist, and tamsulosin even weakly stimulated IP formation. The inhibitory effects of the quinazolines and BE 2254 as well as the stimulatory effect of tamsulosin were equally blocked by SB 216,469 at CAM. At WT in the absence of phenylephrine, tamsulosin did not cause significant stimulation and none of the other compounds caused significant inhibition of basal IP formation. We conclude that alpha1-adrenoceptor antagonsits with a quinazoline structure exhibit greater efficacy as inverse agonists than those without.
More Related Videos
Related Concept Videos
 01:28
Drug-Receptor Interaction: Antagonist
Antagonists can be classified as competitive or noncompetitive based on their...
 01:30
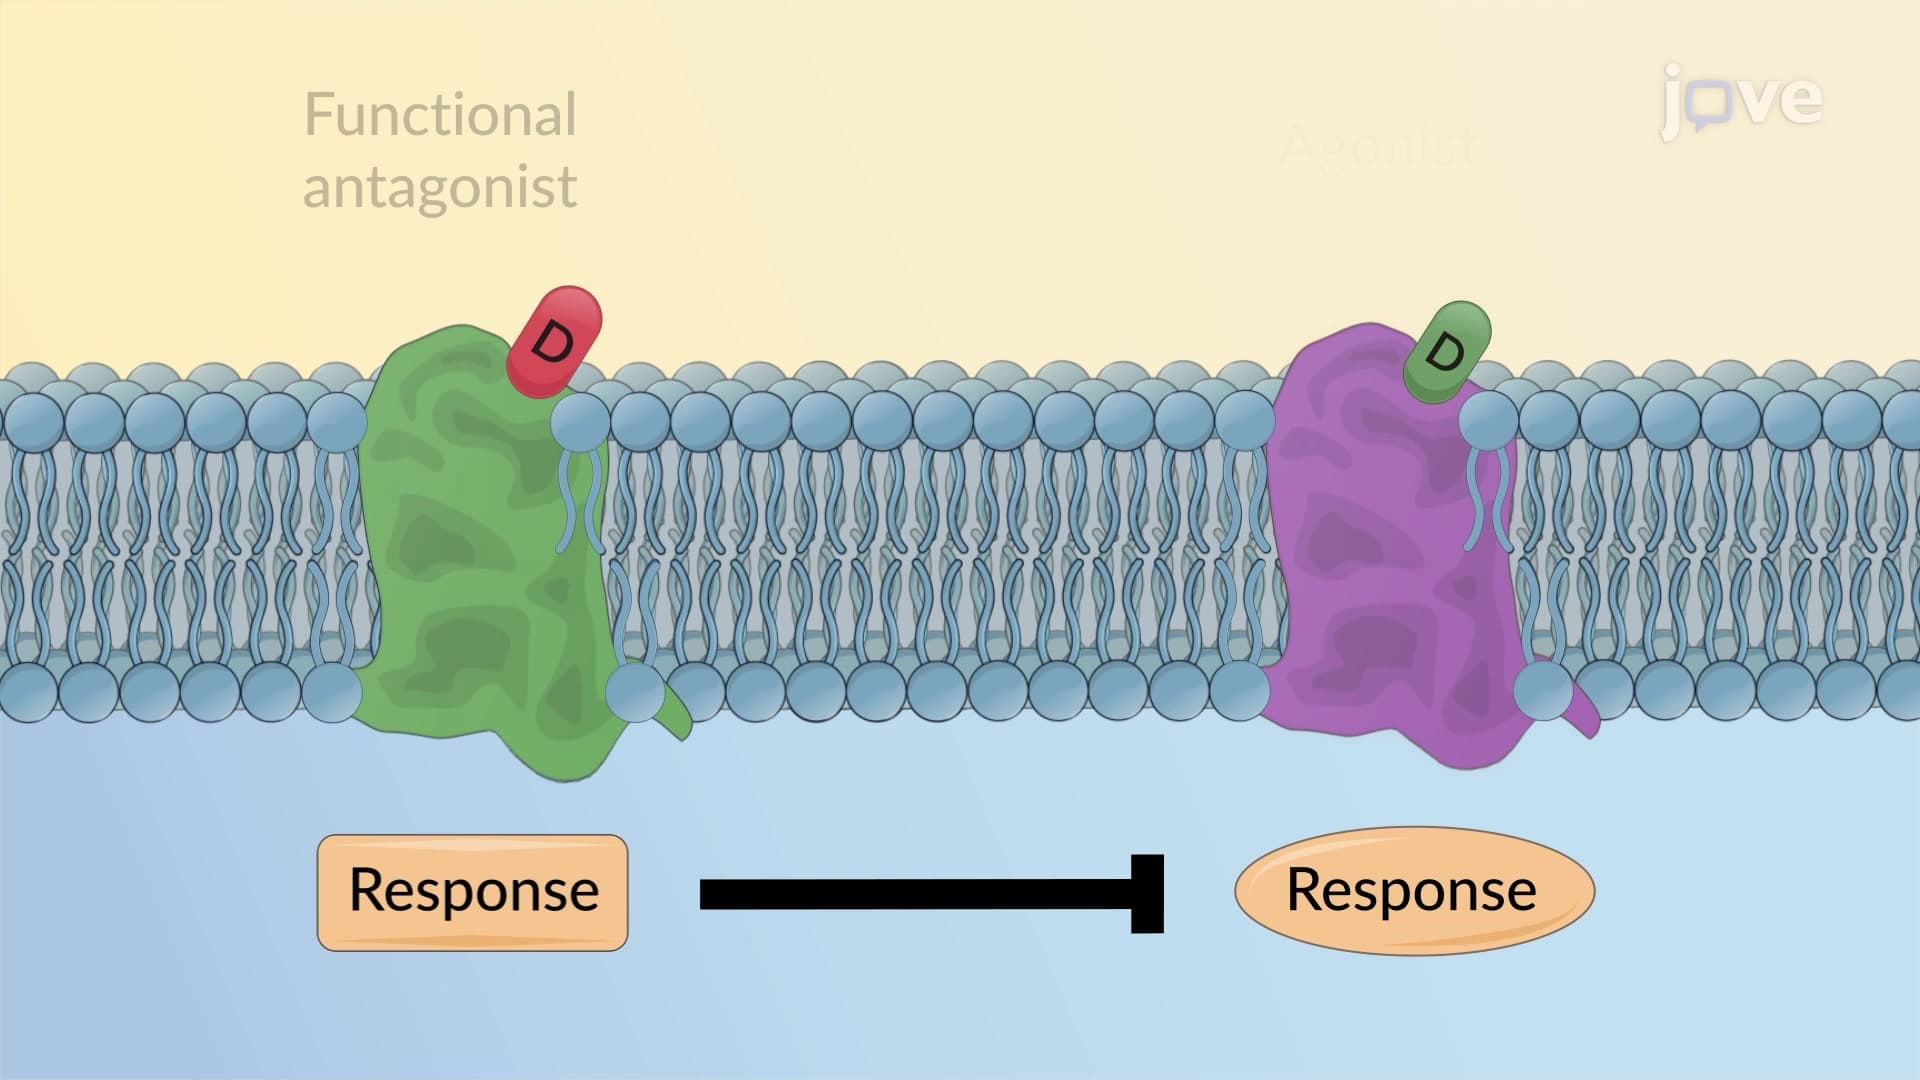
Combined Effects of Drugs: Antagonism
The most common type is receptor antagonism, where one drug acts as an antagonist to block the effects of another drug by...
 01:31
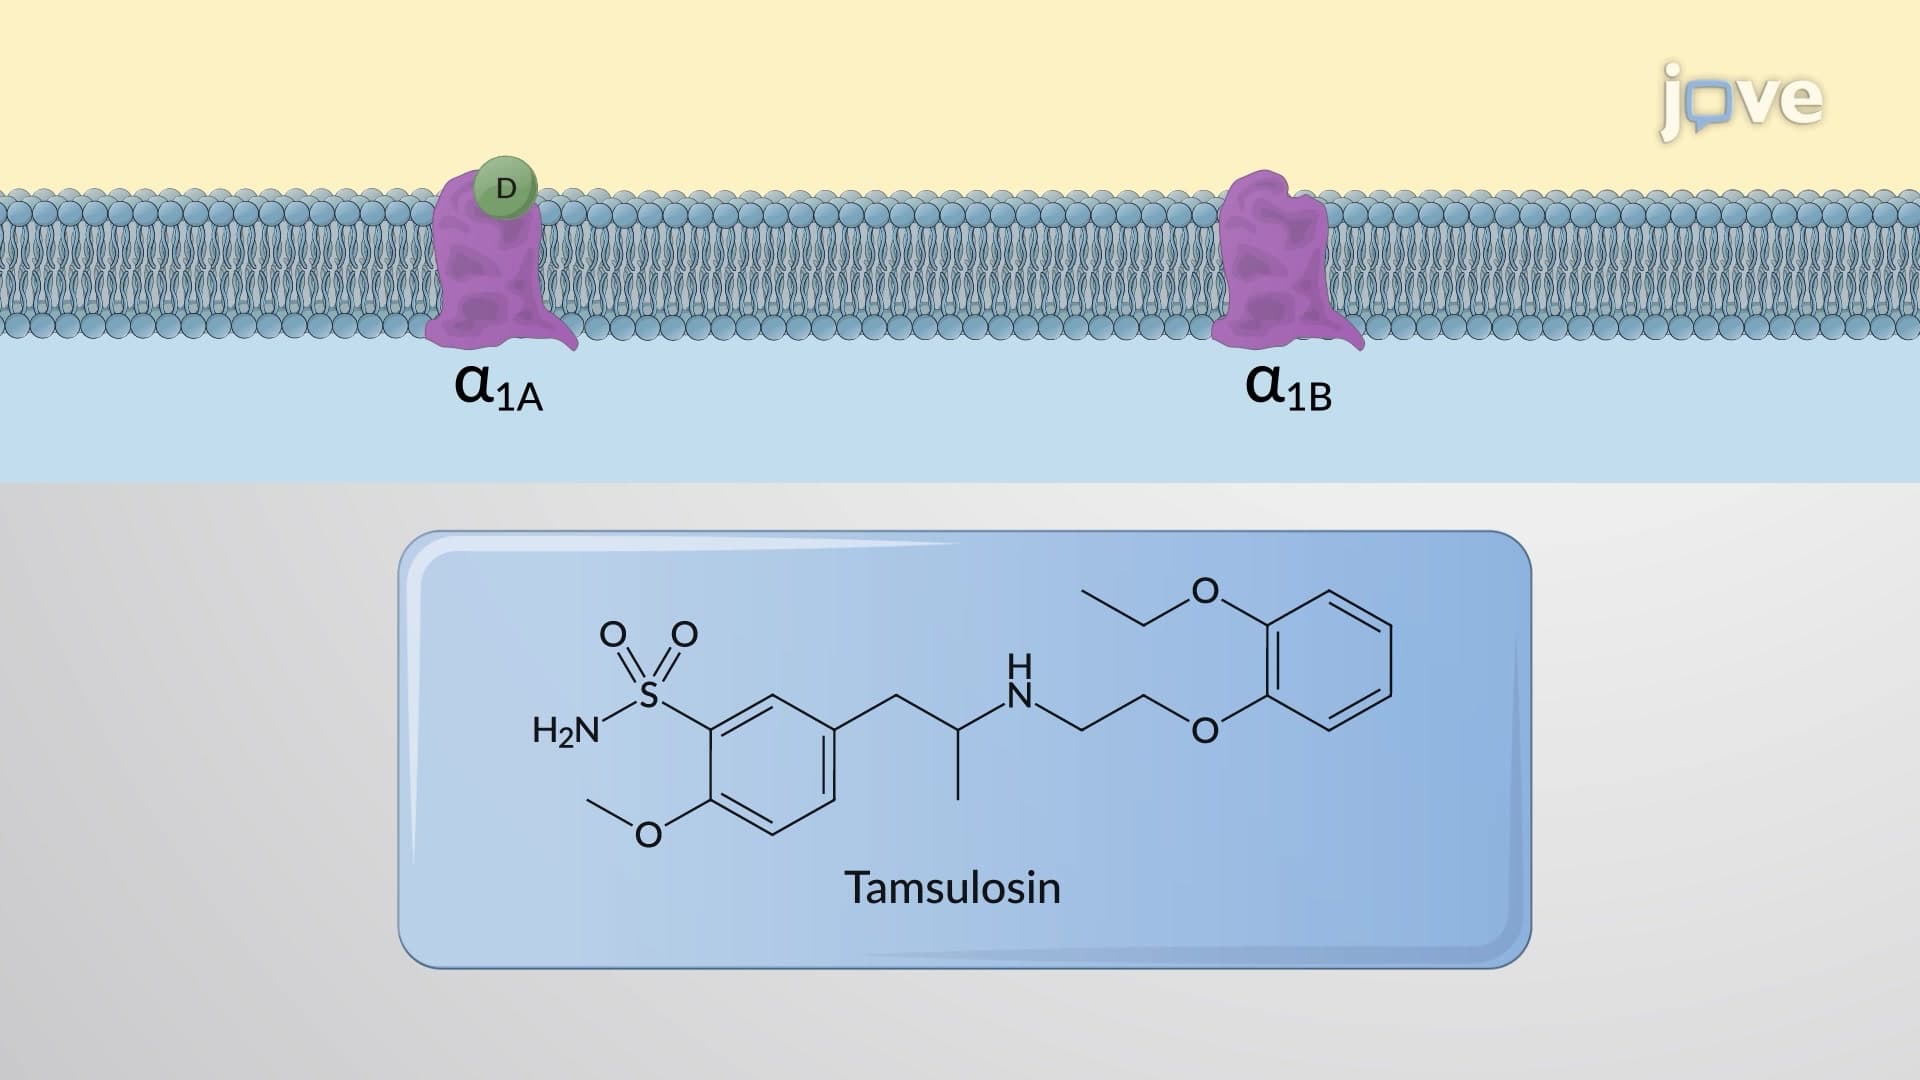
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase C—inositol-1,4,5-trisphosphate...
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:17
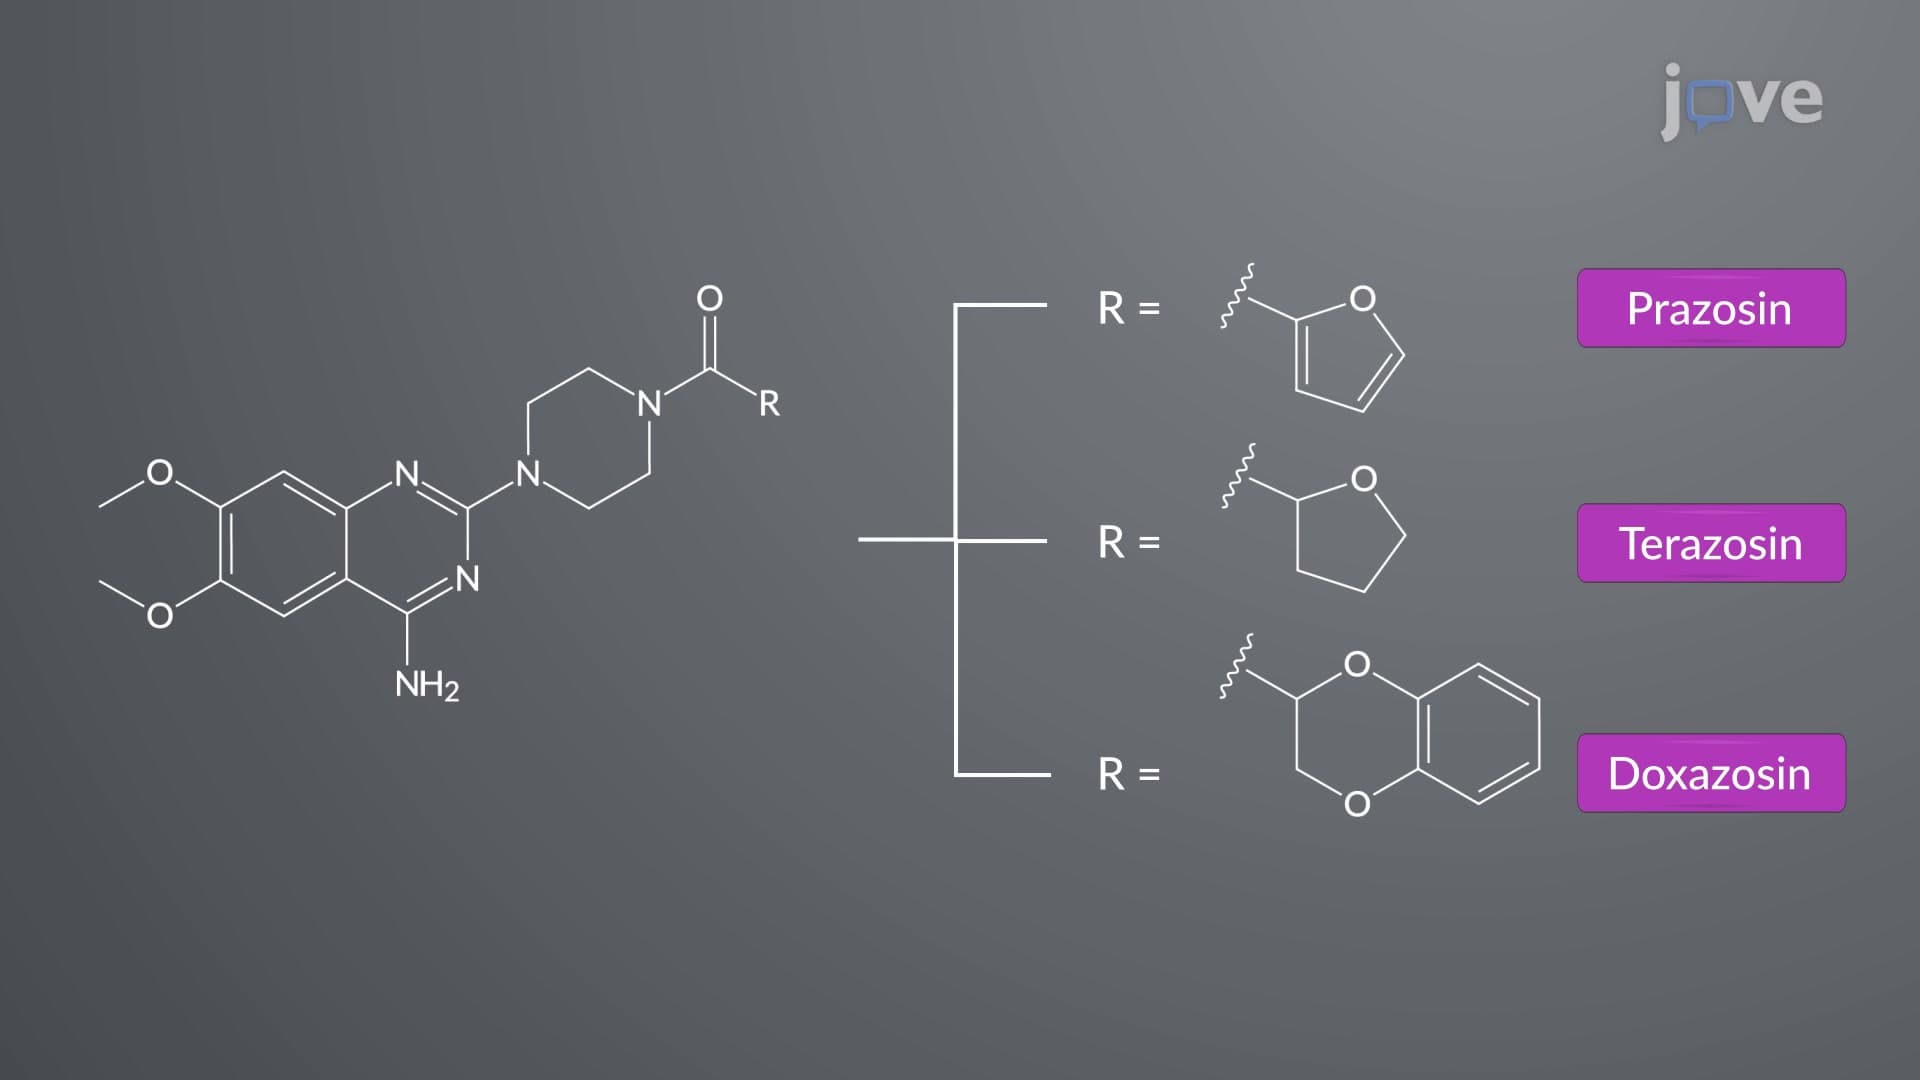
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...
 01:22
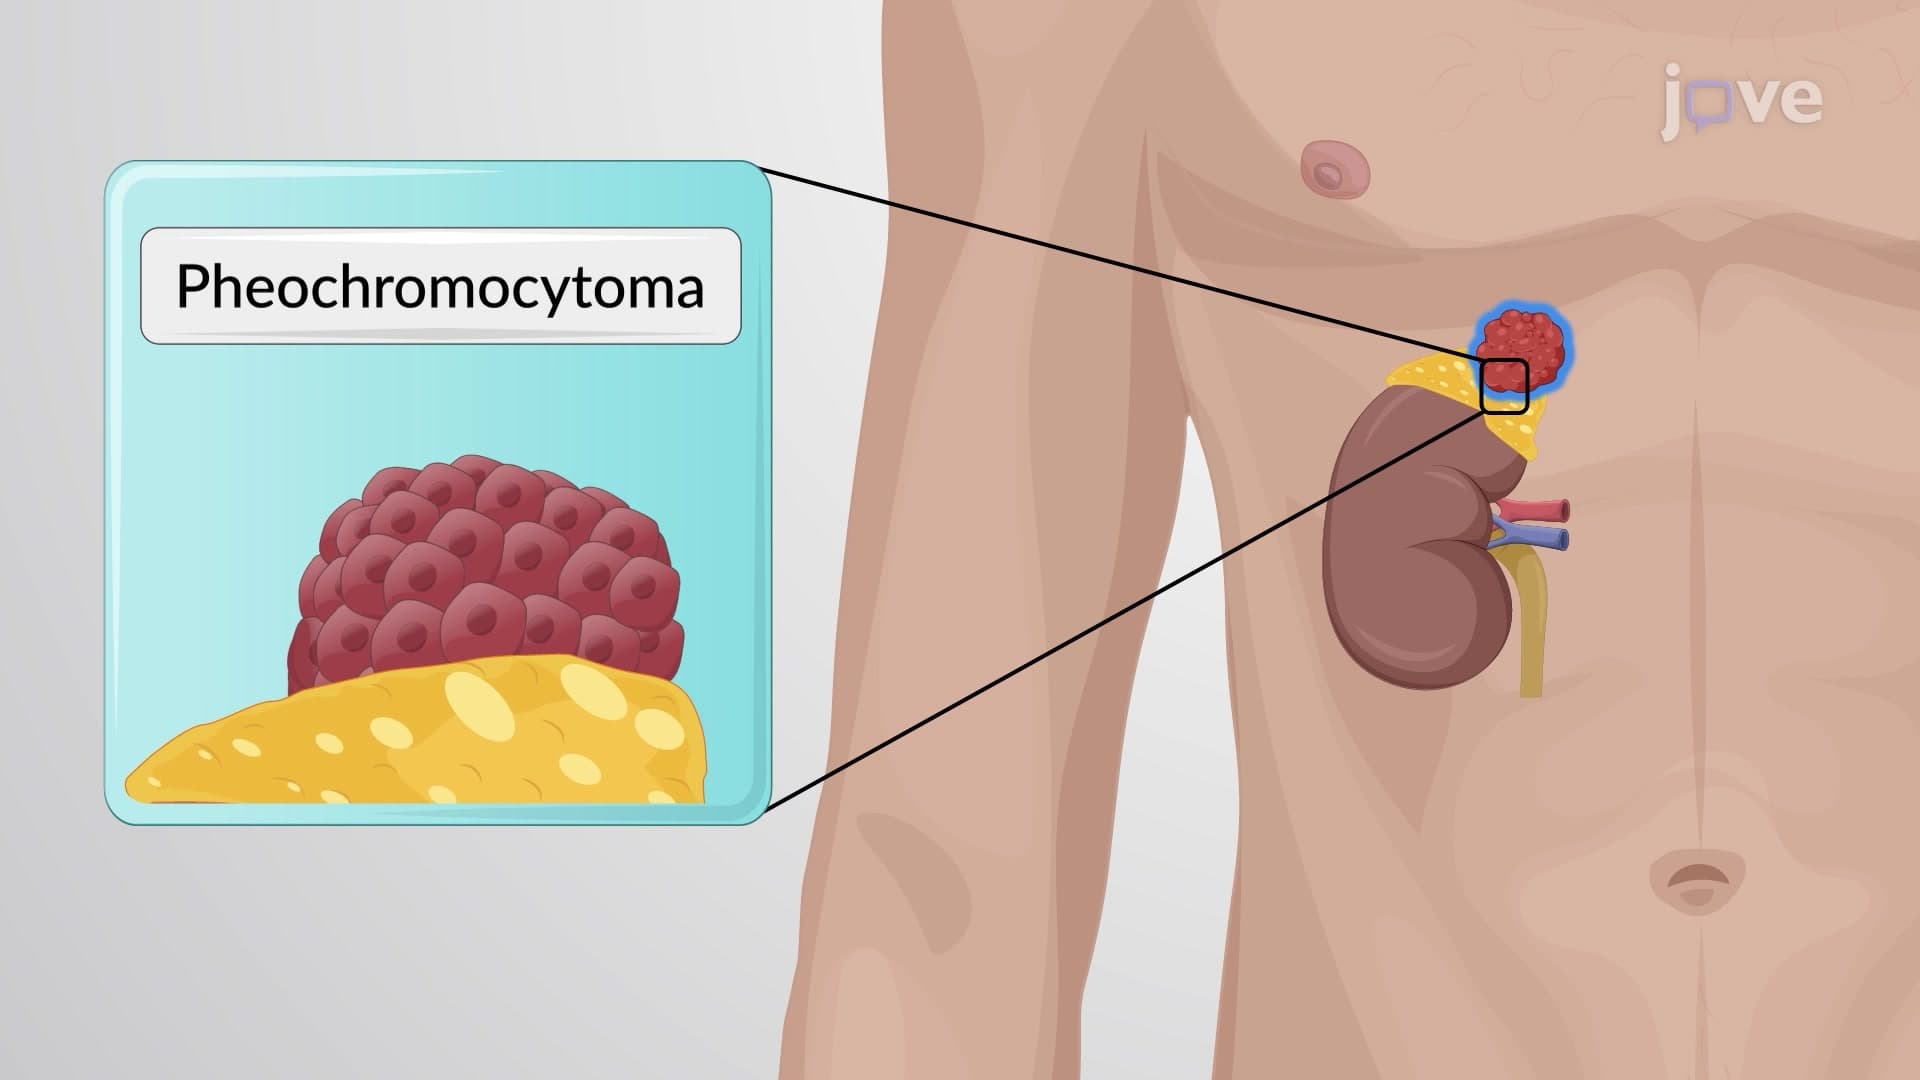
Adrenergic Antagonists: Pharmacological Actions of ɑ-Receptor Blockers
α1-blockers: These drugs inhibit α1-adrenoceptors on smooth muscle cells, resulting in vasodilation. This vasodilation lowers blood pressure, making α1-blockers valuable in treating hypertension. Additionally, α1-blockers effectively address urinary obstruction...