A computational study of the effectiveness of the frontier molecular orbital formalism in predicting conformational isomerism in (p-RC6H4NC)2W(dppe)2
Related Experiment Video
Updated: Oct 9, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
N L Wagner1, J M Kloss, K L Murphy
1Department of Chemistry, University of Wisconsin--Milwaukee, 53211, USA.

Abstract:
Ab initio electronic structure calculations on a series of ligands, p-RC6H4NC:, indicate, that the energy of the LUMO correlates with the electron-withdrawing/donating capabilities of the substituent group, which determines the relative pi-acidity of the ligand. Depending on the nature of the para substituent group on the aryl isocyanide ligand, bis(aryl isocyanide) complexes of tungsten-containing bulky bidentate arylphosphine ligands adopt either cis or trans conformations. The frontier molecular orbital formalism predicts that strong pi-acids, which contain electron-withdrawing groups, tend to polarize sufficient charge density away from the metal center to effect the formation of the sterically less favorable but electronically stabilized cis conformer. Density functional theory calculations on similar complexes containing phosphines which do not impose severe steric contraints indicate that the balance between steric and electronic stabilization can be effectively predicted by comparing the relative energies of the ligand LUMOs.
Related Concept Videos
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 02:33
Stereoisomerism of Cyclic Compounds
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
 02:18
Conformations of Ethane and Propane
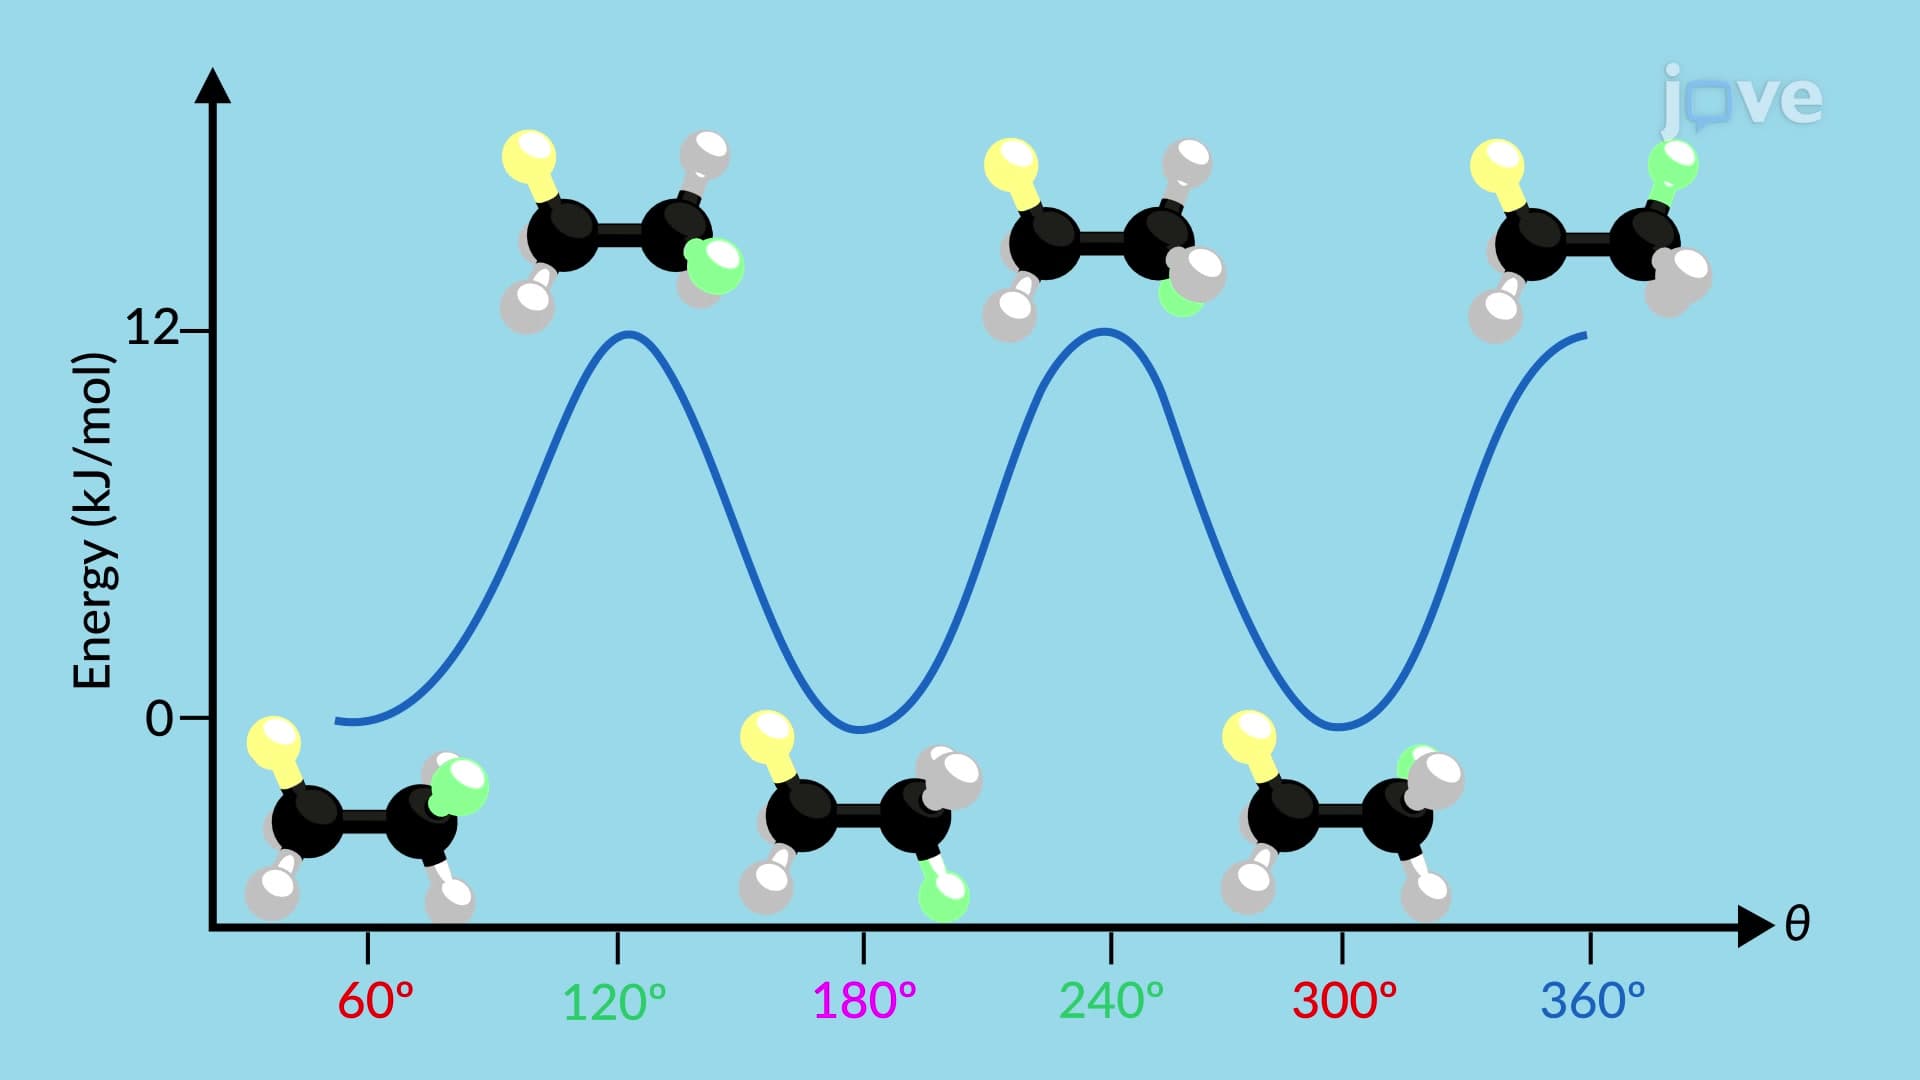
Staggered conformation is a low energy and more stable conformation with the C-H bonds on the front carbon placed at 60°dihedral angles relative to the C-H bonds on the back carbon, leading to a reduced torsional strain. In staggered ethane, the...
 02:02
Chair Conformation of Cyclohexane
The hydrogen atoms linked to carbons are arranged in two different axial and equatorial orientations to achieve this staggered...
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal tetrahedral value,...