Related Experiment Video
Updated: Oct 9, 2026
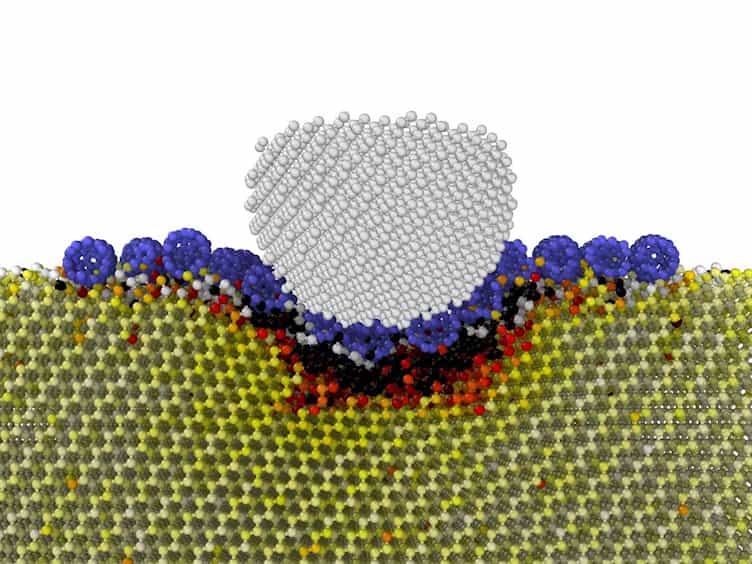

Probing C84-embedded Si Substrate Using Scanning Probe Microscopy and Molecular Dynamics
Published on: September 28, 2016
Evaluation of Spartan semi-empirical molecular modeling software for calculations of molecules on surfaces: CO
1Department of Chemistry, University of Northern Iowa, Cedar Falls 50614-0423, USA.

Abstract:
This paper reports the use of the PM3(tm) semi-empirical method in the Spartan molecular modeling software to optimize geometries and calculate vibrational frequencies for increasingly complex transition metal- and carbon monoxide (CO)-containing systems, culminating in calculations of CO adsorbed on a Ni(111) surface. Mononuclear and dinuclear transition metal carbonyl molecular species were used to establish the level of accuracy that could be expected for vibrational frequencies to provide a context for the results from the adsorbed molecule calculations. One to four CO molecules adsorbed on the (111) face of a 22-atom-nickel crystal were then modeled, and the accuracy of the adsorption geometry and vibrational frequency was evaluated. The calculated CO stretching vibrational frequencies were within 8% larger than the gas phase experimental values for the molecular species and were approximately 10% larger than the range of experimental values for CO on the nickel surface. The geometry optimization predicted that the CO molecules on the Ni(111) surface occupy three-fold hollow sites with no preference for sites over Ni atoms, in agreement with recent structural data and other theoretical calculations. The software was less successful in calculating the CO bond angle to the surface and the distance of the CO molecules from the surface, but the calculation did produce a reasonable distance between CO molecules on the surface. In general, the PM3(tm) method in Spartan shows promise for predicting adsorption sites and vibrational frequencies of molecules on metal surfaces.
More Related Videos
 07:53
Analysis of Complex Molecules and Their Reactions on Surfaces by Means of Cluster-Induced Desorption/Ionization Mass Spectrometry
Published on: March 1, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
