Related Experiment Video
Updated: Oct 9, 2026
Measurement of Ultrafast Vibrational Coherences in Polyatomic Radical Cations with Strong-Field Adiabatic Ionization
Published on: August 6, 2018
Quantum dynamics of the dissociation of H2 on Cu(100): dependence of the site-reactivity on initial rovibrational
D A McCormack1, G J Kroes, R A Olsen
1Leiden Institute of Chemistry, Gorlaeus Laboratories, Leiden University, P.O. Box 9502, 2300 RA Leiden, The Netherlands.

Abstract:
We perform six-dimensional (6D) quantum wavepacket calculations for H2 dissociatively adsorbing on Cu(100) from a variety of rovibrational initial states. The calculations are performed on a new potential energy surface (PES), the construction of which is also detailed. Reaction probabilities are in good agreement with experimental findings. Using a new flux analysis method, we calculate the reaction probability density as a function of surface site and collision energy, for a variety of initial states. This approach is used to study the effects of rotation and vibration on reaction at specific surface sites. The results are explained in terms of characteristics of the PES and intrinsically dynamic effects. An important observation is that, even at low collision energies, reaction does not necessarily proceed predominantly in the region of the minimum potential barrier, but can occur almost exclusively at a site with a higher barrier. This suggests that experimental control of initial conditions could be used to selectively induce reaction at particular surface sites. Our predictions for site-reactivity could be tested using contemporary experimental methods: The calculations predict that, for reacting molecules, there will be a dependence of the quadrupole alignment of j on the incident vibrational state, v. This is a direct result of PES topography in the vicinity of the preferred reaction sites of v = 0 and v = 1 molecules. Invoking detailed balance, evidence for this difference in preferred reaction site of v = 0 and 1 molecules could be obtained through associative desorption experiments.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 06:53
Magnetometric Characterization of Intermediates in the Solid-State Electrochemistry of Redox-Active Metal-Organic Frameworks
Published on: June 9, 2023
Related Concept Videos
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to the...
 02:13
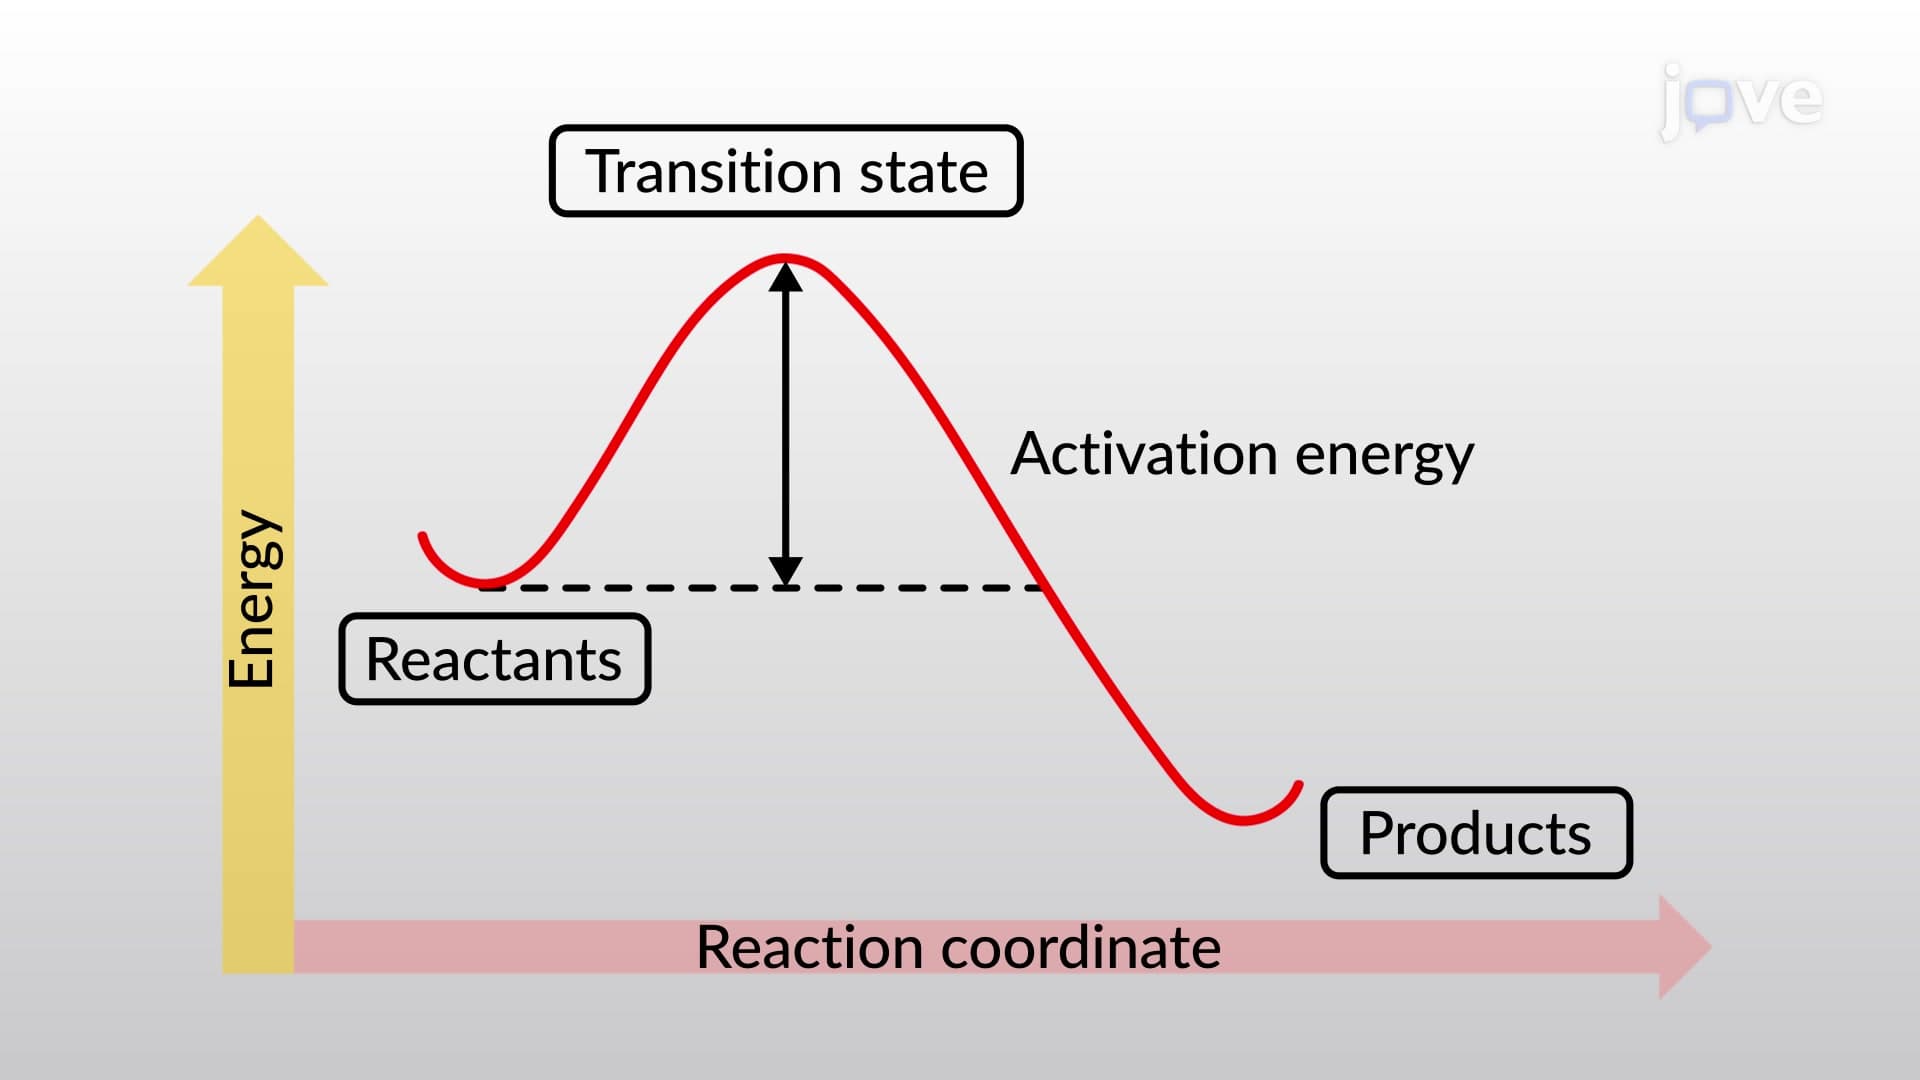
Bond Dissociation Energy and Activation Energy
 03:51
Molecular Orbital Theory II
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:19
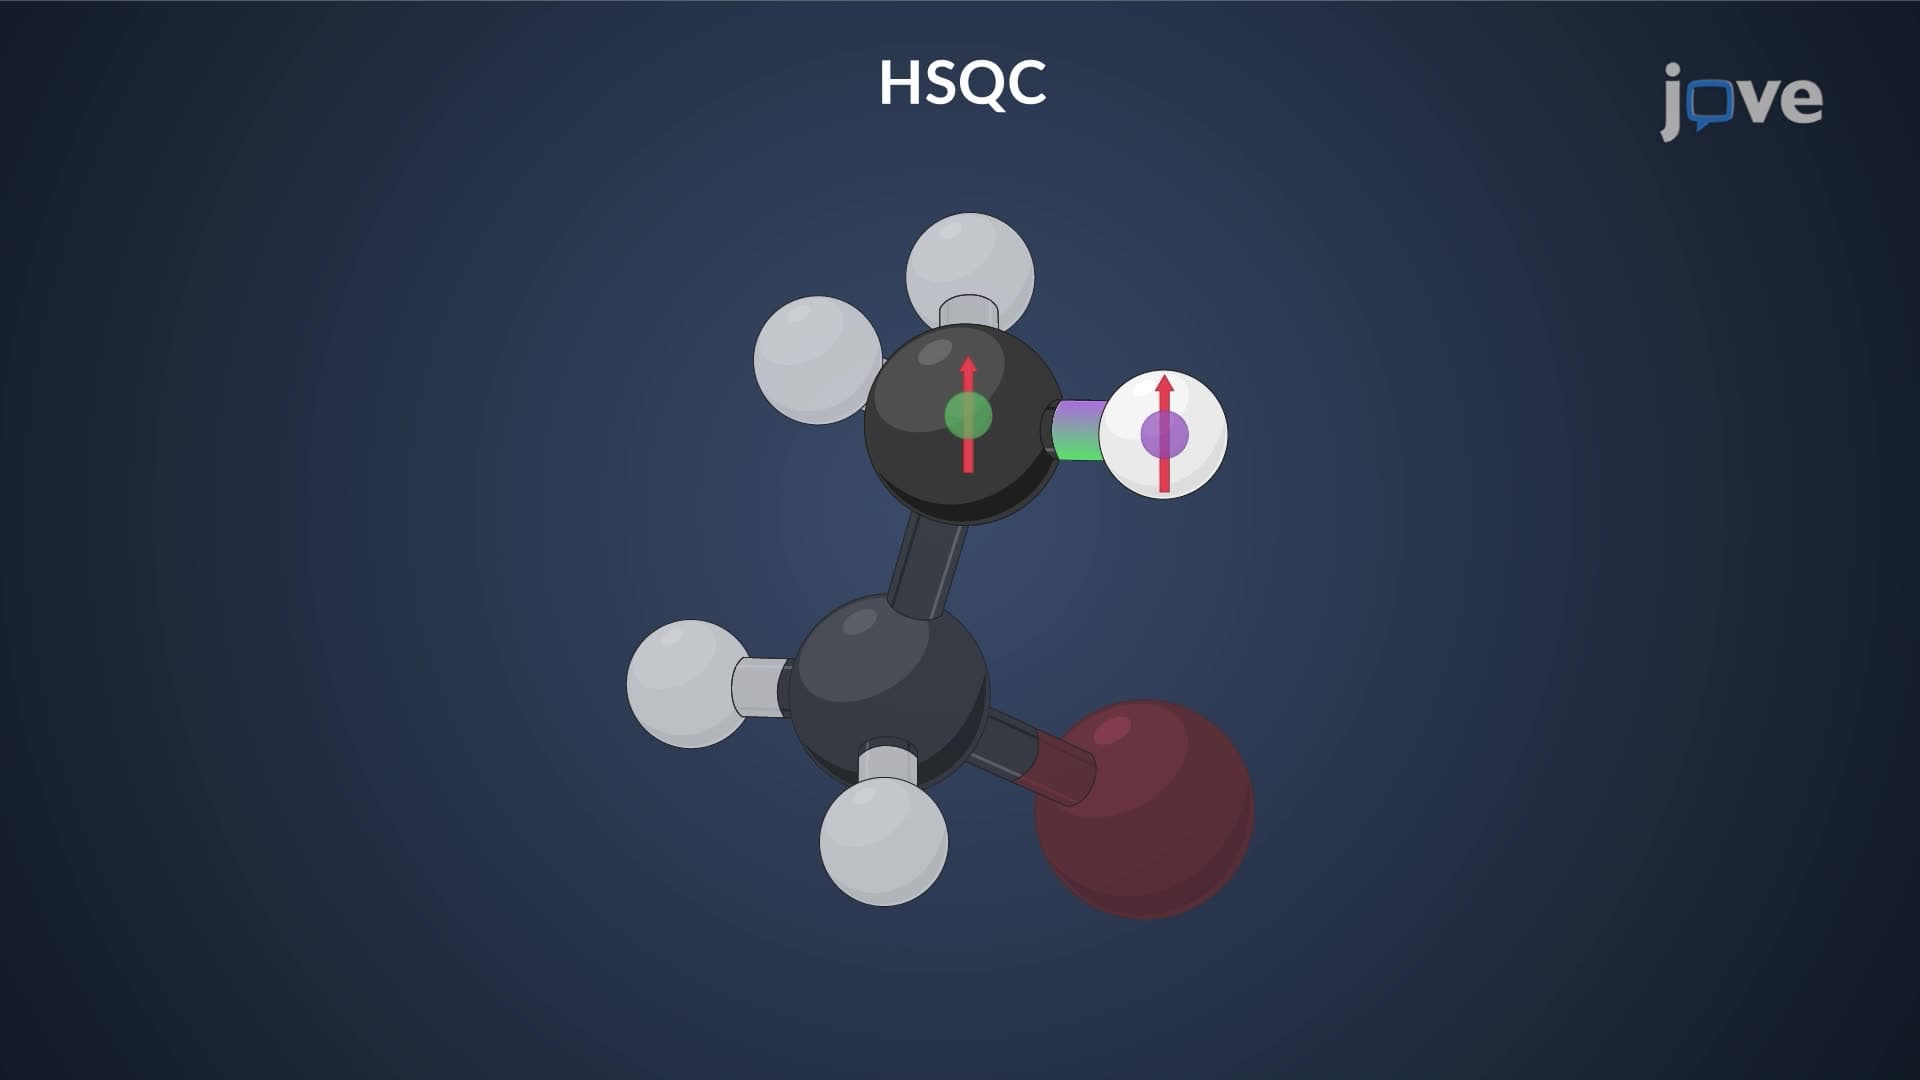
2D NMR: Heteronuclear Single-Quantum Correlation Spectroscopy (HSQC)
 02:49
Bond Energies and Bond Lengths