Related Experiment Video
Updated: Oct 9, 2026
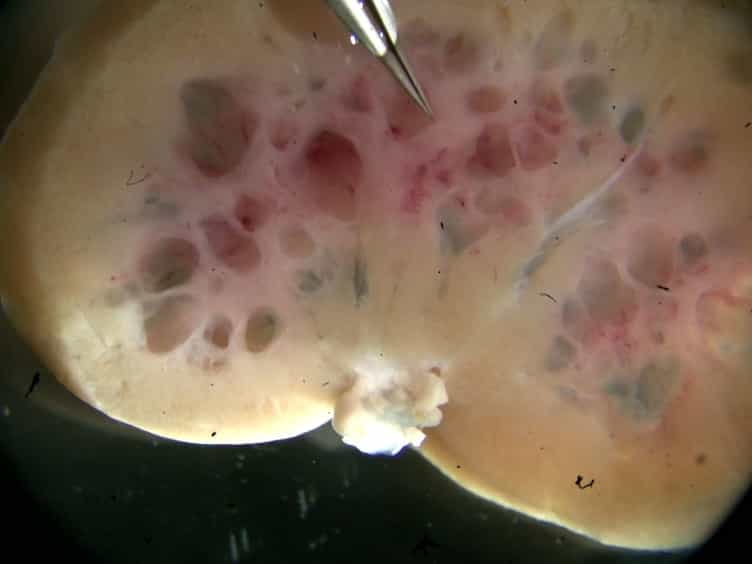

Implementing Patch Clamp and Live Fluorescence Microscopy to Monitor Functional Properties of Freshly Isolated PKD Epithelium
Published on: September 1, 2015
Role of CFTR in autosomal recessive polycystic kidney disease
Koichi Nakanishi1, William E Sweeney1, Katherine Macrae Dell1
1Rainbow Center for Childhood PKD, Department of Pediatrics, Rainbow Babies and Children's Hospital and Case Western Reserve University, Cleveland, Ohio.

Abstract:
An extensive body of in vitro data implicates epithelial chloride secretion, mediated through cystic fibrosis transmembrane conductance regulator (CFTR) protein, in generating or maintaining fluid filled cysts in MDCK cells and in human autosomal dominant polycystic kidney disease (ADPKD). In contrast, few studies have addressed the pathophysiology of fluid secretion in cyst formation and enlargement in autosomal recessive polycystic kidney disease (ARPKD). Murine models of targeted disruptions or deletions of specific genes have created opportunities to examine the role of individual gene products in normal development and/or disease pathophysiology. The creation of a murine model of CF, which lacks functional CFTR protein, provides the opportunity to determine whether CFTR activity is required for renal cyst formation in vivo. Therefore, this study sought to determine whether renal cyst formation could be prevented by genetic complementation of the BPK murine model of ARPKD with the CFTR knockout mouse. The results of this study reveal that in animals that are homozygous for the cystic gene (bpk), the lack of functional CFTR protein on the apical surface of cystic epithelium does not provide protection against cyst growth and subsequent decline in renal function. Double mutant mice (bpk -/-; cftr -/-) developed massively enlarged kidneys and died, on average, 7 d earlier than cystic, non-CF mice (bpk -/-; cftr +/+/-). This suggests fundamental differences in the mechanisms of transtubular fluid secretion in animal models of ARPKD compared with ADPKD.
More Related Videos
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:10
Nephrons
 01:25
Chronic Kidney Disease I: Introduction
 02:21
Pedigree Analysis
 01:24
Renal Tubule and Collecting Duct
Proximal Convoluted Tubule (PCT):
The PCT is the initial segment of the renal tubule, extending from the Bowman's capsule that encloses the glomerulus. Its convoluted structure and microvilli-lined cells increase the surface area for reabsorption. The PCT reabsorbs glucose, amino acids, sodium, and water from the filtrate, ensuring essential...