Related Experiment Video
Updated: Jul 22, 2026
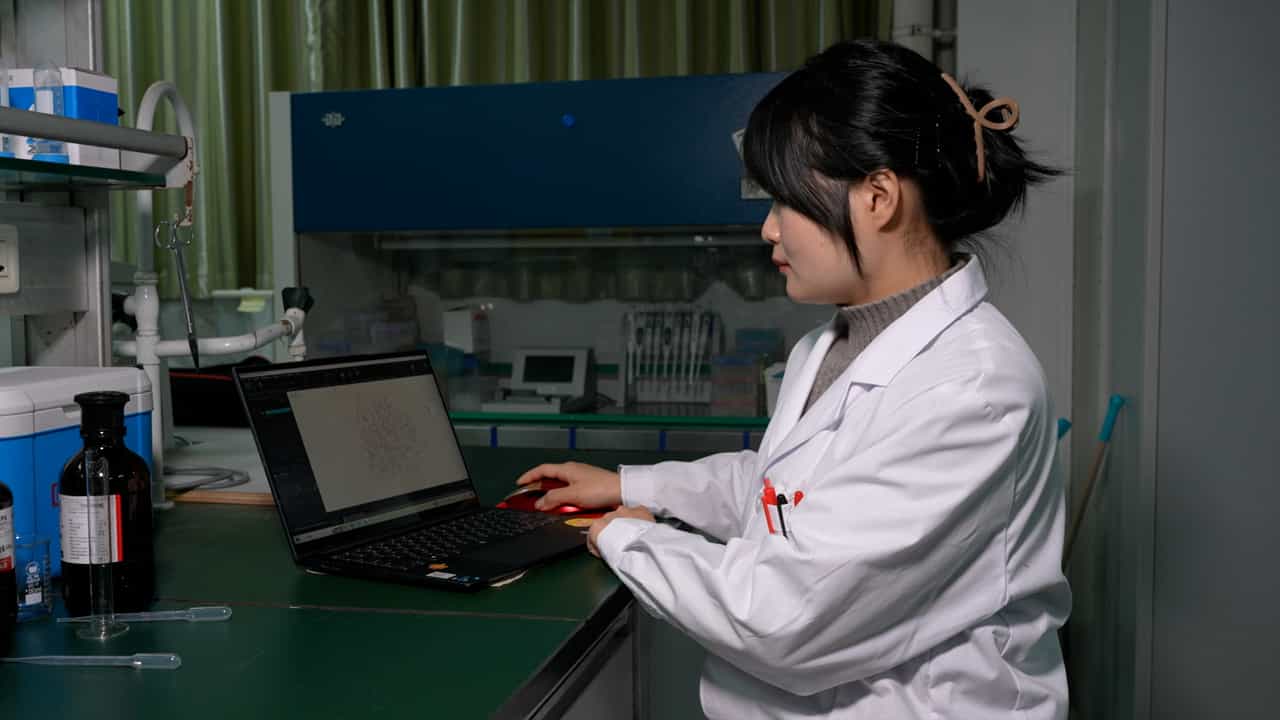
Protein Target Prediction and Validation of Small Molecule Compound
Published on: February 23, 2024
Ligand-protein inverse docking and its potential use in the computer search of protein targets of a small molecule
1Department of Computational Science, National University of Singapore, Blk S17, Level 7, 3 Science Drive 2, Singapore 117543. yzchen@cz3.nus.edu.sg

Abstract:
Ligand-protein docking has been developed and used in facilitating new drug discoveries. In this approach, docking single or multiple small molecules to a receptor site is attempted to find putative ligands. A number of studies have shown that docking algorithms are capable of finding ligands and binding conformations at a receptor site close to experimentally determined structures. These algorithms are expected to be equally applicable to the identification of multiple proteins to which a small molecule can bind or weakly bind. We introduce a ligand-protein inverse-docking approach for finding potential protein targets of a small molecule by the computer-automated docking search of a protein cavity database. This database is developed from protein structures in the Protein Data Bank (PDB). Docking is conducted with a procedure involving multiple-conformer shape-matching alignment of a molecule to a cavity followed by molecular-mechanics torsion optimization and energy minimization on both the molecule and the protein residues at the binding region. Scoring is conducted by the evaluation of molecular-mechanics energy and, when applicable, by the further analysis of binding competitiveness against other ligands that bind to the same receptor site in at least one PDB entry. Testing results on two therapeutic agents, 4H-tamoxifen and vitamin E, showed that 50% of the computer-identified potential protein targets were implicated or confirmed by experiments. The application of this approach may facilitate the prediction of unknown and secondary therapeutic target proteins and those related to the side effects and toxicity of a drug or drug candidate. Proteins 2001;43:217-226.
More Related Videos
 10:01
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
 14:34
A Bilingual Computational Workflow for Identifying Potential PLK1 Inhibitors in American Sign Language and English
Published on: April 3, 2026
Related Concept Videos
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:04
Protein-protein Interfaces
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 00:49
Ligand Binding and Linkage
 02:26
Protein Networks
These interactions can be represented through maps depicting protein-protein interaction networks, represented as nodes and edges. Nodes are circles that are representative of a protein,...
02:40Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...