Related Experiment Video
Updated: Oct 7, 2026
Synthesizing Amino Acids Modified with Reactive Carbonyls in Silico to Assess Structural Effects Using Molecular Dynamics Simulations
Published on: April 26, 2024
Ab initio and molecular mechanics (MM3) calculations of protonated-neutral diamine hydrogen bonds
J B Sorensen1, A H Lewin, J P Bowen
1The Computational Center for Molecular Structure and Design, Department of Chemistry, University of Georgia, Athens, Georgia 30602, USA.

Abstract:
Ab initio calculations of cation-neutral diamine complexes have been carried out at the MP2/6-311+G** level. The geometry and energetics of the charge-reinforced hydrogen bond are analyzed with respect to the alkyl substitution of both the protonated and neutral nitrogen atoms, and these results have been used to improve the quality of the MM3(2000) force field. In addition, specialized hydrogen bond parameters optimized for MM3(2000) are presented. These parameters allow very accurate gas-phase modeling of the charge-neutral diamine environment. Molecular mechanics calculations can model effectively protonated amine-neutral amine hydrogen bonds in the gas phase and solution (continuum dielectric) through a combination of charge-dipole interactions and explicit hydrogen-bonding terms.
More Related Videos


Related Concept Videos
 01:21
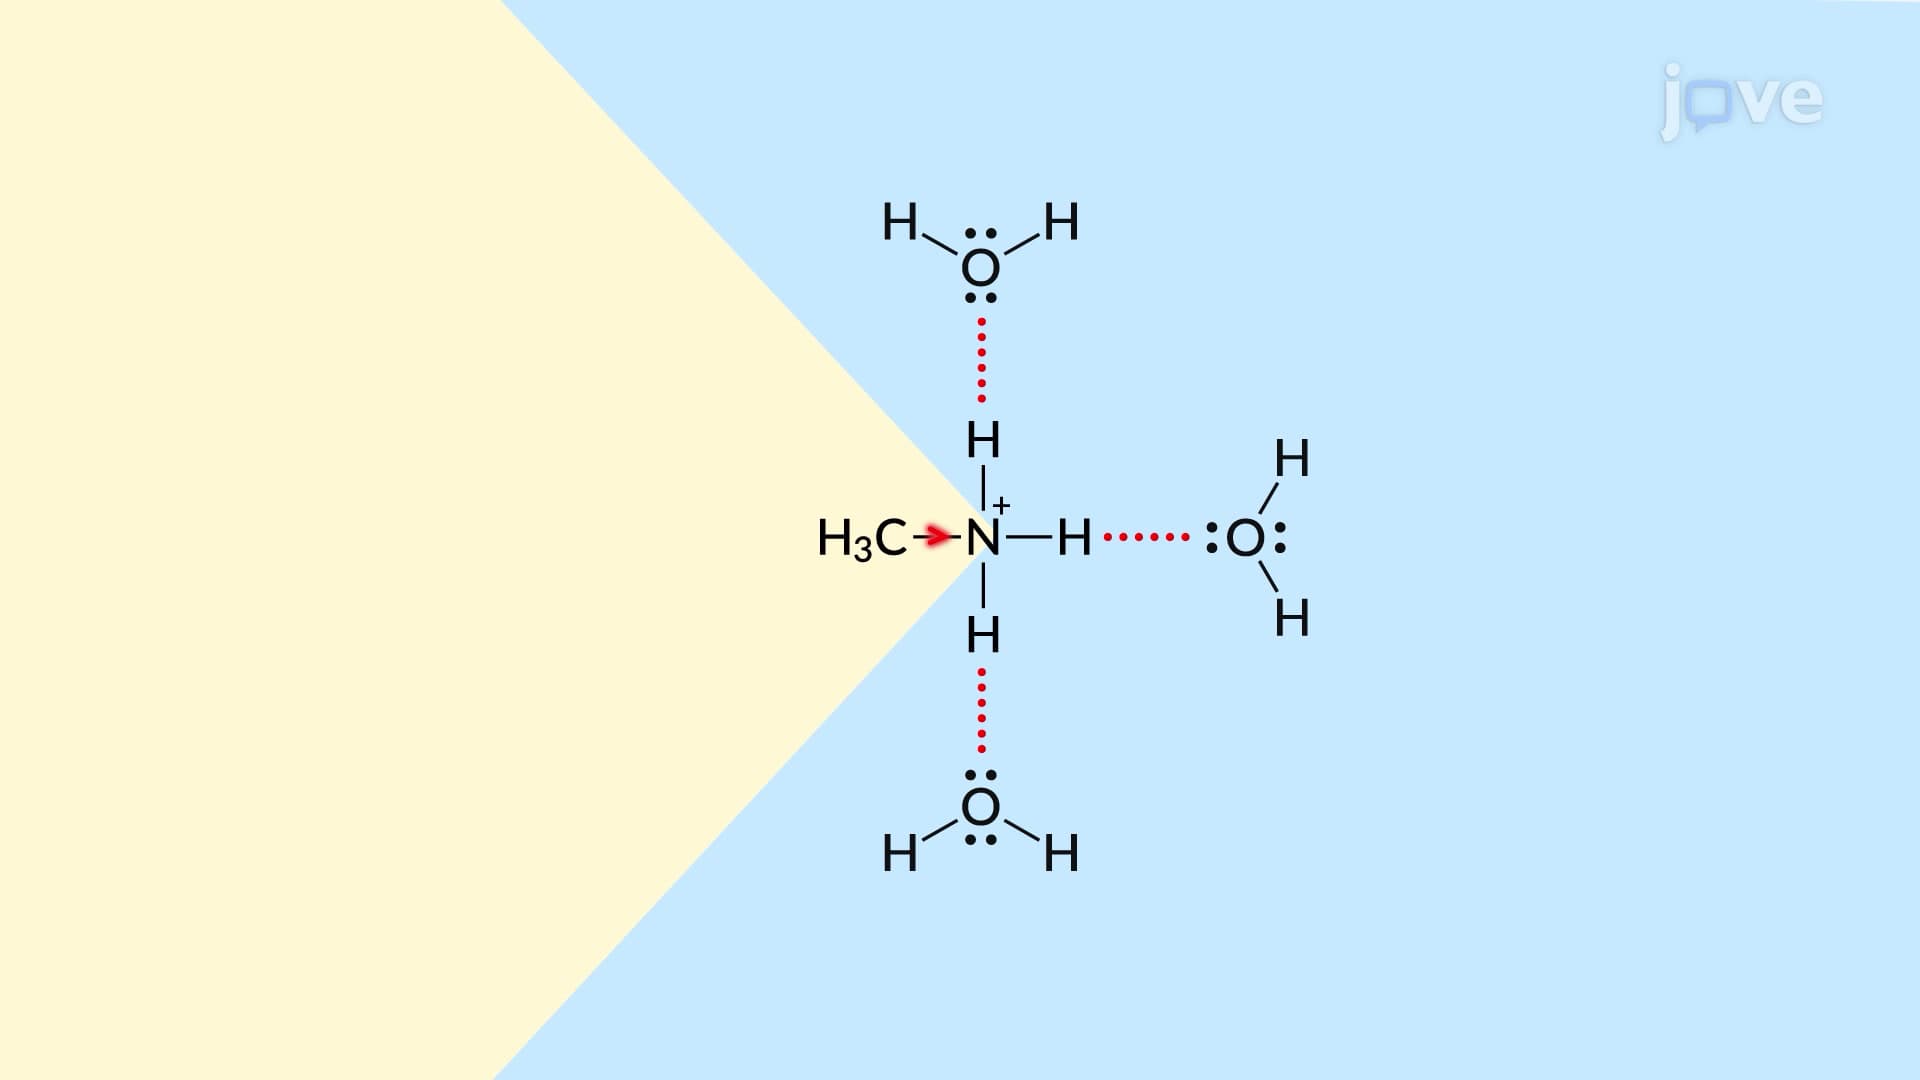
Basicity of Aliphatic Amines
To measure the basicity of amines, two conventions are generally used. The first defines Kb as the basicity constant for the deprotonation reaction of water by the amine, as presented in Figure 1. Conventionally, lower Kb indicates higher...
 01:23
Aldehydes and Ketones with Amines: Imine Formation Mechanism
Imines are formed under mildly acidic conditions. A pH of 4.5 is ideal for the reaction.
If the pH is low or the solution is too acidic, the reaction slows down in the...
 01:26
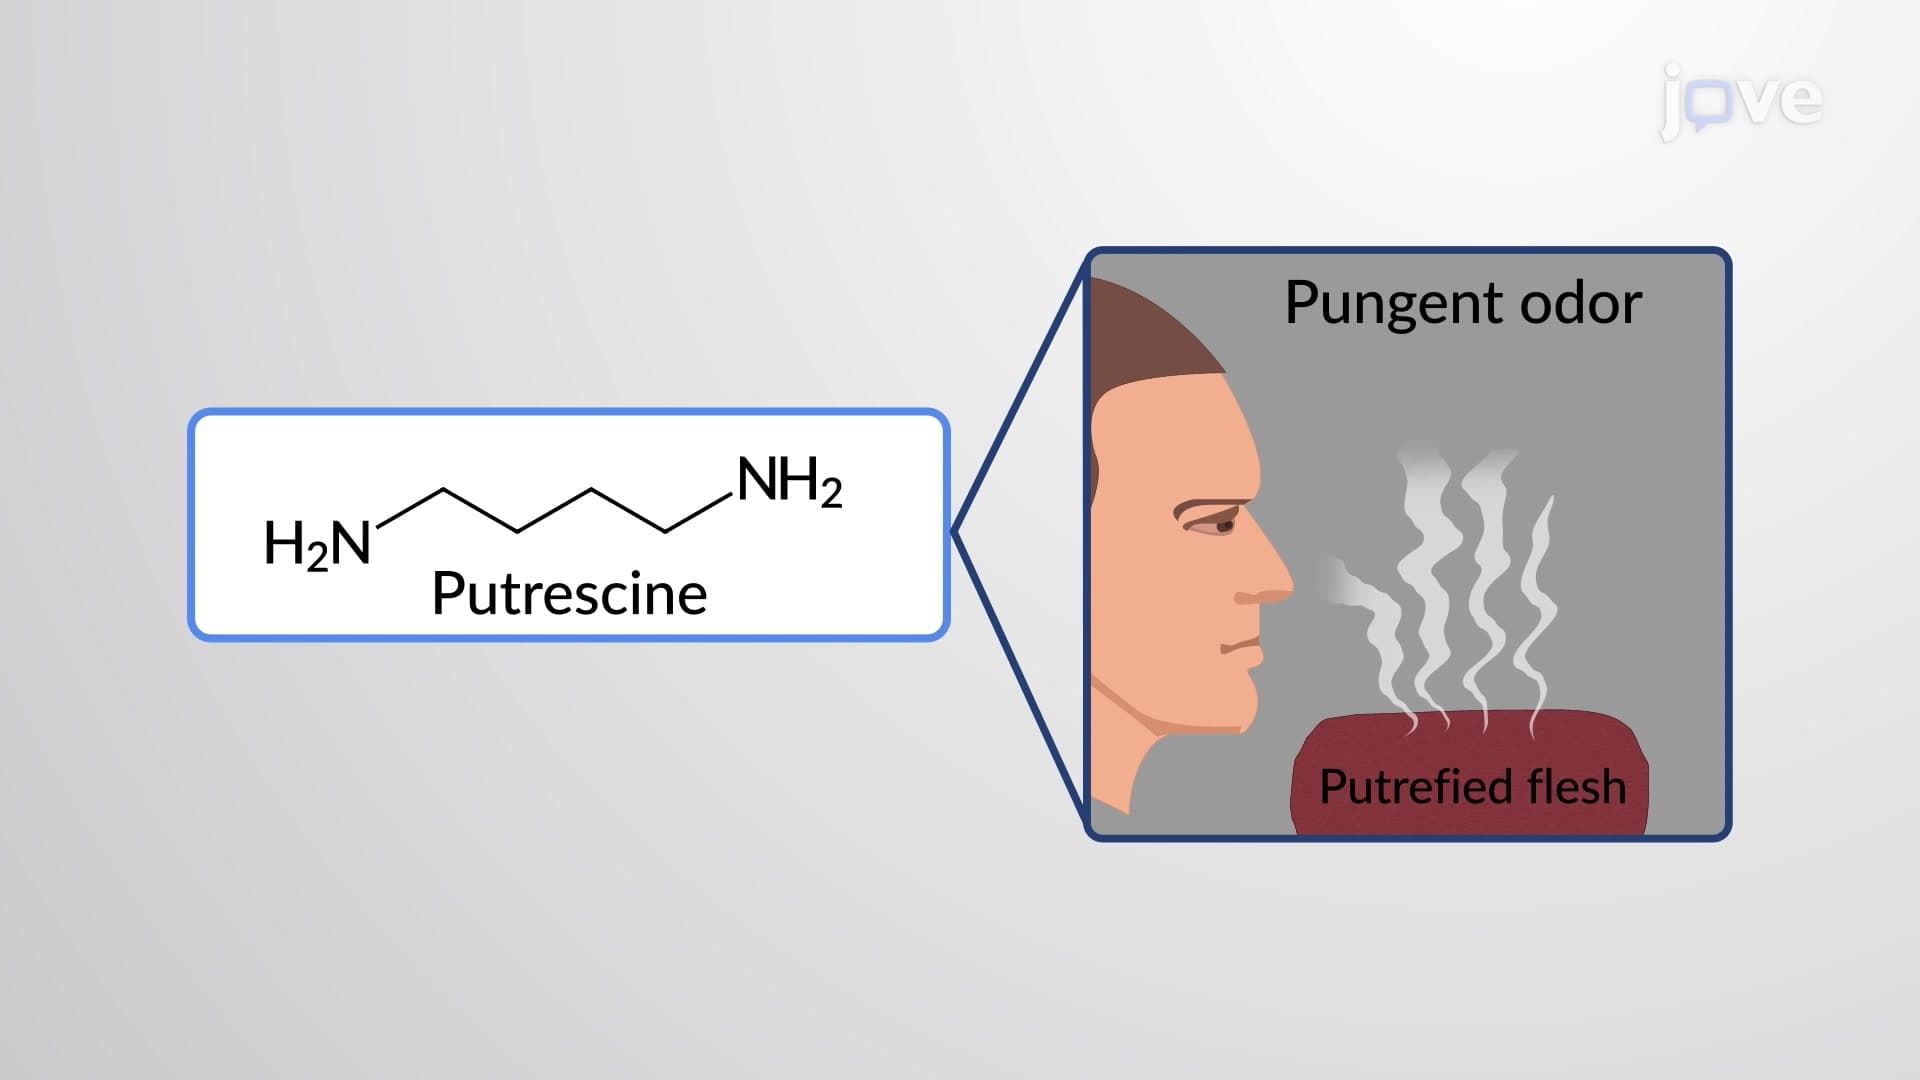
Physical Properties of Amines
 01:04
Hydrogen Bonds
 00:26
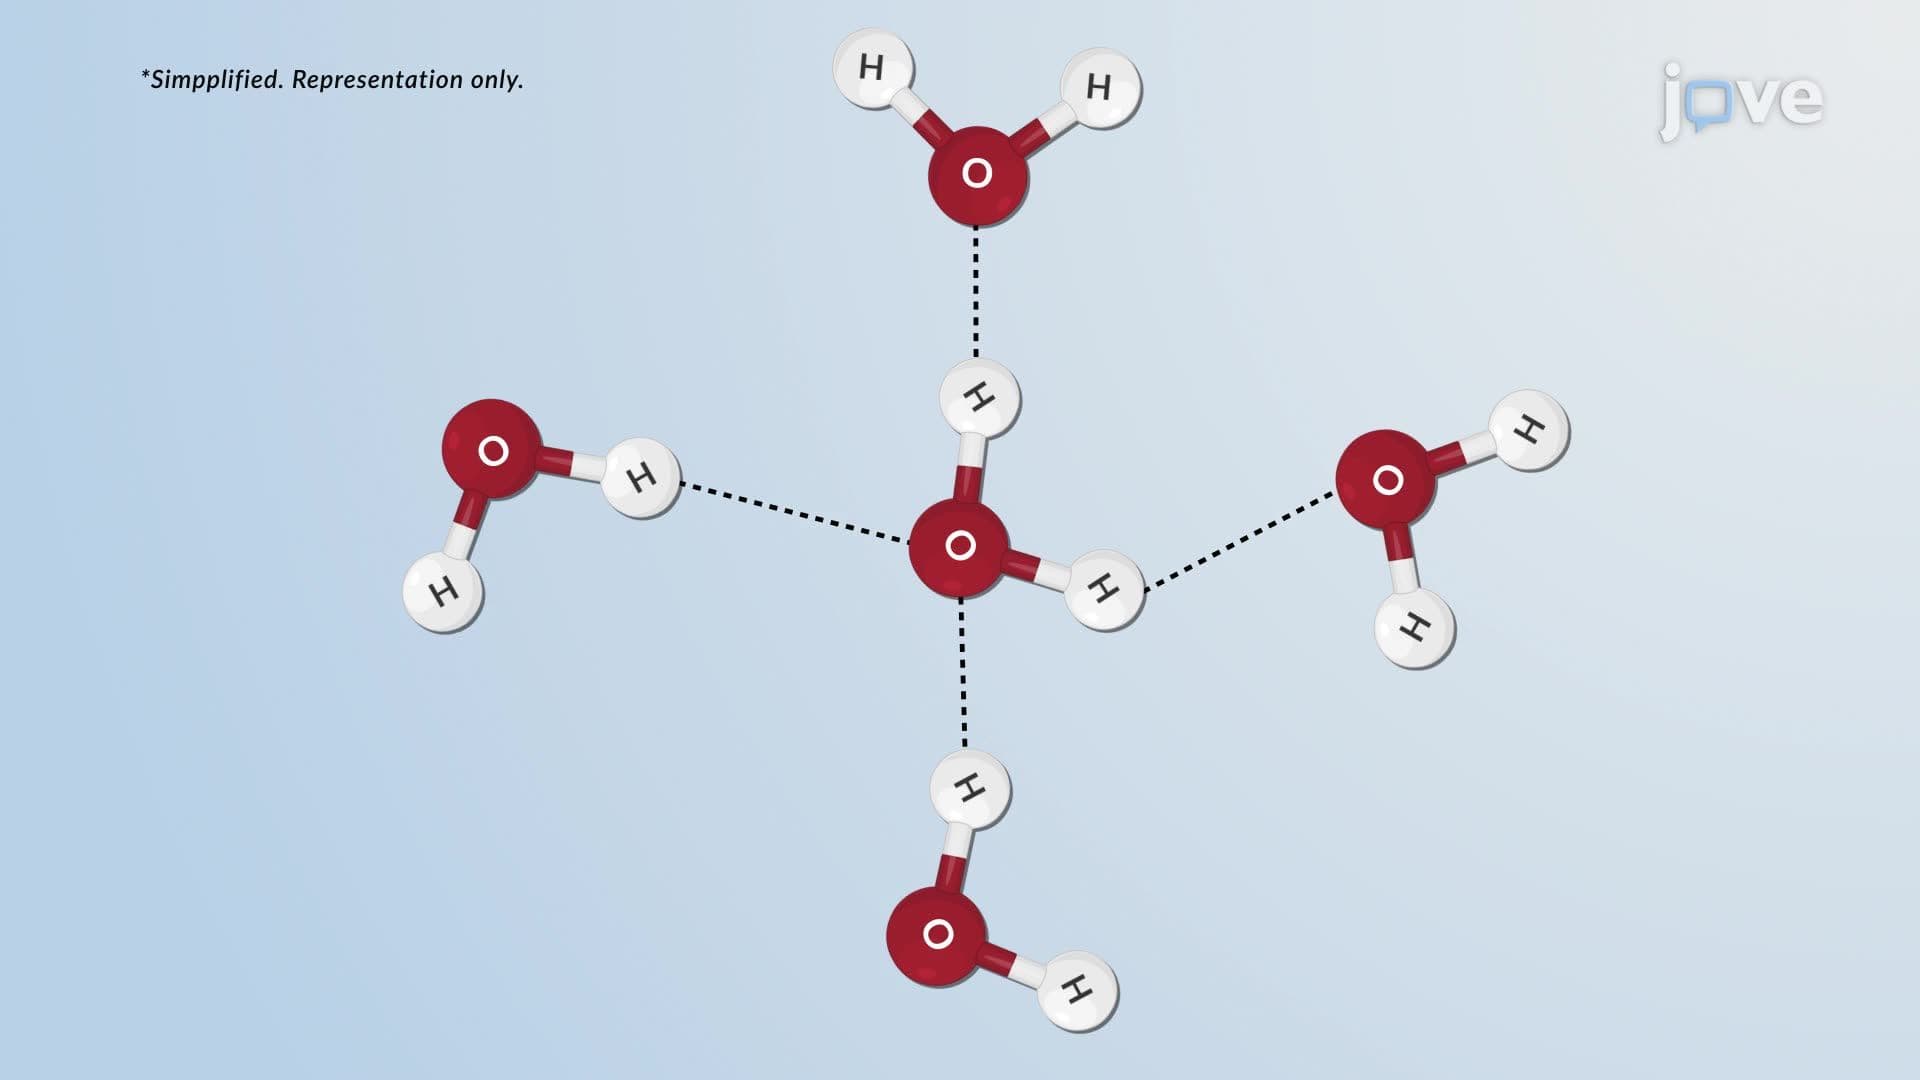
Hydrogen Bonds
 03:38
Polyprotic Acids