Related Experiment Video
Updated: Aug 6, 2026
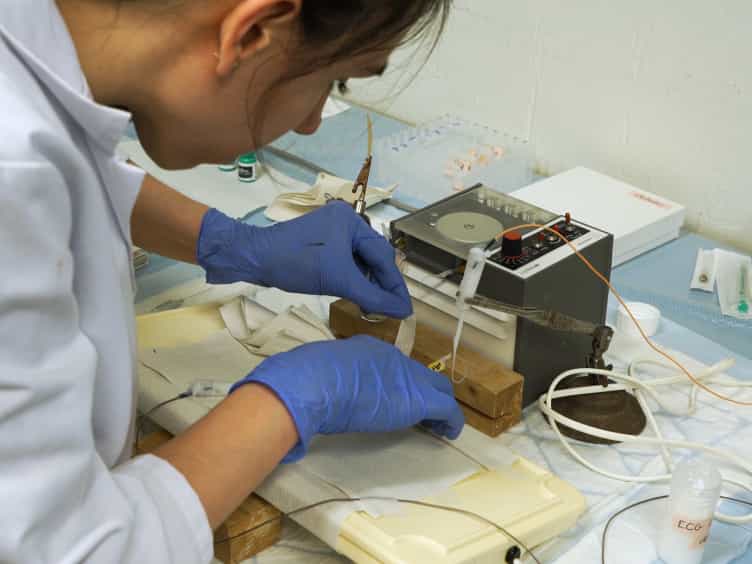

Nasal Potential Difference to Quantify Trans-epithelial Ion Transport in Mice
Published on: July 4, 2018
Evidence that systemic gentamicin suppresses premature stop mutations in patients with cystic fibrosis
1Department of Pediatrics, University of Alabama at Birmingham, USA. jclancy@peds.uab.edu

Abstract:
Here we report the effects of gentamicin treatment on cystic fibrosis transmembrane regulator (CFTR) production and function in CF airway cells and patients with CF with premature stop mutations. Using immunocytochemical and functional [6-methoxy-N- (3-sulfopropyl) quinolinium (SPQ)-based] techniques, ex vivo exposure of airway cells from stop mutation CF patients led to the identification of surface-localized CFTR in a dose-dependent fashion. Next, five patients with CF with stop mutations and five CF control subjects were treated with parenteral gentamicin for 1 wk, and underwent repeated in vivo measures of CFTR function (nasal potential difference [PD] measurements and sweat chloride [Cl(-)] testing). During the treatment period, the number of nasal PD readings in the direction of Cl(-) secretion was increased approximately 3-fold in the stop mutation patient group compared with controls (p < 0.001), and four of five stop mutation patients with CF had at least one reading during gentamicin treatment with a Cl(-) secretory response of more than -5 mV (hyperpolarized). A response of this magnitude was not seen in any of the CF control subjects (p < 0.05). In an independent series of experiments designed to test the ability of repeat nasal PDs to detect wild-type CFTR function, evidence of Cl(-) secretion was seen in 88% of control (non-CF) nasal PDs, and 71% were more than -5 mV hyperpolarized. Together, these results suggest that gentamicin treatment can suppress premature stop mutations in airway cells from patients with CF, and produce small increases in CFTR Cl(-) conductance (as measured by the nasal PD) in vivo.
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:15
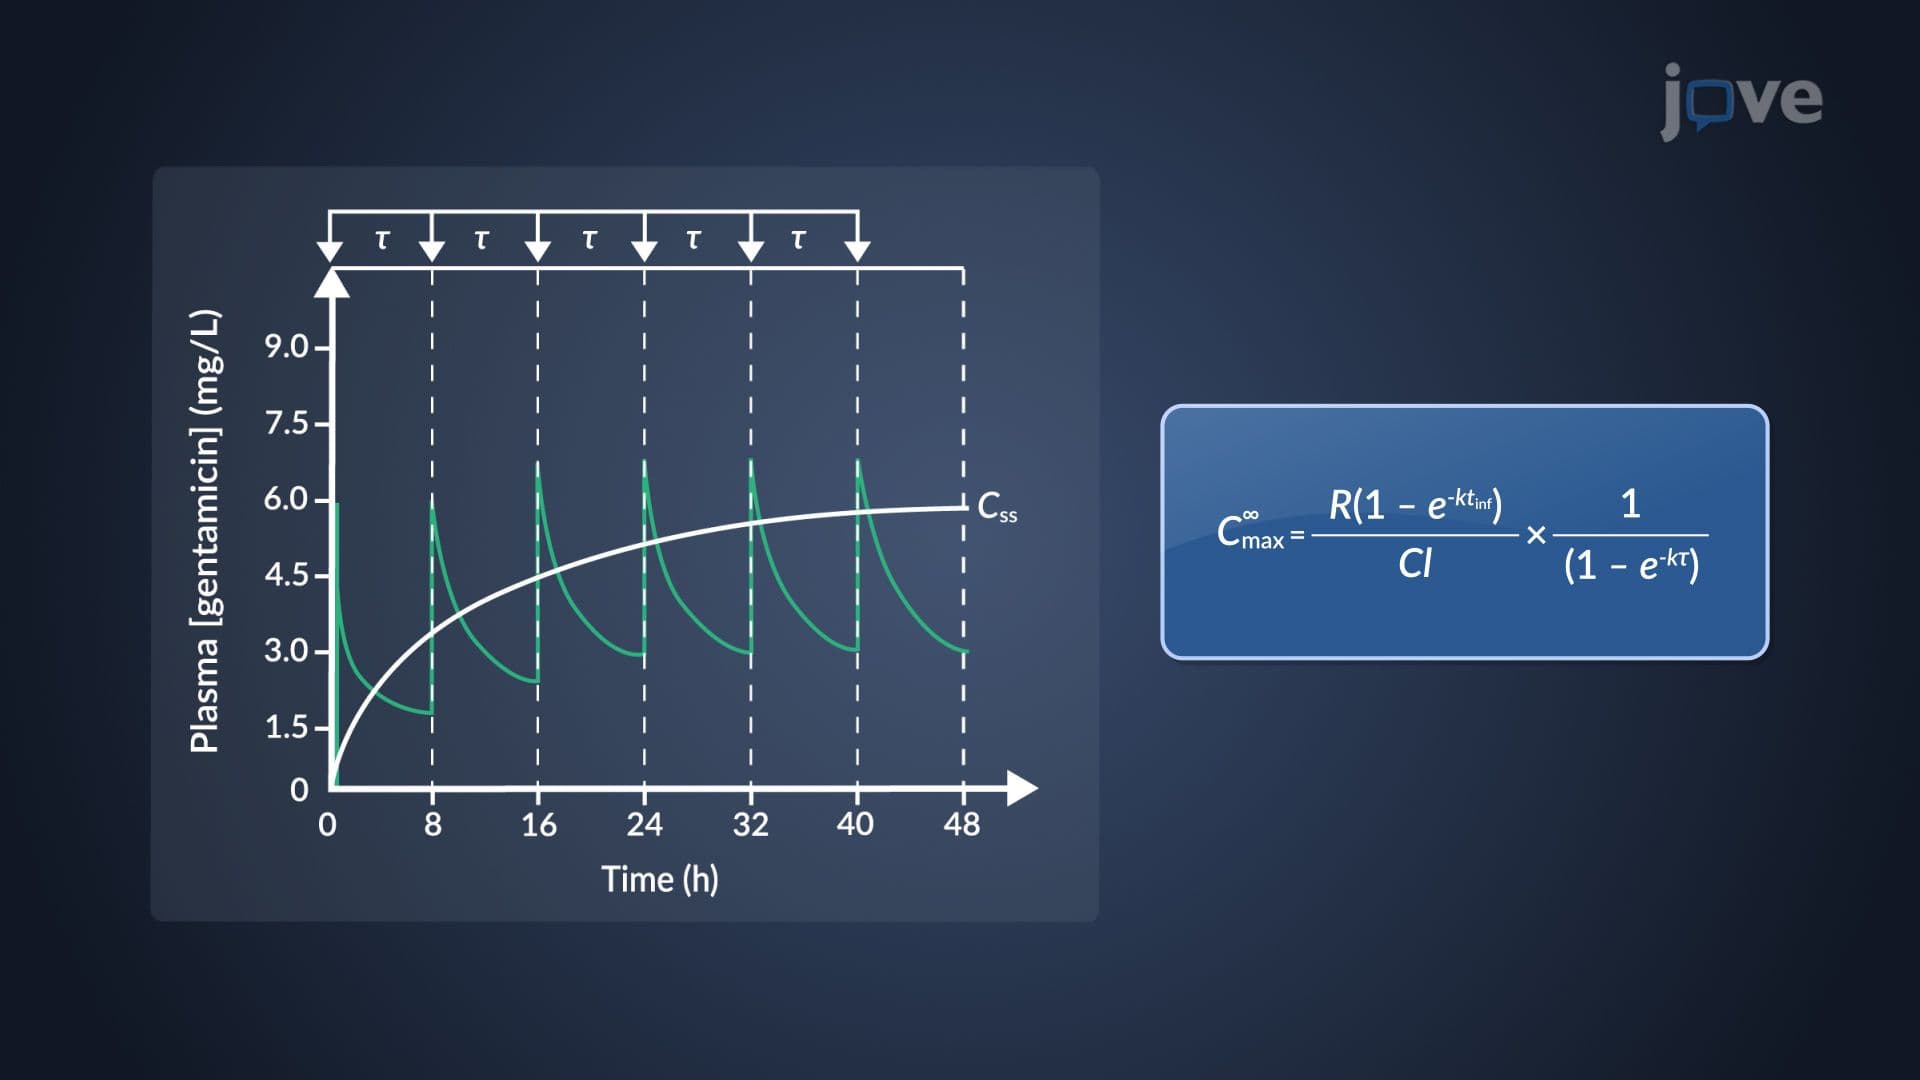
Determination of Multiple Dosing Parameters: Steady-State, Minimum and Maximum Concentrations
 01:25
Mechanism of Antibiotic Resistance in MRSA