Related Experiment Video
Updated: Aug 15, 2026

Structural Studies of Macromolecules in Solution using Small Angle X-Ray Scattering
Published on: November 5, 2018
Aspirin. An ab initio quantum-mechanical study of conformational preferences and of neighboring group interactions
1Department of Chemistry, University of Missouri-Columbia, Columbia, Missouri 65211, USA. glaserr@missouri.edu

Abstract:
The potential energy surface of acetylsalicylic acid, aspirin, has been explored at the RHF/6-31G* and B3LYP/6-31G* levels, and single-point calculations were performed at levels up to B3LYP/6-311G**//B3LYP/6-31G*. All conformational isomers have been located, the thermochemical functions have been computed, and relative energies and free enthalpies were determined. The conformational space of aspirin is spanned by three internal coordinates, and these are the carboxylic acid C-O conformation (s-trans preferred by about 7 kcal/mol), the C-COOH conformation (Z preferred unless there are H-bonding opportunities), and the ester C-O conformation (s-trans preferred by about 4 kcal/mol). There are nine aspirin isomers since one of the conformers realizes hydrogen-bonding structure isomerism as well. Neighboring group interactions are discussed with reference to the intrinsic properties of benzoic acid and phenyl acetate. The intrinsic conformational preference energies for benzoic acid and phenyl acetate are not additive. The acid s-trans preference energies differ by as much as 9 kcal/mol depending on the Ph-COOH and ester conformations. Similarly, the E-preference energies about the Ph-COOH bond vary by as much as 6 kcal/mol depending on the ester conformation. The structural discussion suggests an overall ortho repulsion between the functional groups in all aspirin isomers including the intramolecularly hydrogen-bonded isomers. The isodesmic reaction between the most stable conformers of benzoic acid and phenyl acetate to form aspirin and benzene is found to be endothermic by 2.7 kcal/mol and provides compelling evidence for and a quantitative measure of ortho repulsion. The ortho repulsion of 2.7 kcal/mol is a lower limit, and the ortho repulsion can increase to as much as 6 kcal/mol in some aspirin isomers.

Related Concept Videos
 02:37
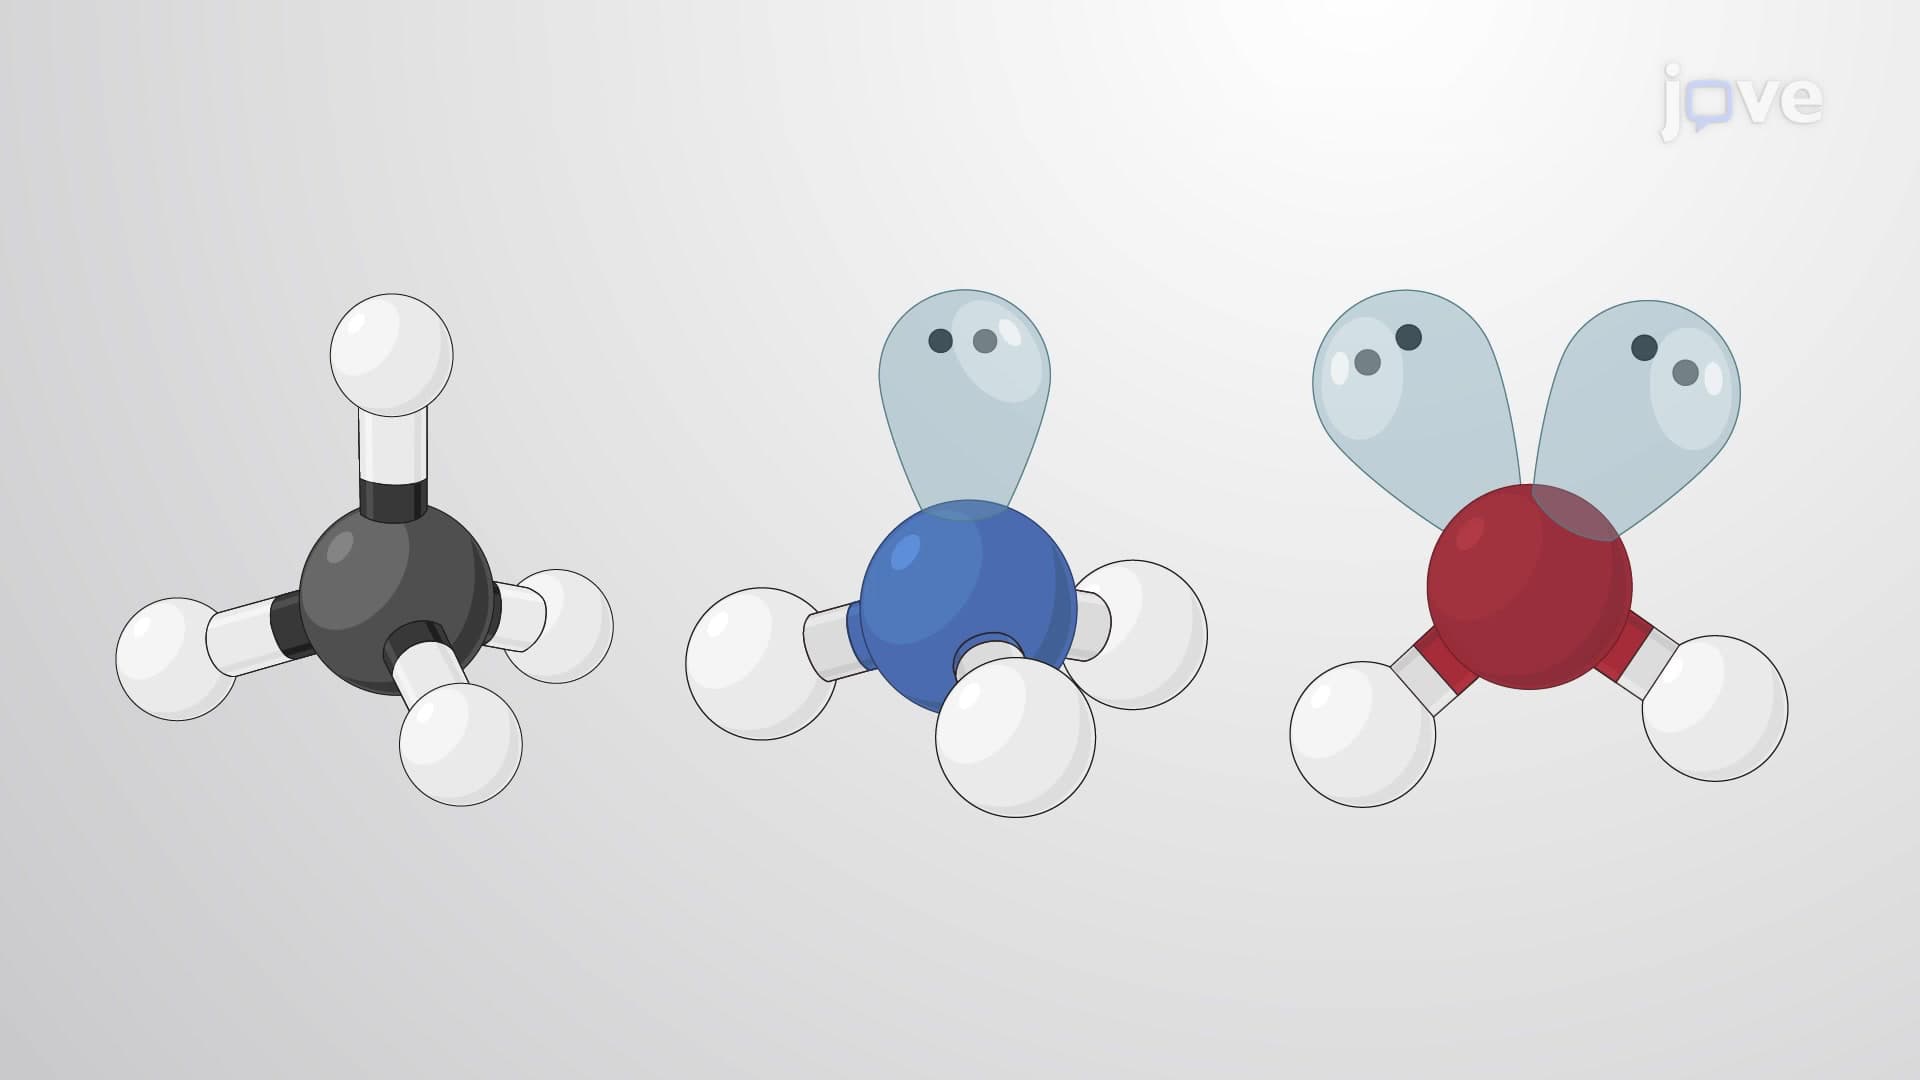
VSEPR Theory
 02:20
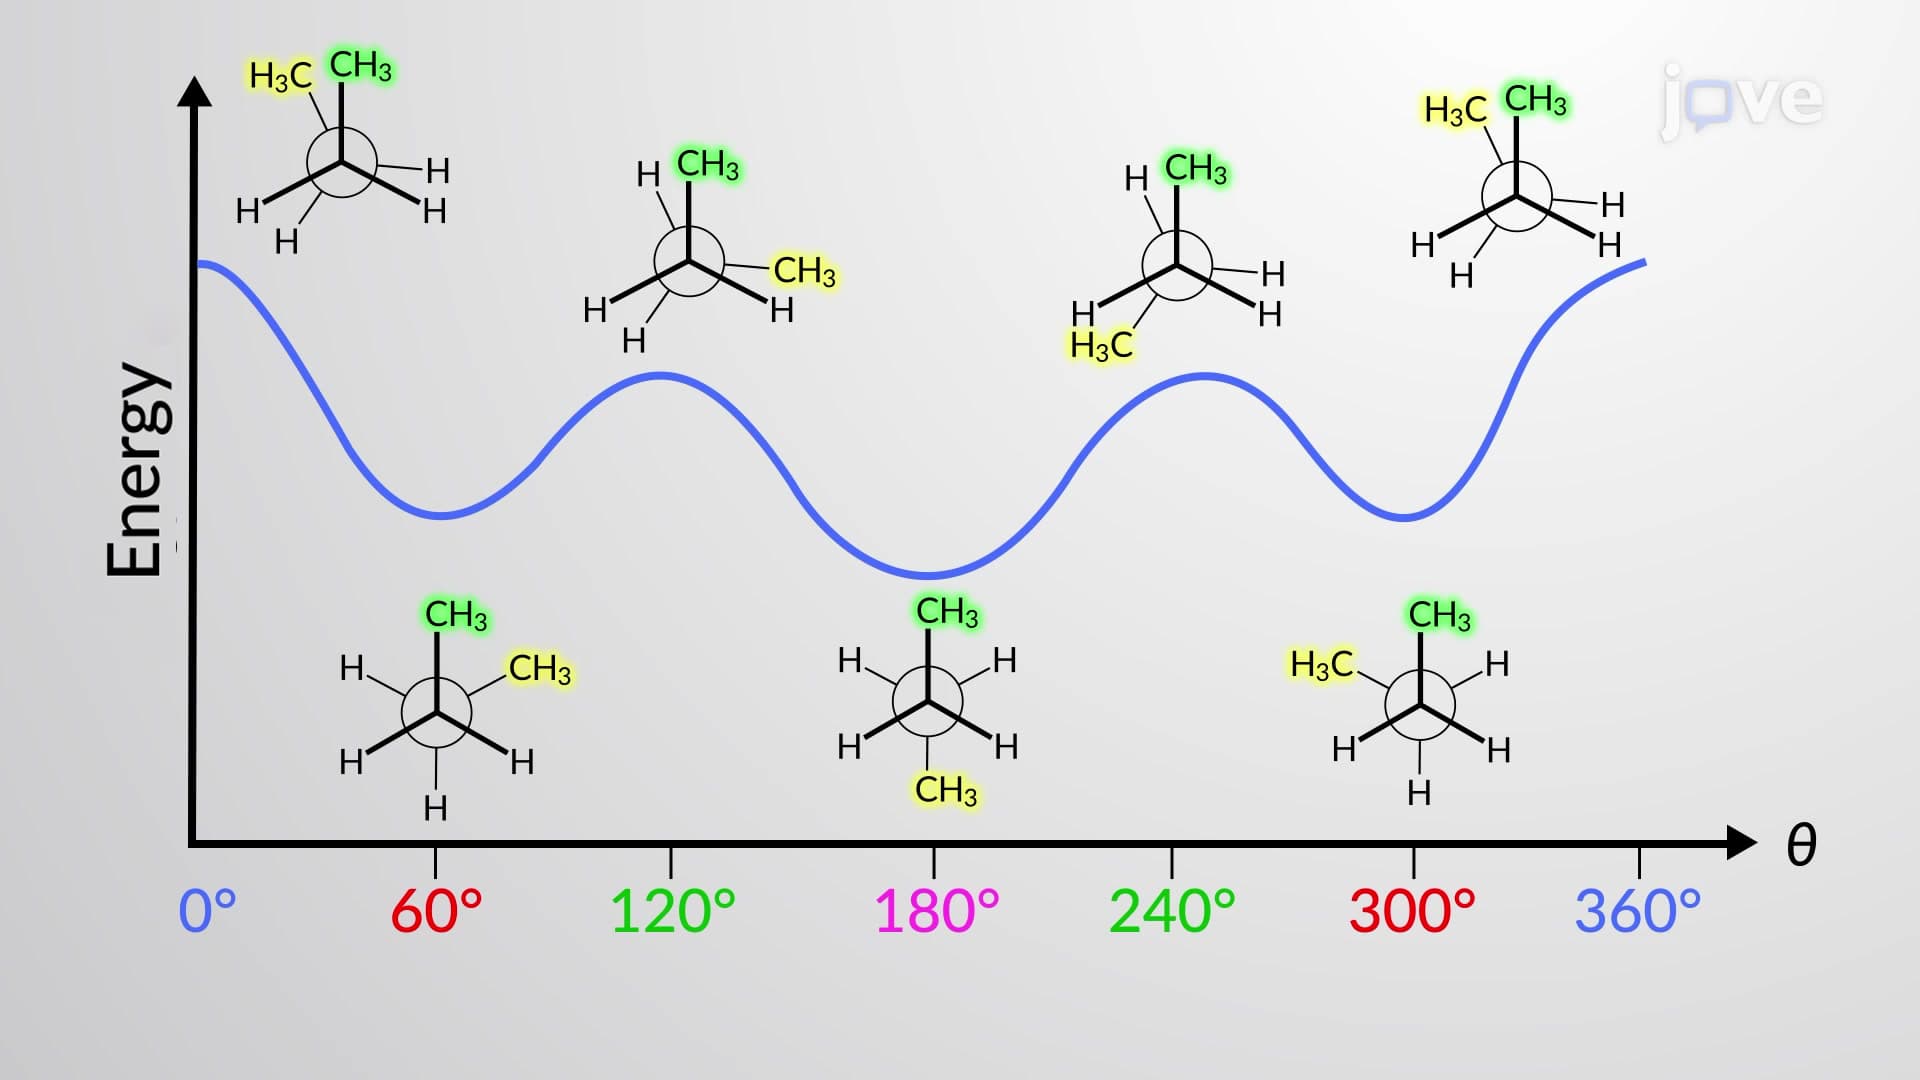
Conformations of Butane
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal tetrahedral value,...
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:34
Woodward–Hoffmann Selection Rules and Microscopic Reversibility