Related Experiment Video
Updated: Oct 5, 2026
Standardized Measurement of Nasal Membrane Transepithelial Potential Difference (NPD)
Published on: September 13, 2018
Increased nitrotyrosine in exhaled breath condensate in cystic fibrosis
B Balint1, S A Kharitonov, T Hanazawa
1Dept of Thoracic Medicine, Imperial College School of Medicine at the National Heart and Lung Institute, London, UK.

Abstract:
Exhaled nitric oxide (ENO), a marker of inflammation in airway diseases is decreased in cystic fibrosis (CF) patients, perhaps because nitric oxide (NO) is metabolized to oxidative end-products. A stable product, 3-nitrotyrosine, may indicate local formation of reactive nitrogen species. Whether NO metabolites in exhaled breath condensate may be increased in CF patients was investigated. The fractional concentration of ENO (Feno), nitrotyrosine and oxides of nitrogen in exhaled breath condensate from 36 stable CF patients were compared to 14 normal subjects using an enzyme immunoassay and fluorescence assay. Nitrotyrosine levels in breath condensate were increased significantly in stable CF patients, compared with normal subjects (25.3 +/- 1.5 versus 6.3 +/- 0.8 ng x mL(-1), p<0.0001). There was an inverse correlation between the levels of nitrotyrosine and the severity of lung disease. Feno levels were significantly lower in CF patients than in normal subjects (4.4 +/- 0.3 versus 5.6 +/- 0.4 (parts per billion), p<0.05). No correlation was found between nitrotyrosine and Feno levels in CF. There was no significant difference in the levels of nitrite and nitrate between CF patients and normals. The elevation in nitrotyrosine may reflect increased formation of reactive nitrogen species such as peroxynitrite or direct nitration by granulocyte peroxidases, indicating increased oxidative stress in airways of cystic fibrosis patients.
More Related Videos
 09:19
Capturing Actively Produced Microbial Volatile Organic Compounds from Human-Associated Samples with Vacuum-Assisted Sorbent Extraction
Published on: June 1, 2022
 06:57
The WinCF Model - An Inexpensive and Tractable Microcosm of a Mucus Plugged Bronchiole to Study the Microbiology of Lung Infections
Published on: May 8, 2017
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:24
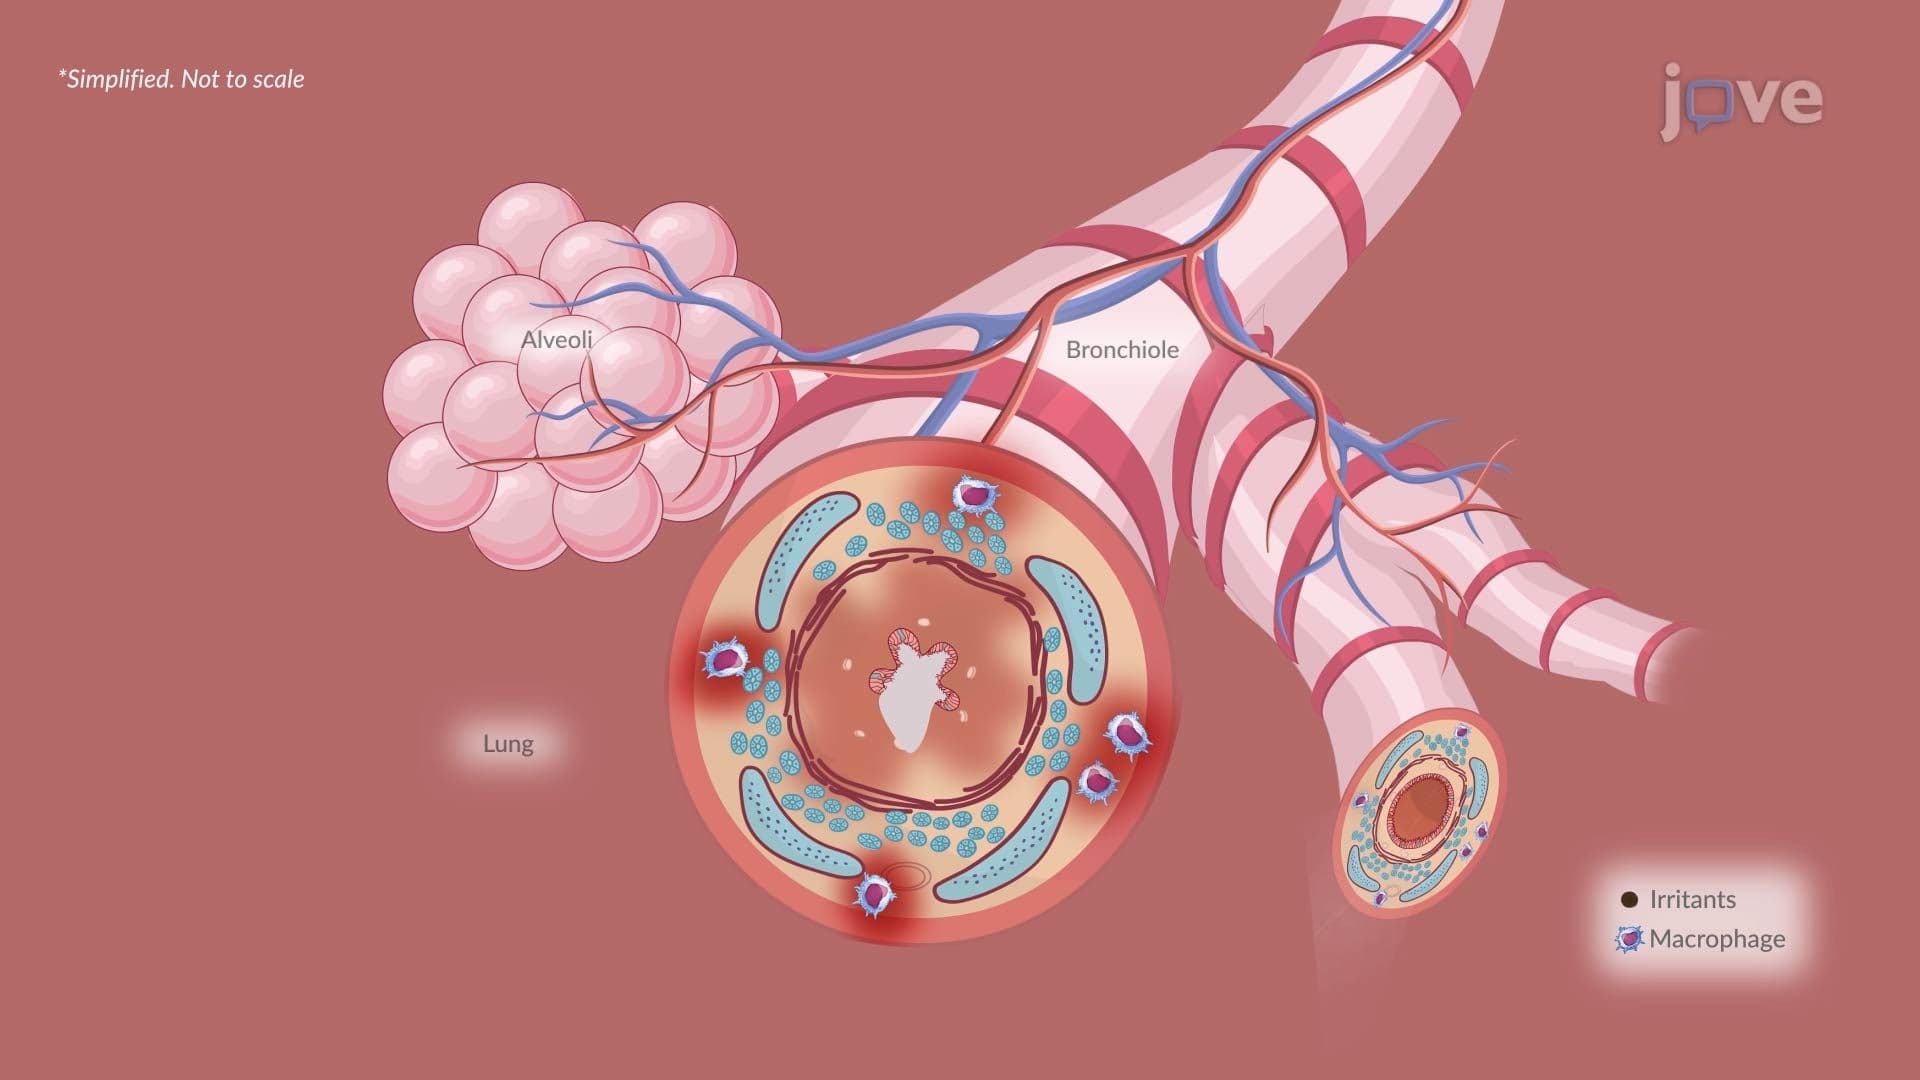
Chronic Obstructive Pulmonary Disease III: Chronic Bronchitis Features
 01:20
Chronic Obstructive Pulmonary Disease-II: Pathophysiology
Chronic Inflammation