Ab initio predictions on the rotational spectra of carbon-chain carbene molecules

S A Maluendes1, A D McLean

  • 1Molecular Research Institute, Palo Alto, CA 94304, USA.

Chemical Physics Letters
|December 18, 1992
PubMed

Related Concept Videos

Chair Conformation of Cyclohexane02:02

Chair Conformation of Cyclohexane

The chair conformation is the most stable form of cyclohexane due to the absence of angle and torsional strain. The absence of angle strain is a result of cyclohexane’s bond angle being very close to the ideal tetrahedral bond angle of 109.5° in its chair conformer. Similarly, the torsional strain is also absent owing to the perfectly staggered arrangement of bonds.
The hydrogen atoms linked to carbons are arranged in two different axial and equatorial orientations to achieve this staggered...
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution00:52

¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
At room temperature, the chair conformer of cyclohexane undergoes rapid ring flipping between two equivalent chair conformers at a rate of approximately 105 times per second. These two chair conformers are in equilibrium. The rapid ring flipping results in the interconversion of the axial proton to an equatorial proton and an equatorial to the axial proton. Such interconversions are too rapid and cannot be detected on the NMR timescale. Hence, the NMR spectrometer cannot distinguish between the...
Spectroscopy of Carboxylic Acid Derivatives01:26

Spectroscopy of Carboxylic Acid Derivatives

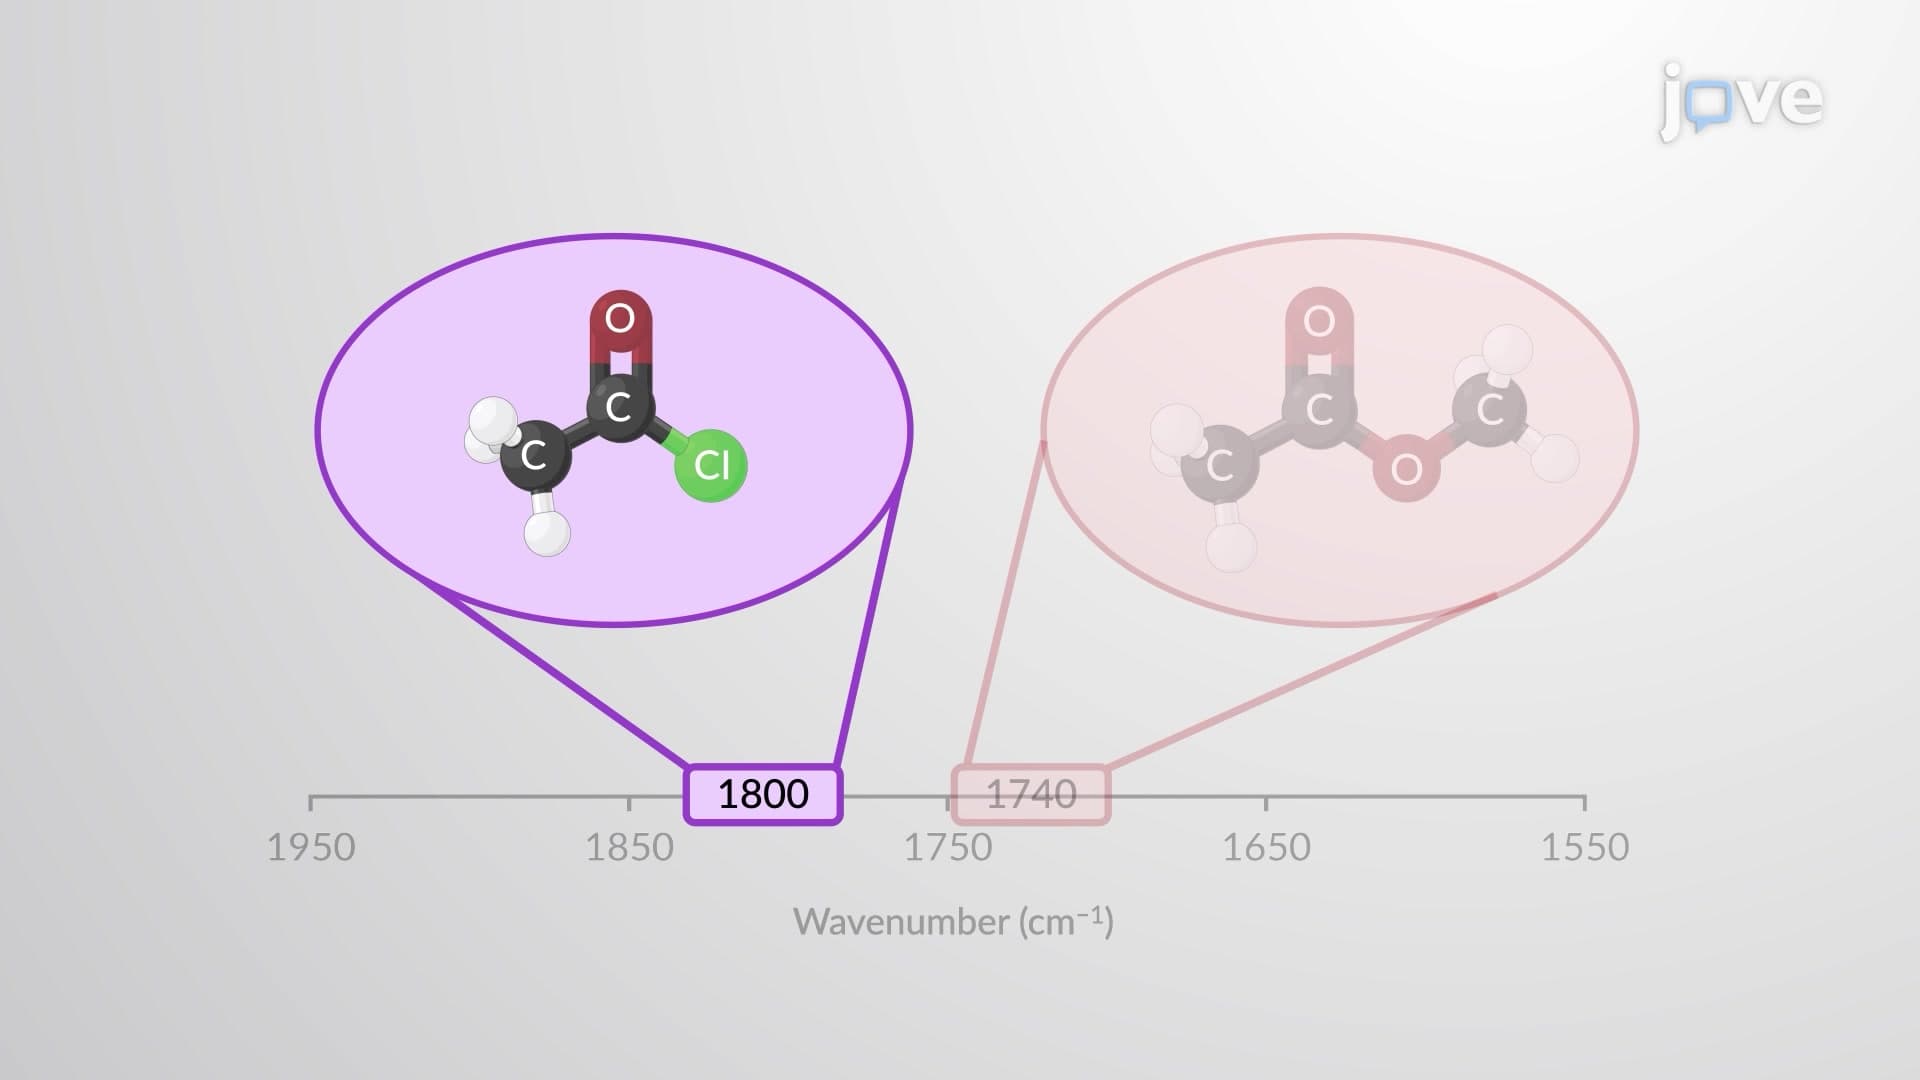
Infrared spectroscopy is primarily used to determine the types of bonds and functional groups. In carboxylic acid derivatives, a typical carbonyl bond absorption is observed around 1650–1850 cm−1. For esters, the absorption is recorded at around 1740 cm−1, while acid halides show the absorption at about 1800 cm−1. Another acid derivative, the acid anhydrides, exhibit two carbonyl absorption around 1760 cm−1 and 1820 cm−1, arising from the symmetrical and unsymmetrical carbonyl vibration.
In the...
NMR Spectroscopy of Aromatic Compounds01:14

NMR Spectroscopy of Aromatic Compounds

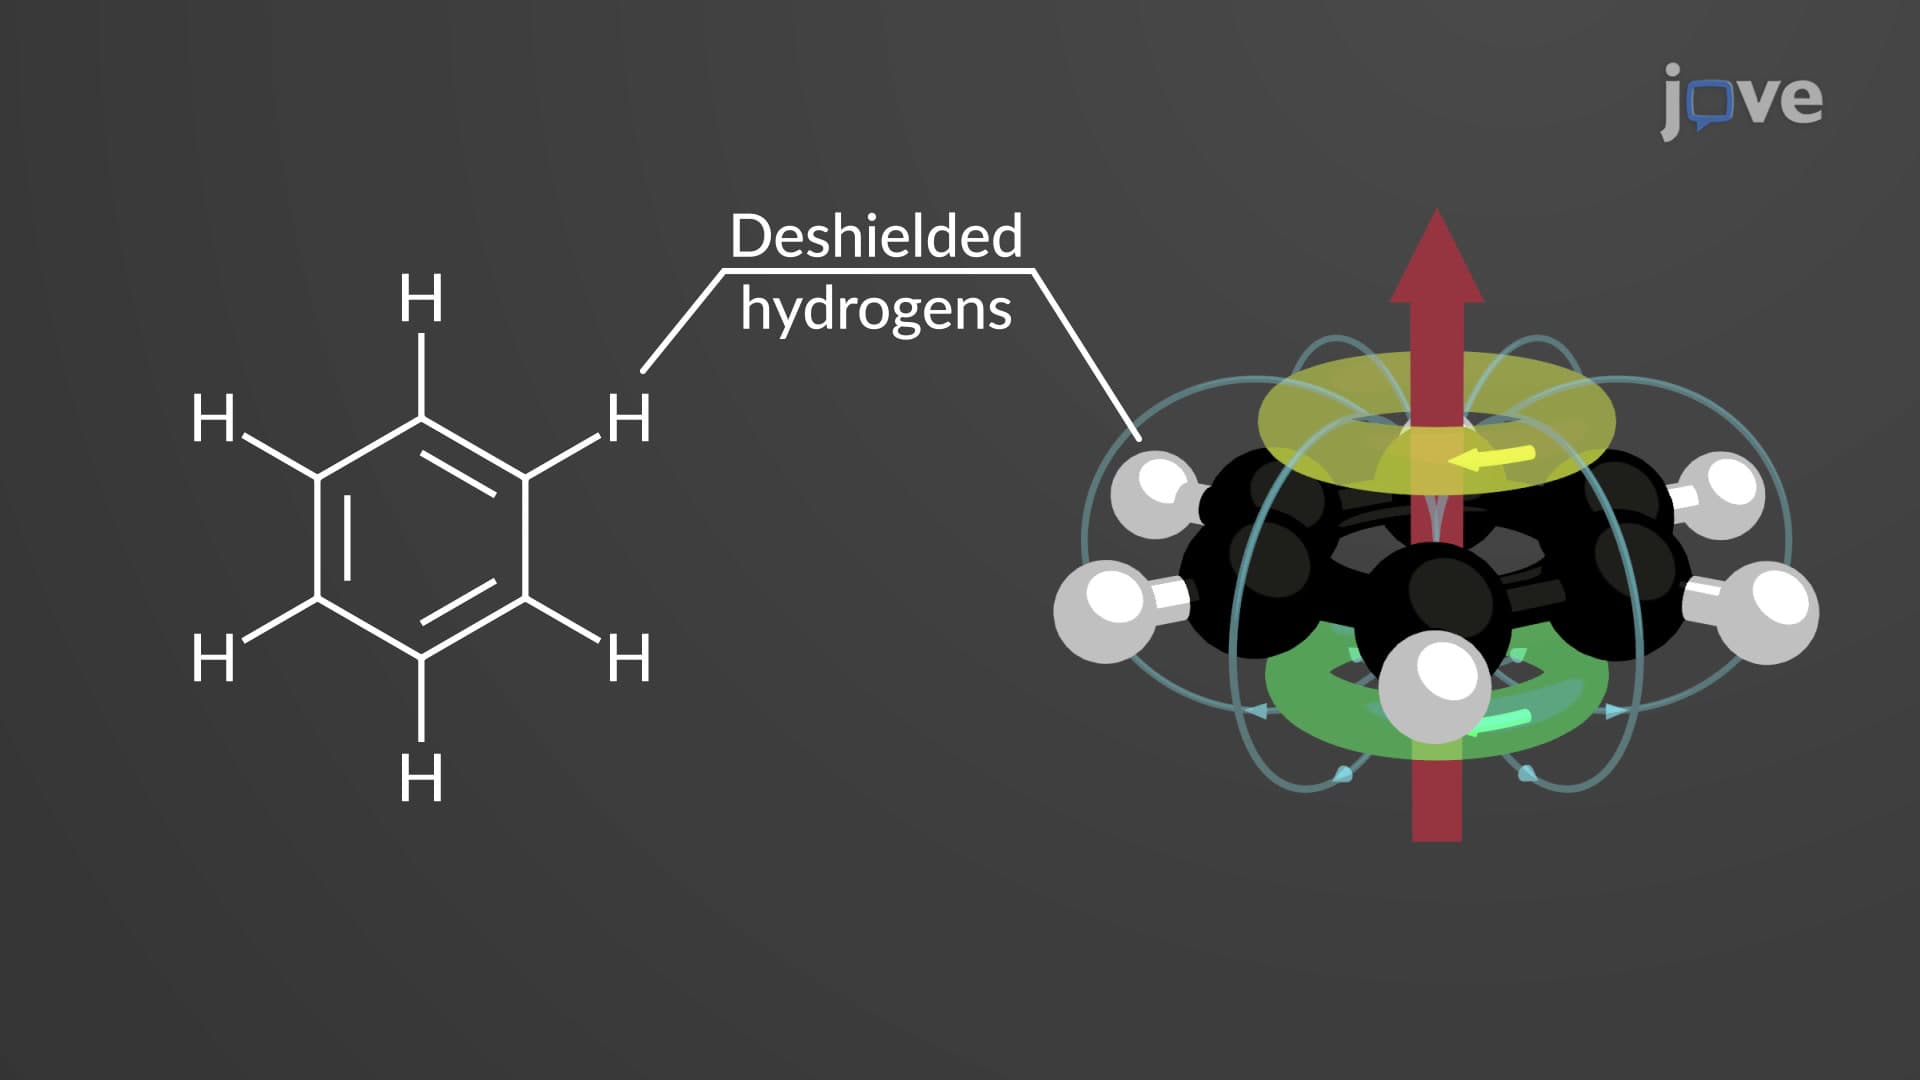
Aromatic compounds can be identified or analyzed using proton NMR and carbon‐13 NMR. Typically, aromatic hydrogens or hydrogens directly bonded to the aromatic rings are strongly deshielded by the aromatic ring current. Therefore, they absorb in the range of 6.5–8.0 ppm in proton NMR spectra. For instance, aromatic hydrogens directly bonded to the benzene ring absorb at 7.3 ppm. However, aromatic hydrogens of larger rings absorb farther upfield or downfield than the ideal range. Consider...
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR01:15

¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR

The axial and equatorial protons in cyclohexane can be distinguished by performing a variable-temperature NMR experiment. In this process, except for one proton, the remaining eleven protons are replaced by deuterium. The deuterium substitution avoids the possible peak splitting caused by the spin-spin coupling between the adjacent protons. The remaining proton flips between the axial and equatorial positions.
UV–Vis Spectroscopy: Woodward–Fieser Rules01:29

UV–Vis Spectroscopy: Woodward–Fieser Rules

UV–Visible absorption spectra of conjugated dienes arise from the lowest energy π → π* transitions. The light-absorbing part of the molecule is called the chromophore, and the substituents directly attached to the chromophore are called auxochromes. A strong correlation exists between the absorption maxima, λmax, and the structure of a conjugated π system. The Woodward–Fieser rules predict the value of λmax for a given structure by adding the contributions...