Related Experiment Video
Updated: Oct 4, 2026

Mass Spectrometry and Luminogenic-based Approaches to Characterize Phase I Metabolic Competency of In Vitro Cell Cultures
Published on: March 28, 2017
alpha-Naphthoflavone acts as activator and reversible or irreversible inhibitor of rabbit microsomal CYP3A6
L Boek-Dohalská1, P Hodek, M Sulc
1Department of Biochemistry, Faculty of Natural Sciences, Charles University, Albertov 2030, 128 40 2, Prague, Czech Republic.

Abstract:
This report describes the effect of alpha-naphthoflavone (alpha-NF), a known substrate, inhibitor and activator of several cytochromes P450 (CYP), on rabbit CYP3A6. Hepatic microsomes of rabbit pretreated with rifampicine (RIF), enriched with CYP3A6, as well as purified CYP3A6 reconstituted with isolated NADPH:CYP reductase were used as enzymatic systems in this study. The data from difference spectroscopy experiments showed that alpha-NF does yield a type I binding spectrum. This compound is oxidized by microsomal CYP3A6 into two metabolites (5,6-epoxide and trans-7,8-dihydrodiol). While alpha-NF is a substrate of CYP3A6, it also acts as an enzyme modulator. Under the conditions used, stimulation of 17beta-estradiol 2-hydroxylation by alpha-NF was observed. In contrast, this compound reversibly inhibited N-demethylation of erythromycin and tamoxifen, competitively with respect to these substrates, having the K(i) values of 51.5 and 18.0 microM, respectively. Moreover, alpha-NF was found to be an effective inactivator of progesterone and testosterone 6beta-hydroxylation catalyzed by CYP3A6 in RIF-microsomes. In addition, time- and concentration-dependent inactivation of human CYP3A4-mediated 6beta-hydroxylation of testosterone by alpha-NF, was determined. The inactivation of CYP3A6 followed pseudo-first-order kinetics and was dependent on both NADPH and alpha-NF. The concentrations required for half-maximal inactivation (K(i)) were 80.1 and 108.5 microM and the times required for half of the enzyme to be inactivated were 10.0 and 11.9 min for 6beta-hydroxylation of progesterone and testosterone, respectively. The loss of the enzyme activity was not recovered following dialysis, while 90% of the ability to form a reduced CO complex remained. This indicates the binding of alpha-NF to a CYP apoprotein molecule rather than to a heme moiety. Protection from inactivation was seen in the presence of all tested CYP3A substrates. Progesterone and testosterone protected CYP3A6 against inactivation competitively with respect to inactivator, erythromycin non-competitively and 17beta-estradiol showed a mixed type of protection. Here, we described for the first time that alpha-NF is capable of irreversible inhibition of microsomal rabbit CYP3A6 and human CYP3A4. The obtained results strongly suggest that the CYP3A active center contains at least two and probably three distinct binding sites for substrates.
More Related Videos
 10:51
Reverse Yeast Two-hybrid System to Identify Mammalian Nuclear Receptor Residues that Interact with Ligands and/or Antagonists
Published on: November 15, 2013
 09:32
Light-mediated Reversible Modulation of the Mitogen-activated Protein Kinase Pathway during Cell Differentiation and Xenopus Embryonic Development
Published on: June 15, 2017
Related Concept Videos
 01:31
Transducer Mechanism: Nuclear Receptors
About 48 different soluble family members of nuclear receptors are identified that can be divided into two main classes:
 01:28
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes
 01:22
Adrenergic Antagonists: Pharmacological Actions of ɑ-Receptor Blockers
α1-blockers: These drugs inhibit α1-adrenoceptors on smooth muscle cells, resulting in vasodilation. This vasodilation lowers blood pressure, making α1-blockers valuable in treating hypertension. Additionally, α1-blockers effectively address urinary obstruction...
 01:26
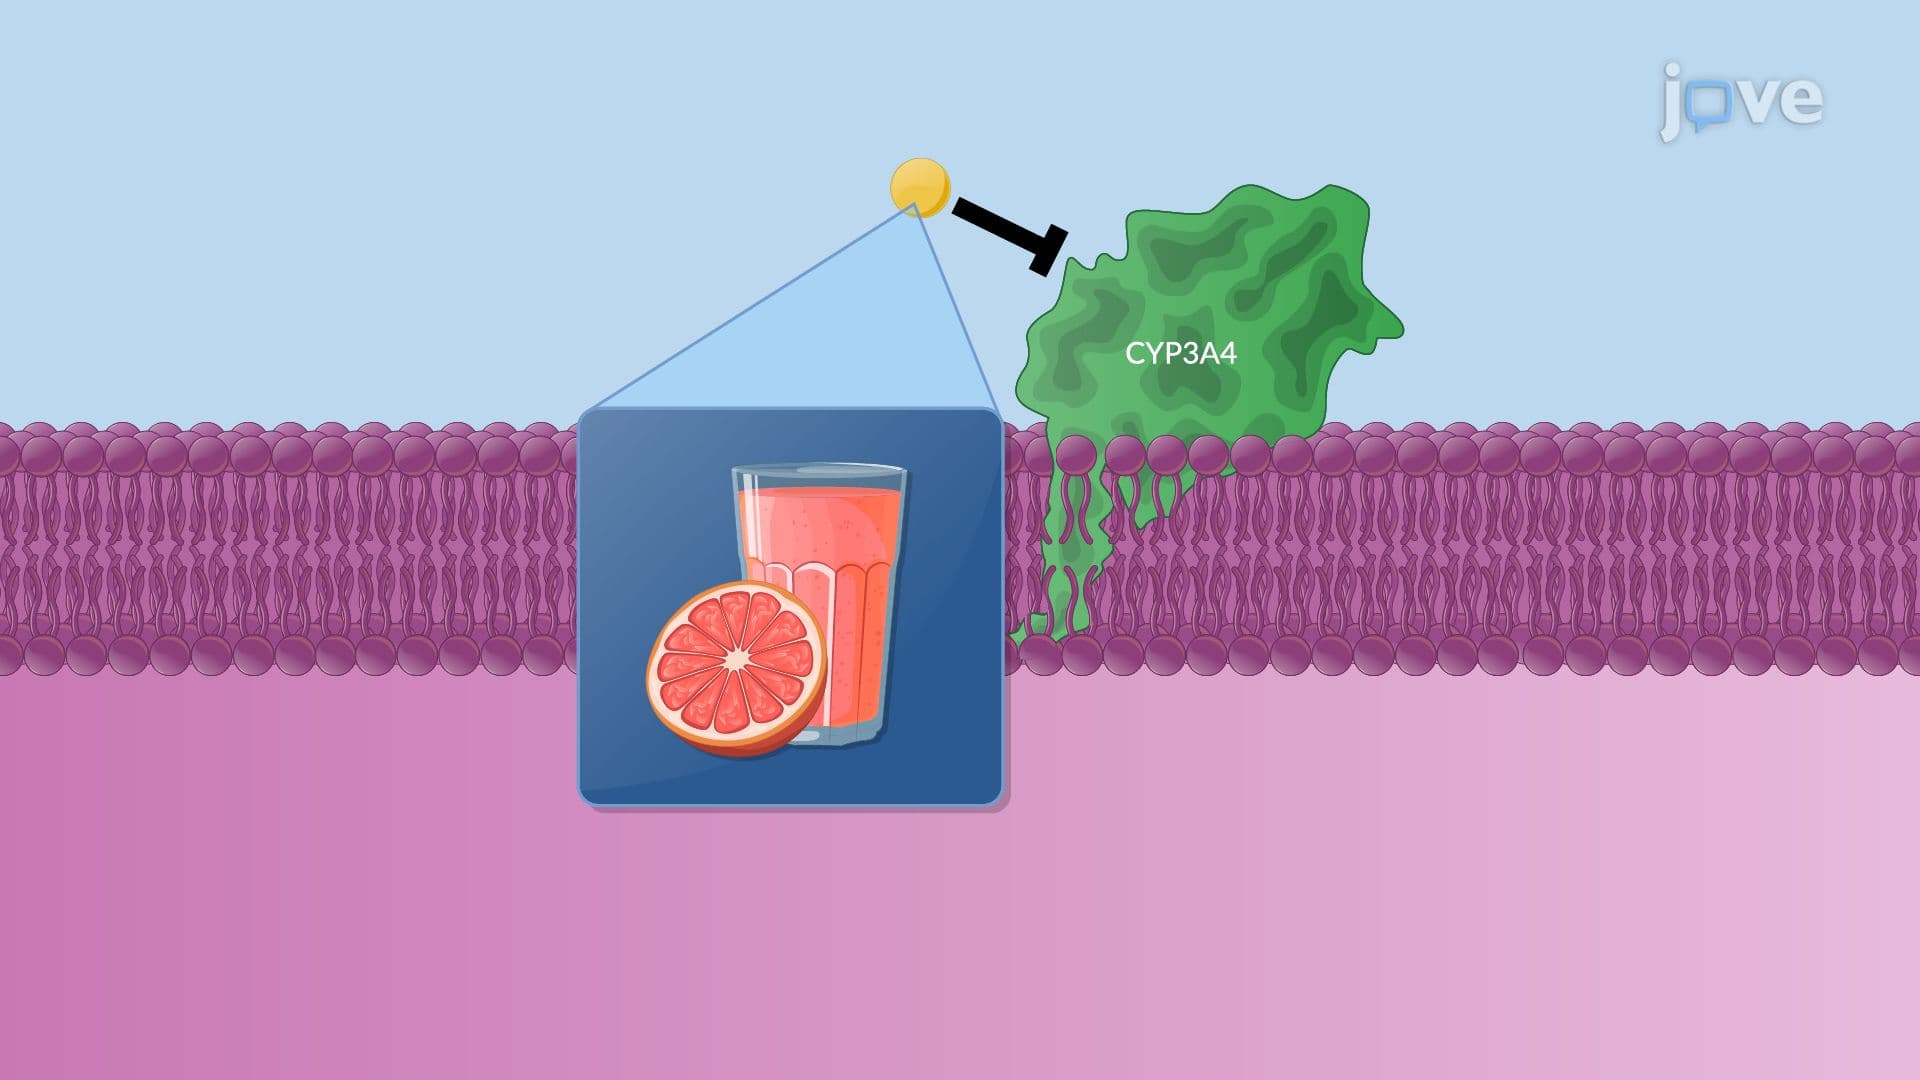
Pharmacokinetics: Drug–Food and Drug–Viral Interactions
 01:17
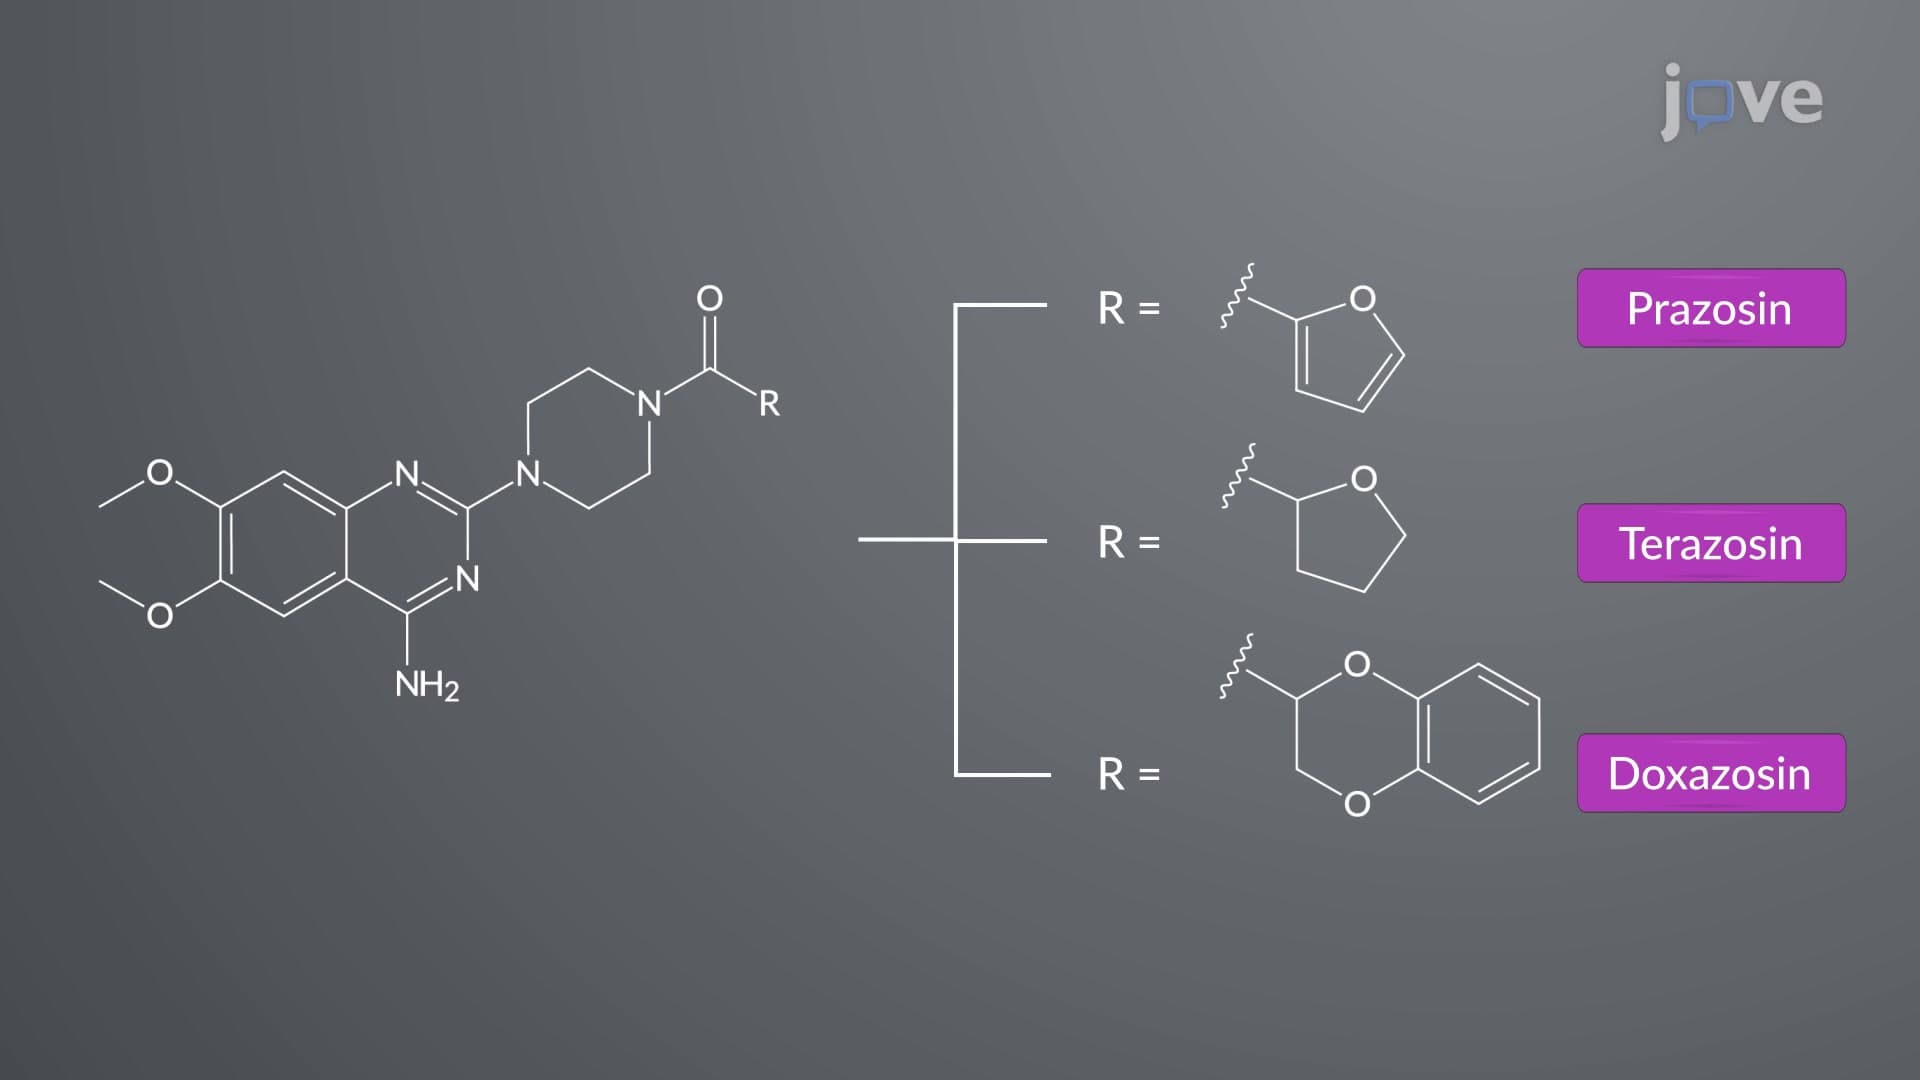
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...