Related Experiment Video
Updated: Oct 3, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Theoretical Modeling of Water Exchange on [Pd(H(2)O)(4)](2+), [Pt(H(2)O)(4)](2+), and trans-[PtCl(2)(H(2)O)(2)]
Robert J. Deeth1, Lars I. Elding
1Inorganic Computational Chemistry Group, Department of Chemistry, University of Warwick, Coventry CV4 7AL, U.K., and Inorganic Chemistry 1, Chemical Center, Lund University, P.O. Box 124, S-221 00 Lund, Sweden.

Abstract:
Density functional theory is applied to modeling the exchange in aqueous solution of H(2)O on [Pd(H(2)O)(4)](2+), [Pt(H(2)O)(4)](2+), and trans-[PtCl(2)(H(2)O)(2)]. Optimized structures for the starting molecules are reported together with trigonal bipyramidal (tbp) systems relevant to an associative mechanism. While a rigorous tbp geometry cannot by symmetry be the actual transition state, it appears that the energy differences between model tbp structures and the actual transition states are small. Ground state geometries calculated via the local density approximation (LDA) for [Pd(H(2)O)(4)](2+) and relativistically corrected LDA for the Pt complexes are in good agreement with available experimental data. Nonlocal gradient corrections to the LDA lead to relatively inferior structures. The computed structures for analogous Pd and Pt species are very similar. The equatorial M-OH(2) bonds of all the LDA-optimized tbp structures are predicted to expand by 0.25-0.30 Å, while the axial bonds change little relative to the planar precursors. This bond stretching in the transition state counteracts the decrease in partial molar volume caused by coordination of the entering water molecule and can explain qualitatively the small and closely similar volumes of activation observed. The relatively higher activation enthalpies of the Pt species can be traced to the relativistic correction of the total energies while the absolute DeltaH() values for exchange on [Pd(H(2)O)(4)](2+) and [Pt(H(2)O)(4)](2+) are reproduced using relativistically corrected LDA energies and a simple Born model for hydration. The validity of the latter is confirmed via some simple atomistic molecular mechanics estimates of the relative hydration enthalpies of [Pd(H(2)O)(4)](2+) and [Pd(H(2)O)(5)](2+). The computed DeltaH() values are 57, 92, and 103 kJ/mol compared to experimental values of 50(2), 90(2), and 100(2) kJ/mol for [Pd(H(2)O)(4)](2+), [Pt(H(2)O)(4)](2+), and trans-[PtCl(2)(H(2)O)(2)], respectively. The calculated activation enthalpy for a hypothetical dissociative water exchange at [Pd(H(2)O)(4)](2+) is 199 kJ/mol. A qualitative analysis of the modeling procedure, the relative hydration enthalpies, and the zero-point and finite temperature corrections yields an estimated uncertainty for the theoretical activation enthalpies of about 15 kJ/mol.
Related Concept Videos
 02:58
Balancing Redox Equations
 02:42
Aqueous Solutions and Heats of Hydration
When ionic compounds dissolve in water, the ions in the solid separate and disperse uniformly throughout the solution because water molecules surround and solvate the ions, reducing the strong electrostatic forces between them. This process...
 01:17
Ion Exchange
 01:16
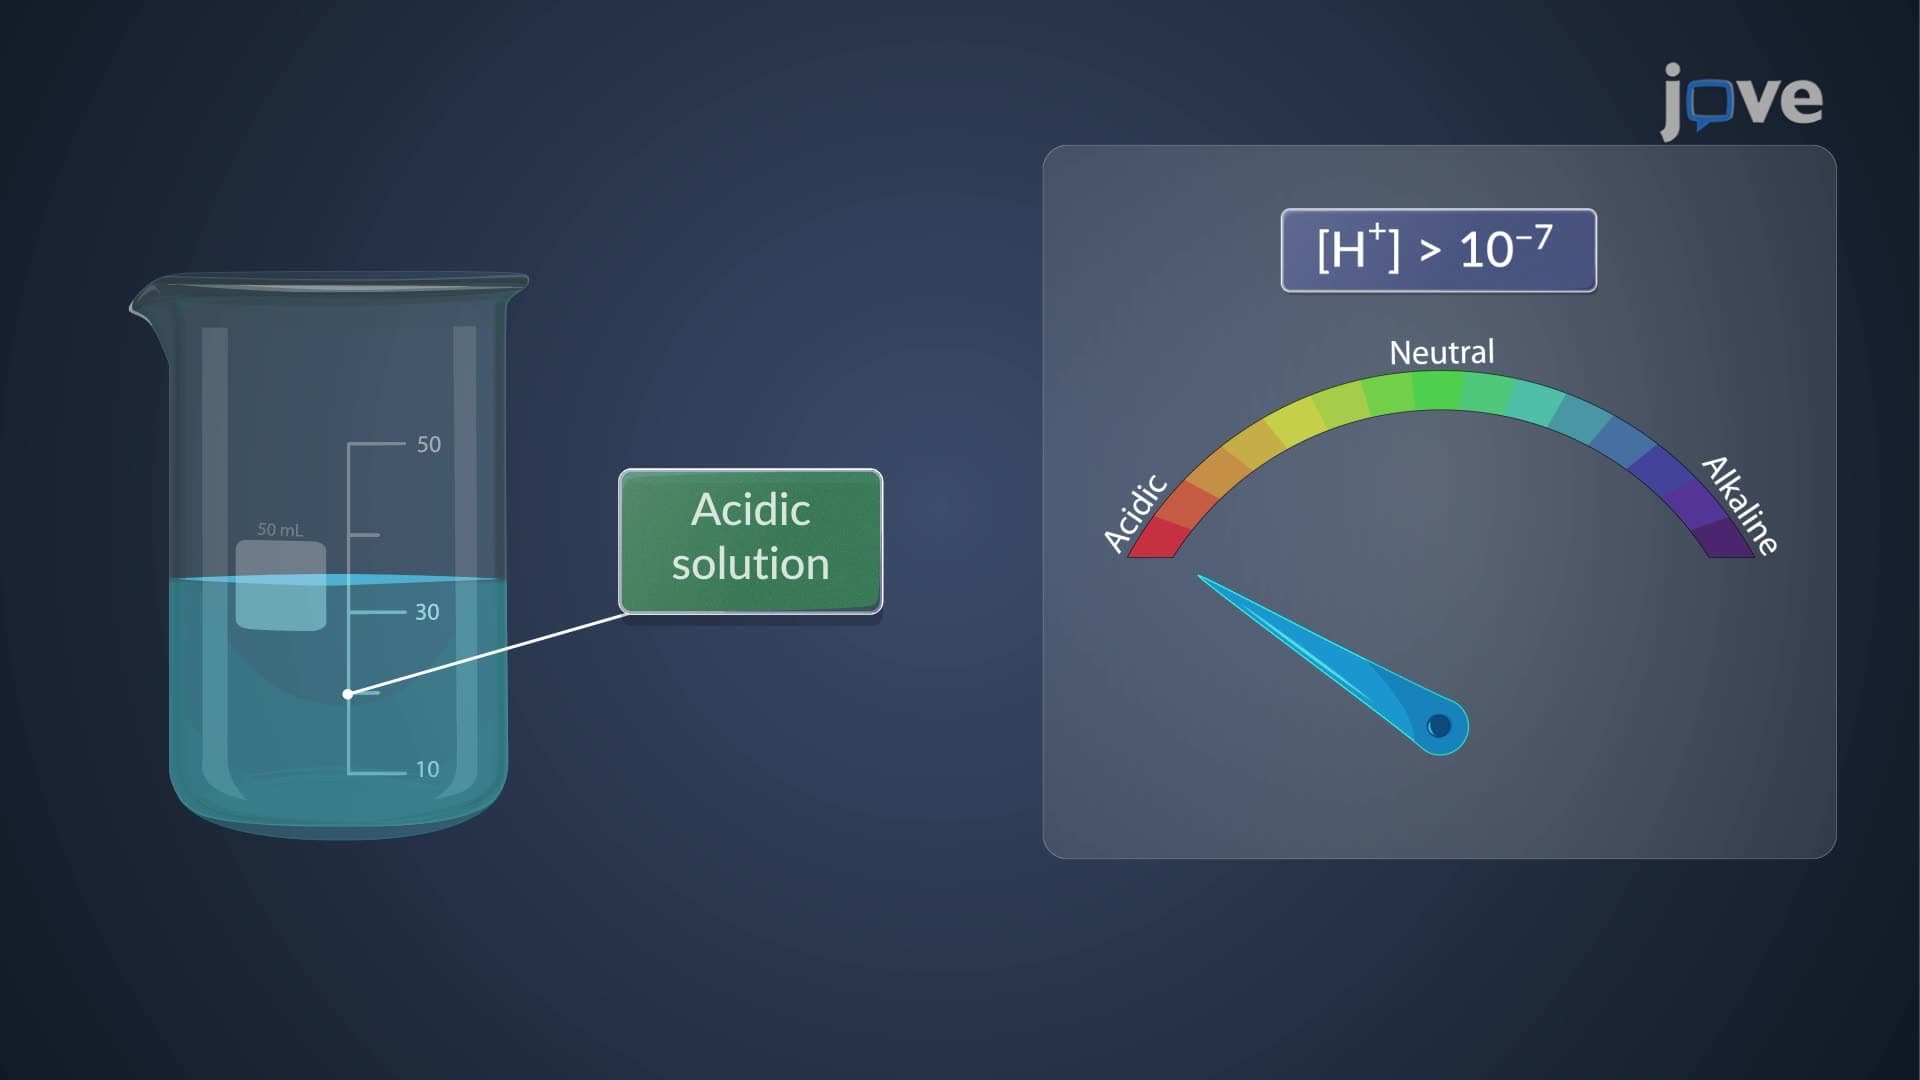
Solubility Equilibria: Ionic Product of Water
The ionic product of water varies with temperature, and its value is 1.0 x 10⁻¹⁴ at standard experimental conditions. Per Le Chatelier's...
 03:13
Intermolecular Forces
 02:30
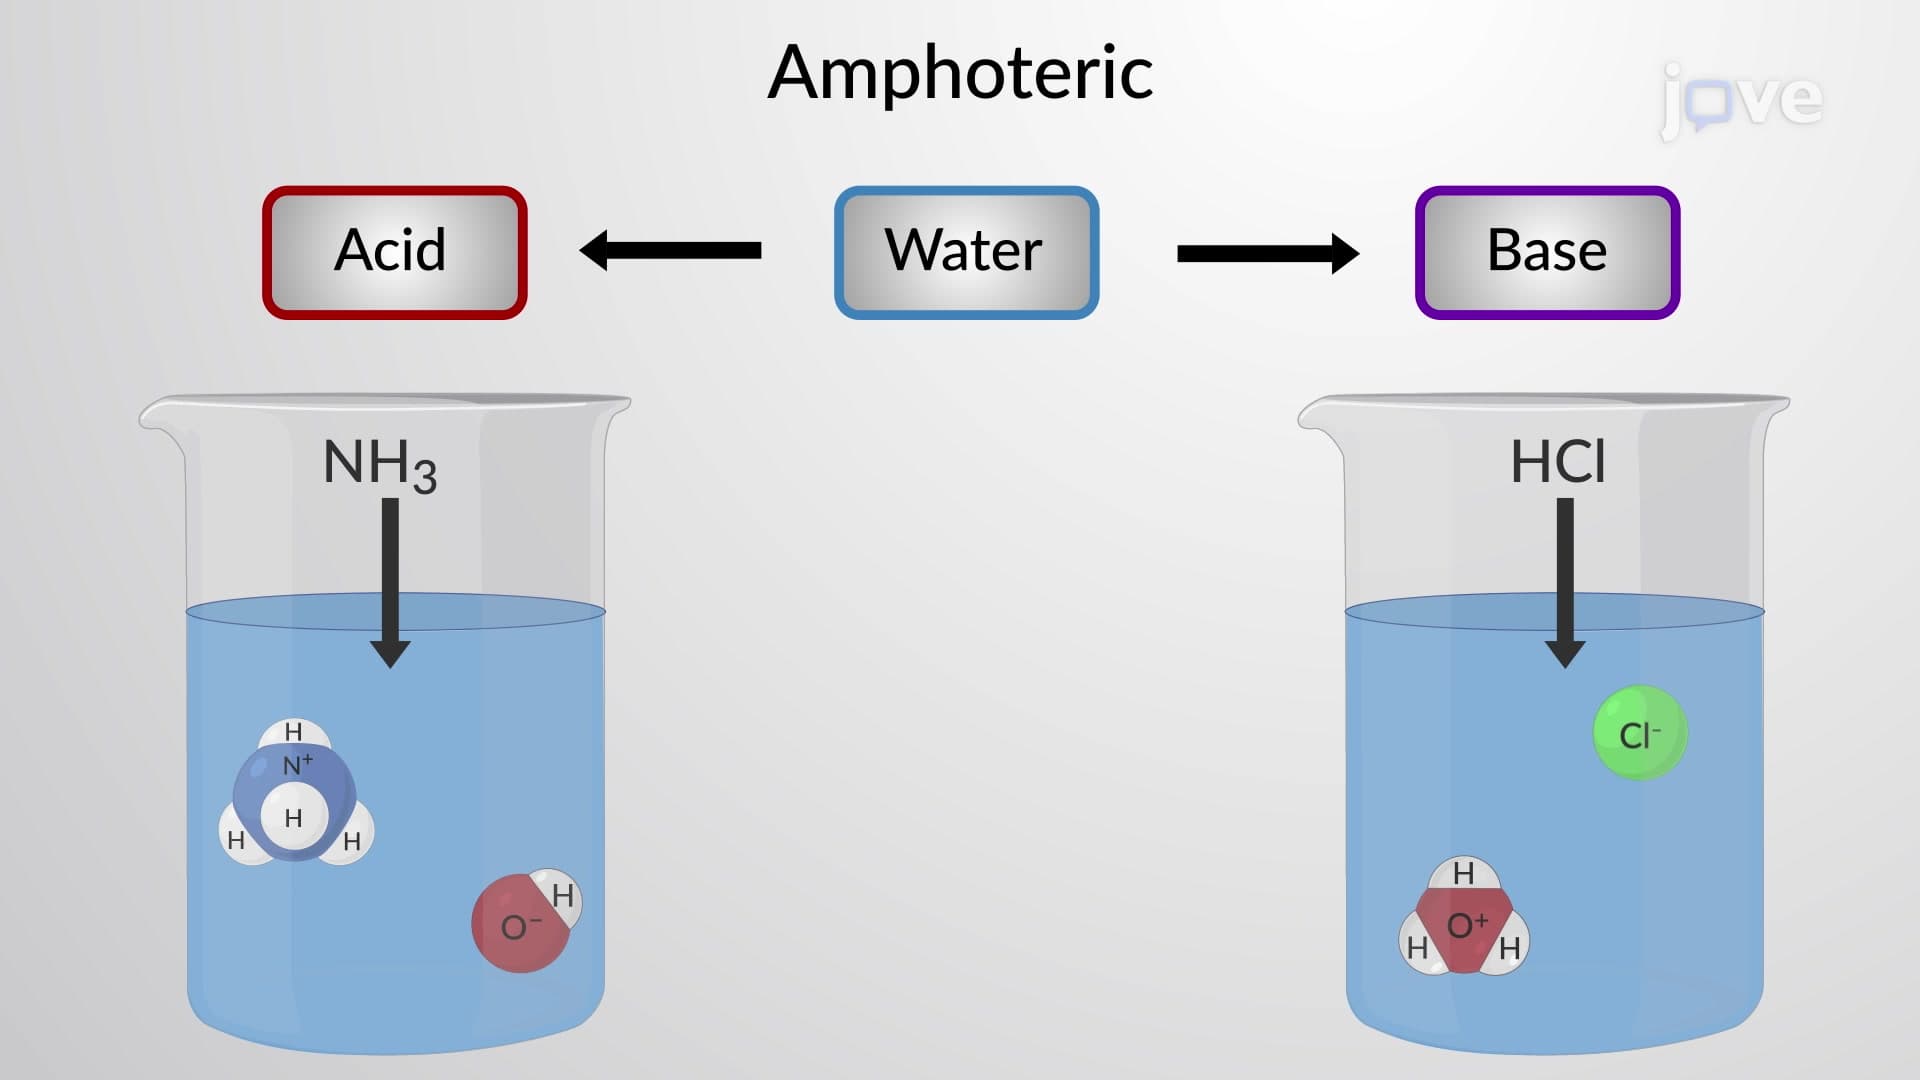
Water: A Bronsted-Lowry Acid and Base