Related Experiment Video
Updated: Jul 5, 2026

Isolating Free Carbenes, their Mixed Dimers and Organic Radicals
Published on: April 19, 2019
Model studies of hydrogen atom addition and abstraction processes involving ortho-, meta-, and para-benzynes
1Department of Chemistry, Indiana University, Bloomington, Indiana 47405-7102, USA.

Abstract:
H-atom addition and abstraction processes involving ortho-, meta-, and para-benzyne have been investigated by multiconfigurational self-consistent field methods. The H(A) + H(B)...H(C) reaction (where r(BC) is adjusted to mimic the appropriate singlet-triplet energy gap) is shown to effectively model H-atom addition to benzyne. The doublet multiconfiguration wave functions are shown to mix the "singlet" and "triplet" valence bond structures of H(B)...H(C) along the reaction coordinate; however, the extent of mixing is dependent on the singlet-triplet energy gap (DeltaE(ST)) of the H(B)...H(C) diradical. Early in the reaction, the ground-state wave function is essentially the "singlet" VB function, yet it gains significant "triplet" VB character along the reaction coordinate that allows H(A)-H(B) bond formation. Conversely, the wave function of the first excited state is predominantly the "triplet" VB configuration early in the reaction coordinate, but gains "singlet" VB character when the H-atom is close to a radical center. As a result, the potential energy surface (PES) for H-atom addition to triplet H(B)...H(C) diradical is repulsive! The H3 model predicts, in agreement with the actual calculations on benzyne, that the singlet diradical electrons are not coupled strongly enough to give rise to an activation barrier associated with C-H bond formation. Moreover, this model predicts that the PES for H-atom addition to triplet benzyne will be characterized by a repulsive curve early in the reaction coordinate, followed by a potential avoided crossing with the (pi)1(sigma*)1 state of the phenyl radical. In contrast to H-atom addition, large activation barriers characterize the abstraction process in both the singlet ground state and first triplet state. In the ground state, this barrier results from the weakly avoided crossing of the dominant VB configurations in the ground-state singlet (S0) and first excited singlet (S1) because of the large energy gap between S0 and S1 early in the reaction coordinate. Because the S1 state is best described as the combination of the triplet X-H bond and the triplet H(B)...H(C) spin couplings, the activation barrier along the S0 abstraction PES will have much less dependence on the DeltaE(ST) of H(B)...H(C) than previously speculated. For similar reasons, the T1 potential surface is quite comparable to the S0 PES.
Related Concept Videos
 02:00
Molecular Models
 03:51
Molecular Orbital Theory II
 01:07
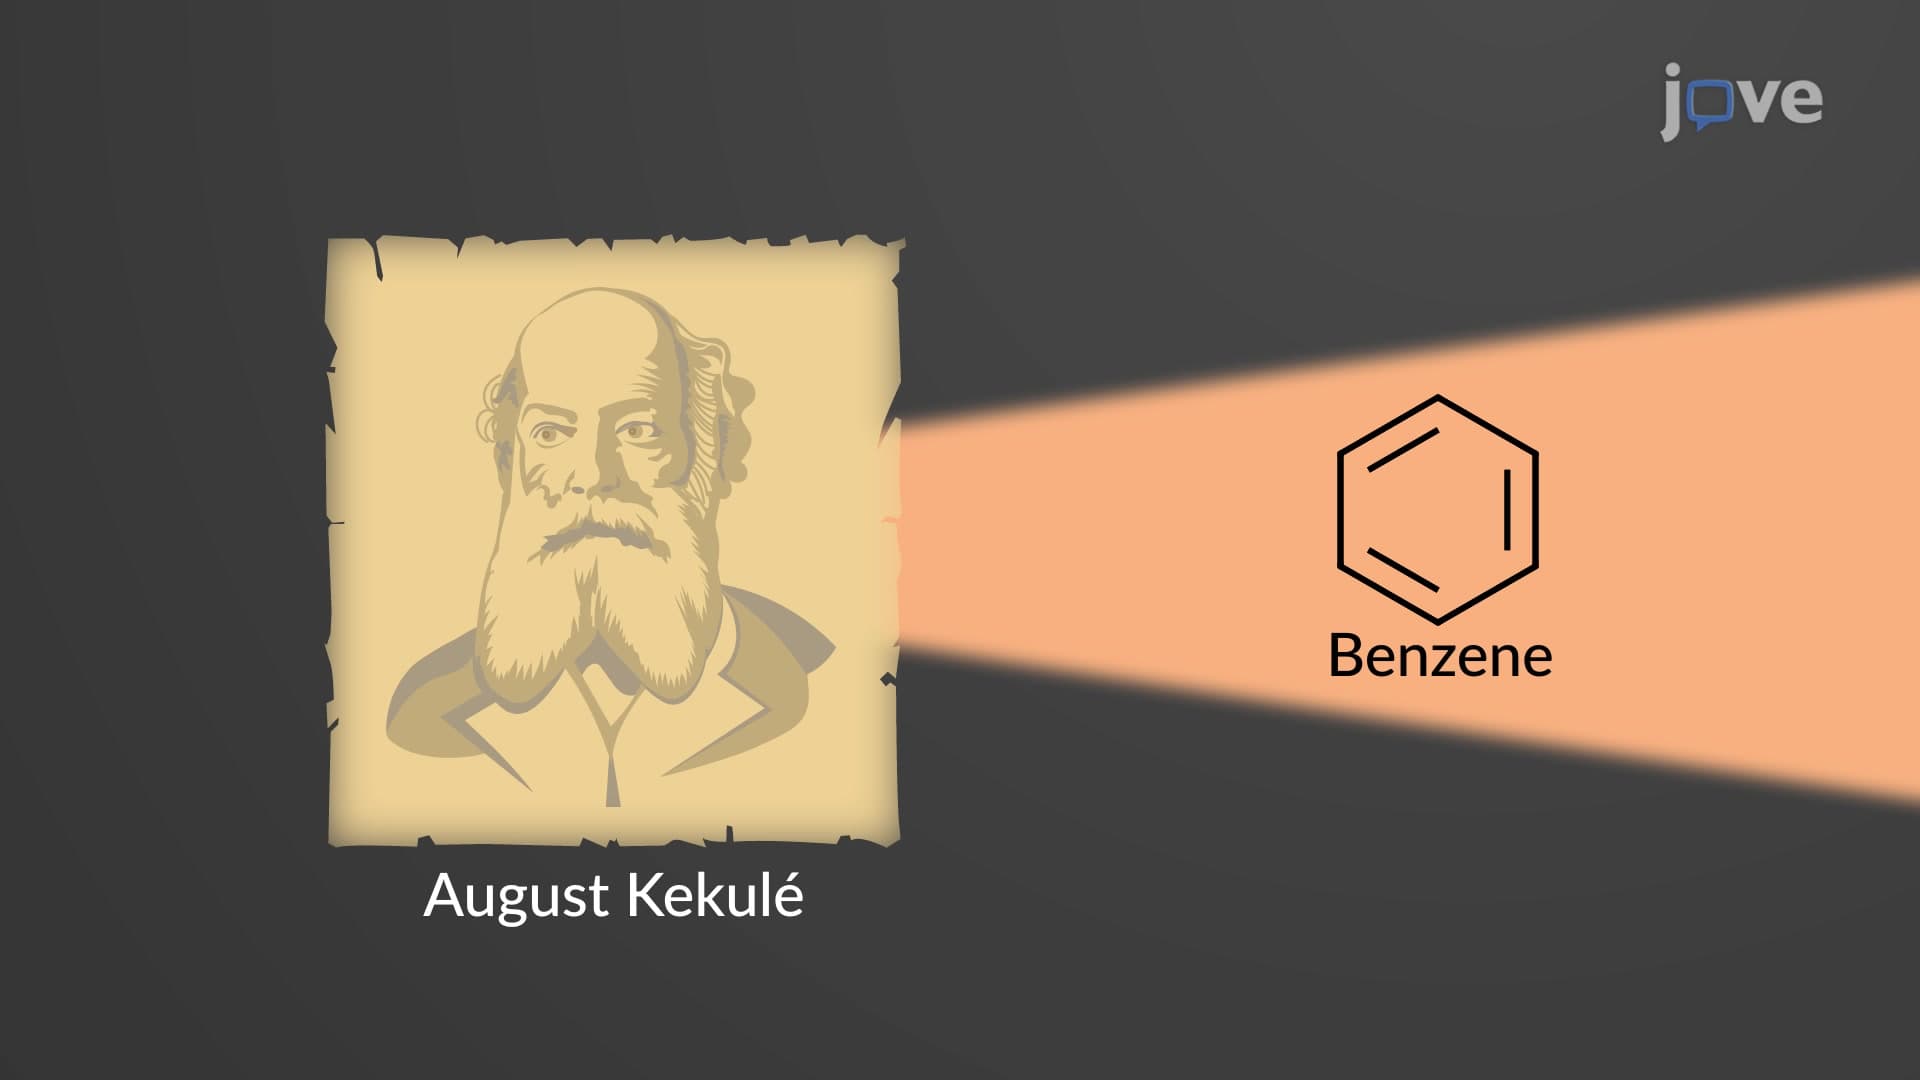
Structure of Benzene: Kekulé Model
He proposed that benzene has a cyclic structure of six carbon atoms attached to one hydrogen atom each, with three alternating pi bonds.
 01:18
Structure of Benzene: Molecular Orbital Model
 01:11
Nomenclature of Aromatic Compounds with Multiple Substituents
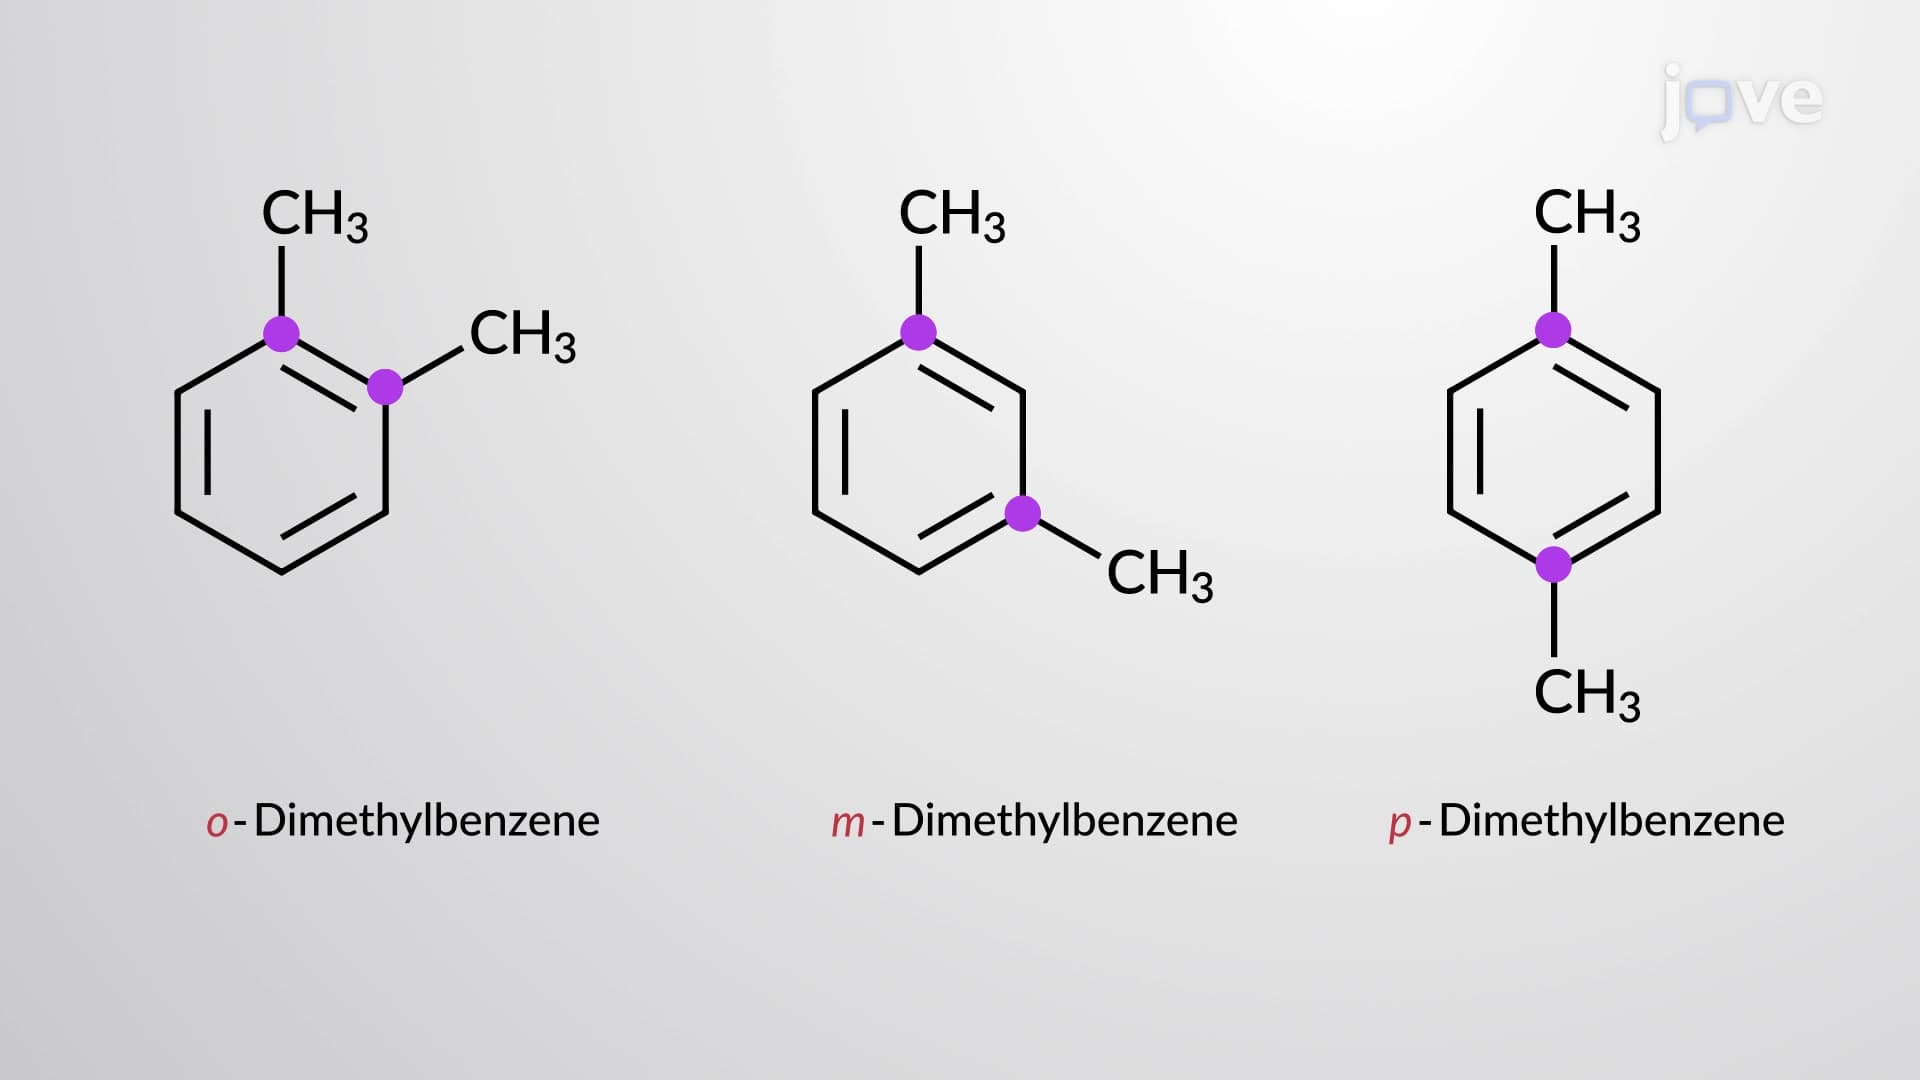
For disubstituted benzene derivatives, with two groups attached to the benzene ring, three constitutional isomers are possible. For example, consider dimethyl benzene, often called xylene, where the second methyl group can be substituted at the second, third, or fourth carbon. The relative position of the substituents is represented by prefixes ortho, meta, or...
 01:37
NMR Spectroscopy of Benzene Derivatives