
Pathogenesis of primary defects in mitochondrial ATP synthesis
E A Schon1, S Santra, F Pallotti
1Columbia University College of Physicians and Surgeons, New York, NY, USA. pas3@columbia.edu

Abstract:
Maternally inherited mutations in the mtDNA-encoded ATPase 6 subunit of complex V (ATP synthase) of the respiratory chain/oxidative phosphorylation system are responsible for a subgroup of severe and often-fatal disorders characterized predominantly by lesions in the brain, particularly in the striatum. These include NARP (neuropathy, ataxia, and retinitis pigmentosa), MILS (maternally inherited Leigh syndrome), and FBSN (familial bilateral striatal necrosis). Of the five known pathogenic mutations causing these disorders, four are located at two codons (156 and 217), each of which can suffer mutations converting a conserved leucine to either an arginine or a proline. Based on the accumulating data on both the structure of ATP synthase and the mechanism by which rotary catalysis couples proton flow to ATP synthesis, we propose a model that may help explain why mutations at codons 156 and 217 are pathogenic.
Related Concept Videos
 02:59
Animal Mitochondrial Genetics
 01:37
Mitochondria
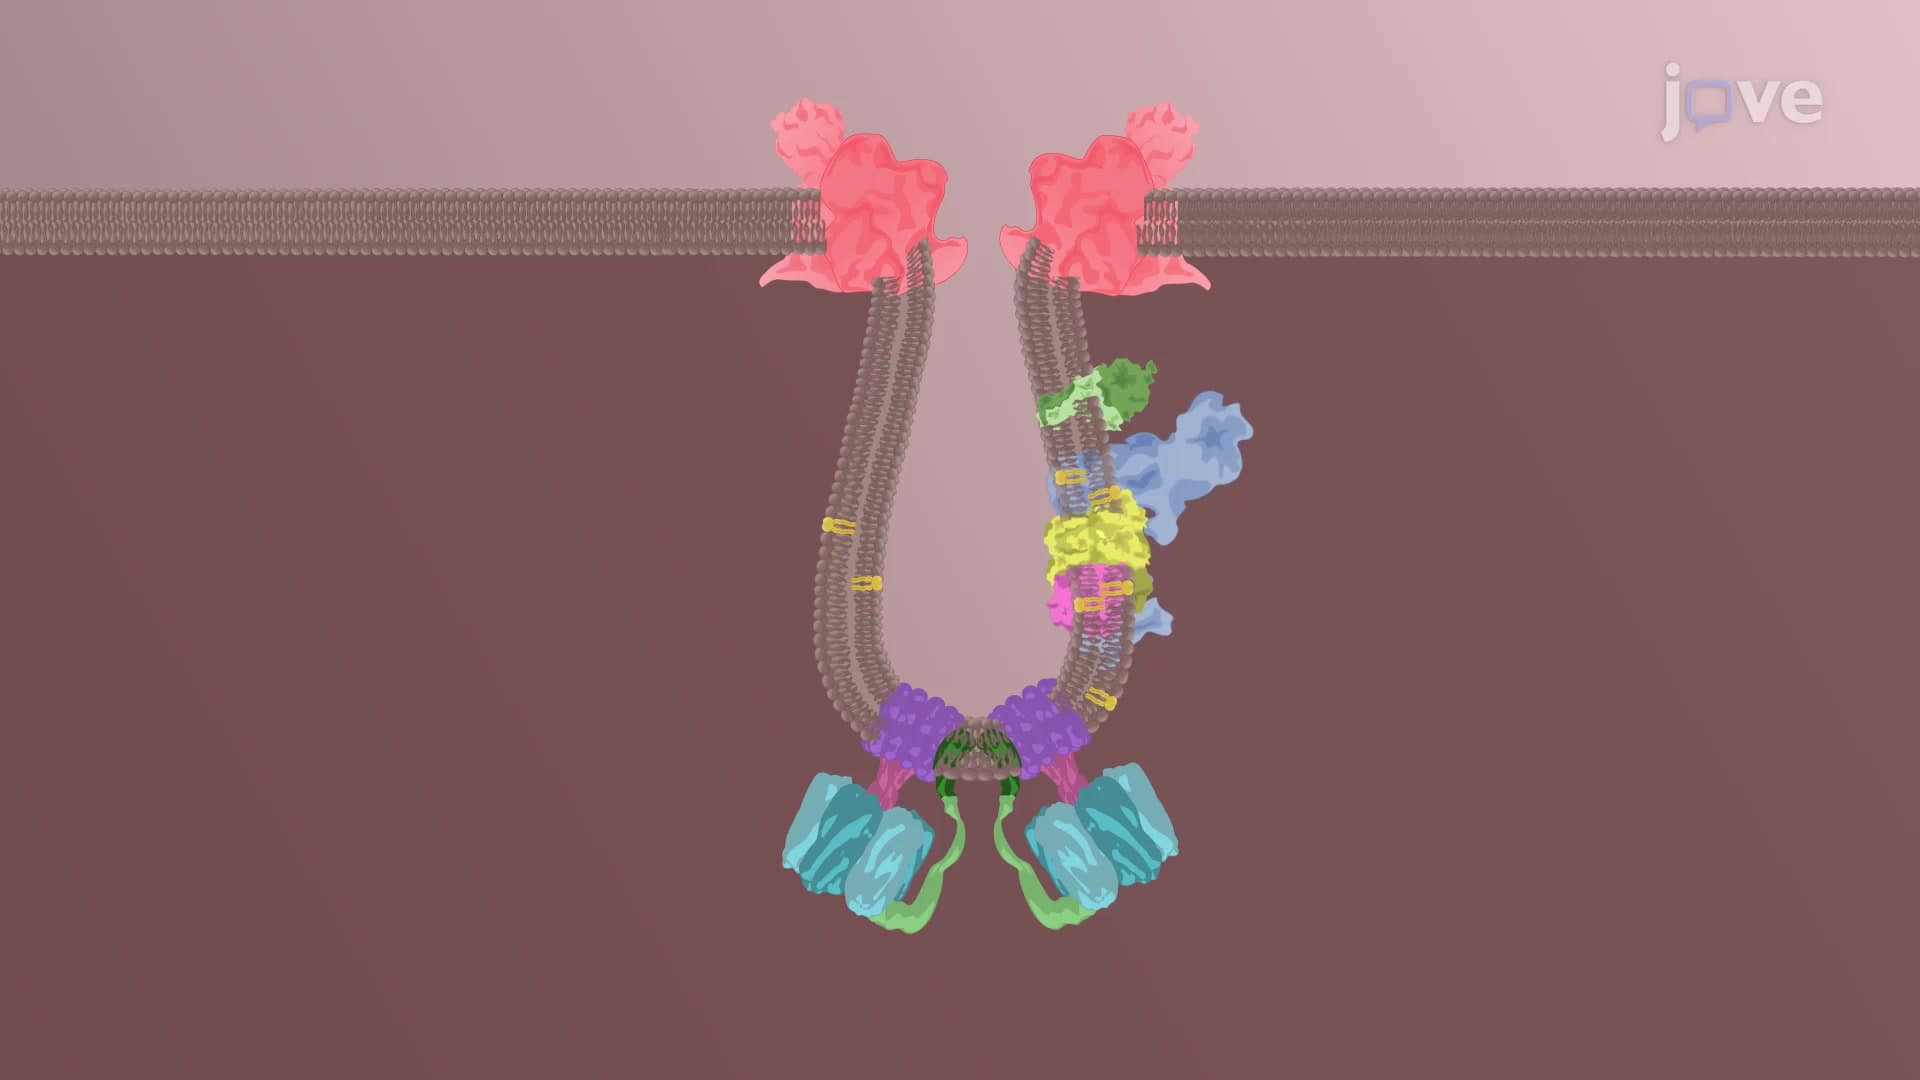 01:28
The Inner Mitochondrial Membrane
 01:48
ATP Synthase: Mechanism
 01:18
ATP Synthase: Structure
 01:19
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...