Related Experiment Video
Updated: Aug 10, 2026
Analyzing Protein Architectures and Protein-Ligand Complexes by Integrative Structural Mass Spectrometry
Published on: October 15, 2018
Hierarchy of simulation models in predicting molecular recognition mechanisms from the binding energy landscapes:
G M Verkhivker1, D Bouzida, D K Gehlhaar
1Agouron Pharmaceuticals, Inc., San Diego, California 92121-1111, USA. verk@agouron.com

Abstract:
Computer simulations using the simplified energy function and simulated tempering dynamics have accurately determined the native structure of the pYVPML, SVLpYTAVQPNE, and SPGEpYVNIEF peptides in the complexes with SH2 domains. Structural and equilibrium aspects of the peptide binding with SH2 domains have been studied by generating temperature-dependent binding free energy landscapes. Once some native peptide-SH2 domain contacts are constrained, the underlying binding free energy profile has the funnel-like shape that leads to a rapid and consistent acquisition of the native structure. The dominant native topology of the peptide-SH2 domain complexes represents an extended peptide conformation with strong specific interactions in the phosphotyrosine pocket and hydrophobic interactions of the peptide residues C-terminal to the pTyr group. The topological features of the peptide-protein interface are primarily determined by the thermodynamically stable phosphotyrosyl group. A diversity of structurally different binding orientations has been observed for the amino-terminal residues to the phosphotyrosine. The dominant native topology for the peptide residues carboxy-terminal to the phosphotyrosine is tolerant to flexibility in this region of the peptide-SH2 domain interface observed in equilibrium simulations. The energy landscape analysis has revealed a broad, entropically favorable topology of the native binding mode for the bound peptides, which is robust to structural perturbations. This could provide an additional positive mechanism underlying tolerance of the SH2 domains to hydrophobic conservative substitutions in the peptide specificity region.
More Related Videos
 06:50
Computational Prediction of Amino Acid Preferences of Potentially Multispecific Peptide-Binding Domains Involved in Protein-Protein Interactions
Published on: January 26, 2024
 10:01
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Related Concept Videos
 02:00
Molecular Models
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:04
Protein-protein Interfaces
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:18
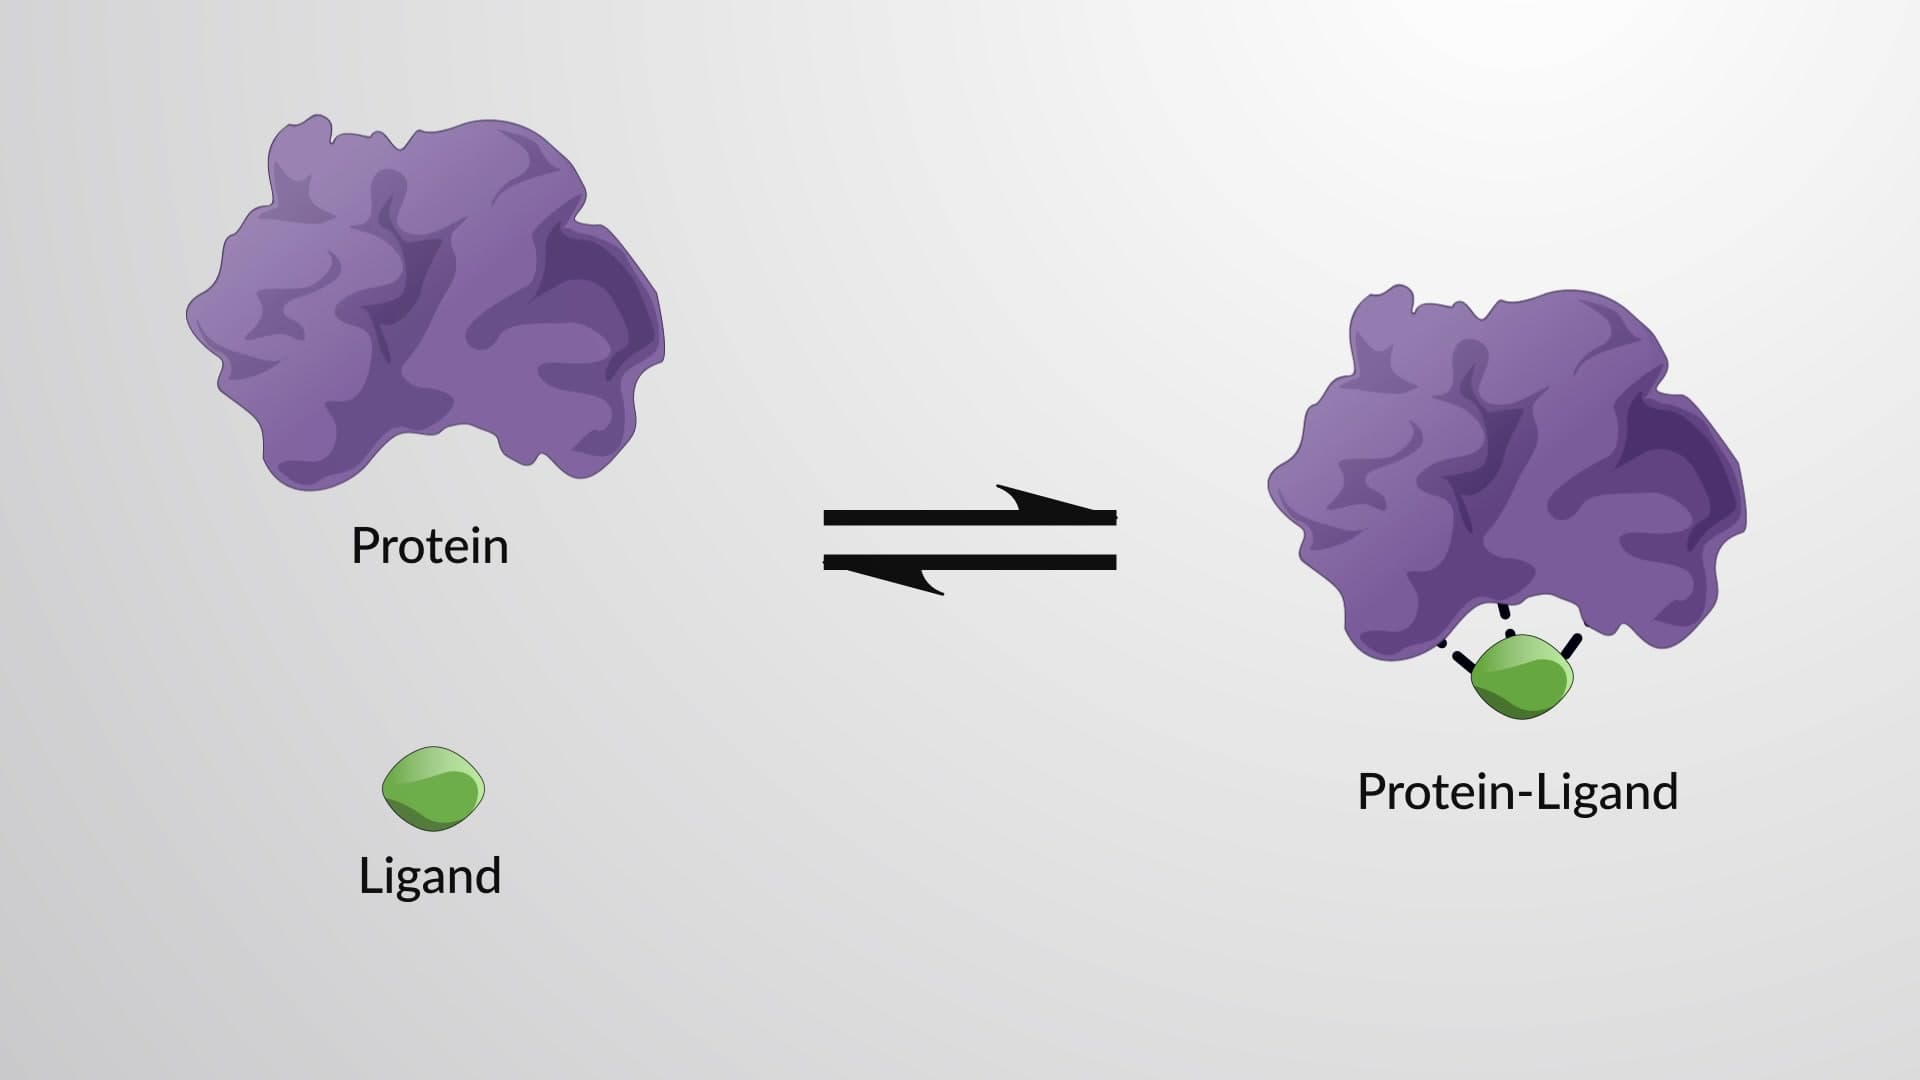
The Equilibrium Binding Constant and Binding Strength
 01:30
Assembly of Signaling Complexes
Interaction domains in cell signaling
Interaction domains recognize exposed features of their binding partners containing post-translationally modified sequences,...