Related Experiment Video
Updated: Oct 2, 2026

Purification of the Cystic Fibrosis Transmembrane Conductance Regulator Protein Expressed in Saccharomyces cerevisiae
Published on: May 10, 2014
Distribution and function of the peptide transporter PEPT2 in normal and cystic fibrosis human lung
D A Groneberg1, P R Eynott, F Döring
1Department of Pediatric Pneumology and Immunology, Charité Campus Virchow, Humboldt University, Augustenburger Platz 1, D-13353 Berlin, Germany.

Background:
Aerosol administration of peptide based drugs has an important role in the treatment of various pulmonary and systemic diseases. The characterisation of pulmonary peptide transport pathways can lead to new strategies in aerosol drug treatment.
Methods:
Immunohistochemistry and ex vivo uptake studies were established to assess the distribution and activity of the beta-lactam transporting high affinity proton coupled peptide transporter PEPT2 in normal and cystic fibrosis human airway tissue.
Results:
PEPT2 immunoreactivity in normal human airways was localised to cells of the tracheal and bronchial epithelium and the endothelium of small vessels. In peripheral lung immunoreactivity was restricted to type II pneumocytes. In sections of cystic fibrosis lung a similar pattern of distribution was obtained with signals localised to endothelial cells, airway epithelium, and type II pneumocytes. Functional ex vivo uptake studies with fresh lung specimens led to an uptake of the fluorophore conjugated dipeptide derivative D-Ala-L-Lys-AMCA into bronchial epithelial cells and type II pneumocytes. This uptake was competitively inhibited by dipeptides and cephalosporins but not ACE inhibitors, indicating a substrate specificity as described for PEPT2.
Conclusions:
These findings provide evidence for the expression and function of the peptide transporter PEPT2 in the normal and cystic fibrosis human respiratory tract and suggest that PEPT2 is likely to play a role in the transport of pulmonary peptides and peptidomimetics.
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:06
Carrier-Mediated Transport
Active transport involves two types of membrane-spanning transporters: uptake and efflux. Uptake transporters are expressed in the small...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:23
Transcellular Transport of Solutes
 01:31
Membrane Transporters
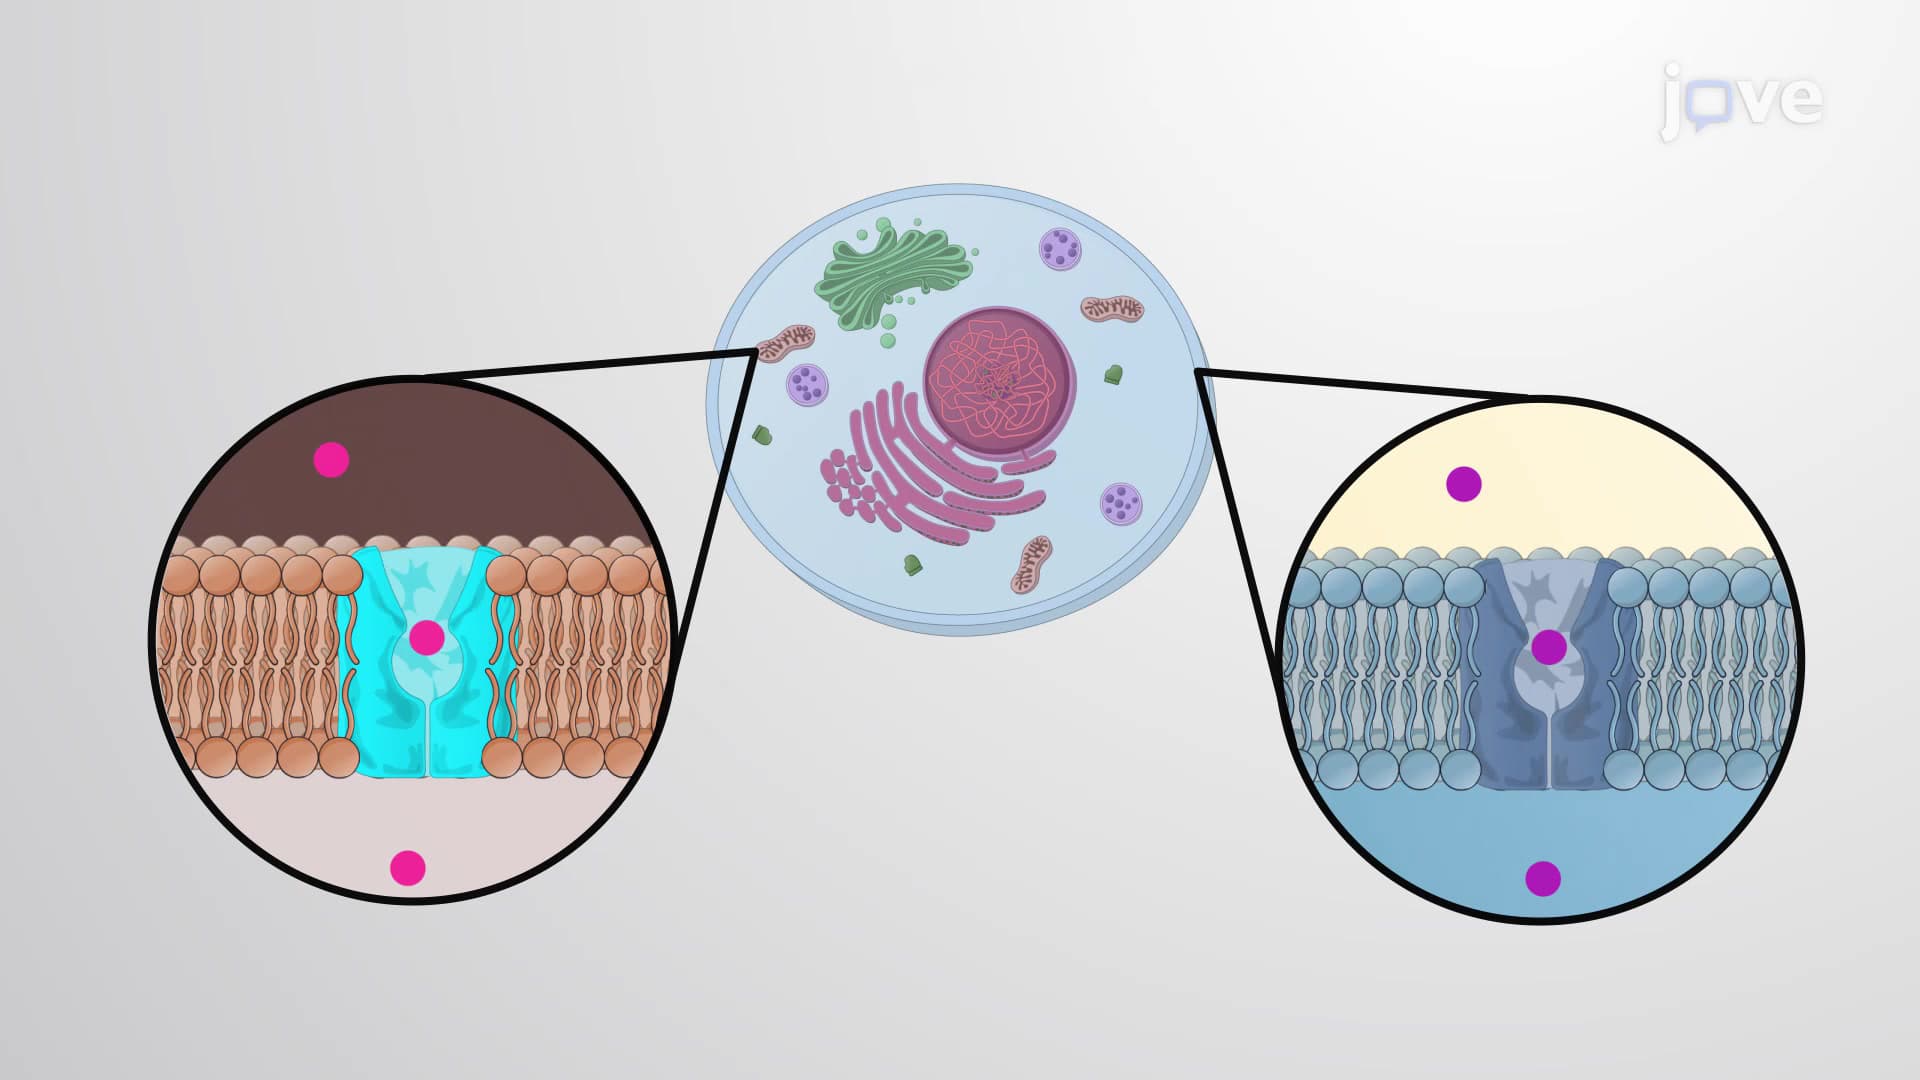
Transporters are mainly composed of alpha-helices, built from bundles of ten or more helices traversing the plasma membrane. The solute-binding sites are located midway, where some of the helices are broken or distorted, making space for the binding site through...
 01:44
The Significance of Membrane Transport
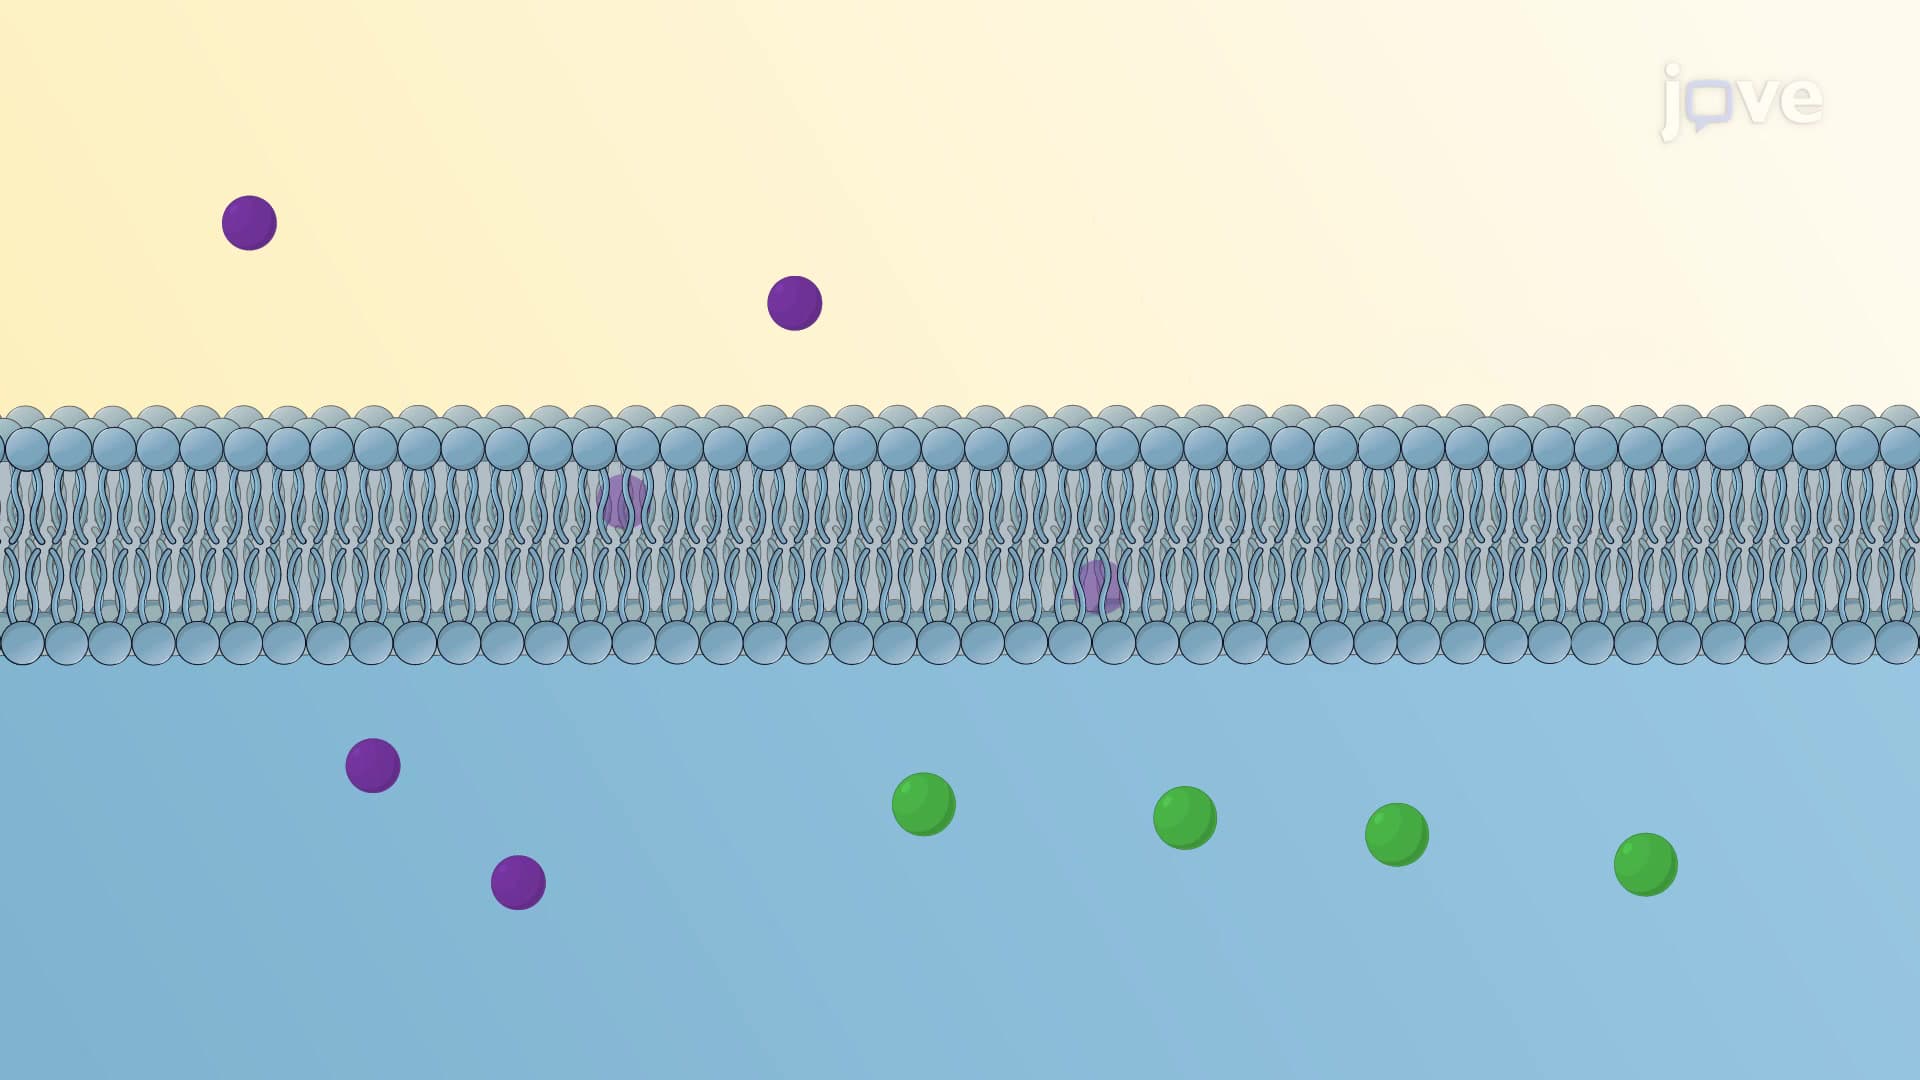
Transporters facilitate either an active or passive movement of solutes. They can allow a single-molecule transport down its...