Related Experiment Video
Updated: Sep 21, 2026

Measuring Biomolecular DSC Profiles with Thermolabile Ligands to Rapidly Characterize Folding and Binding Interactions
Published on: November 21, 2017
Influence of the ORM1 phenotypes on serum unbound concentration and protein binding of quinidine
Jin Heng Li1, Jing Qiu Xu, Xiao Mei Cao
1Department of Clinical Pharmacology, Jinling Hospital, Clinical School of Medical College, Nanjing University, 305 East Zhongshan Road, 210002, Nanjing, PR China. ljhengyr@jlonline.com

Background:
Only unbound or free drug in plasma can be transported to its site of action. The fraction of unbound drug in plasma varies widely for highly bound drugs among individuals. The genetic polymorphism of orosomucoid (ORM) could be related to the interindividual variability in plasma binding of basic drugs, as ORM is the transport protein for these drugs in plasma. The ORM is a major binding protein in plasma for various basic drugs and is coded by two loci, ORM1 and ORM2, which are closely linked on chromosome 9q31-->34.1. ORM1 locus is highly polymorphic and the ORM2 locus is monomorphic in most population.
Methods:
Twenty-eight healthy volunteers were selected with three ORM1 phenotypes, containing homozygotes ORM1 F1 (n=10) and ORM1 S (n=8), and heterozygote ORM1 F1S (n=10), identified by isoelectric focusing on polyacrylamide gels followed immunoblotting after desialylation of sera. After a single oral dose of quinidine 200 mg, serum total (HPLC) and unbound concentrations in ultrafiltrate (ultrafiltration/HPLC) were determined, and the pharmacokinetic parameters and protein binding rate were calculated.
Results:
Serum concentrations of ORM (553.8-573.2 mg/l) and albumin proteins (57.5-58.4 mg/l) were similar in the three groups (P>0.05). Unbound quinidine concentration in ORM1 F1 phenotype subjects was higher than that in ORM1 S and ORM1 F1S phenotype; the free drug percentage for the subjects with ORM1 F1 phenotype (19.79%) was twice as high as that with ORM1 S phenotype (10.96%) (P<0.01) at 24 h after administration of oral quinidine when the state of disposition equilibrium occurred. The elimination t(1/2) values and the other pharmacokinetic parameters of quinidine were not affected by the different ORM1 phenotypes.
Conclusions:
Different ORM1 phenotypes may affect the disposition of quinidine, a basic drug, rather than its hepatic metabolism and elimination. The functional heterogeneity of ORM1 could be responsible for the differences in plasma binding of quinidine. Therefore, monitoring of the unbound quinidine concentration would be important for the patients with different ORM1 phenotypes who are treated with quinidine.
Related Concept Videos
 01:29
Drug Distribution: Plasma Protein Binding
 01:29
Factors Affecting Protein-Drug Binding: Patient-Related Factors
Age stands as a key determinant in protein-drug binding. Neonates, characterized by low albumin content, experience heightened concentrations of unbound drugs such as phenytoin and...
 01:09
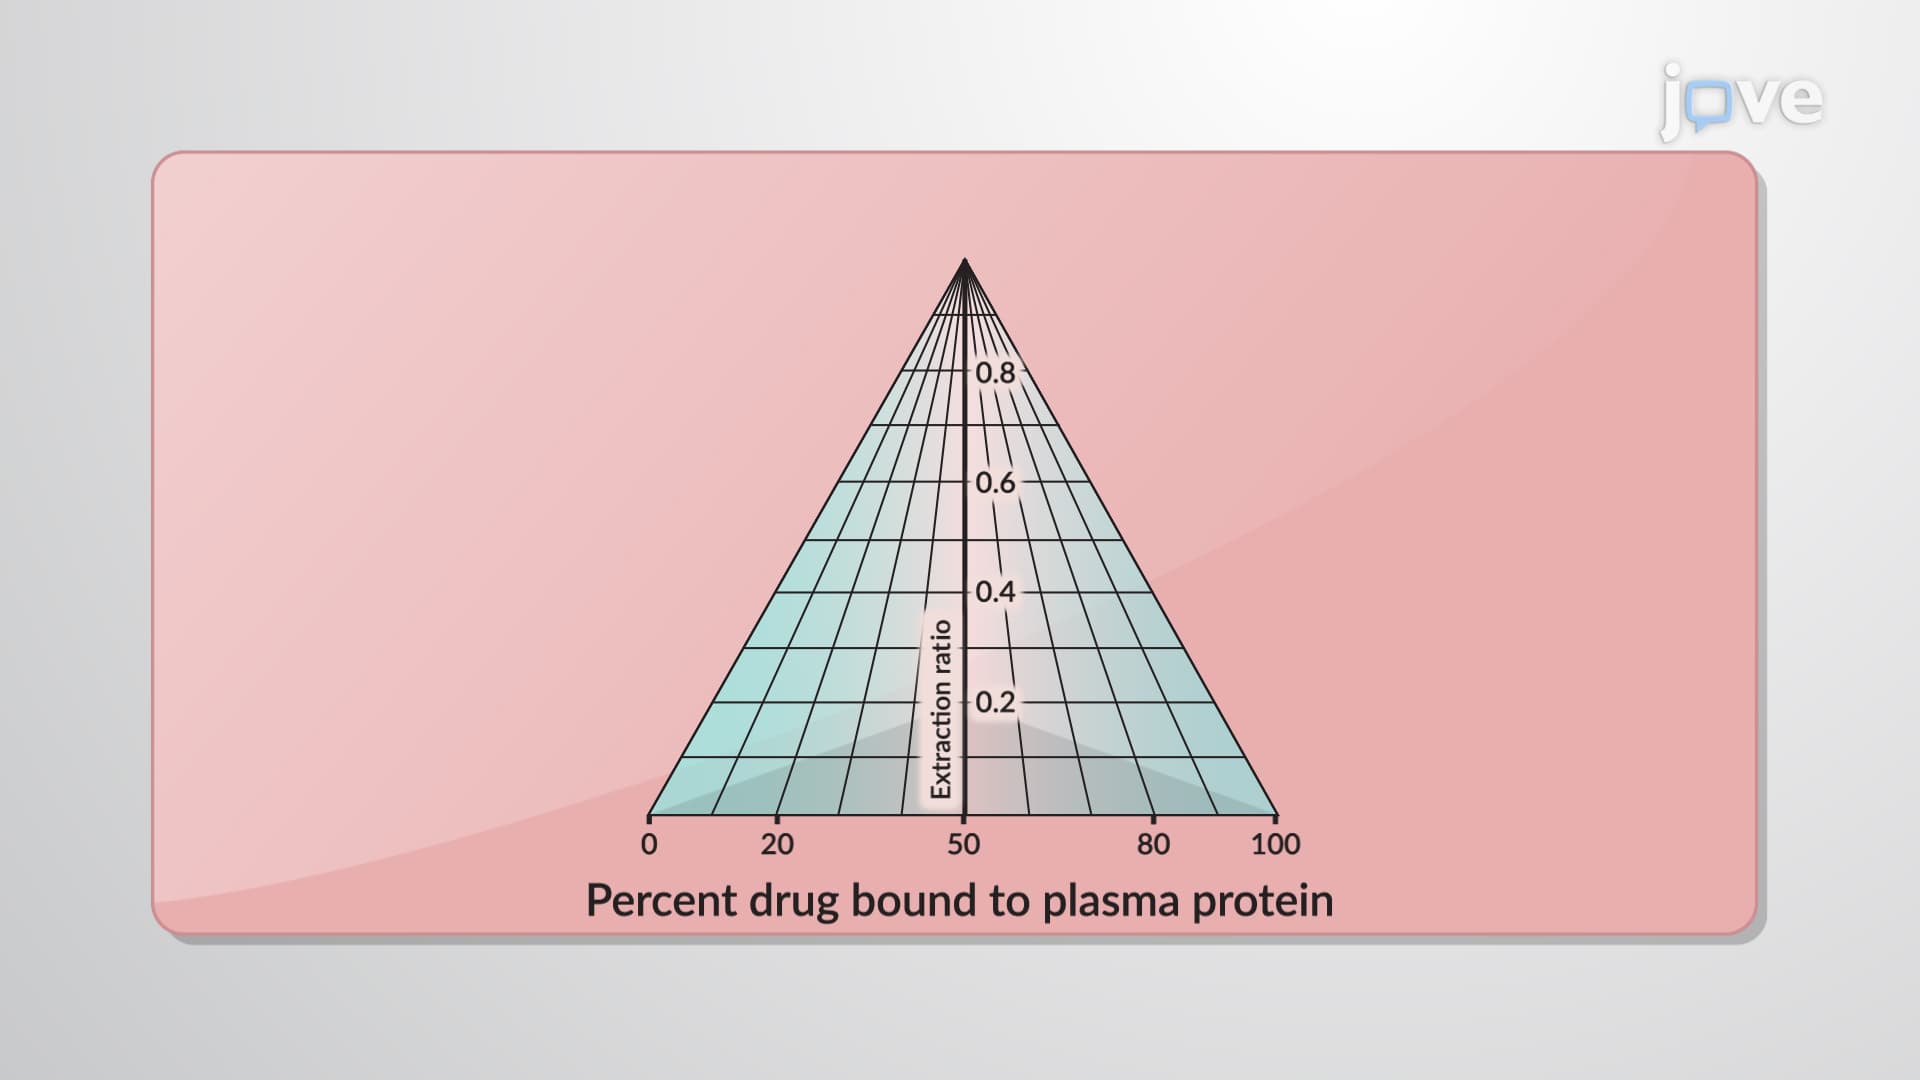
Hepatic Drug Clearance: Effect of Protein Binding
For low-extraction-ratio drugs that are less than 80% protein-bound, minor changes in protein binding...
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 01:29
Pharmacogenetic Phenotypes: Alterations in Pharmacokinetics, Drug Targets and Biologic Milieu
 01:15
Pharmacodynamic Models: Linear Concentration–Effect Model