Structure solution of the basic decagonal Al-Co-Ni phase by the atomic surfaces modelling method
Related Experiment Video
Updated: Oct 2, 2026
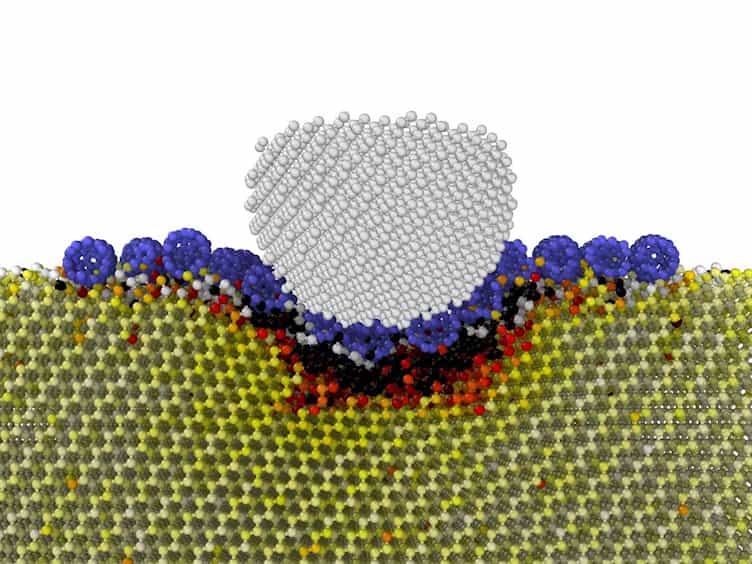
Probing C84-embedded Si Substrate Using Scanning Probe Microscopy and Molecular Dynamics
Published on: September 28, 2016
Antonio Cervellino1, Torsten Haibach, Walter Steurer
1Laboratory of Crystallography, Swiss Federal Institute of Technology (ETHZ) and University of Zurich, Zurich, Switzerland.

Abstract:
The atomic surfaces modelling technique has been used to solve the structure of the basic Ni-rich Al-Co-Ni decagonal phase. Formula Al70.6Co6.7Ni22.7, space group P10, five-dimensional unit-cell parameters: d1 = d4 = 4.752 (3) A, d2 = d3 = 3.360 (2) A, d5 = 8.1710 (2) A; alpha12 = alpha34 = 69.295 degrees, alpha13 = alpha24 = 45 degrees, alpha14 = 41.410 degrees, alpha23 = alphai5 = 90 degrees (i = 1-4), V = 291.2 (7) A5; D(x) = 3.887 Mg x m(-3). Refinement based on /F/; 2767 unique reflections (/F/ > 0), 749 parameters, R = 0.17, wR = 0.06. Describing the structure of quasicrystals embedded in n-dimensional superspace in principle takes advantage of n-dimensional periodicity to select the minimal set of degrees of freedom for the structure. The method of modelling of the atomic surfaces yielded the first fully detailed structure solution of this phase. Comparison with numerous former, less accurate models confirms several features already derived, but adds a new essential insight of the structure and its complexity. The atoms fill the space forming recurrent structure motifs, which we will (generically) refer to as clusters. However, no unique cluster exists, although differences are small. Each cluster shows a high degree of structural disorder. This gives rise to a large configurational entropy, as much as expected in a phase which is stable at high temperature. On the other side, the cluster spatial arrangement is perfectly quasiperiodic. These considerations, corroborated by analysis of the structural relationship with neighbouring periodic phases, strongly suggest the existence of a non-local, long-range interaction term in the total energy which may be essential to the stability.
Related Concept Videos
 02:37
Metallic Solids
All metallic solids exhibit high thermal and electrical conductivity, metallic luster, and malleability. Many...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:42
Ionic Crystal Structures
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 02:06
Newman Projections
The organic molecules rotate across the single bonds leading to numerous temporary three-dimensional structures of varying energy known as conformers.
 02:22
Structures of Solids
 02:33
Lattice Centering and Coordination Number
Types of Unit Cells
Imagine taking a large number of identical...