Related Experiment Video
Updated: Oct 2, 2026
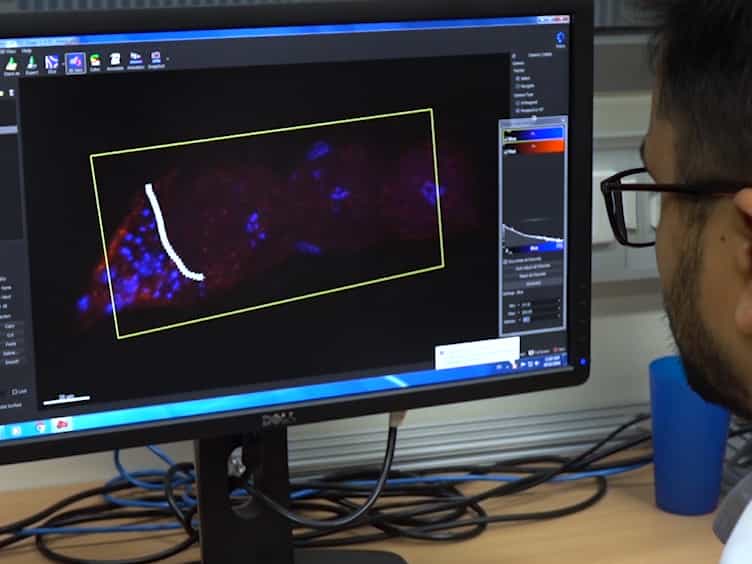

Computational Analysis of the Caenorhabditis elegans Germline to Study the Distribution of Nuclei, Proteins, and the Cytoskeleton
Published on: April 19, 2018
The Caenorhabditis elegans mucolipin-like gene cup-5 is essential for viability and regulates lysosomes in multiple
Bradley M Hersh1, Erika Hartwieg, H Robert Horvitz
1Howard Hughes Medical Institute, Department of Biology, 68-425, Massachusetts Institute of Technology, Cambridge, MA 02139, USA.

Abstract:
The misregulation of programmed cell death, or apoptosis, contributes to the pathogenesis of many diseases. We used Nomarski microscopy to screen for mutants containing refractile cell corpses in a C. elegans strain in which all programmed cell death is blocked and such corpses are absent. We isolated a mutant strain that accumulates refractile bodies resembling irregular cell corpses. We rescued this mutant phenotype with the C. elegans mucolipidosis type IV (ML-IV) homolog, the recently identified cup-5 (coelomocyte-uptake defective) gene. ML-IV is a human autosomal recessive lysosomal storage disease characterized by psychomotor retardation and ophthalmological abnormalities. Our null mutations in cup-5 cause maternal-effect lethality. In addition, cup-5 mutants contain excess lysosomes in many and possibly all cell types and contain lamellar structures similar to those observed in ML-IV cell lines. The human ML-IV gene is capable of rescuing both the maternal-effect lethality and the lysosome-accumulation abnormality of cup-5 mutants. cup-5 mutants seem to contain excess apoptotic cells as detected by staining with terminal deoxynucleotidyltransferase-mediated dUTP nick end labeling. We suggest that the increased apoptosis seen in cup-5 mutants is a secondary consequence of the lysosomal defect, and that abnormalities in apoptosis may be associated with human lysosomal storage disorders.
Insights
Researchers identified a C. elegans gene, cup-5, linked to lysosomal storage diseases like mucolipidosis type IV (ML-IV). cup-5 mutants show increased cell death and lysosome accumulation, suggesting a connection between lysosomal defects and apoptosis in disease.
Area of Science:
- Cell Biology
- Genetics
- Developmental Biology
Background:
- Misregulation of programmed cell death (apoptosis) is implicated in various diseases.
- Mucolipidosis type IV (ML-IV) is a human lysosomal storage disorder with neurological and ocular symptoms.
Purpose of the Study:
- To identify genes involved in apoptosis regulation using a C. elegans screening method.
- To investigate the function of the C. elegans cup-5 gene, a homolog of the human ML-IV gene.
Main Methods:
- Utilized Nomarski microscopy to screen for C. elegans mutants with refractile cell corpses.
- Generated null mutations in the cup-5 gene and performed genetic rescue experiments using the human ML-IV gene.
- Analyzed cup-5 mutants for lysosome accumulation and apoptosis using transmission electron microscopy and TUNEL staining.
Main Results:
- A C. elegans mutant accumulating refractile bodies, resembling cell corpses, was identified.
- The cup-5 gene was found to be responsible for the observed phenotype, and its null mutations caused maternal-effect lethality.
- cup-5 mutants exhibited excessive lysosomes and lamellar structures, characteristic of ML-IV, and also showed increased apoptosis, suggesting a secondary consequence of lysosomal dysfunction.
Conclusions:
- The C. elegans cup-5 gene plays a crucial role in lysosomal function and is essential for development.
- Lysosomal defects, as exemplified by cup-5 mutations, may contribute to increased apoptosis.
- These findings suggest a potential link between lysosomal storage disorders and abnormalities in programmed cell death in humans.
Related Concept Videos
 01:33
Overview of Secretory Vesicles
Various proteins regulate the aggregation of molecules inside the secretory vesicles. Chromogranins...
 01:22
Lysosomal Hydrolases