New space warping method for the simulation of large-scale macromolecular conformational changes
Related Experiment Video
Updated: Oct 2, 2026
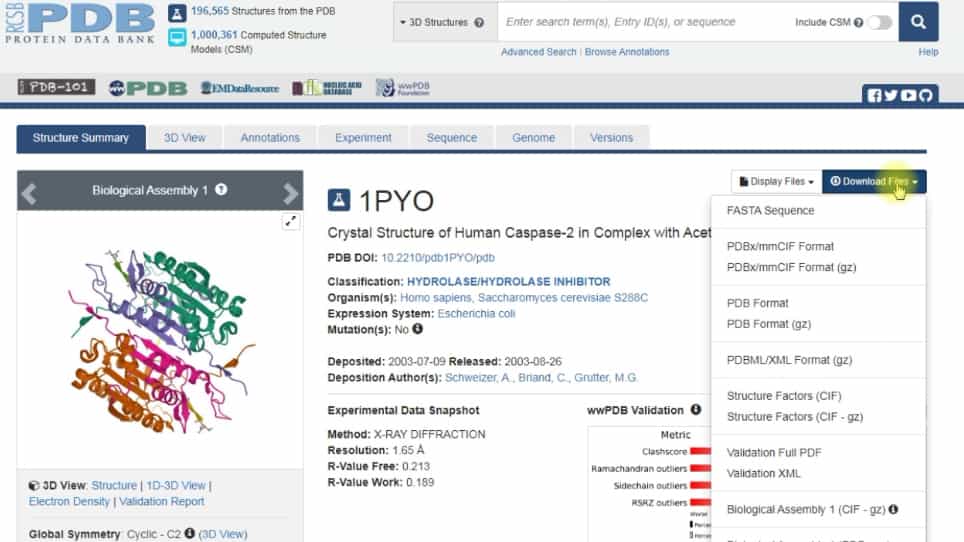
Exploring Caspase Mutations and Post-Translational Modification by Molecular Modeling Approaches
Published on: October 13, 2022
Khuloud Jaqaman1, Peter J Ortoleva
1Department of Chemistry, Indiana University, Bloomington, Indiana 47405, USA.

Abstract:
A space warping method, facilitating the modeling of large-scale conformational changes in mesoscopic systems, is presented. The method uses a set of "global (or collective) coordinates" that capture overall behavior, in conjunction with the set of atomic coordinates. Application of the space warping method to energy minimization is discussed. Several simulations where the method is used to determine the energy minimizing structures of simple central force systems are analyzed. Comparing the results and behavior of the space warping method to simulations involving atomic coordinates only, it is found that the space warping method scales better with system size and also finds lower minima when the potential energy surface has multiple minima. It is shown that the transformation of [Ala16]+ in vacuo from linear to globular is captured efficiently using the space warping method.
More Related Videos
Related Concept Videos
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
 02:06
Newman Projections
The organic molecules rotate across the single bonds leading to numerous temporary three-dimensional structures of varying energy known as conformers.
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal tetrahedral value,...