Related Experiment Video
Updated: Oct 2, 2026


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
E-state modeling of HIV-1 protease inhibitor binding independent of 3D information
Hlaing Hlaing Maw1, Lowell H Hall
1Department of Chemistry, Eastern Nazarene College, Quincy, Massachusetts 02170, USA.

Abstract:
Data for HIV-1 protease inhibitors (in vitro enzyme binding) were used as a training set to develop a QSAR model based on topological descriptors, including two hydrogen E-state indices, along with a molecular connectivity chi and a kappa shape index. A statistically satisfactory four-variable model was obtained for the 32 compounds in the training set, r2 = 0.86, s = 0.60, and q2 = 0.79, without the use of information from 3D geometries or detailed interaction energy calculations. The model was validated through the prediction of 15 compounds in the external test set, yielding a mean absolute error, MAE, = 0.82. Structure interpretation is given for each variable to assist in the design of new compounds. Structure features emphasized in the model include hydrogen bond donating ability, nonpolar groups, skeletal branching, and molecular globularity. On the basis of these statistical criteria, this E-state model may be considered useful for prediction of pIC50 values for new HIV-1 protease inhibitors.
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 01:24
Protein Organization
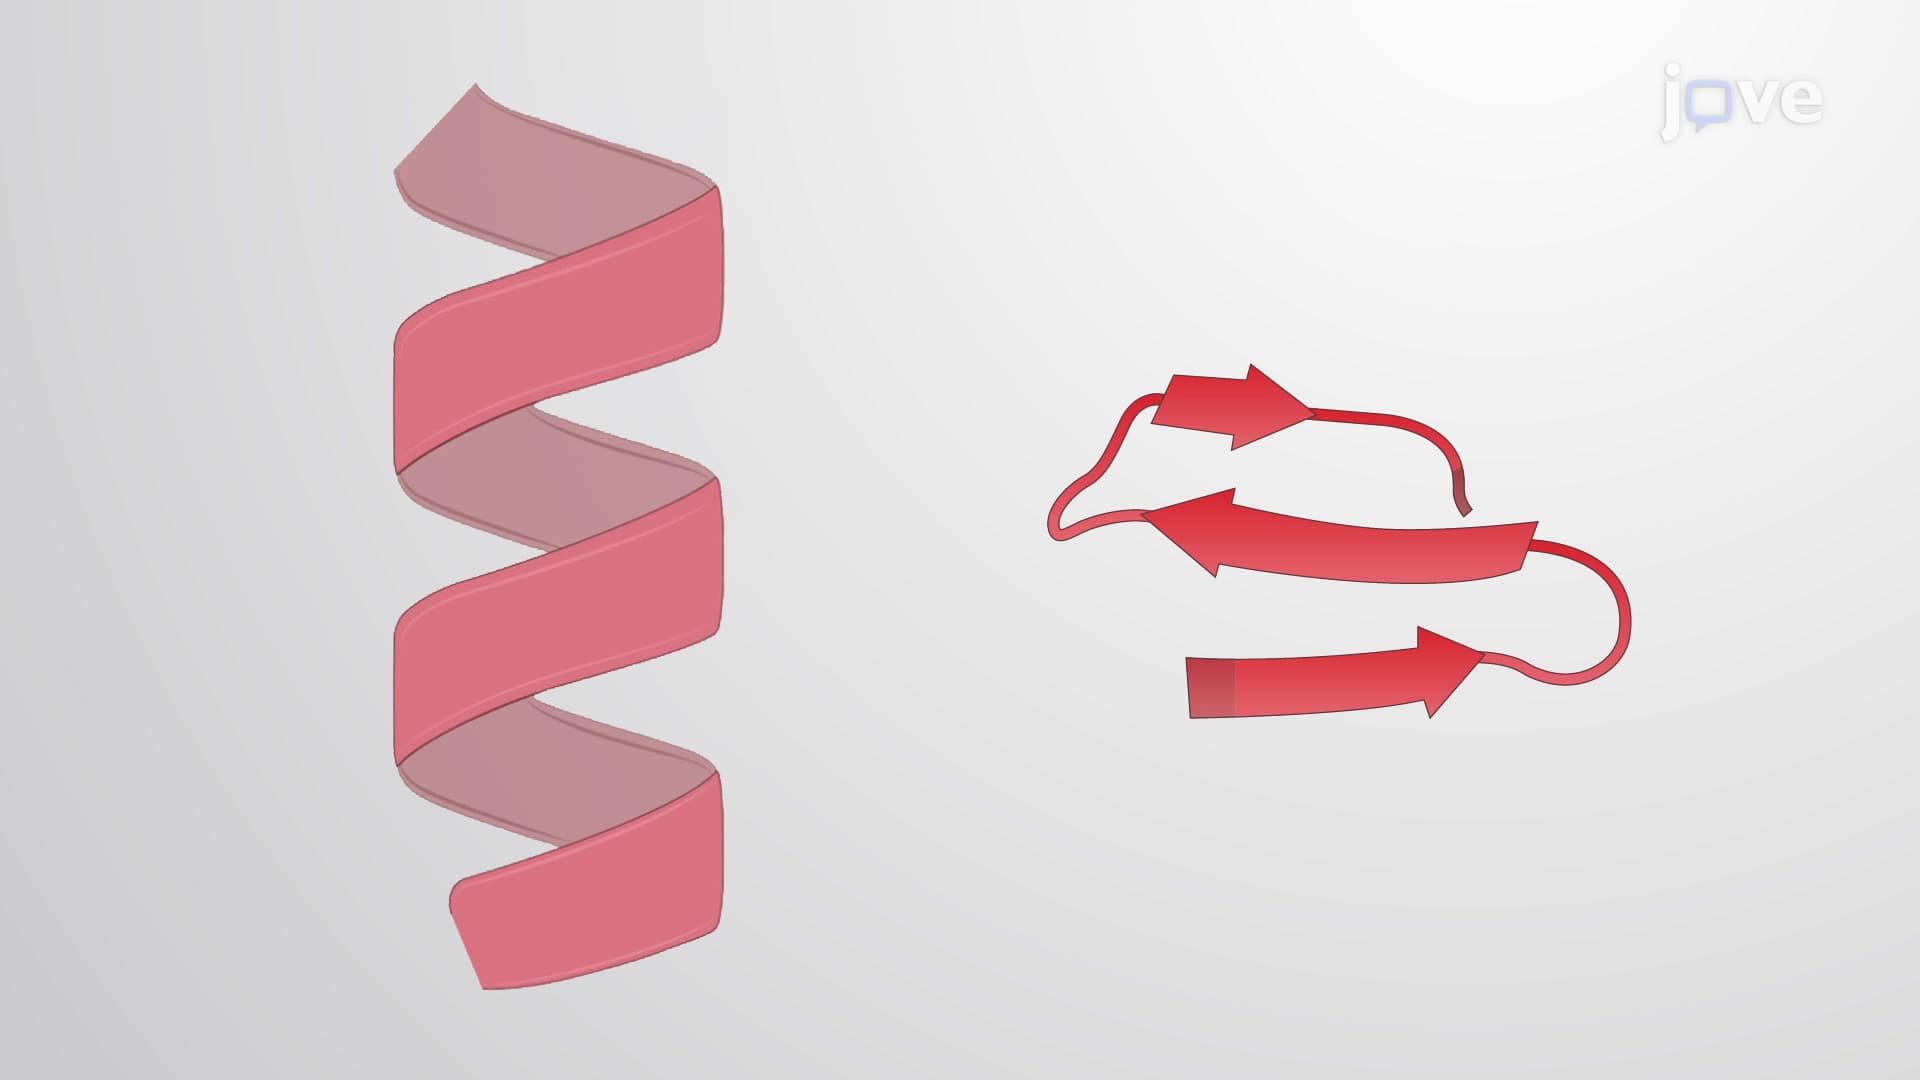
The primary structure of a protein is its amino acid sequence.
 01:19
Inhibitors of Virion Maturation and Assembly