The gene mutated in autosomal recessive polycystic kidney disease encodes a large, receptor-like protein
Related Experiment Video
Updated: Oct 2, 2026
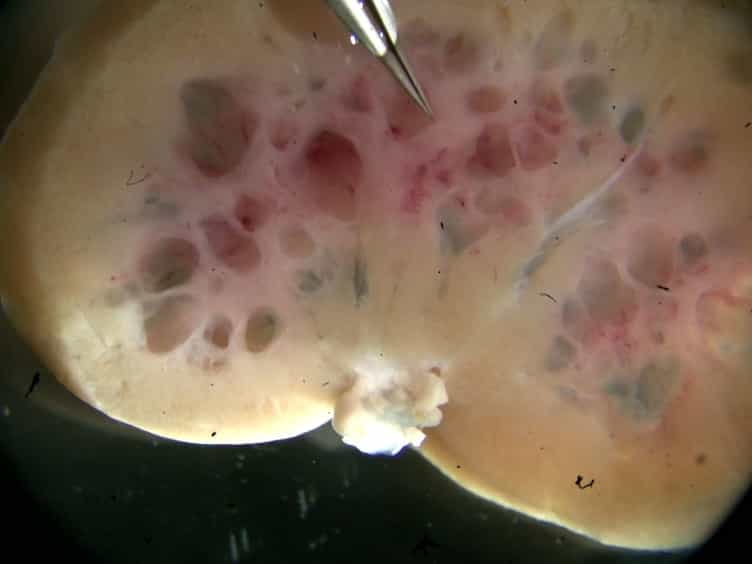
Implementing Patch Clamp and Live Fluorescence Microscopy to Monitor Functional Properties of Freshly Isolated PKD Epithelium
Published on: September 1, 2015
Christopher J Ward1, Marie C Hogan, Sandro Rossetti
1Division of Nephrology, Mayo Clinic, 200 First Street SW, Rochester, Minnesota 55905, USA.

Insights
Researchers identified the PKHD1 gene as a cause of autosomal recessive polycystic kidney disease (ARPKD). This discovery sheds light on the genetic basis of ARPKD, a serious kidney and liver disorder.
Area of Science:
- Genetics
- Molecular Biology
- Pediatric Nephrology
Background:
- Autosomal recessive polycystic kidney disease (ARPKD) is a significant genetic disorder causing kidney and liver dysfunction.
- It is characterized by collecting duct dilation and biliary dysgenesis, leading to substantial morbidity and mortality.
Purpose of the Study:
- To identify the genetic cause of ARPKD.
- To characterize the PKHD1 gene and its encoded protein, fibrocystin, in relation to ARPKD pathogenesis.
Main Methods:
- Genetic analysis of a rat model with recessive polycystic kidney disease.
- Screening of the human ortholog gene, PKHD1, in ARPKD patients.
- Characterization of PKHD1 transcript and protein expression.
Main Results:
- An orthologous relationship was established between the rat locus and the human ARPKD region.
- Mutations in PKHD1 were identified in ARPKD probands, including truncating and missense variants.
- PKHD1 encodes a large protein, fibrocystin, expressed in kidney, liver, and pancreas.
Conclusions:
- PKHD1 is the causative gene for ARPKD.
- Fibrocystin, the protein product of PKHD1, is likely involved in collecting duct and biliary differentiation.
- This finding provides a molecular basis for understanding ARPKD and potential therapeutic targets.
Abstract:
Autosomal recessive polycystic kidney disease (ARPKD) is characterized by dilation of collecting ducts and by biliary dysgenesis and is an important cause of renal- and liver-related morbidity and mortality. Genetic analysis of a rat with recessive polycystic kidney disease revealed an orthologous relationship between the rat locus and the ARPKD region in humans; a candidate gene was identified. A mutation was characterized in the rat and screening the 66 coding exons of the human ortholog (PKHD1) in 14 probands with ARPKD revealed 6 truncating and 12 missense mutations; 8 of the affected individuals were compound heterozygotes. The PKHD1 transcript, approximately 16 kb long, is expressed in adult and fetal kidney, liver and pancreas and is predicted to encode a large novel protein, fibrocystin, with multiple copies of a domain shared with plexins and transcription factors. Fibrocystin may be a receptor protein that acts in collecting-duct and biliary differentiation.
Related Concept Videos
 01:20
Renal Corpuscle
Glomerulus: Structure and Function
The glomerulus is a tiny, intricate network of capillaries located at the beginning of the nephron. It's enveloped by the Bowman's capsule and receives its blood supply from an afferent arteriole, which divides into numerous capillaries...
 01:27
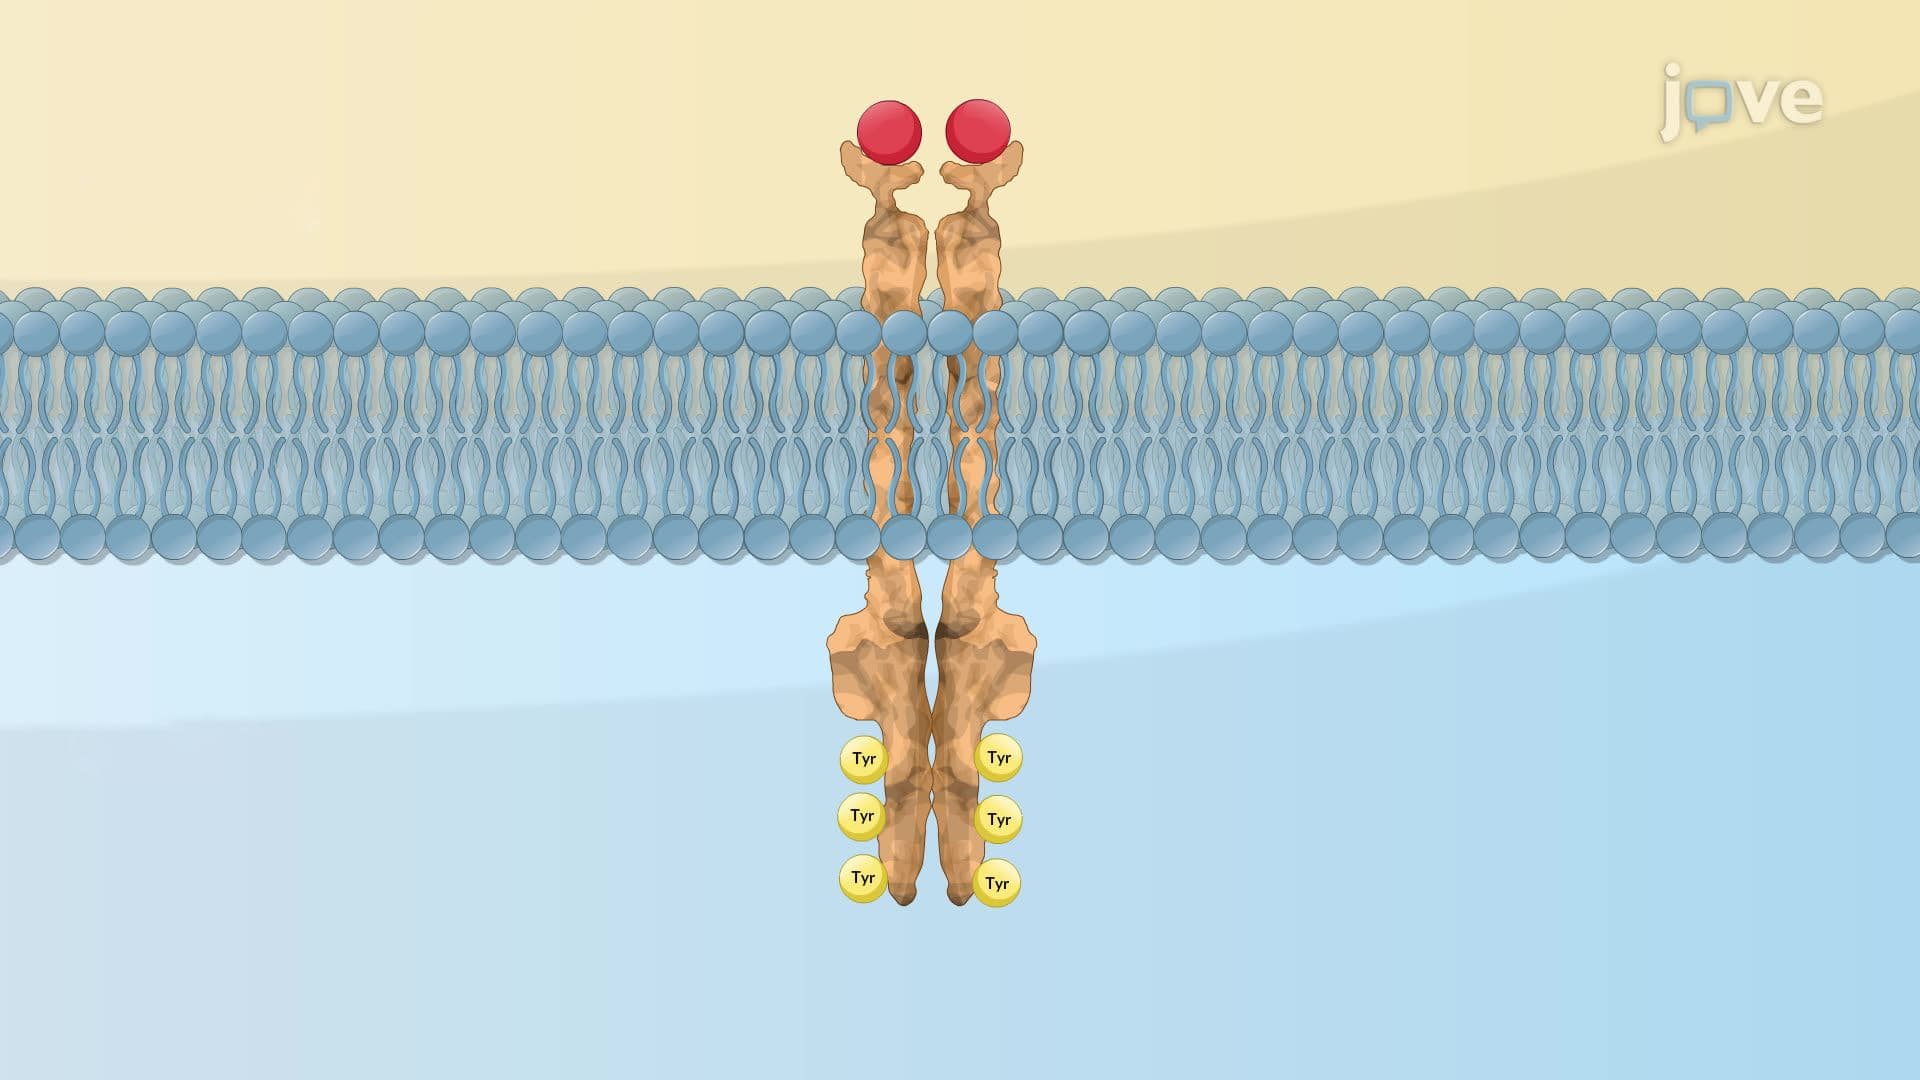
Transducer Mechanism: Enzyme-Linked Receptors
Major types that are helpful drug targets include:
 02:38
The Ras Gene
Ras is a superfamily...
 01:10
Nephrons
 01:14
Rab Proteins
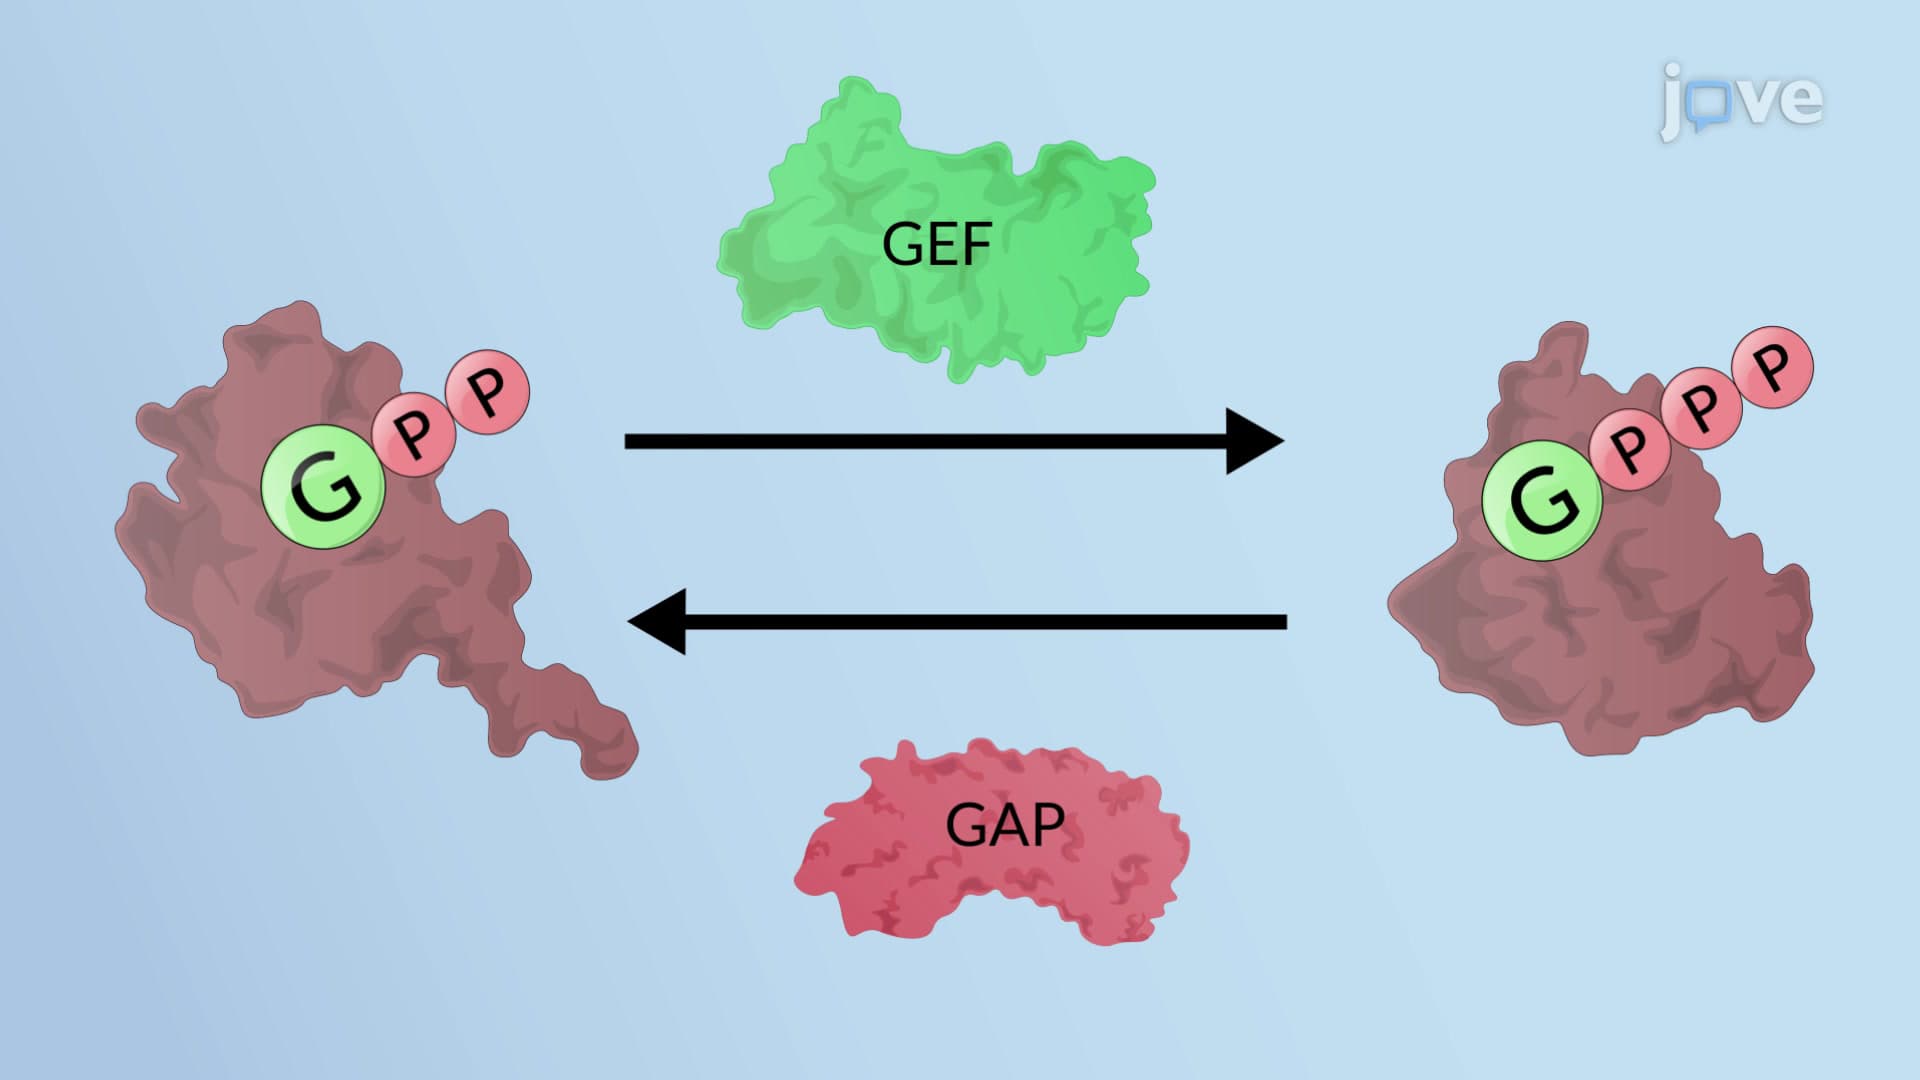
Rab proteins switch between a cytosolic, GDP-bound inactive state and a membrane-anchored, GTP-bound active state. By themselves, Rabs show slow rates of GDP/GTP exchange and GTP hydrolysis. Thus, Rab proteins are considered...
 01:13
Enzyme-linked Receptors