Related Experiment Video
Updated: Oct 1, 2026
Using Phylogenetic Analysis to Investigate Eukaryotic Gene Origin
Published on: August 14, 2018
The accuracy of fast phylogenetic methods for large datasets
Luay Nakhleh1, Bernard M E Moret, Usman Roshan
1Dept. of Computer Sciences, University of Texas, Austin, TX 78712, USA.

Abstract:
Whole-genome phylogenetic studies require various sources of phylogenetic signals to produce an accurate picture of the evolutionary history of a group of genomes. In particular, sequence-based reconstruction will play an important role, especially in resolving more recent events. But using sequences at the level of whole genomes means working with very large amounts of data--large numbers of sequences--as well as large phylogenetic distances, so that reconstruction methods must be both fast and robust as well as accurate. We study the accuracy, convergence rate, and speed of several fast reconstruction methods: neighbor-joining, Weighbor (a weighted version of neighbor-joining), greedy parsimony, and a new phylogenetic reconstruction method based on disk-covering and parsimony search (DCM-NJ + MP). Our study uses extensive simulations based on random birth-death trees, with controlled deviations from ultrametricity. We find that Weighbor, thanks to its sophisticated handling of probabilities, outperforms other methods for short sequences, while our new method is the best choice for sequence lengths above 100. For very large sequence lengths, all four methods have similar accuracy, so that the speed of neighbor-joining and greedy parsimony makes them the two methods of choice.
Related Concept Videos
 02:54
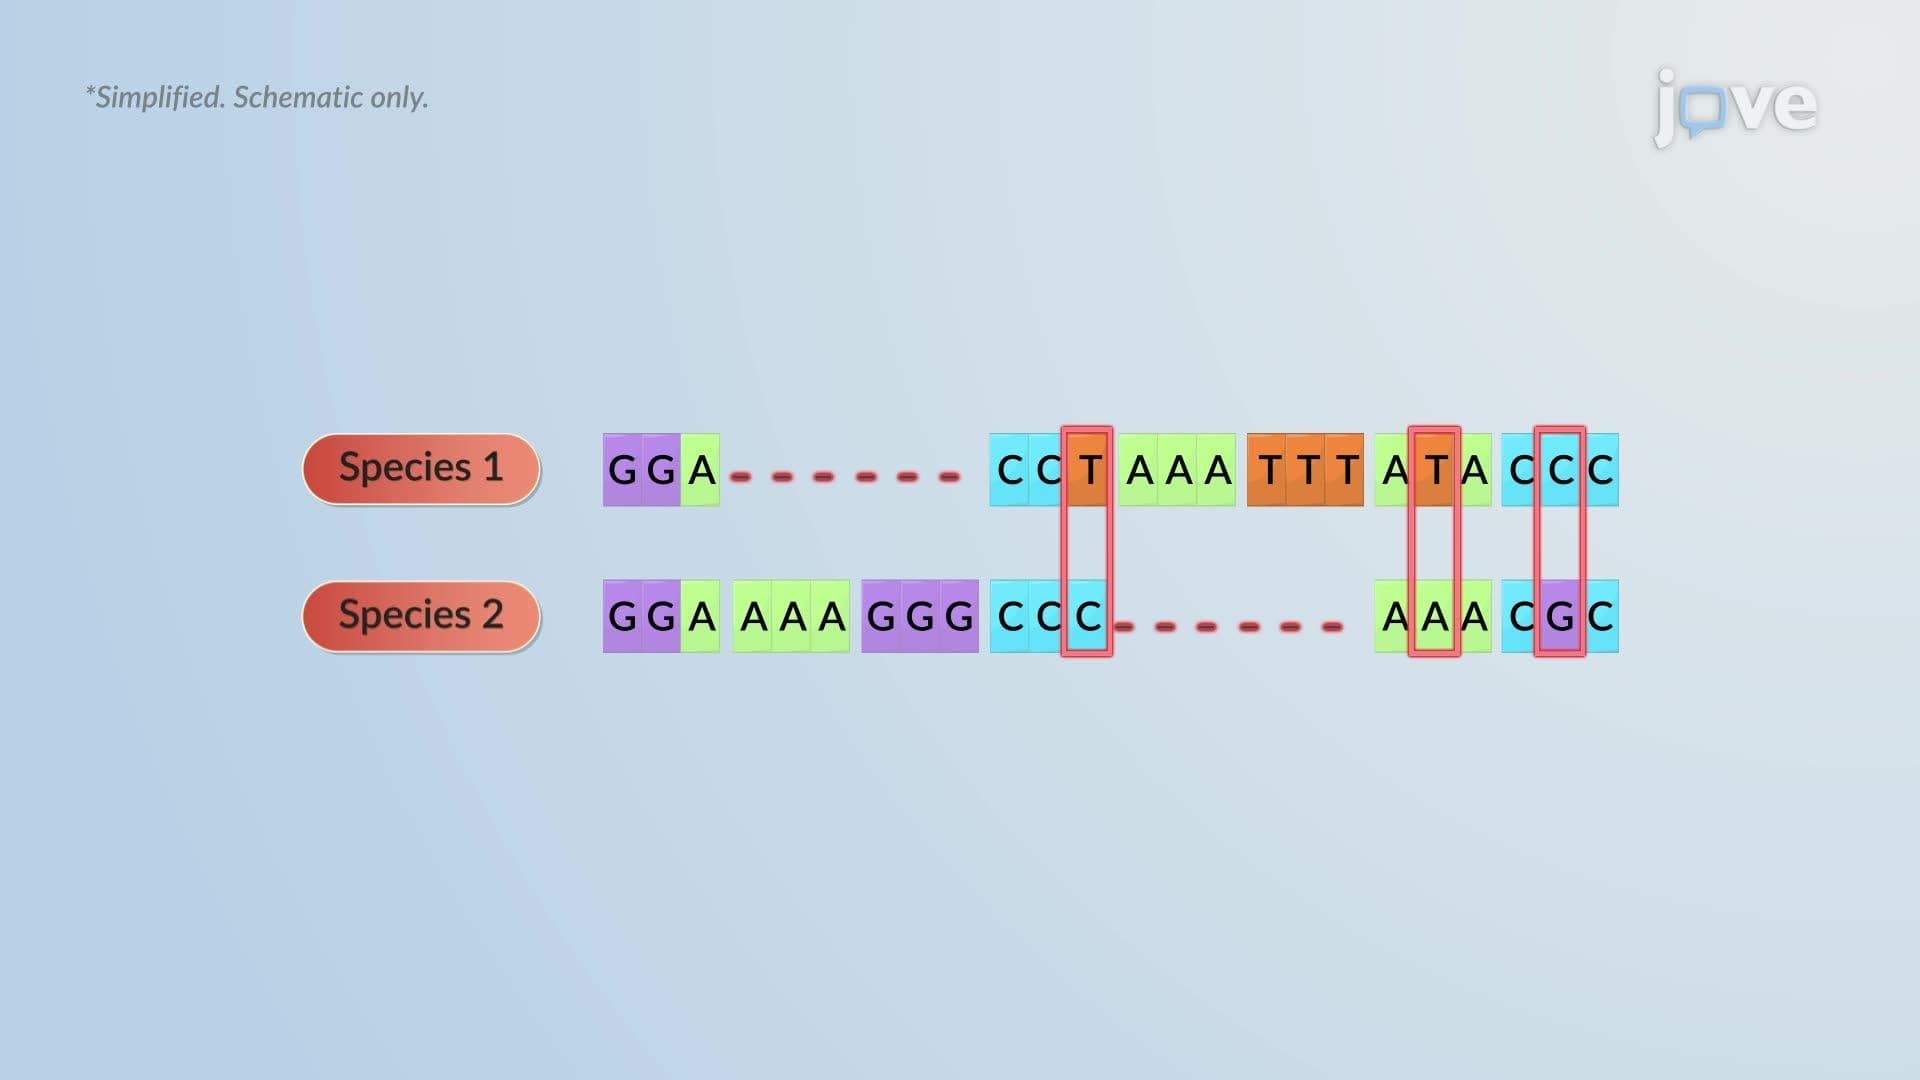
Evolutionary Relationships through Genome Comparisons
 02:05
Gene Evolution - Fast or Slow?
In contrast, regions which code...
02:05Gene Evolution - Fast or Slow?
In contrast, regions which code...
 01:28
Microbial Phylogeny
 03:21
Phylogenetic Trees
03:21Phylogenetic Trees