
Anharmonic vibrational spectra of hydroxylamine and its 15N, 18O, and deuterium substituted analogs
1Institute of Inorganic Chemistry and Metallurgy of Rare Elements, Wrocław University of Technology, Poland. atk@ichn.ch.pwr.wroc.pl

Abstract:
Vibrational self-consistent field (VSCF) and correlation-corrected vibrational self-consistent field (CC-VSCF) methods were used to compute the anharmonic frequencies of fundamentals, overtones, and combination transitions of natural abundance hydroxylamine, 15NH2OH, NH2(18)OH, ND2OD, ND2OH, and NH2OD isotopomers at second order Møller-Plesset perturbation theory (MP2) in basis sets of triple-zeta quality. Frequencies of the fundamental transitions observed in the gas phase spectrum were reproduced by CC-VSCF treatment within 20 cm(-1) in TZV(d,p) and TZV(2d,2p) basis sets, and the change of basis set composition had only minor effect on the frequencies of the computed fundamentals. CC-VSCF computed wave numbers of overtone and combination transitions were typically within 1-40 cm(-1) of the gas phase band positions, except for those resulting from multiple excitations of v2, v3, and v7 fundamentals, because of a strong mutual coupling between these modes. Integral transition intensities calculated at MP2 level closely followed those of experimental spectrum, including intensity decrease in v1, 2v1, 3v1 progression, and 30-fold intensity increase of 2v8 in respect to that of v8 fundamental. The frequency of the OH torsional fundamental was found to be strongly dependent on the mode-mode interaction potential among v9 and v1, v7, v2, v4, v5 modes. Band shifts resulting from 15N, 18O and complete 2H substitutions were reproduced almost quantitatively by CC-VSCF computation in TZV(d,p) basis. Computed anharmonic isotope frequency shifts were different from those obtained in the harmonic approximation and no scaling procedure seemed capable of performing their interchange.
More Related Videos
 06:18
Syntheses, Crystallization, and Spectroscopic Characterization of 3,5-Lutidine N-Oxide Dehydrate
Published on: April 24, 2018
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:19
NMR Spectroscopy Of Amines
 01:08
IR Spectrum Peak Splitting: Symmetric vs Asymmetric Vibrations
 00:48
¹H NMR of Labile Protons: Deuterium (²H) Substitution
 01:14
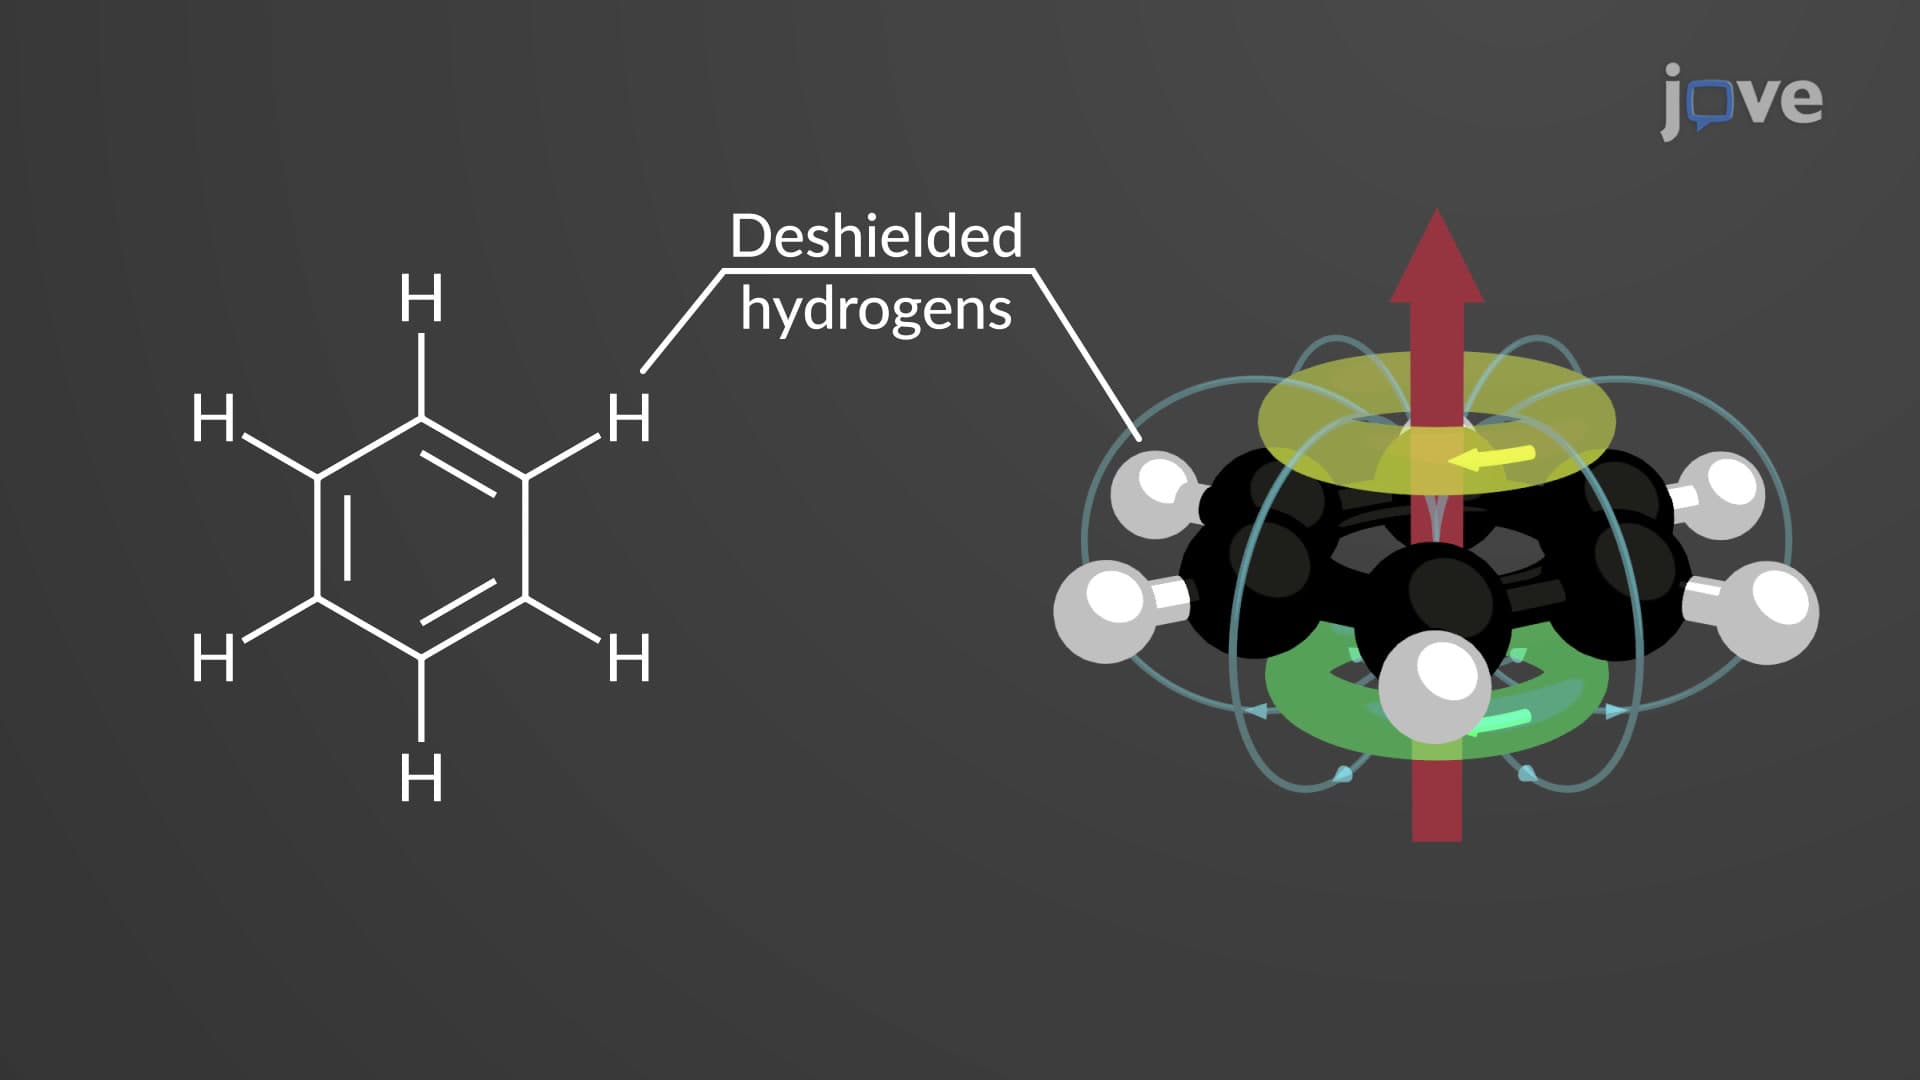
NMR Spectroscopy of Aromatic Compounds
 00:58
¹H NMR Chemical Shift Equivalence: Enantiotopic and Diastereotopic Protons
In chiral compounds such as 2-butanol, replacing the methylene hydrogens at C3 produces a pair of...
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are slanted or...