Related Experiment Video
Updated: Aug 24, 2026
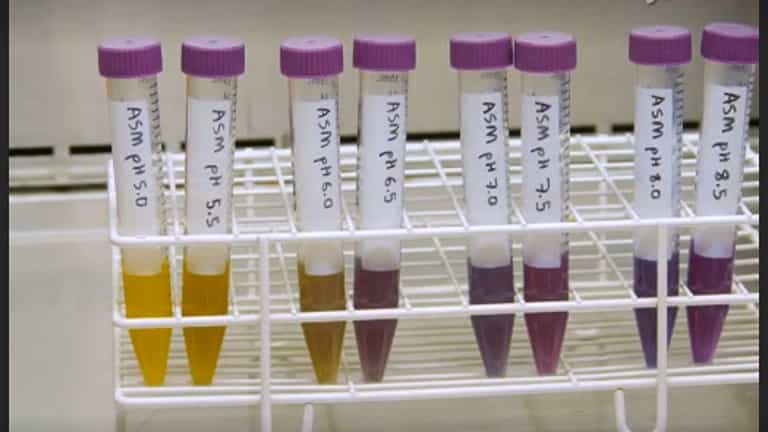

The WinCF Model - An Inexpensive and Tractable Microcosm of a Mucus Plugged Bronchiole to Study the Microbiology of Lung Infections
Published on: May 8, 2017
Clinical findings and lung pathology in children with cystic fibrosis
Refika Hamutcu1, Jon M Rowland, Monica V Horn
1Division of Pediatric Pulmonology, Childrens Hospital Los Angeles and Keck School of Medicine, University of Southern California, Los Angeles, California 90027, USA.

Abstract:
Cystic fibrosis pulmonary disease is assessed by pulmonary function tests, arterial blood gases, and chest X-rays, but the correlation with lung pathology is unknown. We reviewed the clinical findings and lung pathology of 21 cystic fibrosis patients who had lung transplant. Pulmonary function tests, Brasfield scores, arterial blood gases, and age were correlated with lung pathology. All patients had severe Brasfield scores (9.0 +/- 3.2), airways obstruction (FEV1 25.6 +/- 5.6% predicted, FEF(25-75%) 11.0 +/- 4.5% predicted), and hyperinflation (residual volume [RV] 341.8 +/- 75.8% predicted). All patients were hypoxemic (PO2 64.2 +/- 8.2 mm Hg), and 5 of 21 (24%) were hypercapneic (PCO2 > 50 mm Hg). Pulmonary function tests and Brasfield scores were within a narrow range, and did not allow correlation with lung pathology. Small airway density (airways < 2 mm/cm2) decreased with increasing age. There were no differences in small airways inflammation and fibrous narrowing between the hypercapneic and nonhypercapneic patients, but the percent of smallest airways (airways < 0.35 mm) was significantly lower in the hypercapneic group. We conclude that there is significant correlation between airway pathology and increased age and CO2 retention. We speculate that decreased small airway density in older patients and the decreased proportion of smallest airways in hypercapneic patients is caused by increased dilatation of small airways.
More Related Videos
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:20
Chronic Obstructive Pulmonary Disease-II: Pathophysiology
Chronic Inflammation
 01:27
Chronic Obstructive Pulmonary Disease-IV: Assessement and Diagnostic Studies
Medical History
 01:24
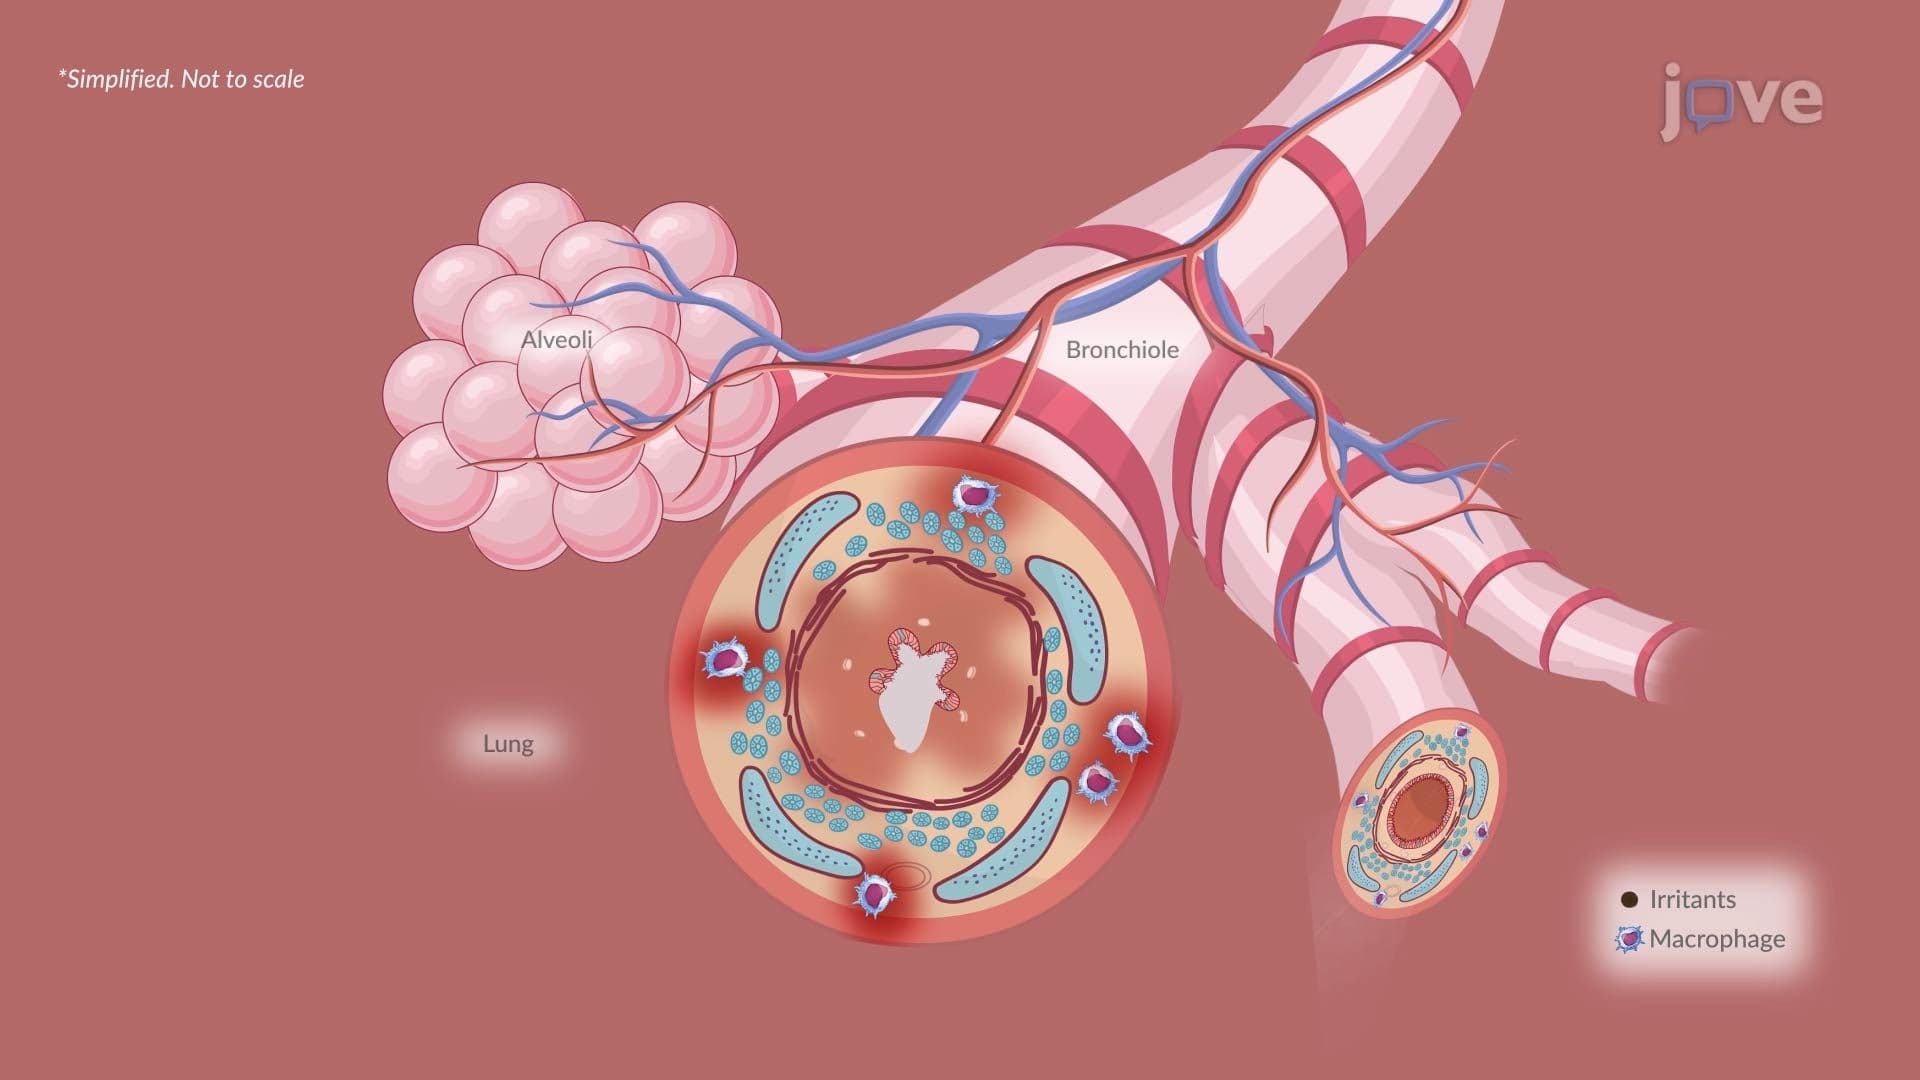
Chronic Obstructive Pulmonary Disease III: Chronic Bronchitis Features
 01:19
Chronic Obstructive Pulmonary Disease IV: Clinical Manifestations