Related Experiment Video
Updated: Jul 27, 2026

Preparation, Purification, and Characterization of Lanthanide Complexes for Use as Contrast Agents for Magnetic Resonance Imaging
Published on: July 21, 2011
Conformational characterization of lanthanide(III)-DOTA complexes by ab initio investigation in vacuo and in aqueous
Ugo Cosentino1, Alessandra Villa, Demetrio Pitea
1Dipartimento di Scienze dell'Ambiente e del Territorio (DISAT), Università degli Studi di Milano-Bicocca, Piazza della Scienza 1, 20126 Milano, Italy. ugo.cosentino@unimib.it

Abstract:
The conformational behavior of four [Ln(DOTA)(H(2)O)](-) systems (Ln = La, Gd, Ho, and Lu) has been characterized by means of ab initio calculations performed in vacuo and in aqueous solution, the latter by using the polarizable continuum model (PCM). Calculated molecular geometries and conformational energies of the [Ln(DOTA)(H(2)O)](-) systems, and interconversion mechanisms, barriers, and (13)C NMR spectra of the [Lu(DOTA)](-) complex are compared with experimental values. For each system, geometry optimizations, performed in vacuo and in solution at the HF/3-21G level and using a 46+4f(n) core electron effective core potential (ECP) for lanthanides, provide two minima corresponding to a square antiprismatic (A) and an inverted antiprismatic (IA) coordination geometry. All the systems are nonacoordinated, with the exception of the IA isomer of the Lu complex that, from in solution calculations, is octacoordinated, in agreement with experimental data. On comparing the in vacuo relative free energies calculated at different theory levels it can be seen that the nonacoordinated species dominates at the beginning of the lanthanide series while the octacoordinated one does so at the end. Furthermore, on passing along the series the IA isomer becomes less and less favored with respect to A and for the Lu complex a stabilization of the IAisomer is observed in solution (but not in vacuo), in agreement with the experimental data. Investigation of the A<-->IA isomerization process in the [Lu(DOTA)](-) system provides two different interconversion mechanisms: a single-step process, involving the simultaneous rotation of the acetate arms, and a multistep path, involving the inversion of the cyclen cycle configuration. While in vacuo the energy barrier for the acetate arm rotation is higher than that involved in the ring inversion (23.1 and 13.1 kcal mol(-)(1) at the B3LYP/6-311G level, respectively), in solution the two mechanisms present comparable barriers (14.7 and 13.5 kcal mol(-)(1)), in fairly good agreement with the experimental values. The NMR shielding constants for the two isomers of the [Lu(DOTA)](-) complex have been calculated by means of the ab initio GIAO and CSGT methods, and using a 46-core-electron ECP for Lu. The calculated (13)C NMR chemical shifts are in close agreement with the experimental values (rms 3.3 ppm, at the HF/6-311G level) and confirm the structural assignment of the two isomers based on experimental NMR spectra in solution. The results demonstrate that our computational approach is able to predict several physicochemical properties of lanthanide complexes, allowing a better characterization of this class of compounds for their application as contrast agents in medical magnetic resonance imaging (MRI).
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 07:24
Hyperspectral Imaging as a Tool to Study Optical Anisotropy in Lanthanide-Based Molecular Single Crystals
Published on: April 14, 2020
Related Concept Videos
 02:42
Valence Bond Theory
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human eye.
 01:07
Ladder Diagrams: Complexation Equilibria
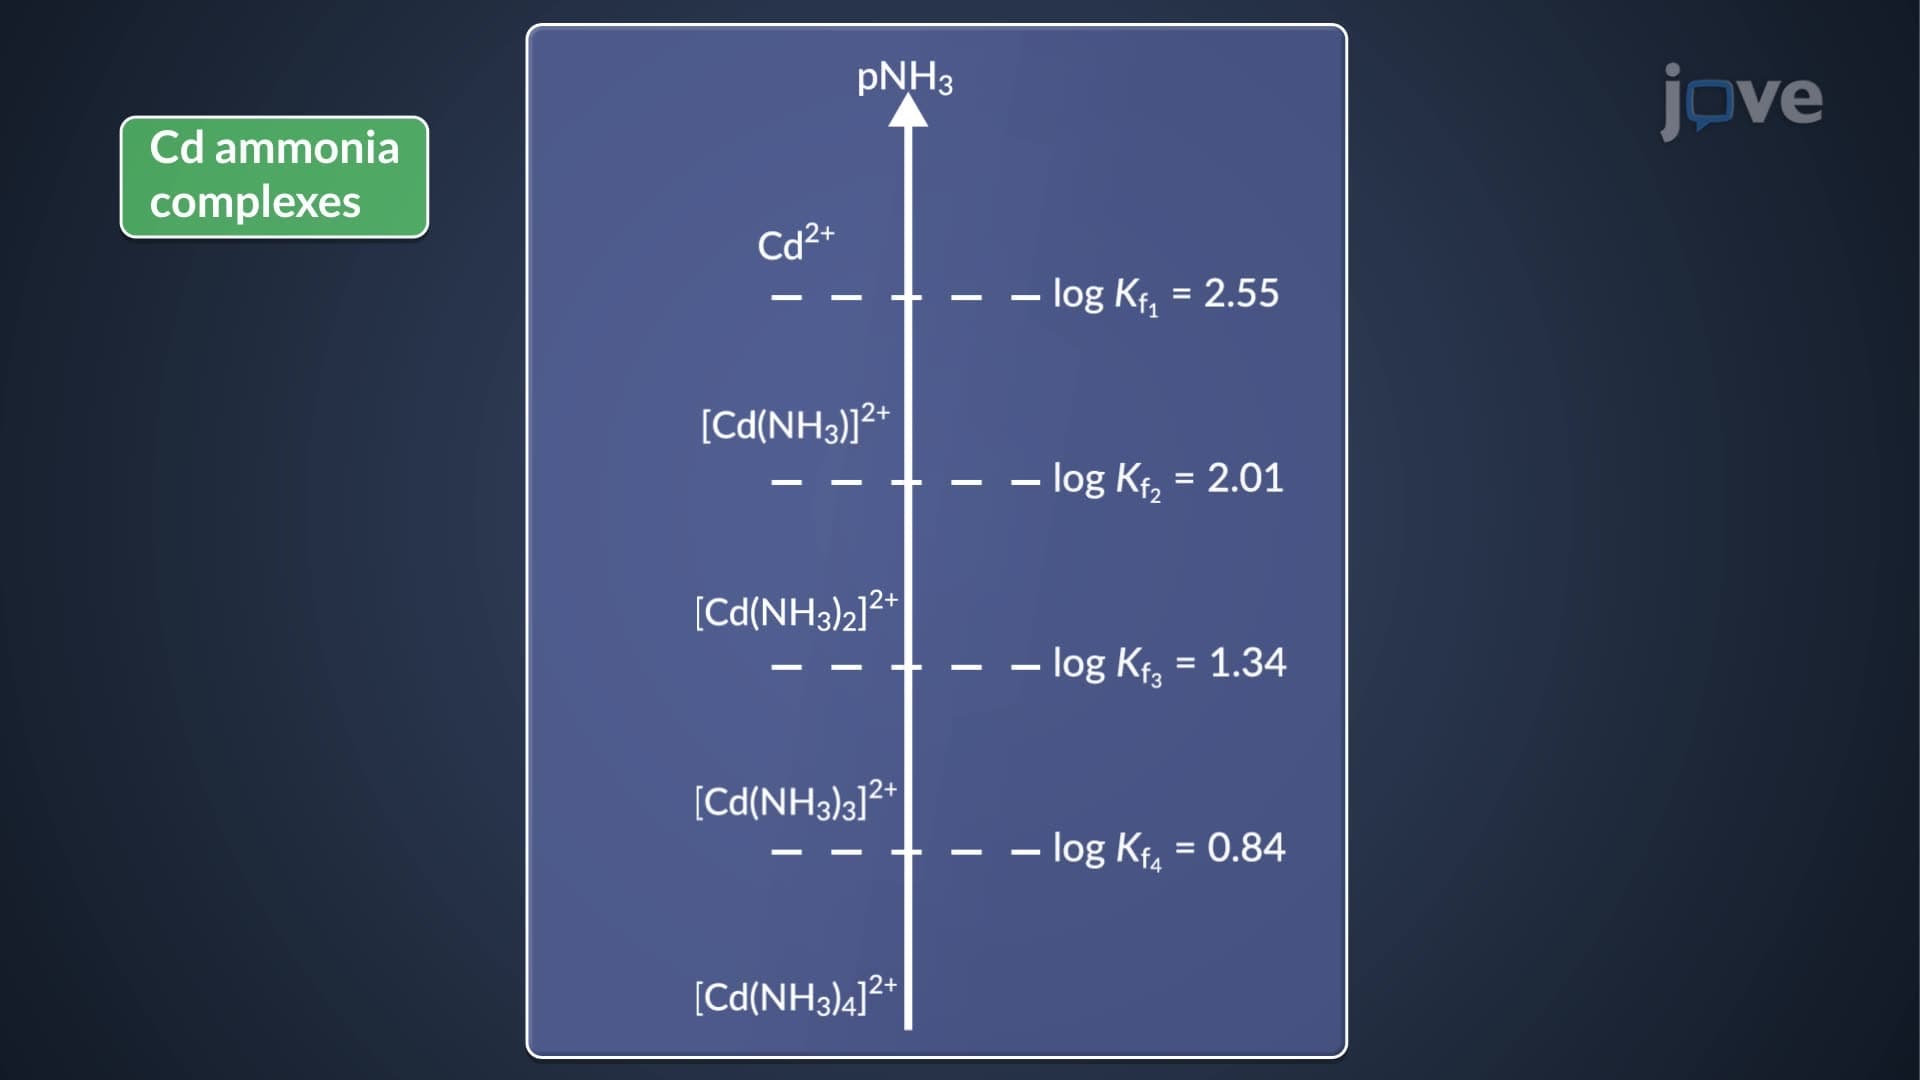
The formation constant, K1, for the formation of Cd(NH3)2+ complex from cadmium and ammonia is 3.55 × 10². Log K1 (i.e. pNH3) is 2.55, and...
 00:43
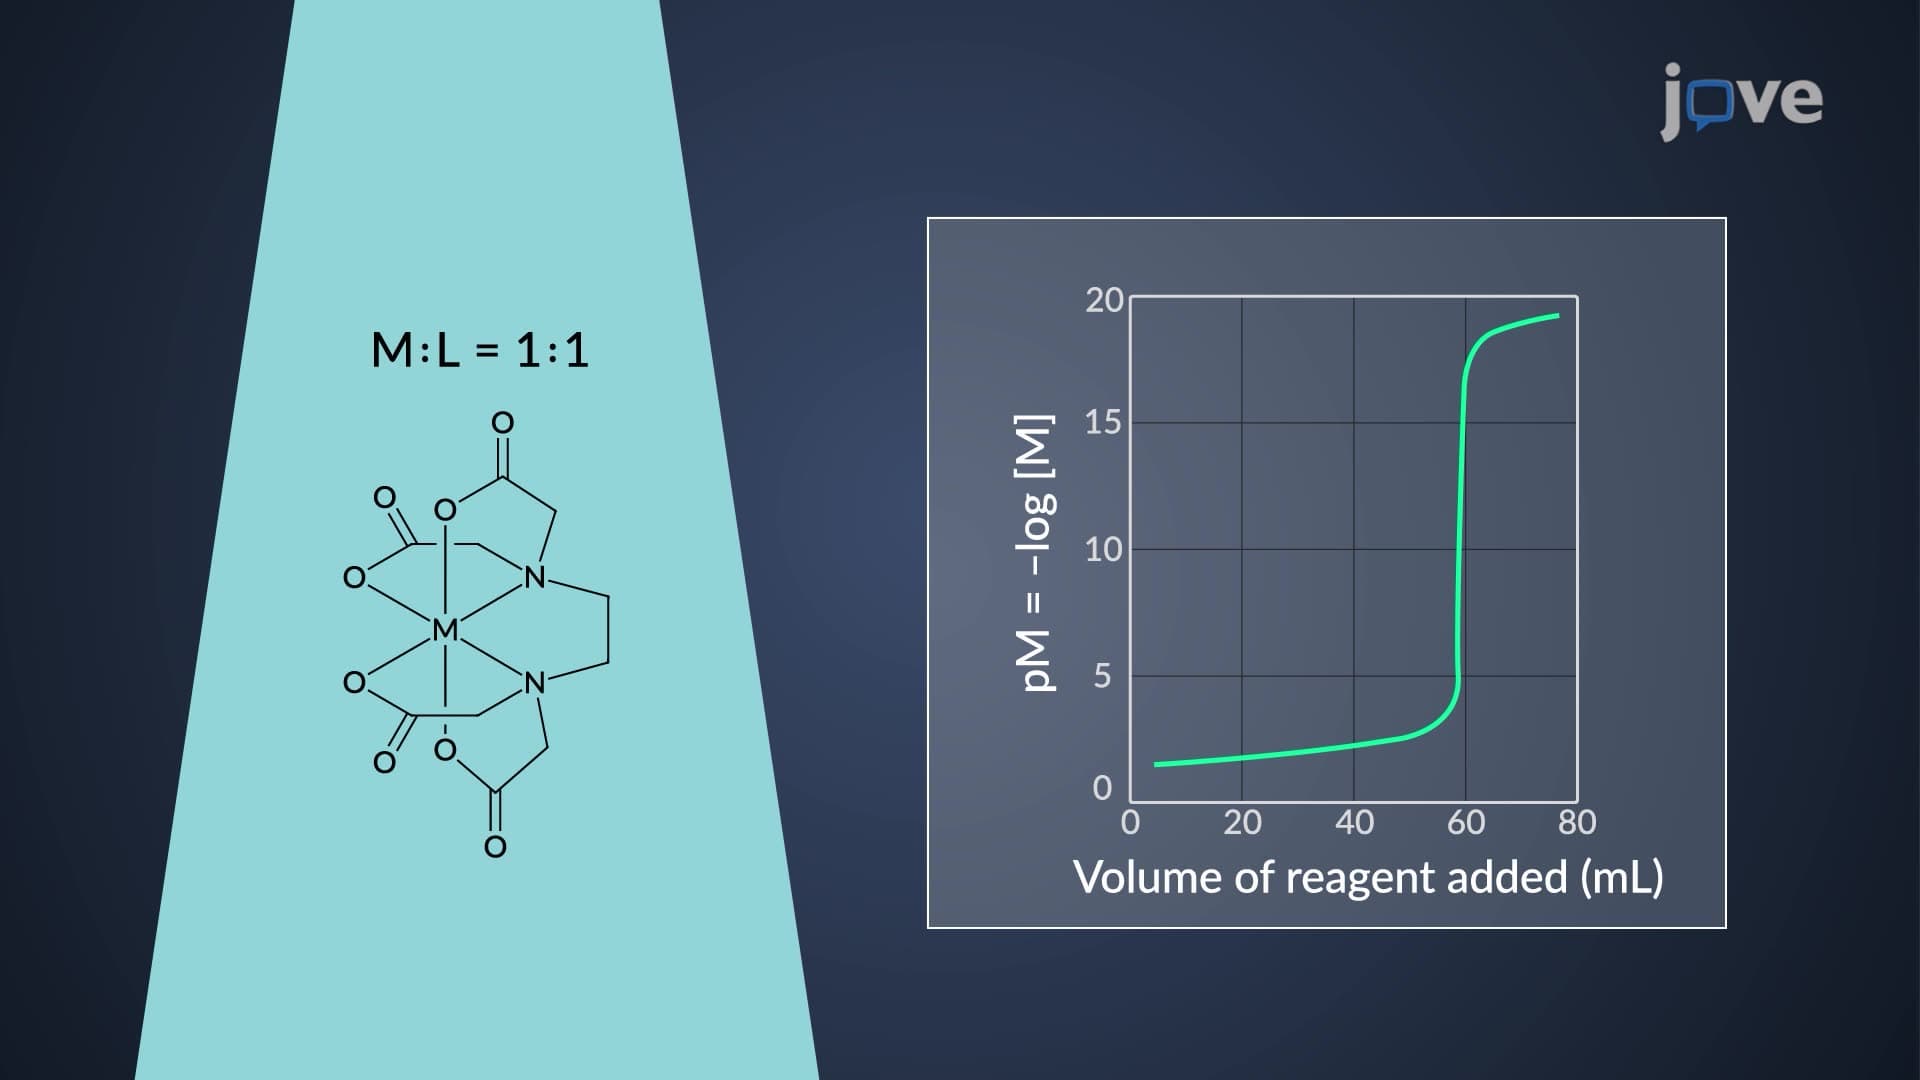
Complexometric Titration: Ligands