Related Experiment Video
Updated: Oct 1, 2026
Voltage-Dependent Potassium Current Recording on H9c2 Cardiomyocytes via the Whole-Cell Patch-Clamp Technique
Published on: November 11, 2022
The nature of the receptor site for the reversible K+ channel blocking by aminopyridines
Camelia Muñoz-Caro1, Alfonso Niño
1Grupo de Química Computacional, Escuela Superior de Informática, Universidad de Castilla-La Mancha, Paseo de la Universidad 4, 13071, Ciudad Real, Spain. quimcom@uclm.es

Abstract:
This work presents a theoretical study aimed at the identification of the receptor site for the blocking of the voltage dependent K+ channels by protonated aminopyridines. Thus, the density functional theory (DFT) at the B3LYP/6-311 G (d,p) level is applied, both in vacuum and in solution, to a series of active (protonated) compounds: 2-aminopyridine, 3-aminopyridine, 4-aminopyridine, 3,4-diaminopyridine, and 4-aminoquinoleine. Analysis of the X-ray structure of the alpha-subunit of the channel shows that charged aminopyridines can interact electrostatically with a glutamic acid residue in the outside of the pore, or through a cation-pi interaction in the inside. To test both possibilities, model complexes are built using as nucleophiles a carboxylic group and an ethylene molecule, respectively. The three-dimensional electrostatic potential distribution of the protonated aminopyridines shows that an approaching nucleophile will be oriented toward the N-H (protonated) bond. Interaction with the carboxylic residue leads to a proton transfer, with the aminopyridine-carboxylic acid linked by a hydrogen bond. The observed breaking of the equivalence of the Laplacian of the charge density, the relative energy variation for the complexes, and the interaction with only one of the carboxylic residues in the fourfold alpha-subunit of the K+ channel are not compatible with the observed in vitro activity variation of aminopyridines. On the other hand, the study on the ethylene complexes shows, in vacuum and solution, a cation-pi interaction, clearly characterized by the atoms in molecules (AIM) theory. The variation of relative energy in solution is very small, but approaches the variation of in vitro activity. Our results, the pharmacophoric characteristics of aminopyridines, and the analysis of the three-dimensional internal structure of the K+ channel alpha-subunit suggest two putative receptor sites. One is formed by the four Thr-Thr-Val chains conforming the entrance to the narrow part of the inner K+ channel. The other is defined by four Thr residues within the pore.
More Related Videos
Related Concept Videos
 01:30
Ligand-Gated Ion Channel Receptor: Gating Mechanism
 01:12
Antiarrhythmic Drugs: Class III Agents as Potassium Channel Blockers
 01:18
Indirect-Acting Cholinergic Agonists: Mechanism of Action
Reversible inhibitors like edrophonium bind to a specific part of the enzyme called the anionic catalytic site. They form noncovalent bonds, which means they are not strongly attached to the enzyme. This creates a temporary and less stable enzyme–inhibitor complex, leading to...
 01:29
Indirect-Acting Cholinergic Agonists: Chemistry and Structure-Activity Relationship
Reversible inhibitors display short to medium durations of action. Short-acting agents include simple alcohols with...
 01:17
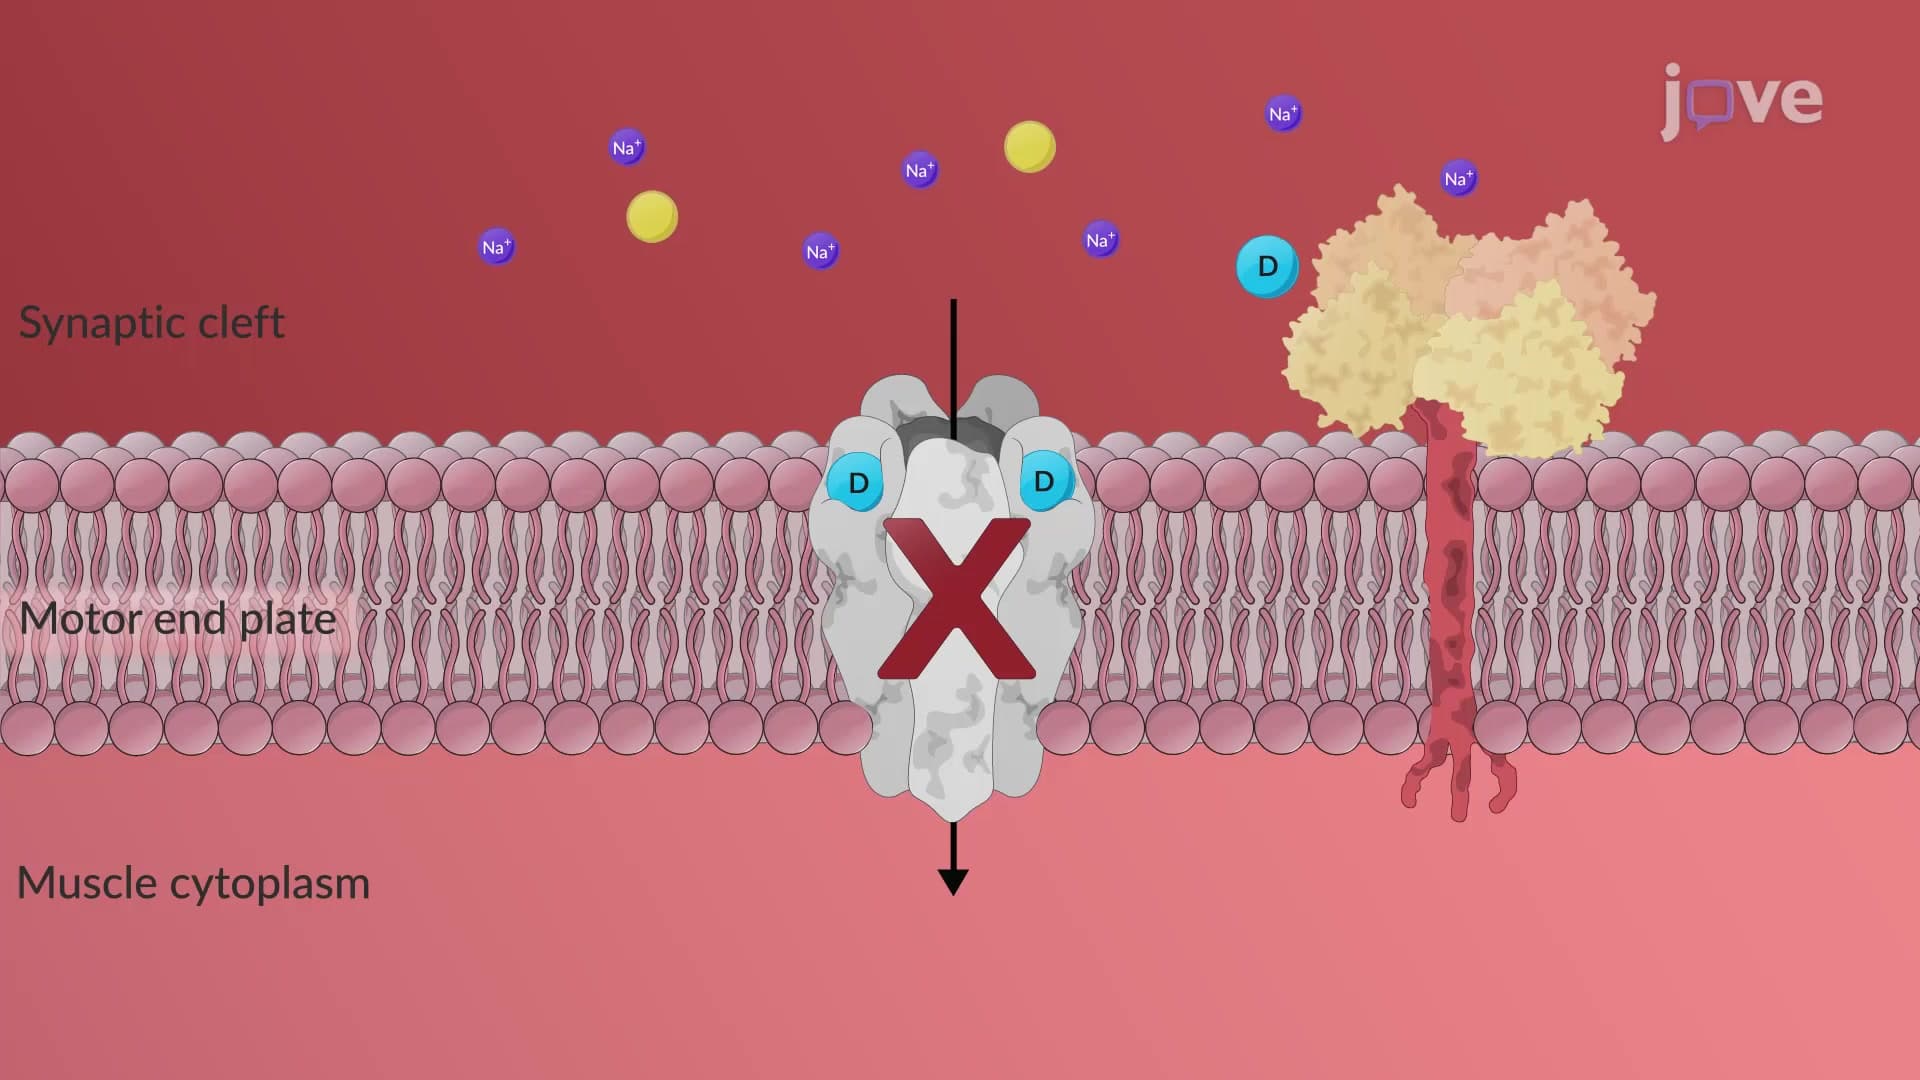
Nondepolarizing (Competitive) Neuromuscular Blockers: Mechanism of Action
Competitive antagonists prevent acetylcholine from binding to its receptor, inhibiting membrane depolarization. Without conformational changes or intrinsic...
 01:22
Adrenergic Antagonists: Pharmacological Actions of ɑ-Receptor Blockers
α1-blockers: These drugs inhibit α1-adrenoceptors on smooth muscle cells, resulting in vasodilation. This vasodilation lowers blood pressure, making α1-blockers valuable in treating hypertension. Additionally, α1-blockers effectively address urinary obstruction...