Related Experiment Video
Updated: Oct 1, 2026
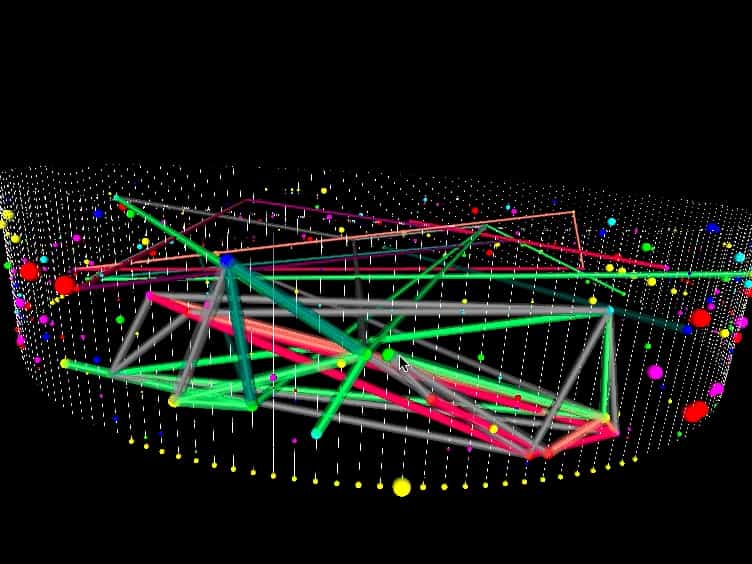

Optimization of Synthetic Proteins: Identification of Interpositional Dependencies Indicating Structurally and/or Functionally Linked Residues
Published on: July 14, 2015
Use of conserved key amino acid positions to morph protein folds
Boojala V B Reddy1, Wilfred W Li, Philip E Bourne
1San Diego Supercomputer Center, University of California-San Diego, 9500 Gilman Drive, La Jolla, CA 92093, USA.

Abstract:
By using three-dimensional (3D) structure alignments and a previously published method to determine Conserved Key Amino Acid Positions (CKAAPs) we propose a theoretical method to design mutations that can be used to morph the protein folds. The original Paracelsus challenge, met by several groups, called for the engineering of a stable but different structure by modifying less than 50% of the amino acid residues. We have used the sequences from the Protein Data Bank (PDB) identifiers 1ROP, and 2CRO, which were previously used in the Paracelsus challenge by those groups, and suggest mutation to CKAAPs to morph the protein fold. The total number of mutations suggested is less than 40% of the starting sequence theoretically improving the challenge results. From secondary structure prediction experiments of the proposed mutant sequence structures, we observe that each of the suggested mutant protein sequences likely folds to a different, non-native potentially stable target structure. These results are an early indicator that analyses using structure alignments leading to CKAAPs of a given structure are of value in protein engineering experiments.
More Related Videos
Related Concept Videos
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
 01:49
Protein Folding
01:49Protein Folding
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
01:49Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to form...