Spin densities in two-component relativistic density functional calculations: noncollinear versus collinear approach
Related Experiment Video
Updated: Aug 18, 2026
Spatial Separation of Molecular Conformers and Clusters
Published on: January 9, 2014
1Technische Universität Berlin, Sekr. C3, Strasse des 17. Juni 115, D-10623 Berlin, Germany. Christoph.vanWullen@TU-Berlin.de

Abstract:
With present day exchange-correlation functionals, accurate results in nonrelativistic open shell density functional calculations can only be obtained if one uses functionals that do not only depend on the electron density but also on the spin density. We consider the common case where such functionals are applied in relativistic density functional calculations. In scalar-relativistic calculations, the spin density can be defined conventionally, but if spin-orbit coupling is taken into account, spin is no longer a good quantum number and it is not clear what the "spin density" is. In many applications, a fixed quantization axis is used to define the spin density ("collinear approach"), but one can also use the length of the local spin magnetization vector without any reference to an external axis ("noncollinear approach"). These two possibilities are compared in this work both by formal analysis and numerical experiments. It is shown that the (nonrelativistic) exchange-correlation functional should be invariant with respect to rotations in spin space, and this only holds for the noncollinear approach. Total energies of open shell species are higher in the collinear approach because less exchange energy is assigned to a given Kohn-Sham reference function. More importantly, the collinear approach breaks rotational symmetry, that is, in molecular calculations one may find different energies for different orientations of the molecule. Data for the first ionization potentials of Tl, Pb, element 113, and element 114, and for the orientation dependence of the total energy of I+2 and PbF indicate that the error introduced by the collinear approximation is approximately 0.1 eV for valence ionization potentials, but can be much larger if highly ionized open shell states are considered. Rotational invariance is broken by the same amount. This clearly indicates that the collinear approach should not be used, as the full treatment is easily implemented and does not introduce much more computational effort.
More Related Videos
 06:34
In Situ Monitoring of Diffusion of Guest Molecules in Porous Media Using Electron Paramagnetic Resonance Imaging
Published on: September 2, 2016
 08:55
Methods of Ex Situ and In Situ Investigations of Structural Transformations: The Case of Crystallization of Metallic Glasses
Published on: June 7, 2018
Related Concept Videos
 03:06
The Pauli Exclusion Principle
 01:08
NMR Spectroscopy: Spin–Spin Coupling
 01:08
Spin–Spin Coupling Constant: Overview
Qualitatively, any spin plus-half nucleus polarizes the spins of its electrons to the minus-half state. Consequently, the paired electron in the hydrogen–carbon bond must have a...
 01:17
Spin–Spin Coupling: One-Bond Coupling
 01:20
Spin–Spin Coupling: Two-Bond Coupling (Geminal Coupling)
The central atom need not be NMR-active because its electrons are affected by the electron polarization of the spin-active atoms. However, spin information is transmitted less effectively than in one-bond coupling, and 2J values are usually weaker than 1J values. The energy of...
 01:22
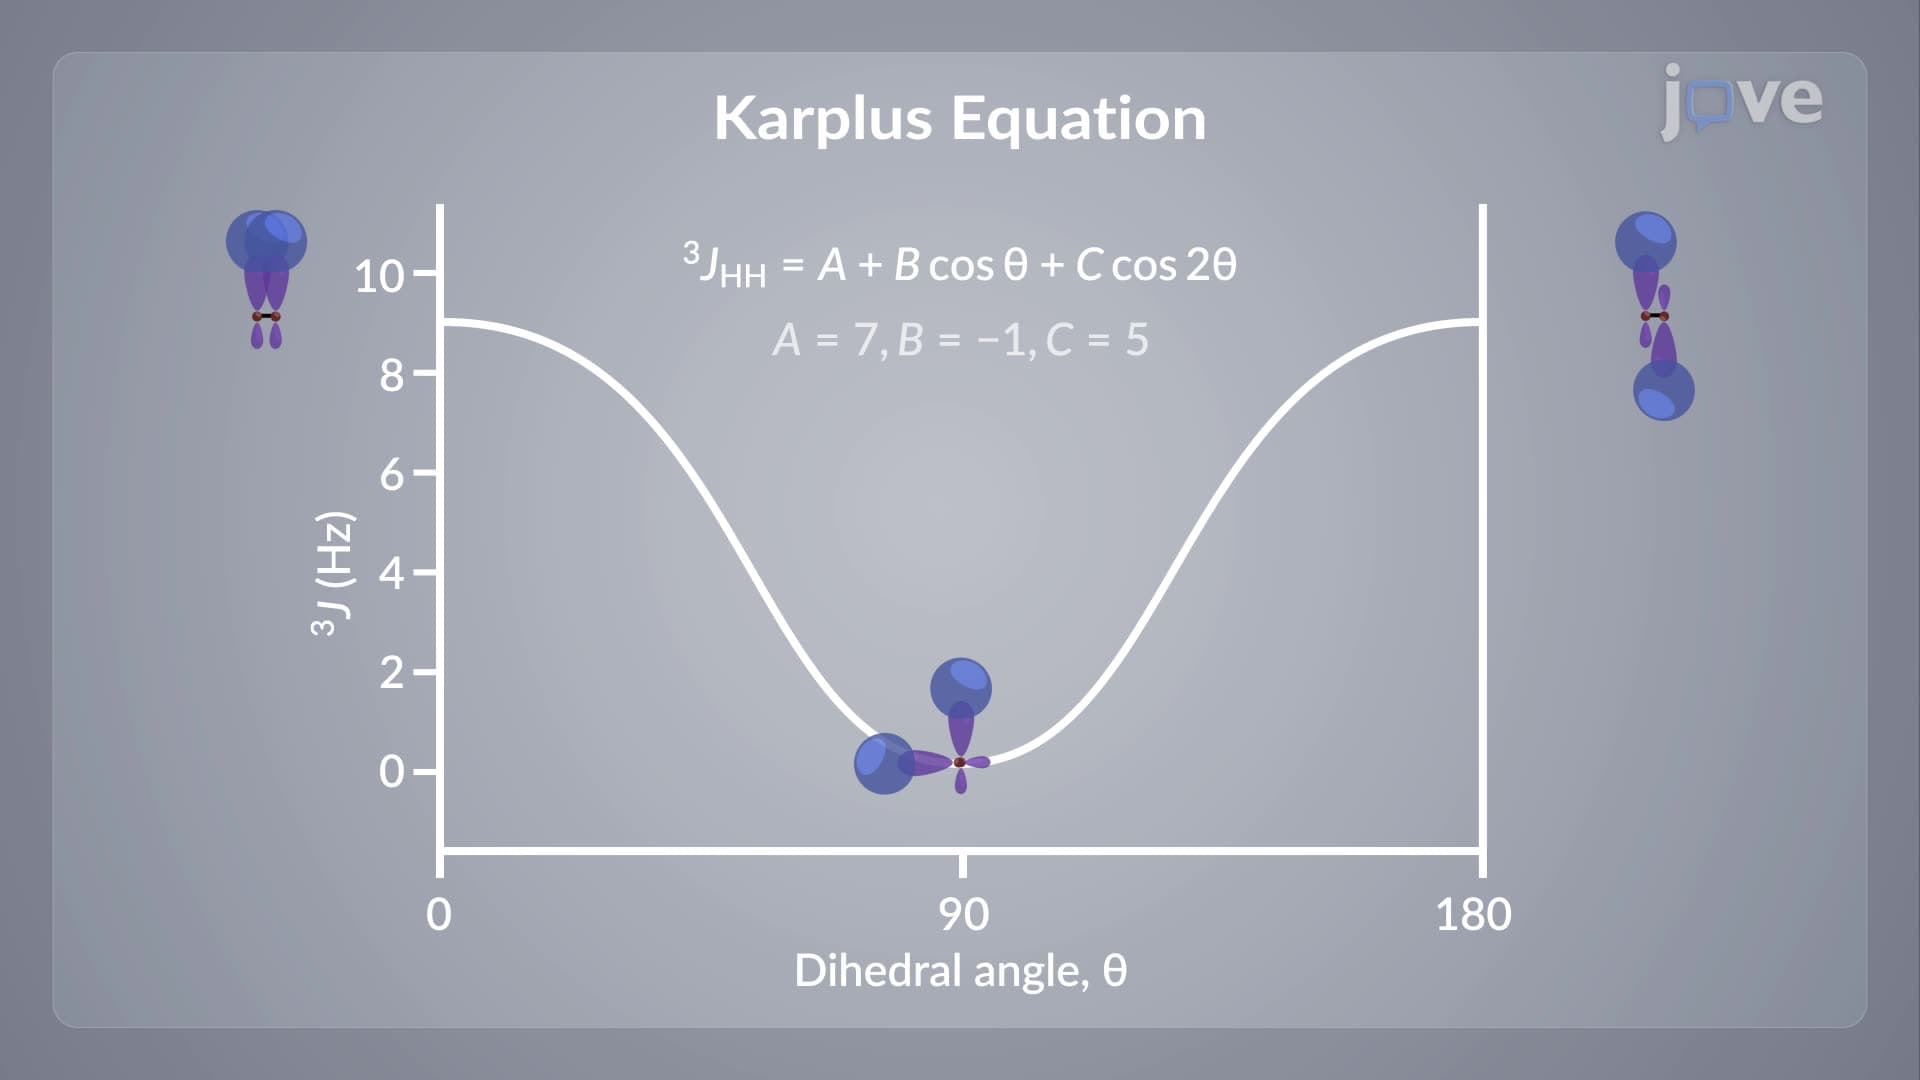
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the involved orbitals. The...