Related Experiment Video
Updated: Aug 2, 2026

A Practical Guide to Phylogenetics for Nonexperts
Published on: February 5, 2014
A fast method for approximating maximum likelihoods of phylogenetic trees from nucleotide sequences
1Department of Biological Sciences, University of New Orleans, New Orleans, Louisiana 70148, USA. jsrbs@uno.edu

Abstract:
We have developed a rapid parsimony method for reconstructing ancestral nucleotide states that allows calculation of initial branch lengths that are good approximations to optimal maximum-likelihood estimates under several commonly used substitution models. Use of these approximate branch lengths (rather than fixed arbitrary values) as starting points significantly reduces the time required for iteration to a solution that maximizes the likelihood of a tree. These branch lengths are close enough to the optimal values that they can be used without further iteration to calculate approximate maximum-likelihood scores that are very close to the "exact" scores found by iteration. Several strategies are described for using these approximate scores to substantially reduce times needed for maximum-likelihood tree searches.
Related Concept Videos
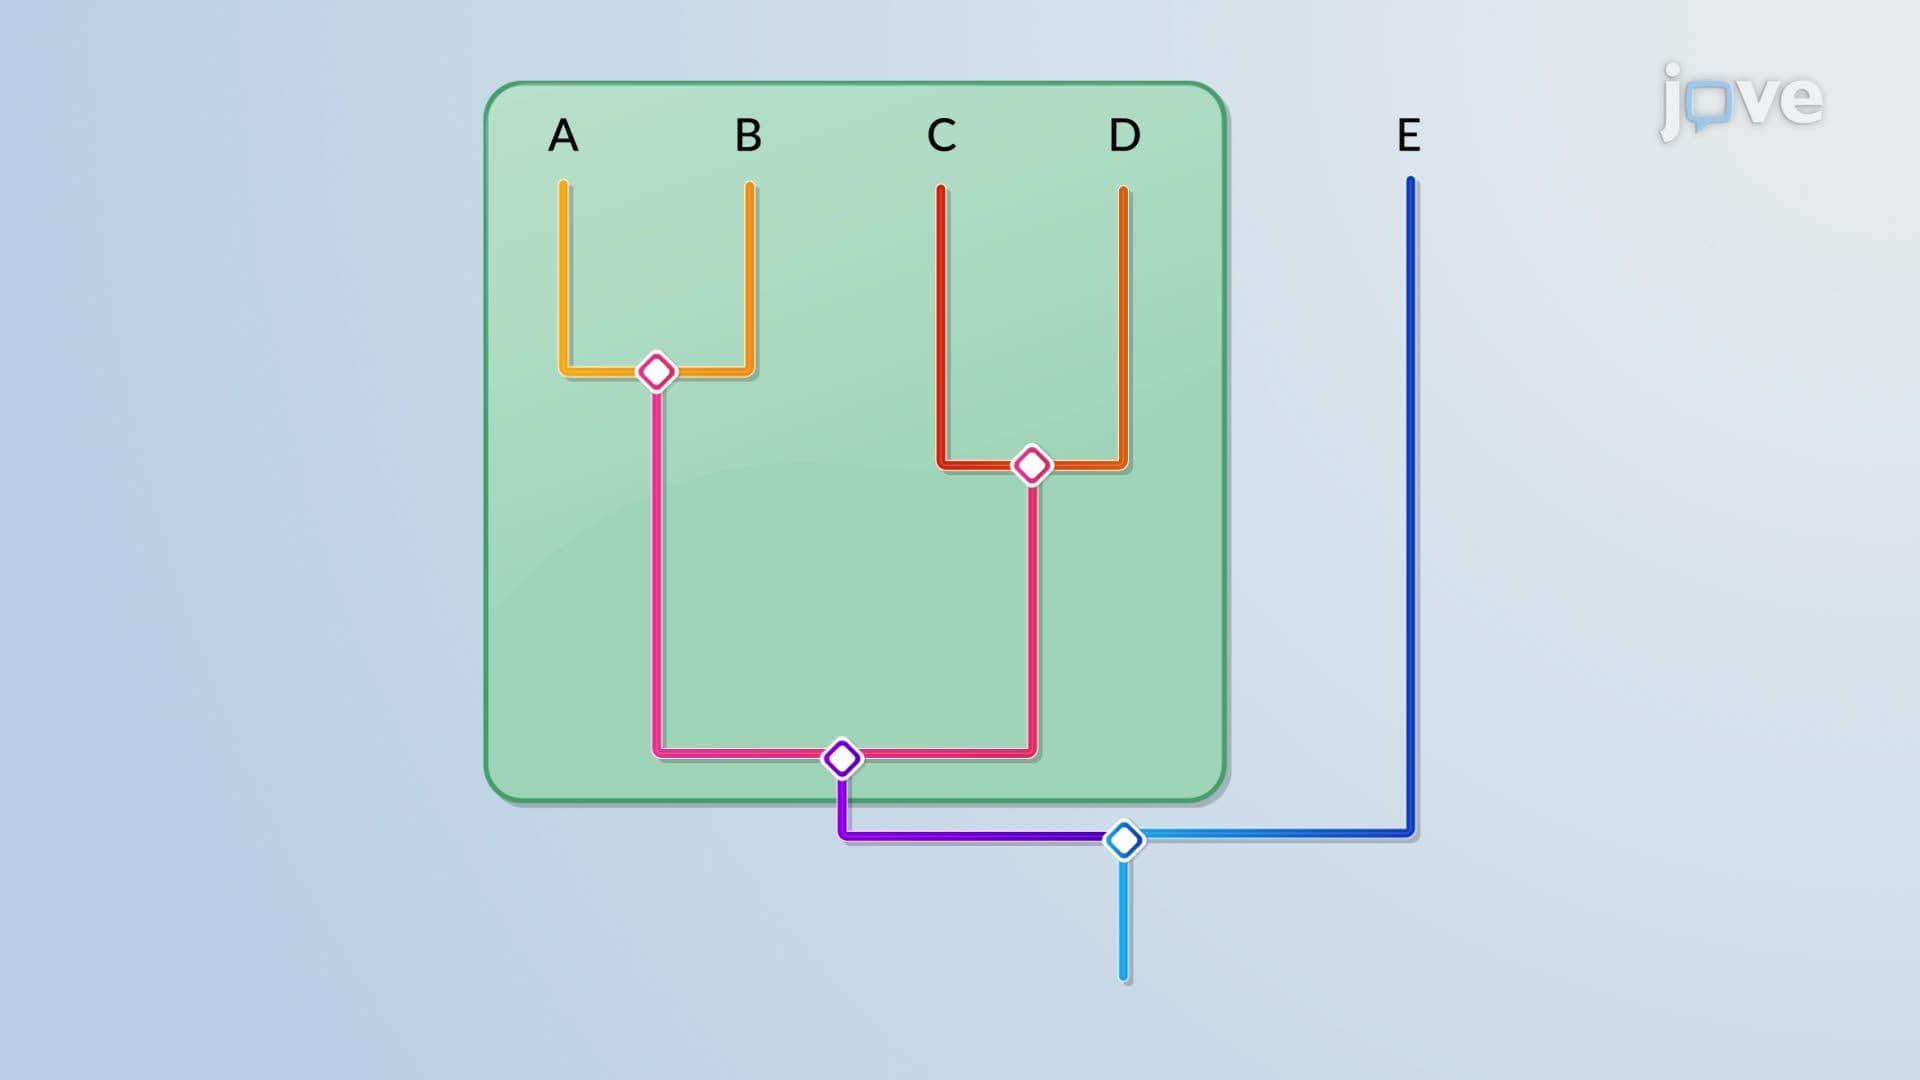 03:21
Phylogenetic Trees
 02:05
Gene Evolution - Fast or Slow?
In contrast, regions which code...
 02:54
Evolutionary Relationships through Genome Comparisons
03:21Phylogenetic Trees
02:05Gene Evolution - Fast or Slow?
In contrast, regions which code...
 01:28
Microbial Phylogeny