Related Experiment Video
Updated: Sep 30, 2026
Functionalized Spirocyclic Heterocycle Synthesis and Cytotoxicity Assay
Published on: February 9, 2021
The effect of precision of molecular orbital descriptors on toxicity modeling of selected pyridines
J R Seward1, M T D Cronin, T W Schultz
1Graduate Program in Ecology and Evolutionary Biology, The University of Tennessee, Knoxville 37996-4500, USA.

Abstract:
The response-surface approach to QSARs attempts to model toxic potency of diverse groups of chemicals while avoiding problems associated with the identification of the mechanism of toxic action or specific chemical class often associated with other approaches. However, while hydrophobicity-dependent, simple regression QSARs derived for congeneric series of organic compounds typically have coefficients of determination greater than 0.90, more heterogeneous multiple regression QSARs exhibit typically 10-15% more unexplained variability. One difference between these approaches is the use of a quantum chemical (QC) descriptor, particularly molecular orbital (MO) energy values such as the energy of the lowest unoccupied molecular orbital (E(LUMO)). The reduced statistical fit exhibited by QSAR models, which include these QC-MO descriptors, could be a result of the variability inherent in the calculation of these descriptors. The present investigation with a structurally and mechanistically diverse set of pyridines revealed that variability is associated with the calculation of the MO descriptor E(LUMO) both between selected Hamiltonians and selected software packages. However, this variability in no way affects the statistical significance of QSARs for toxicity using these values. Specifically, the E(LUMO) values calculated with the PM3 and AM1 Hamiltonians in the two software packages were highly related. There was no relationship between molecular complexity or chemical reactivity and increased differences in individual ELUMO values as described by the standard errors of the mean. Although nine appeared to be the number of calculations, which best minimizes the standard error in energy values relative to computational costs; this minimization did not alter the statistics of the QSARs derived with single vs. mean E(LUMO) values. While the energy of the highest occupied molecular orbital (E(HOMO)) values were not used in the modeling of toxicity, a comparison of these values revealed greater variability between the Hamiltonians and software packages than observed for ELUMO values. Examination of the magnitudes of standard error of the E(HOMO) values in connection to structural features or reactivity likewise revealed no trends.
Related Concept Videos
 01:25
Basicity of Heterocyclic Aromatic Amines
 01:24
Drug Toxicity: Dose-Dependent Reactions
 01:18
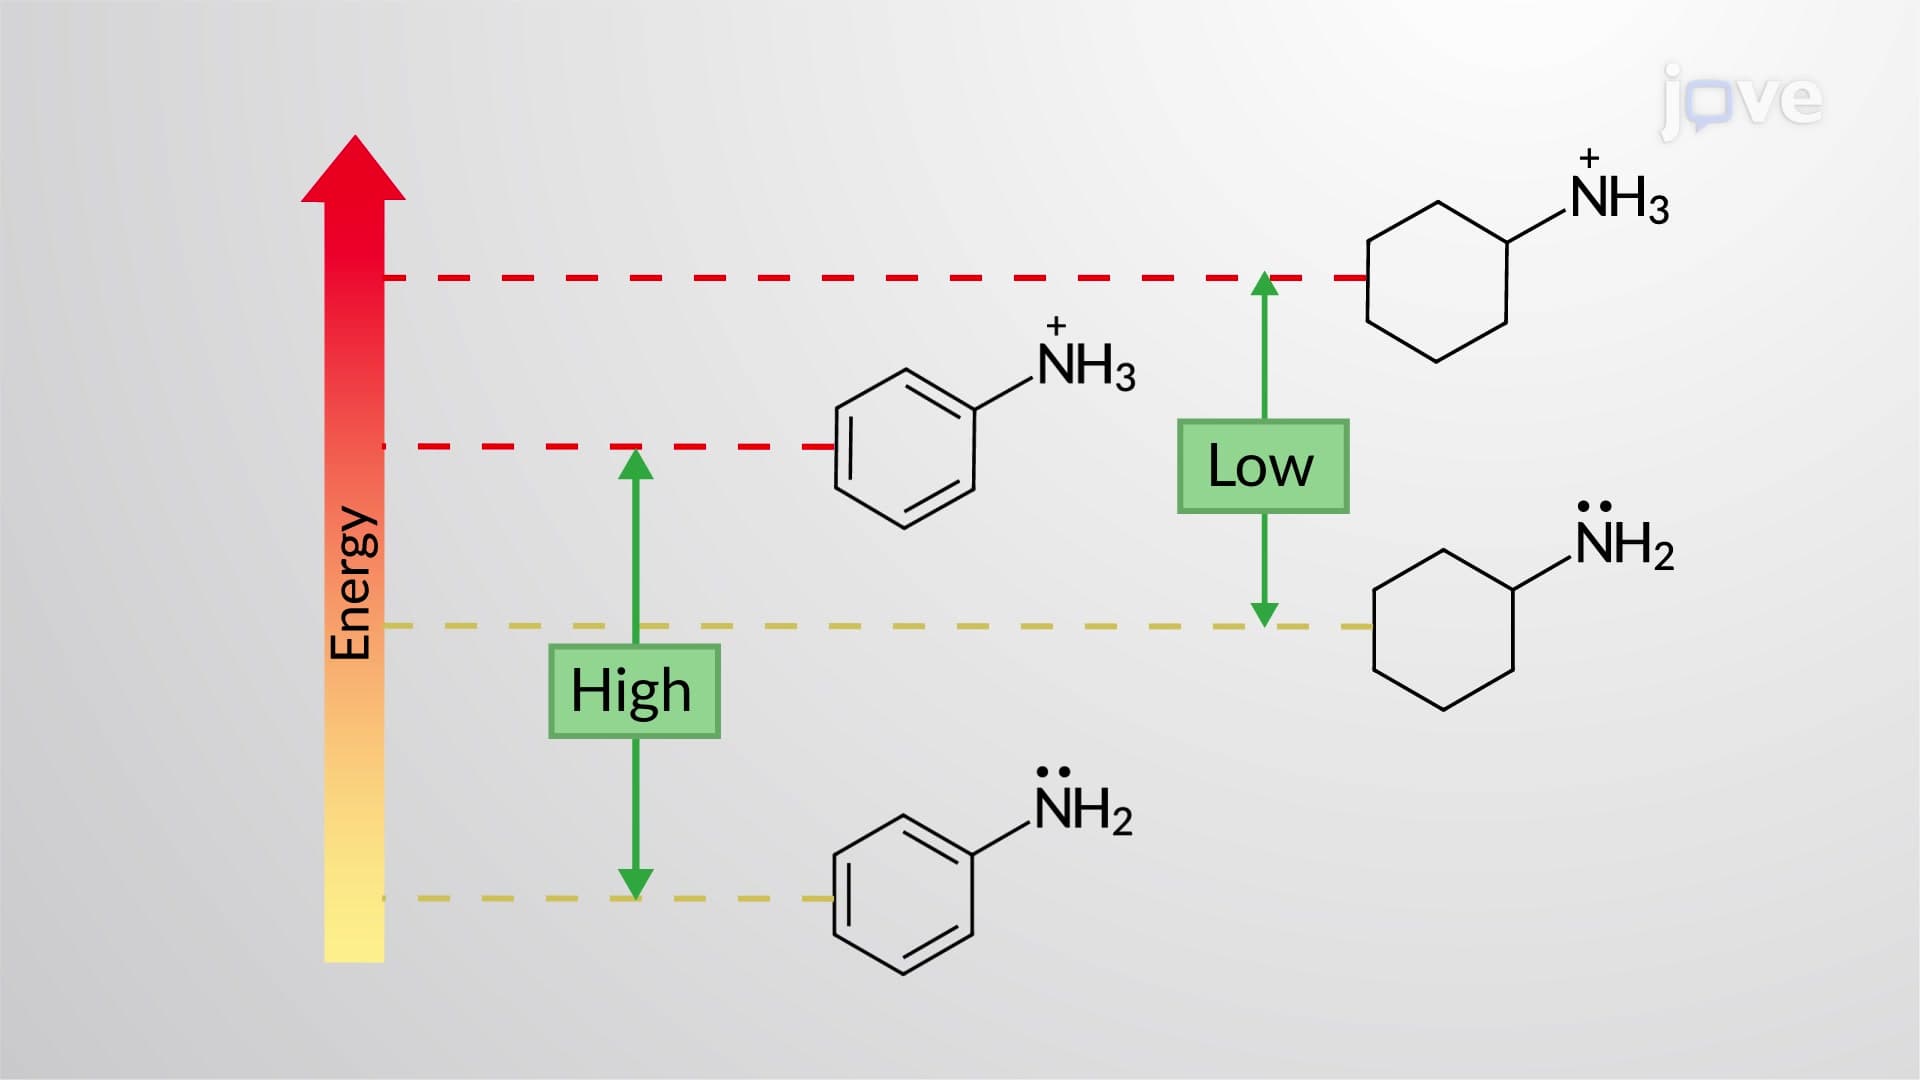
Basicity of Aromatic Amines
 01:26
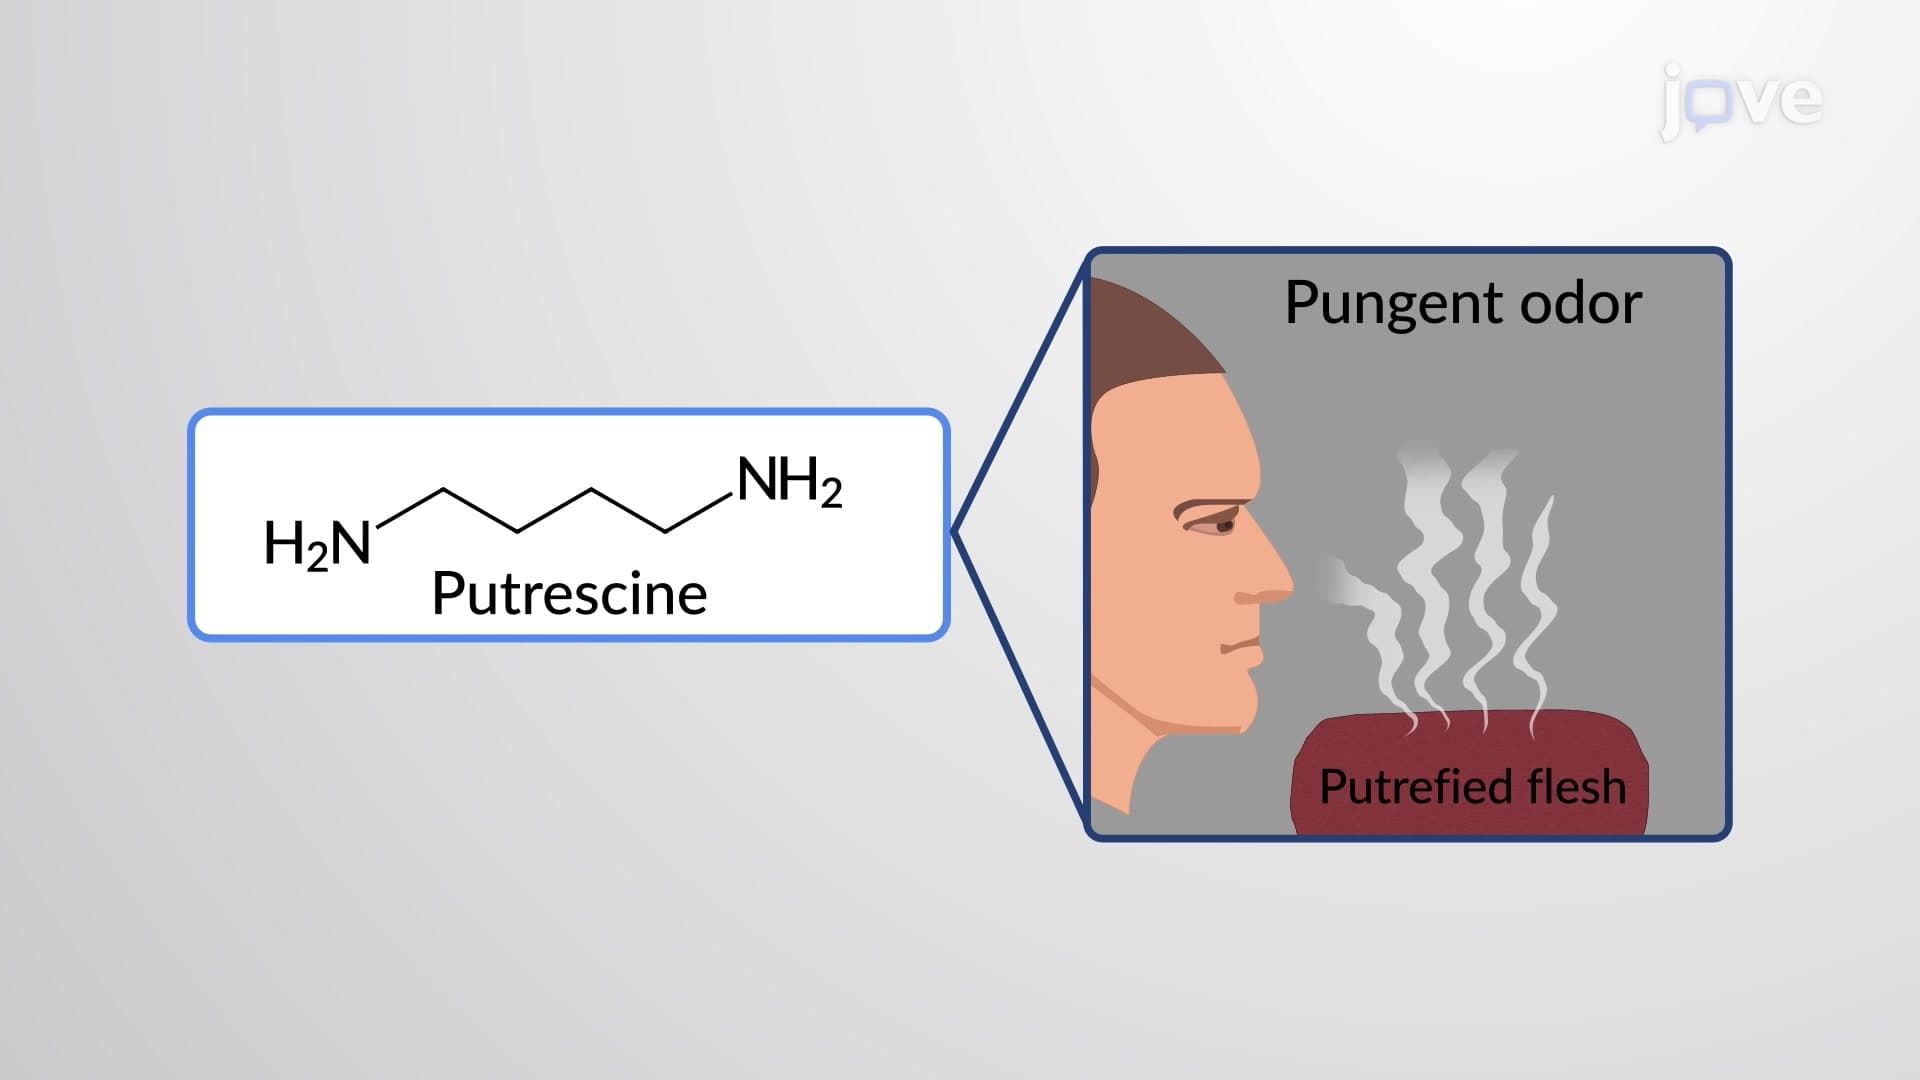
Physical Properties of Amines
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 03:51
Molecular Orbital Theory II