Related Experiment Video
Updated: Aug 9, 2026

Methods of Ex Situ and In Situ Investigations of Structural Transformations: The Case of Crystallization of Metallic Glasses
Published on: June 7, 2018
(57)Fe Mössbauer isomer shifts of heme protein model systems: electronic structure calculations
Yong Zhang1, Junhong Mao, Eric Oldfield
1Department of Chemistry, University of Illinois at Urbana-Champaign, 600 South Mathews Avenue, Urbana, IL 61801, USA.

Abstract:
We report the results of density functional theory (DFT) calculations of the (57)Fe Mössbauer isomer shifts (delta(Fe)) for a series of 24 inorganic, organometallic, and metalloprotein/metalloporphyrin model systems in S = 0, (1)/(2), 1, (3)/(2), 2, and (5)/(2) spin states. We find an excellent correlation between calculation and experiment over the entire 2.34 mm s(-1) range of isomer shifts: a 0.07-0.08 mm s(-1) rms deviation between calculation and experiment (corresponding to 3-4% of the total delta(Fe) range, depending on the functionals used) with R(2) values of 0.973 and 0.981 (p < 0.0001). The best results are obtained by using the hybrid exchange-correlation functional B3LYP, used previously for (57)Fe Mössbauer quadrupole splittings and (57)Fe NMR chemical shifts and chemical shielding anisotropies. The relativistically corrected value of alpha, alpha(rel), converges with the large basis set used in this work, but the exact values vary somewhat with the methods used: -0.253 a(0)(3) mm s(-1) (Hartree-Fock; HF); -0.316 a(0)(3) mm s(-1) (hybrid HF-DFT; B3LYP), or -0.367 a(0)(3) mm s(-1) (pure DFT; BPW91). Both normal and intermediate spin state isomer shifts are well reproduced by the calculations, as is the broad range of delta(Fe) values: from [Fe(VI)O(4)](2-) (-0.90 mm s(-1) expt; -1.01 mm s(-1) calc) to KFe(II)F(3) (1.44 mm s(-1) expt; 1.46 mm s(-1) calc). Molecular orbital analyses of all inorganic solids as well as all organometallic and metalloporphyrin systems studied reveal that there are three major core MO contributions to rho(tot)(0), the total charge density at the iron nucleus (and hence delta(Fe)), that do not vary with changes in chemistry, while the valence MO contributions are highly correlated with delta(Fe) (R(2) = 0.915-0.938, depending on the functionals used), and the correlation between the valence MO contributions and the total MO contribution is even better (R(2) = 0.965-0.976, depending on the functionals used). These results are of general interest since they demonstrate that DFT methods now enable the accurate prediction of delta(Fe) values in inorganic, organometallic, and metalloporphyrin systems in all spin states and over a very wide range of delta(Fe) values with a very small rms error.
Related Concept Videos
 01:18
Electron Orbital Model
 02:18
The Bohr Model
 03:26
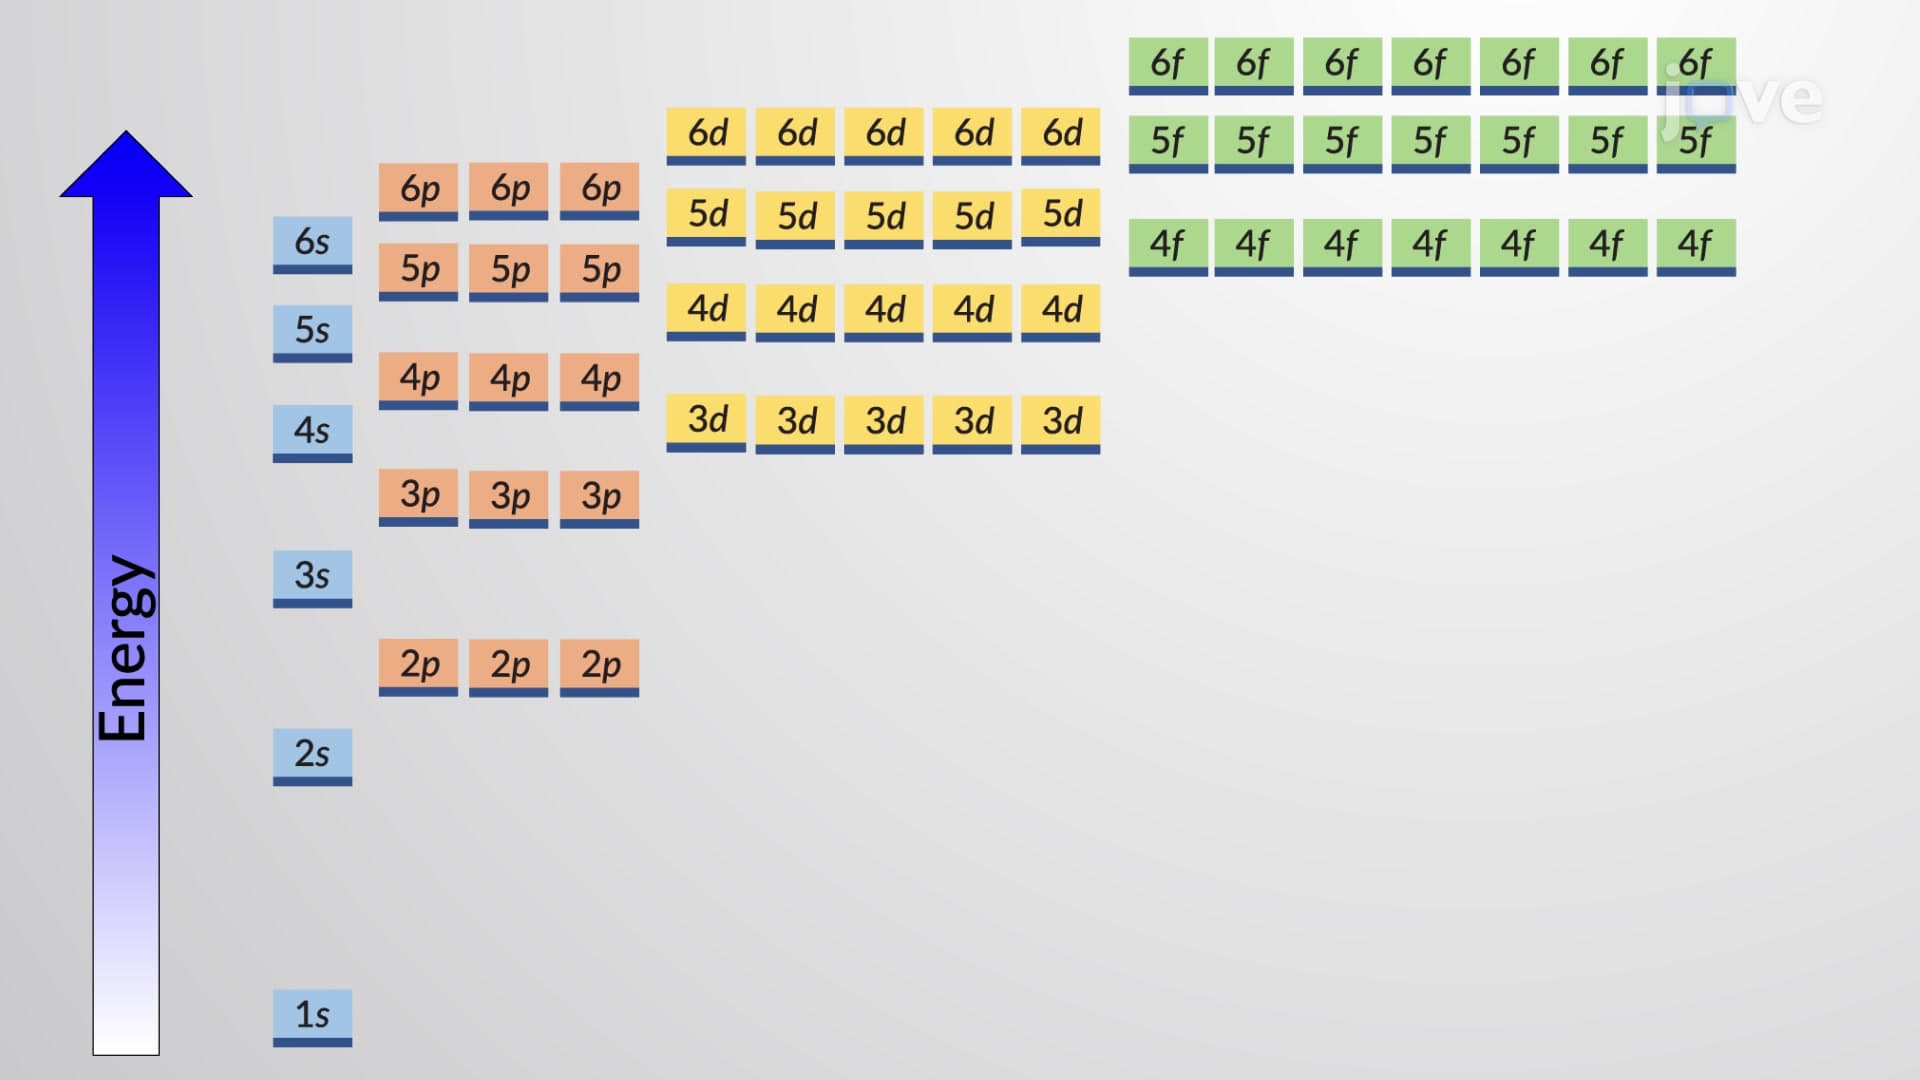
Electron Configuration of Multielectron Atoms
 02:28
Electronic Structure of Atoms
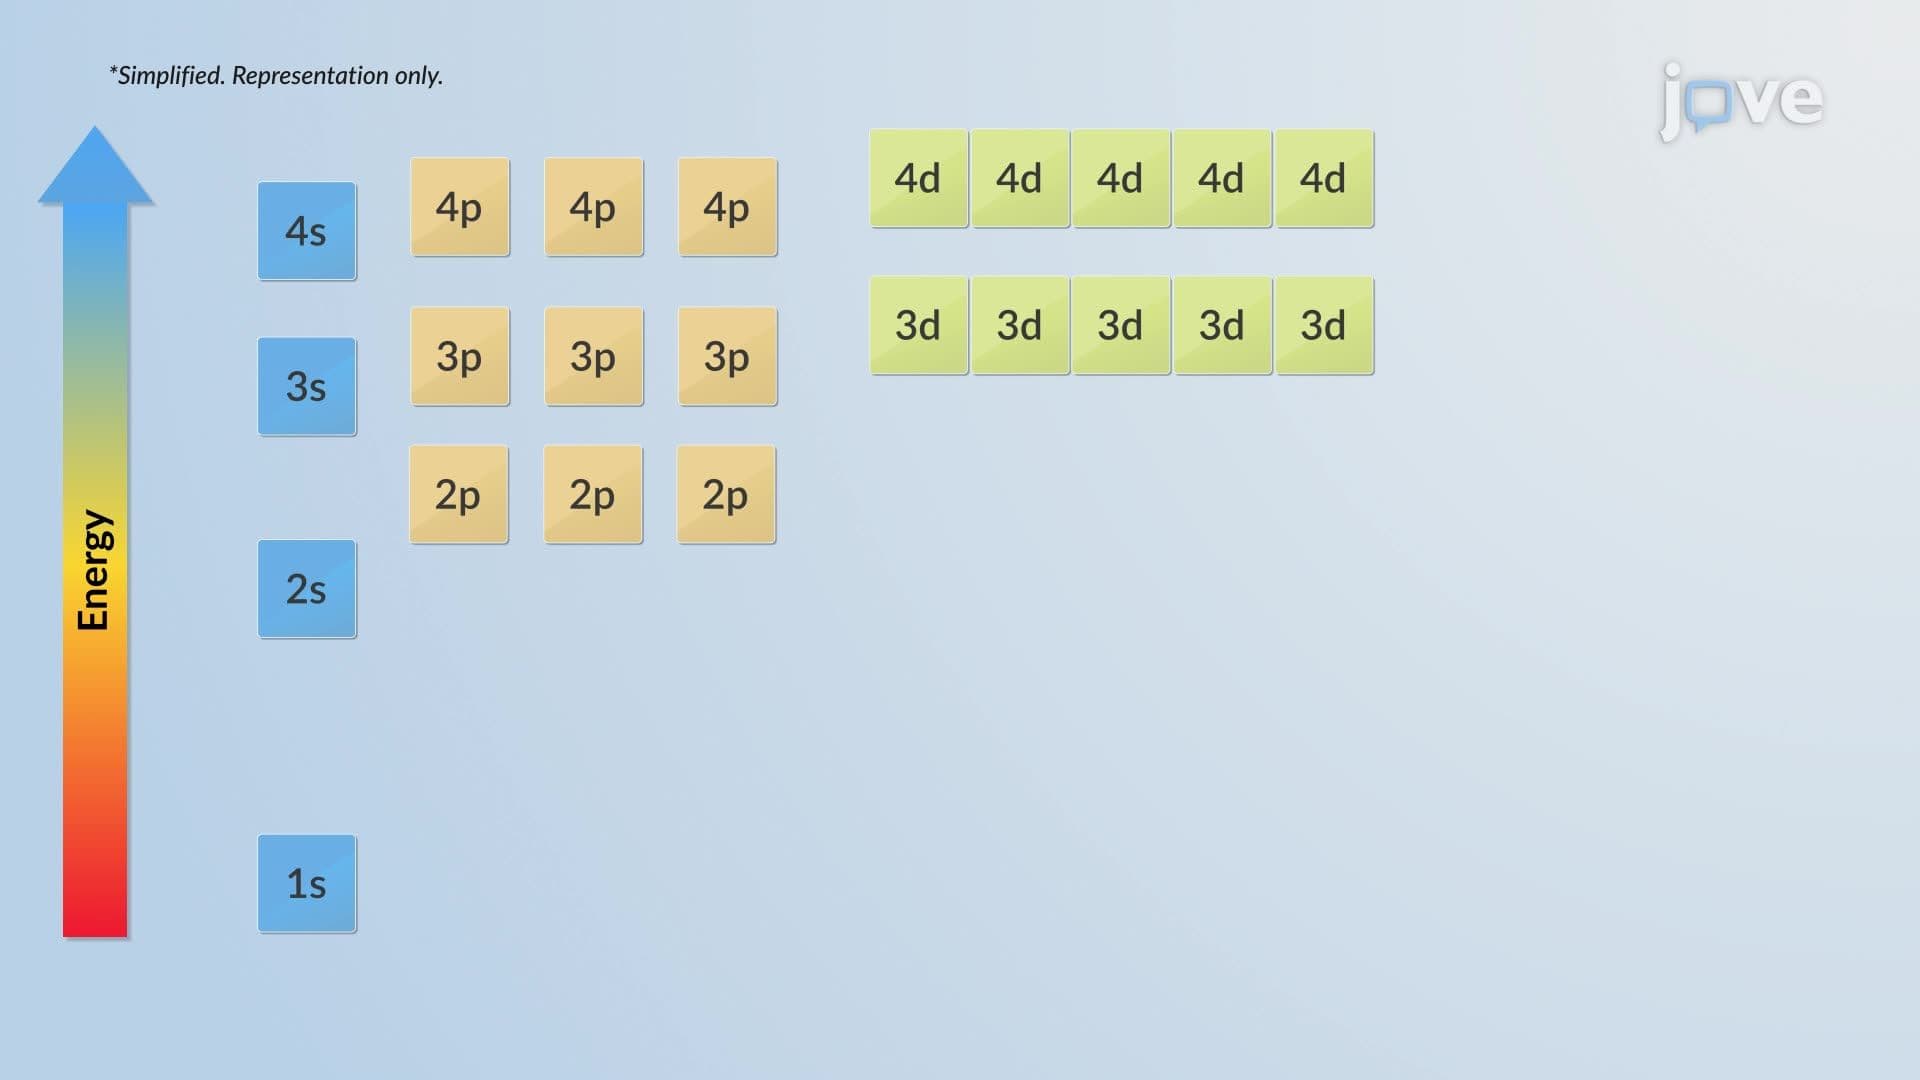
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum numbers: n, l, ml, and...
 01:08
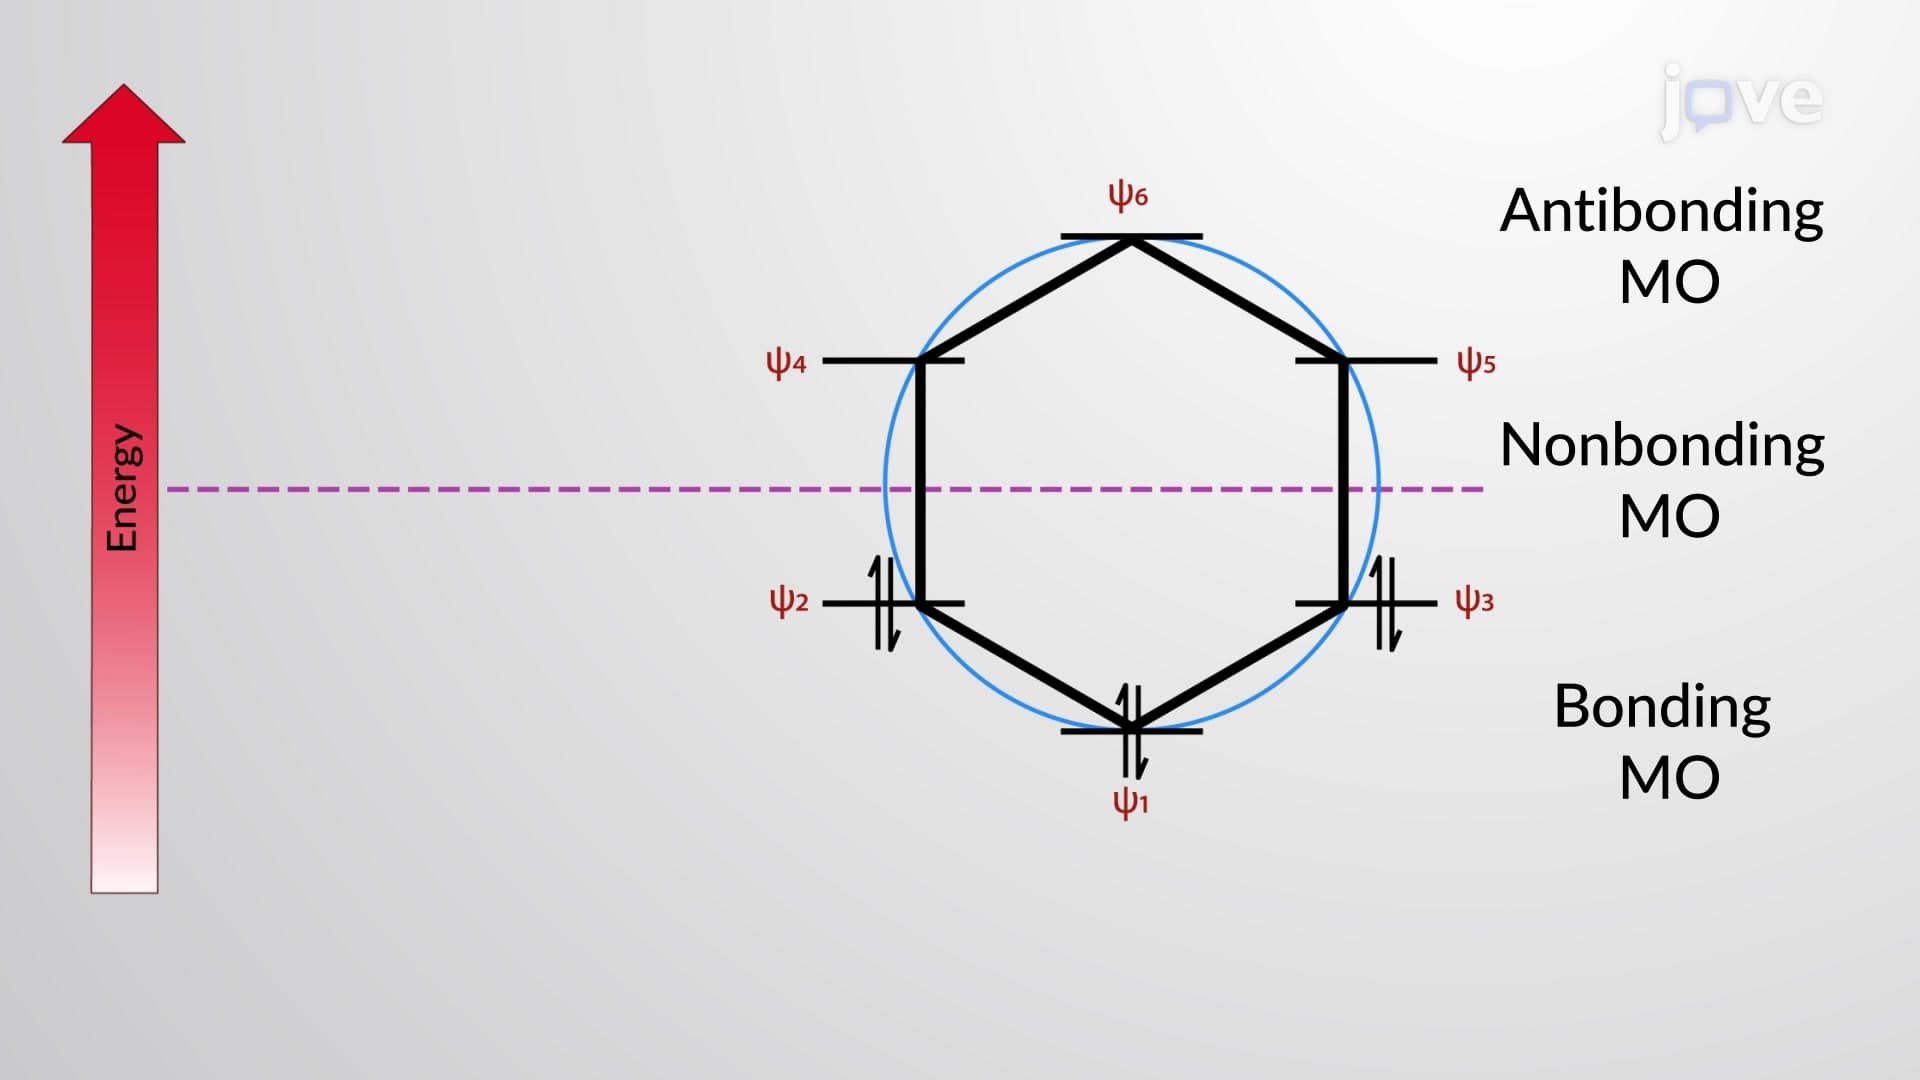
Hückel's Rule Diagram of π MOs: Frost Circle
A Frost circle is constructed by drawing a polygon whose number of edges is equal to the number of carbons of the given cyclic system, with one of the vertices pointing down. Then, a circle is drawn enclosing the polygon so that...
 01:27
π Electron Effects on Chemical Shift: Overview