Related Experiment Video
Updated: Sep 20, 2026
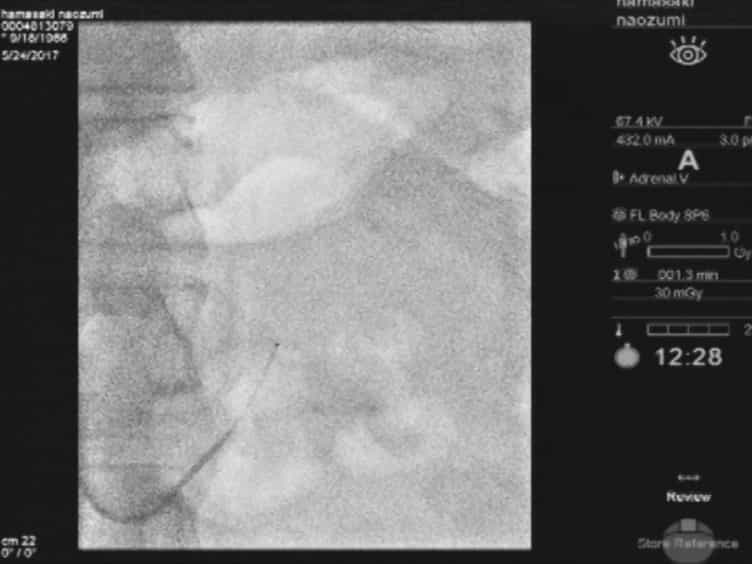
A Novel Method: Super-selective Adrenal Venous Sampling
Published on: September 15, 2017
Early identification of patients with von Hippel-Lindau disease at risk for pheochromocytoma
1Urologic Oncology Branch, National Cancer Institute, 10101 Grosvenor Place, #518, North Bethesda, MD 20852, USA. Maranchj@mail.nih.gov

Abstract:
von Hippel-Lindau disease (VHL) is an autosomal dominant familial syndrome that predisposes to the formation of tumors in multiple organ systems, including adrenal and extra-adrenal pheochromocytomas. However, fewer than 30% of VHL families develop pheochromocytomas. In recent years, this clinical heterogeneity has been correlated with missense mutations. The VHL patient requires vigilant, lifelong biochemical and radiographic screening for pheochromocytoma. Half of VHL pheochromocytomas present bilaterally, and there is a high incidence of recurrence after surgery. Because of the morbidity of bilateral total adrenalectomy with subsequent steroid replacement therapy, the recent therapeutic trend has been toward observation and minimally invasive adrenal-sparing procedures.