Macromolecular prodrugs: X. Kinetics of fenoprofen release from PHEA-fenoprofen conjugate

T van der Merwe1, B Boneschans, B Zorc

  • 1Department of Pharmaceutical Chemistry, Potchefstroom University for CHE, P.B. X6001, Potchefstroom 2520, South Africa.

Related Concept Videos

Drug Metabolism: Phase II Reactions01:14

Drug Metabolism: Phase II Reactions

Phase II reactions are essential for the detoxification and elimination of drugs from the body. These reactions involve the conjugation of parent drugs or their phase I metabolites with endogenous molecules, resulting in more hydrophilic drug conjugates. The primary conjugation reactions in this phase are sulfation and glucuronidation. Both sulfation and glucuronidation typically produce biologically inactive metabolites. However, in some cases involving prodrugs, active metabolites may be...
Nonlinear Pharmacokinetics: Dependence of Elimination Half-Life and Dose Clearance01:23

Nonlinear Pharmacokinetics: Dependence of Elimination Half-Life and Dose Clearance

The elimination half-life and drug clearance of drugs following nonlinear kinetics can vary with dosage. The Michaelis-Menten parameters and drug concentration influence these factors. As the dose increases, the elimination half-life tends to lengthen, resulting in a reduction in clearance and a disproportionately larger area under the curve. The total clearance can be derived from the Michaelis-Menten equation for drugs following a one-compartment model.
A study on guinea pigs examined the...
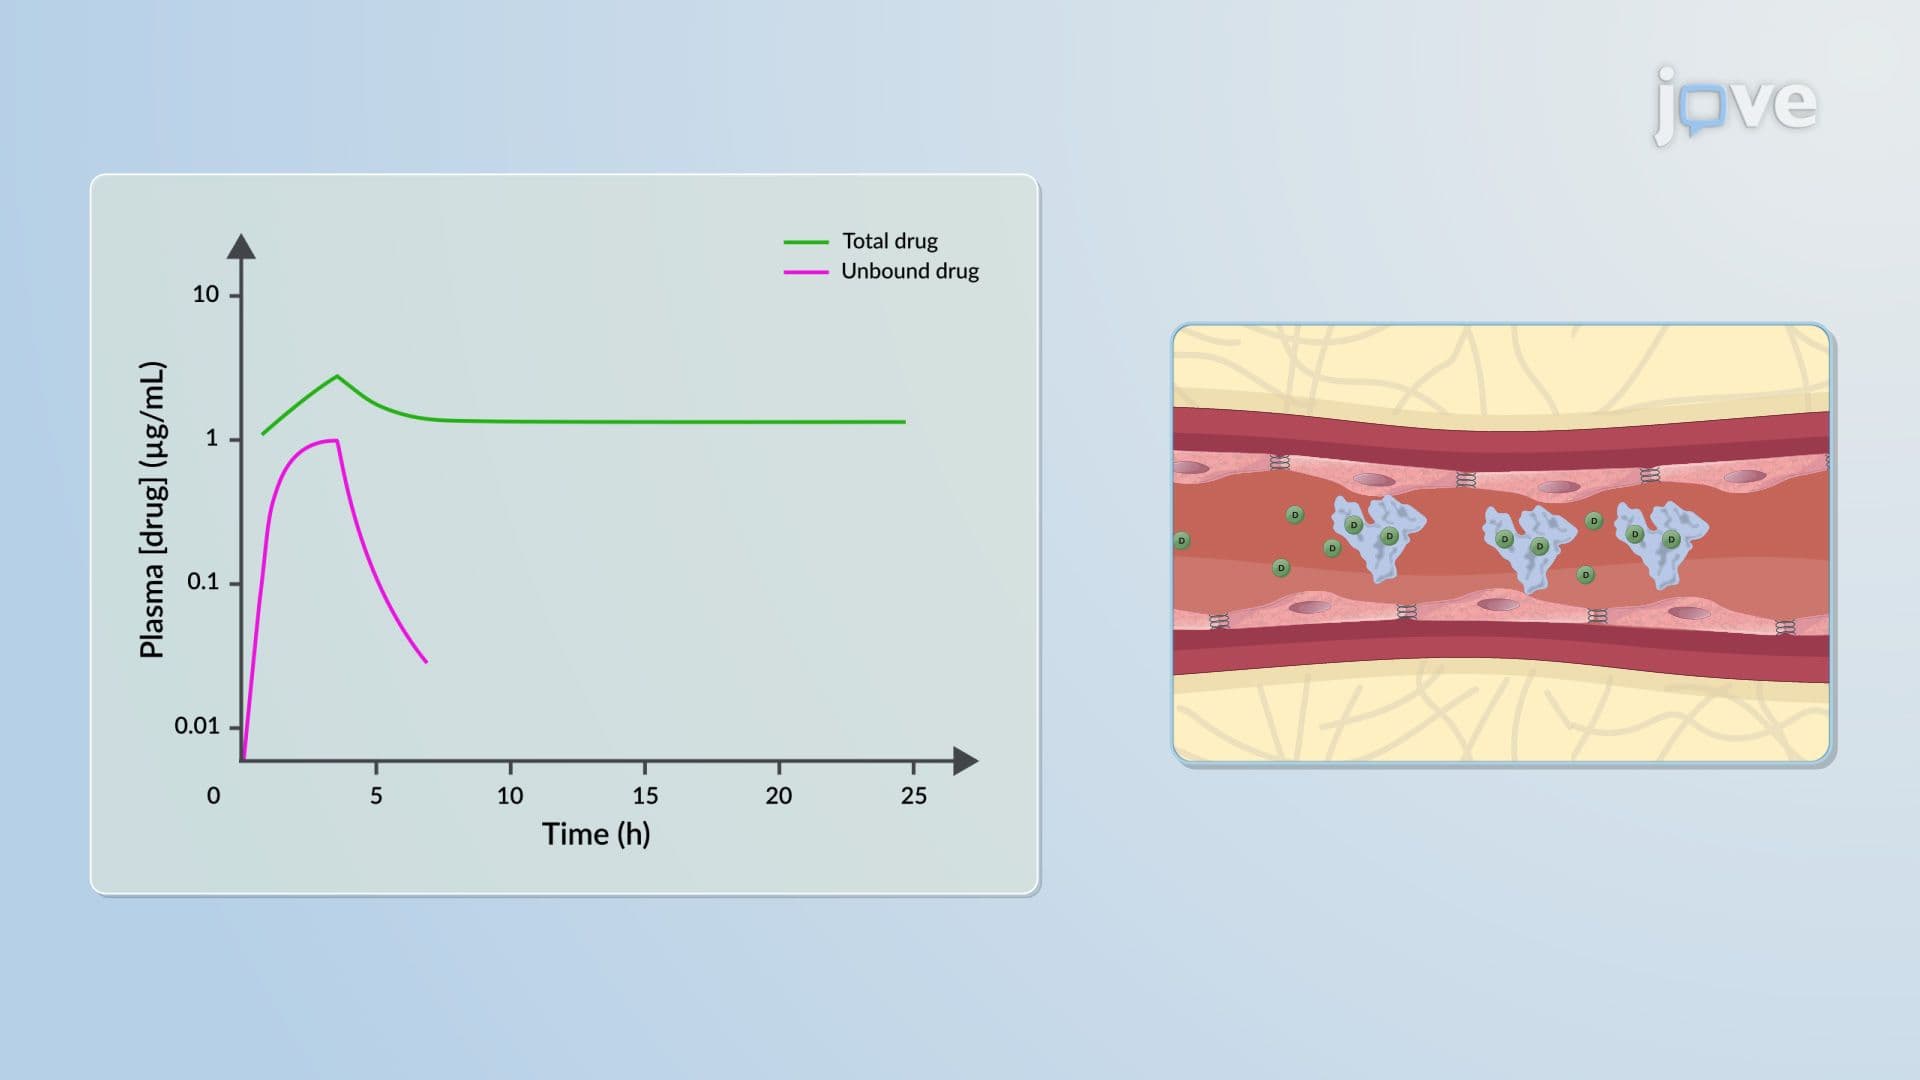
Nonlinear Pharmacokinetics: Bioavailability and Protein-Drug Binding01:22

Nonlinear Pharmacokinetics: Bioavailability and Protein-Drug Binding

When a drug follows nonlinear pharmacokinetics, its bioavailability, the amount of the drug that reaches the systemic circulation, can change with different doses. This is due to the presence of a saturable pathway. The pathway becomes saturated as the drug concentration increases, decreasing the absorption rate. Consequently, the drug's bioavailability may be lower than expected at higher doses.
To quantify the extent of bioavailability, pharmacologists often use a parameter called .
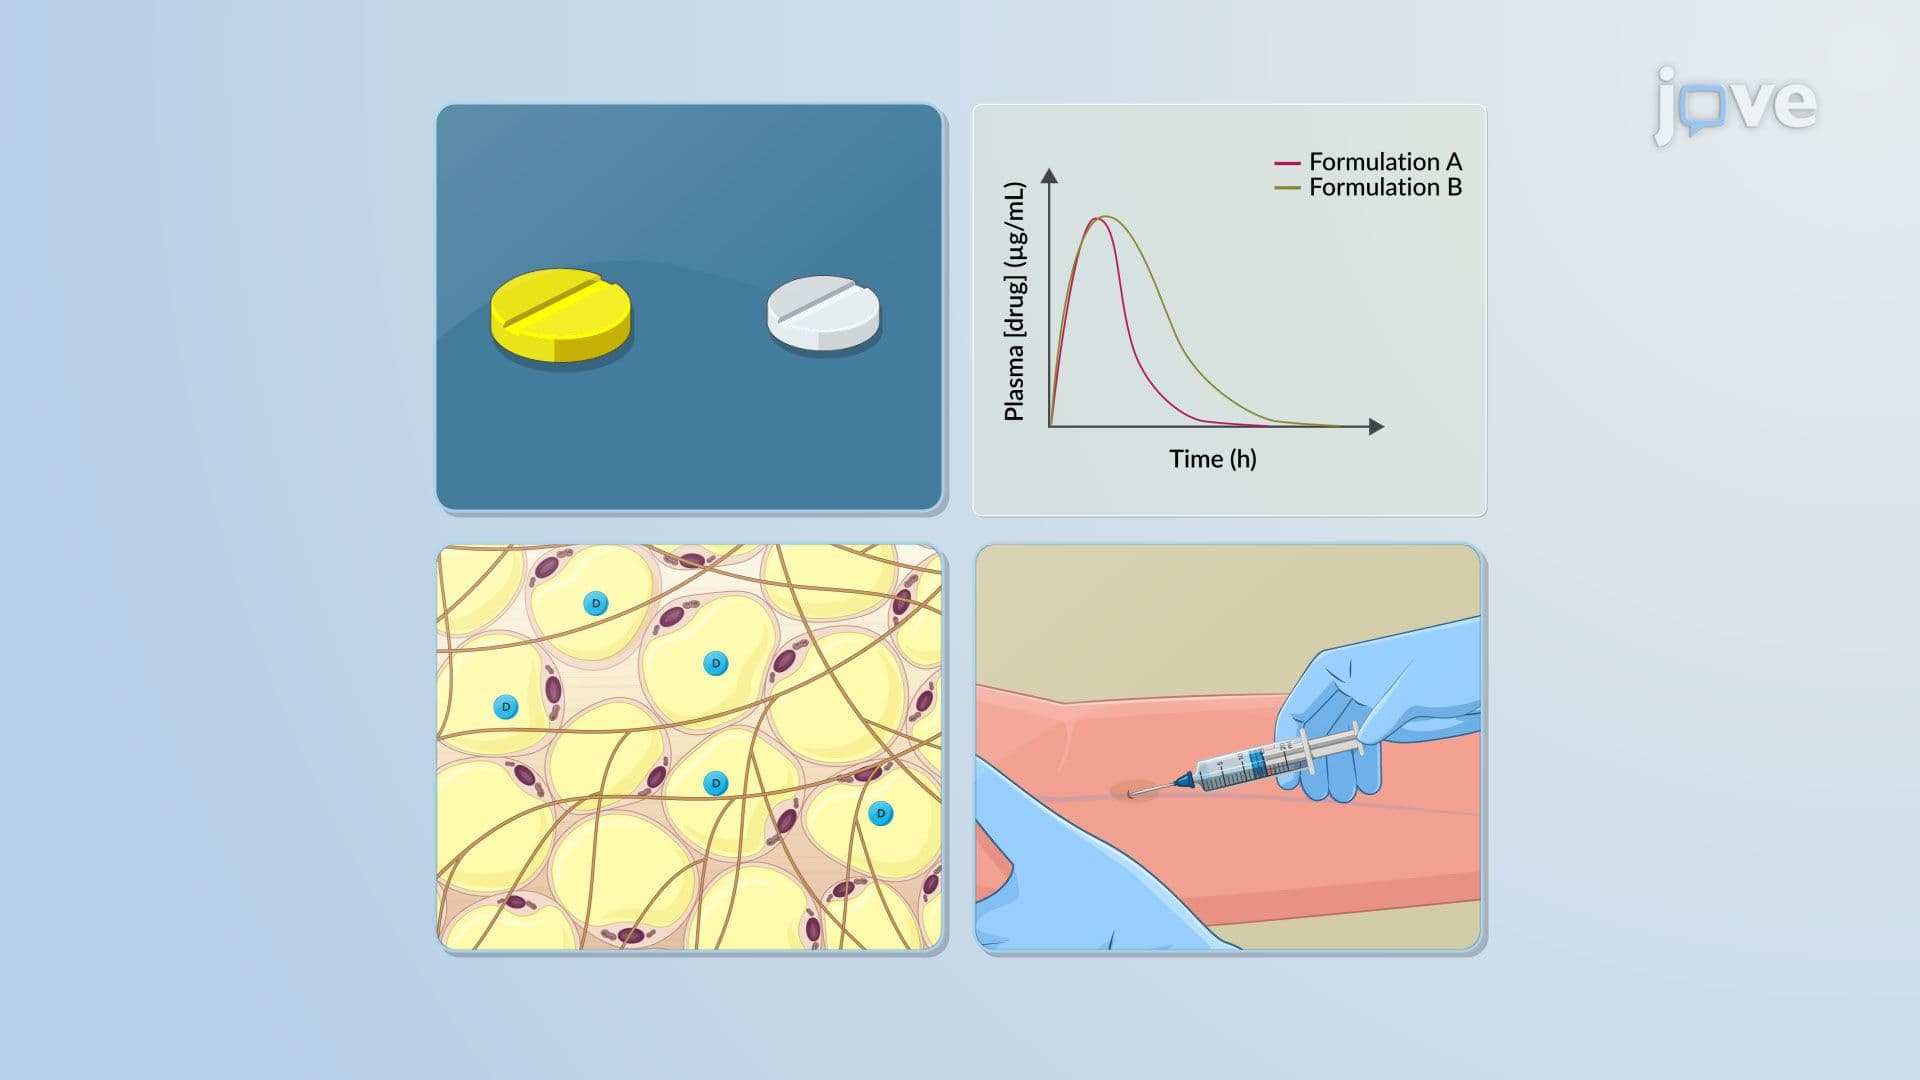
Pharmacokinetic–Pharmacodynamic Relationship: Duration of Dose-Effect Relationship01:14

Pharmacokinetic–Pharmacodynamic Relationship: Duration of Dose-Effect Relationship
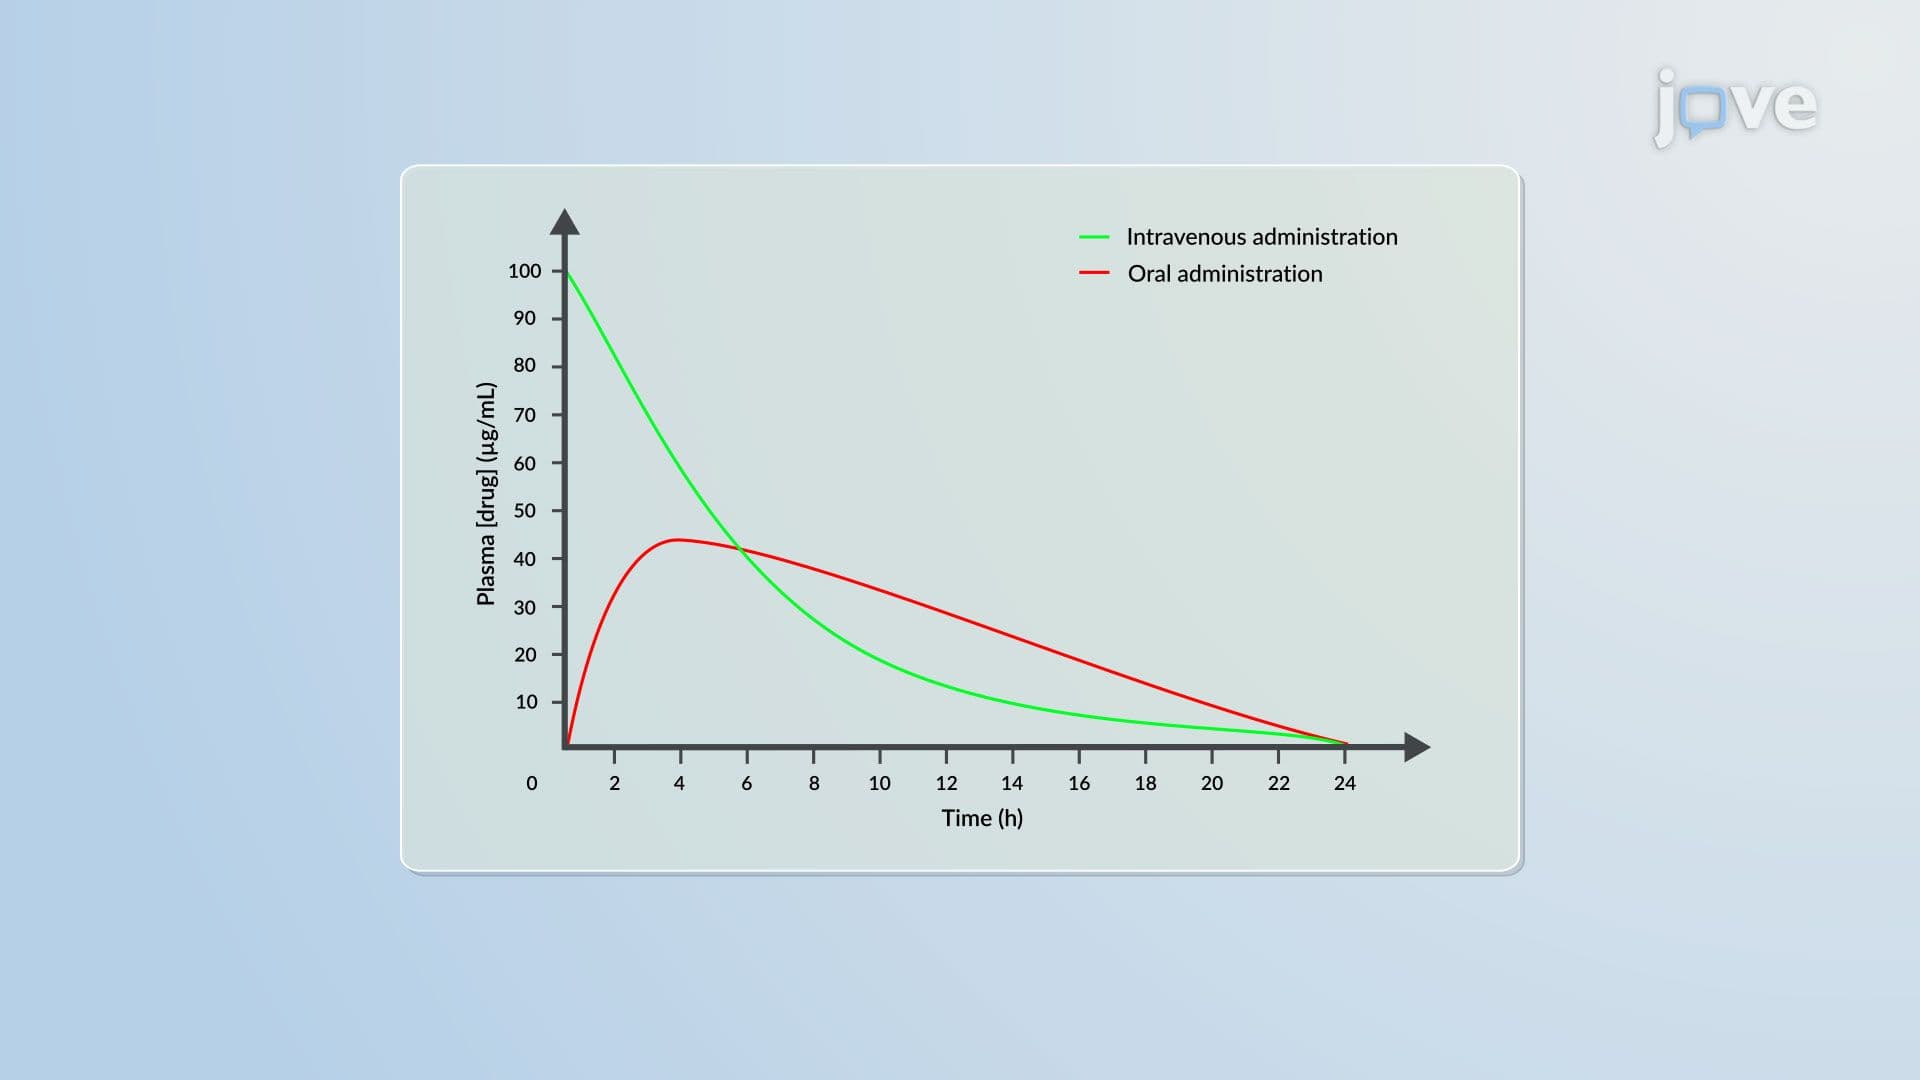
For drugs producing a quantal response, onset occurs when plasma concentration reaches a minimum effective level (Cmin). The drug's action duration depends on how long the plasma concentration remains above Cmin.Two primary factors influence this duration: dose size and the rate of drug removal from the action site. Both depend on the drug's redistribution to poorly perfused tissues and elimination processes. A larger dose promotes rapid onset and prolongs the effect's duration.Consider a...
Pharmacokinetic–Pharmacodynamic Relationship: Problems01:24

Pharmacokinetic–Pharmacodynamic Relationship: Problems
The empirical approach to drug therapy optimization relies on correlating pharmacological response with administered dosage. Such an approach can be costly, time-consuming, and often yields poor correlation due to variables like formulation factors and drug elimination characteristics. A more precise approach correlates response with plasma drug concentration or the amount of drug in the body, rather than dosage. This is achieved through pharmacokinetic-pharmacodynamic (PK/PD) modeling, which...
Pharmacokinetic–Pharmacodynamic Relationship: Model Components01:14

Pharmacokinetic–Pharmacodynamic Relationship: Model Components

Pharmacokinetic-pharmacodynamic (PK–PD) modeling is essential in drug development and clinical pharmacology. It provides a quantitative framework to predict drug behavior and response over time. This approach integrates pharmacokinetics (PK), which describes the drug's absorption, distribution, metabolism, and excretion, with pharmacodynamics (PD), which characterizes the drug’s biological effects and mechanisms of action.The disposition kinetics of a drug determine its plasma...