Related Experiment Video
Updated: Aug 4, 2026

A Practical Guide to Phylogenetics for Nonexperts
Published on: February 5, 2014
Multiple data sets, congruence, and hypothesis testing for the phylogeny of basal groups of the lizard genus
O Flores-Villa1, K M Kjer, M Benabib
1Department of Zoology and M. L. Bean Museum, Brigham Young University, Provo, Utah 84602, USA. ofv@hp.fciencias.unam.mx

Abstract:
Several data partitions, including nuclear and mitochondrial gene sequences, chromosomes, isoenzymes, and morphological characters, were used to propose a new phylogeny and to test previously published hypotheses about the phylogenetic positions of basal clades of the lizard genus Sceloporus and the relationship of Sceloporus to the former genus "Sator". In accord with earlier studies, our results grouped "Sator" as internal to Sceloporus, and both support a hypothesis of transgulfian vicariance for the origin of the former genus "Sator" on islands in the Sea of Cortez. Robustness of support for internal nodes in our best tree was established though widely used indices (bootstrap proportions, decay values) but also through congruence among independent data partitions. Several deep nodes in the tree recovered by several methods, including equally weighted and differentially weighted parsimony and maximum likelihood models, are only weakly supported by the traditional indices. This methodological concordance is taken as evidence for insensitivity of the deep structure of the topology to alternative assumptions.
Related Concept Videos
 01:23
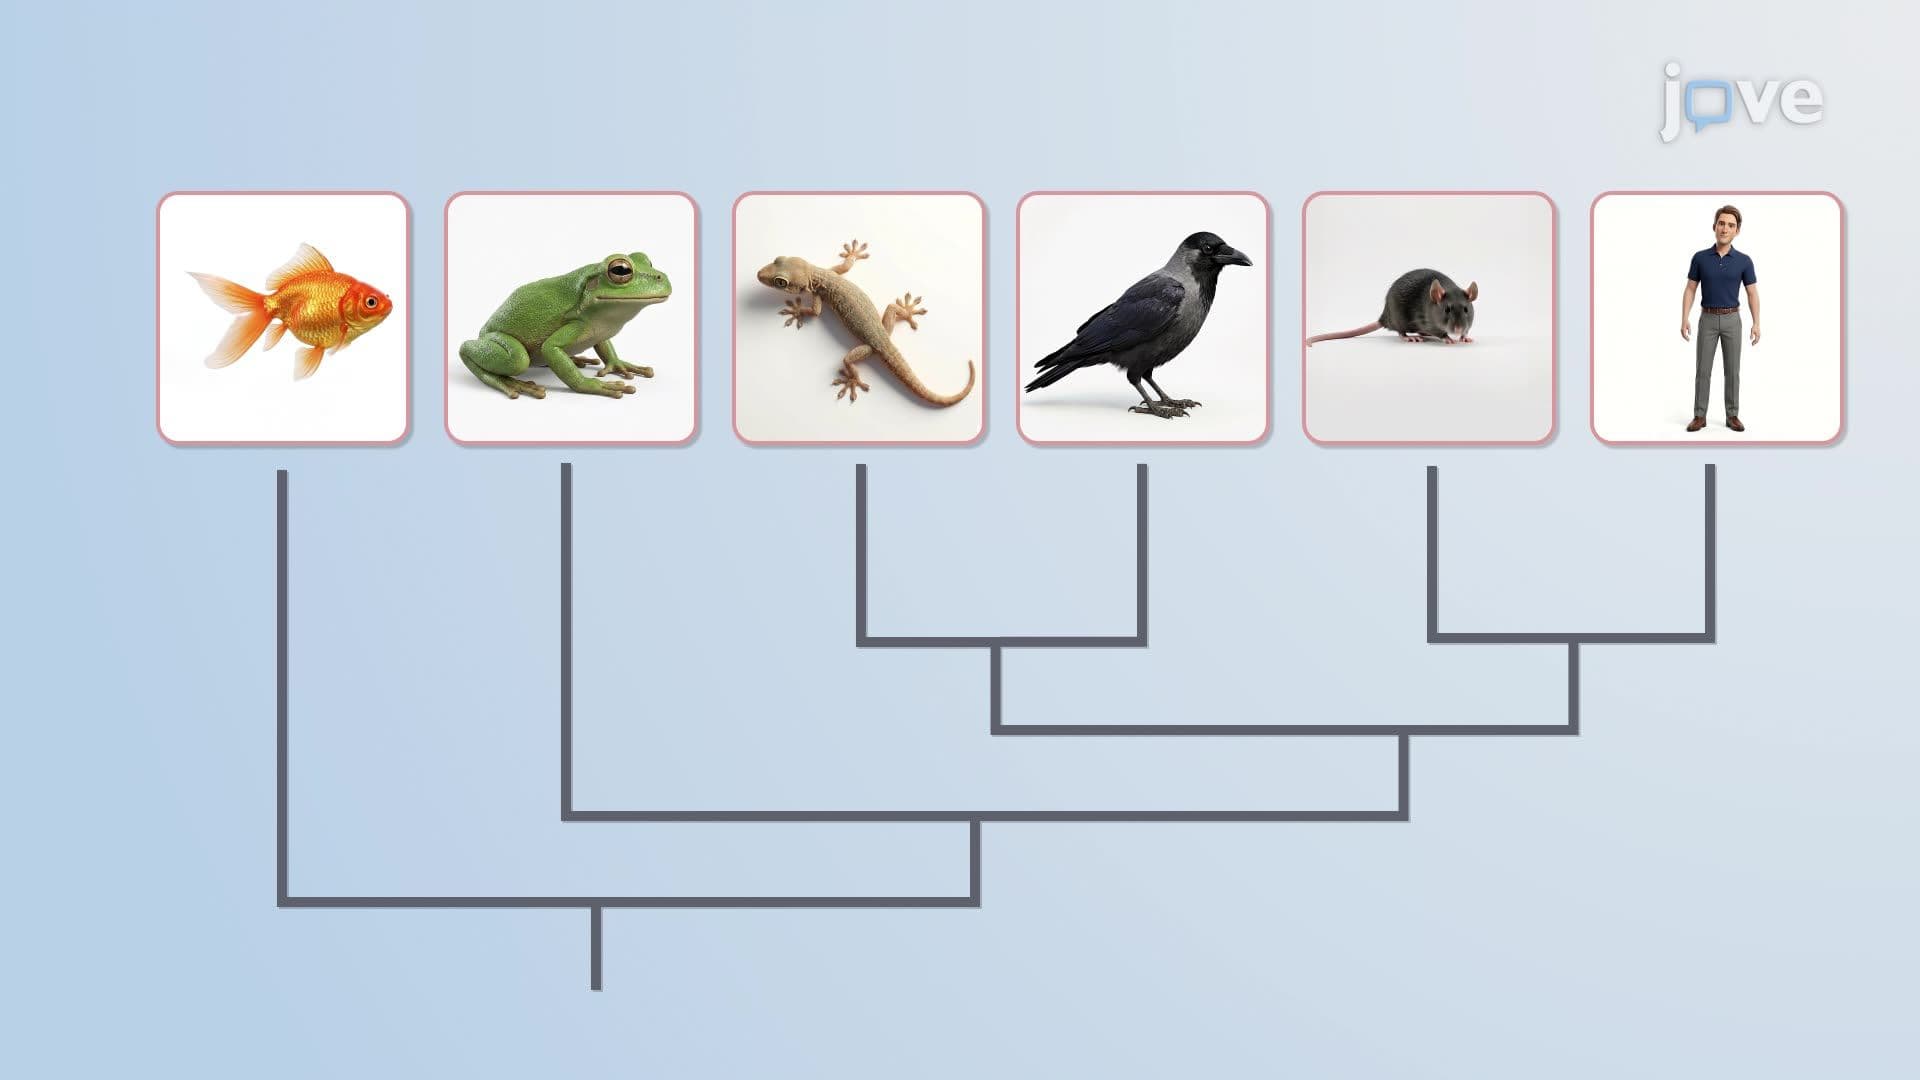
Phylogeny
 01:54
Convergent Evolution
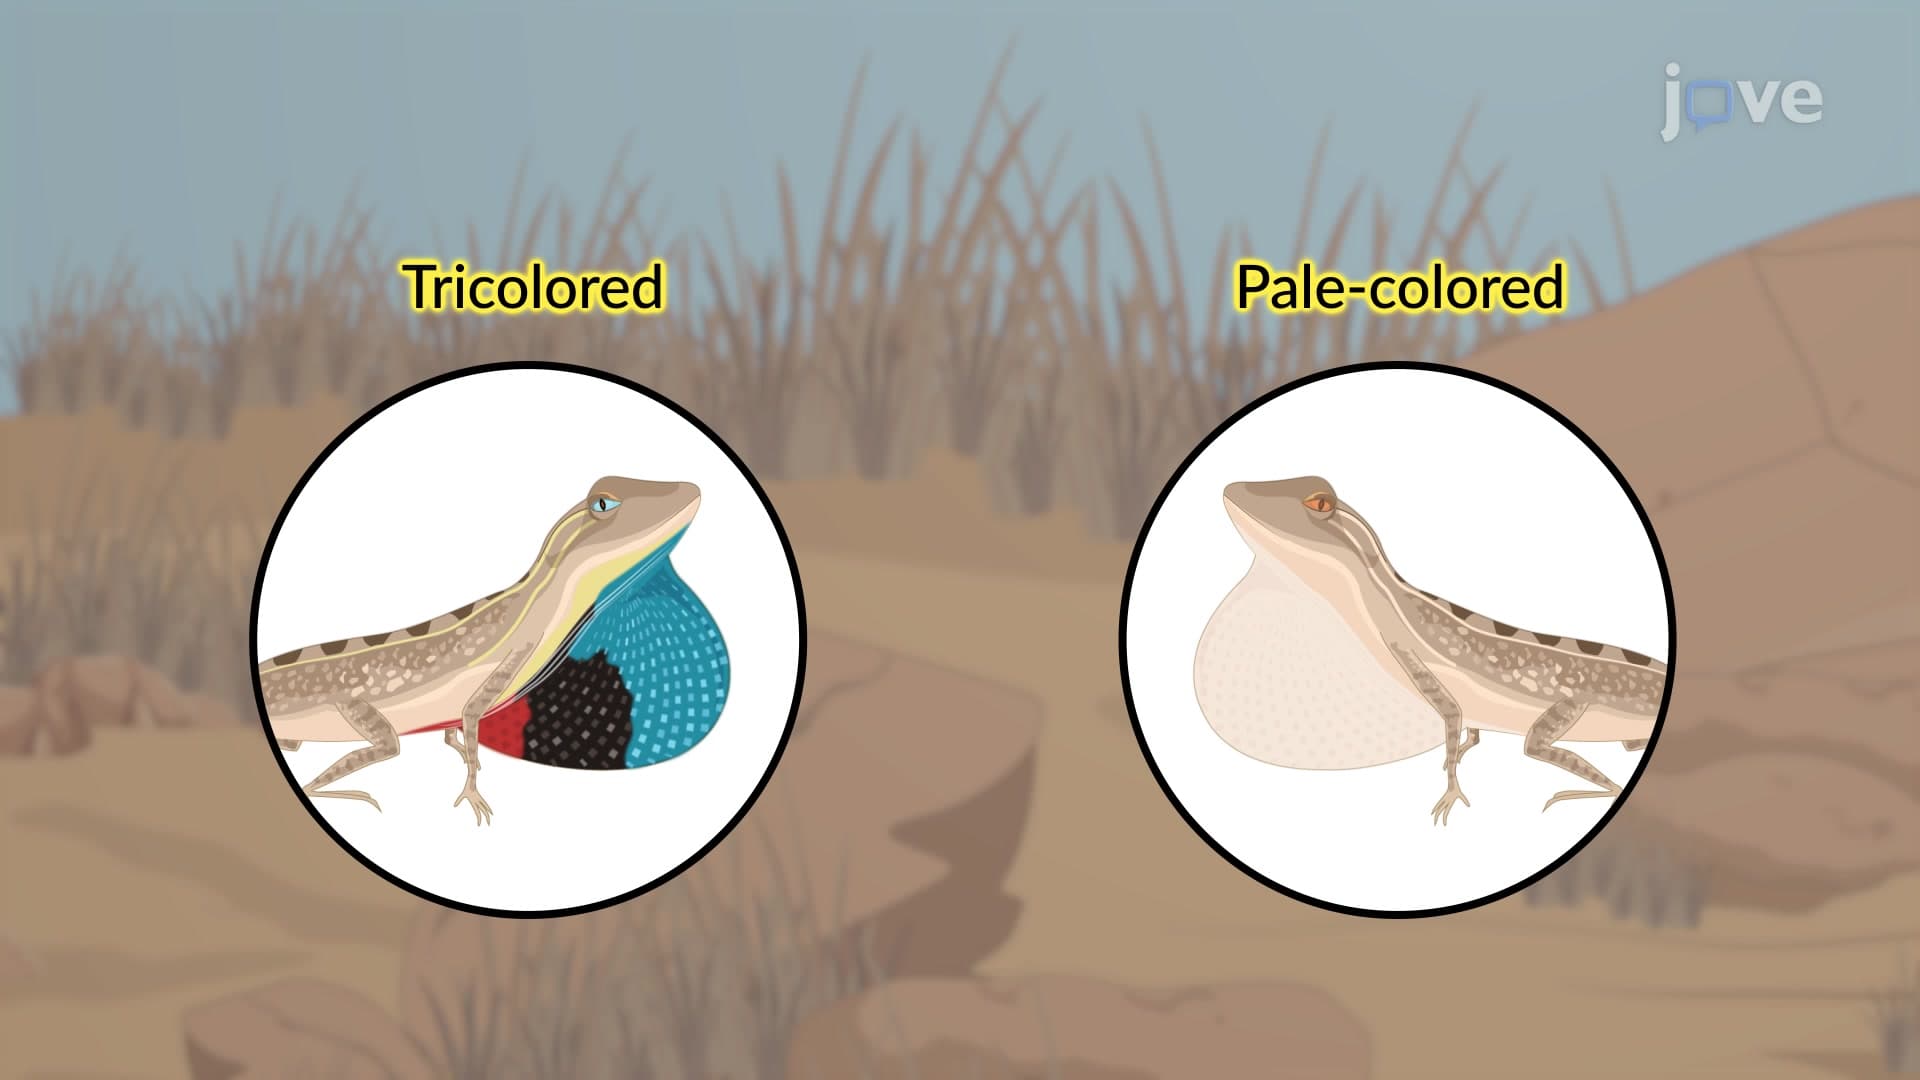
 02:54
Evolutionary Relationships through Genome Comparisons
 02:31
Synteny and Evolution
Around 80 million years ago, the human and mice lineages diverged from the common ancestor. During the course of evolution, the ancestral chromosome underwent...
 01:21
Testing a Claim about Mean: Unknown Population SD
Estimating a population mean requires the samples to be approximately normally distributed. The data should be collected from the randomly selected samples having no sampling bias. There is no specific requirement for sample size. But if the sample size is less than 30, and we don't know the population standard deviation, a different approach is used; instead...
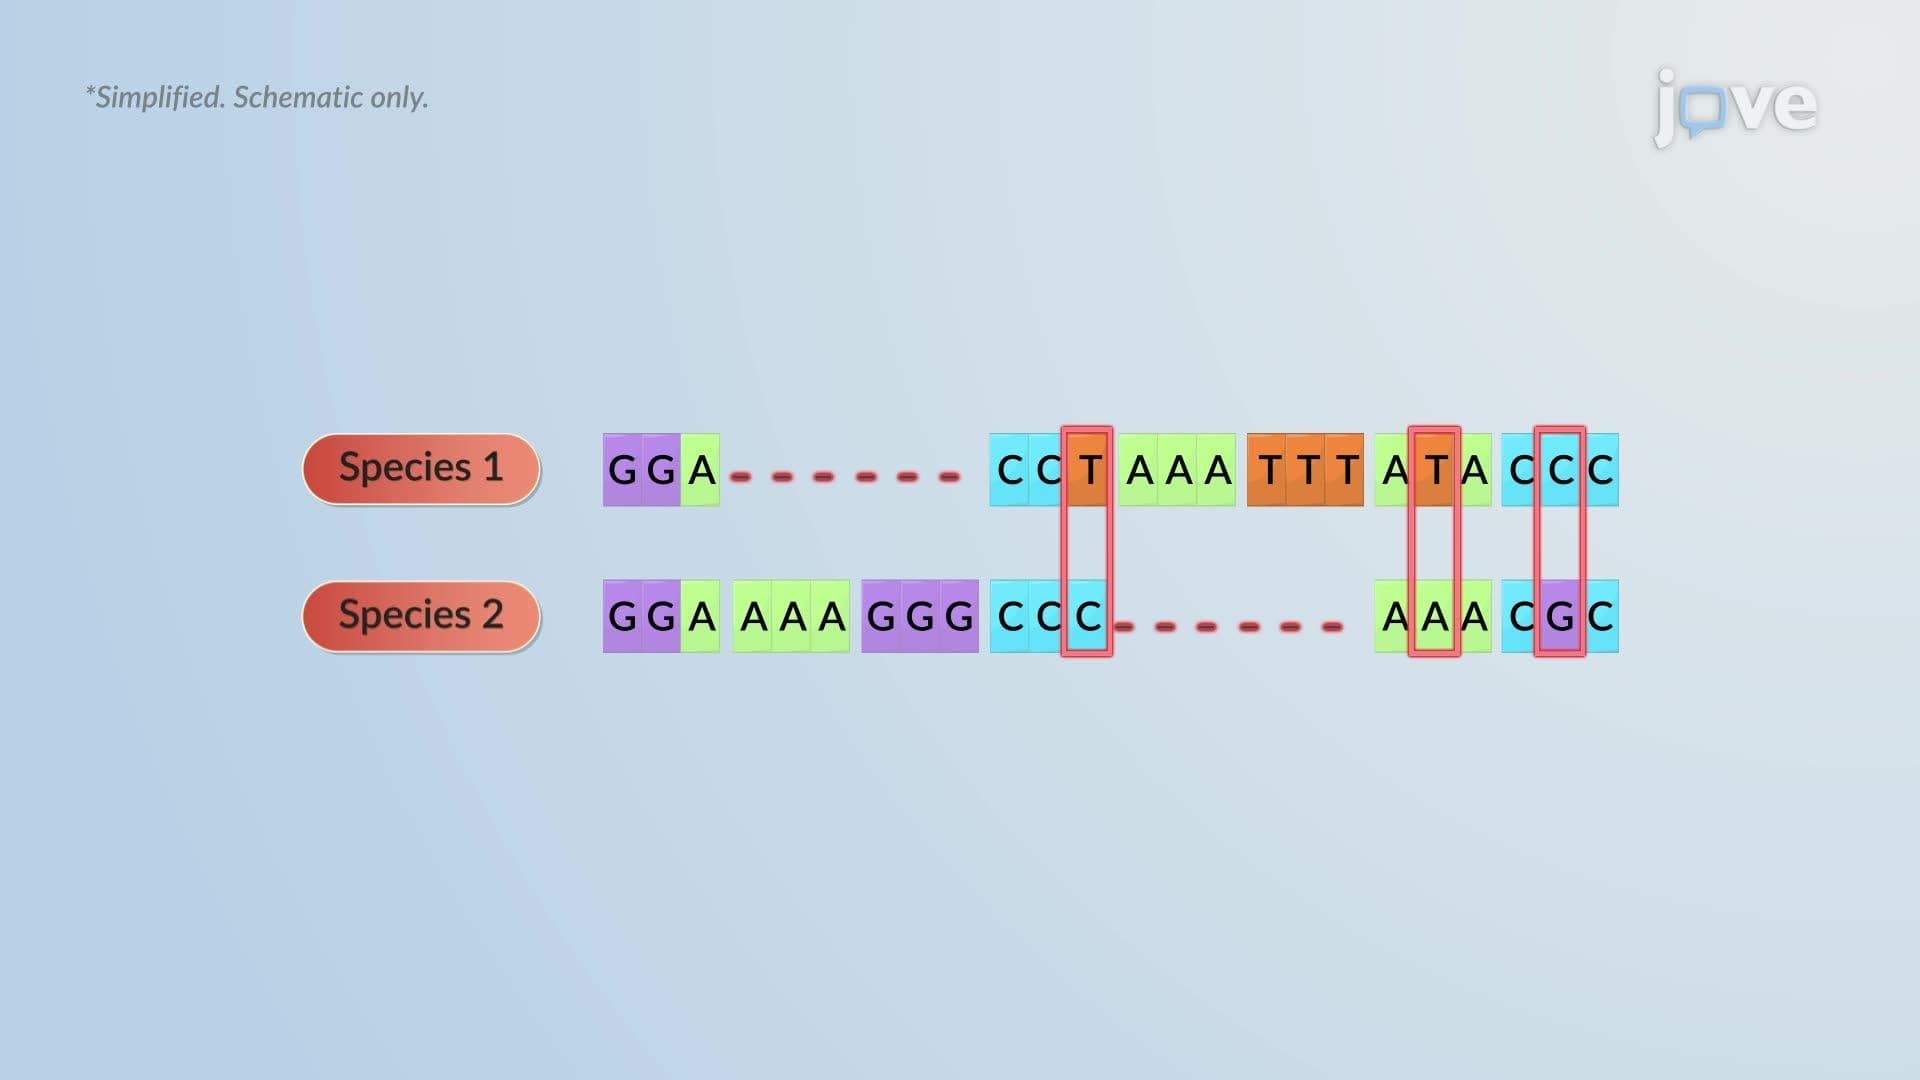 01:28
Microbial Phylogeny