Related Experiment Video
Updated: Sep 30, 2026
Visualizing and Quantifying Pharmaceutical Compounds within Skin using Coherent Raman Scattering Imaging
Published on: November 24, 2021
Quantitative structure-activity relationships (QSARs) for the prediction of skin permeation of exogenous chemicals
Hiren Patel1, Wil ten Berge, Mark T D Cronin
1School of Pharmacy and Chemistry, Liverpool John Moores University, UK.

Abstract:
Quantitative structure-activity relationships (QSARs) for the skin permeability coefficients of 158 compounds through excised human skin in vitro have been developed. A number of compounds, including hydrocortisone derivatives, were removed from the dataset as reported permeability data for these compounds was considered to be in error. QSARs developed for the dataset with the outliers removed included terms for hydrophobicity, molecular size, and hydrogen bonding. These descriptors provided an excellent fit to the data (r2 = 0.90), are easily calculated from molecular structure, and are mechanistically interpretable. Further analyses of the dataset indicated that good QSARs could be developed utilising hydrophobicity and molecular size alone, with molecular volume and molecular weight providing good quantification of molecular size.
More Related Videos
 00:05
In Silico Modeling Method for Computational Aquatic Toxicology of Endocrine Disruptors: A Software-Based Approach Using QSAR Toolbox
Published on: August 28, 2019
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 01:12
Two-Compartment Open Model: Extravascular Administration
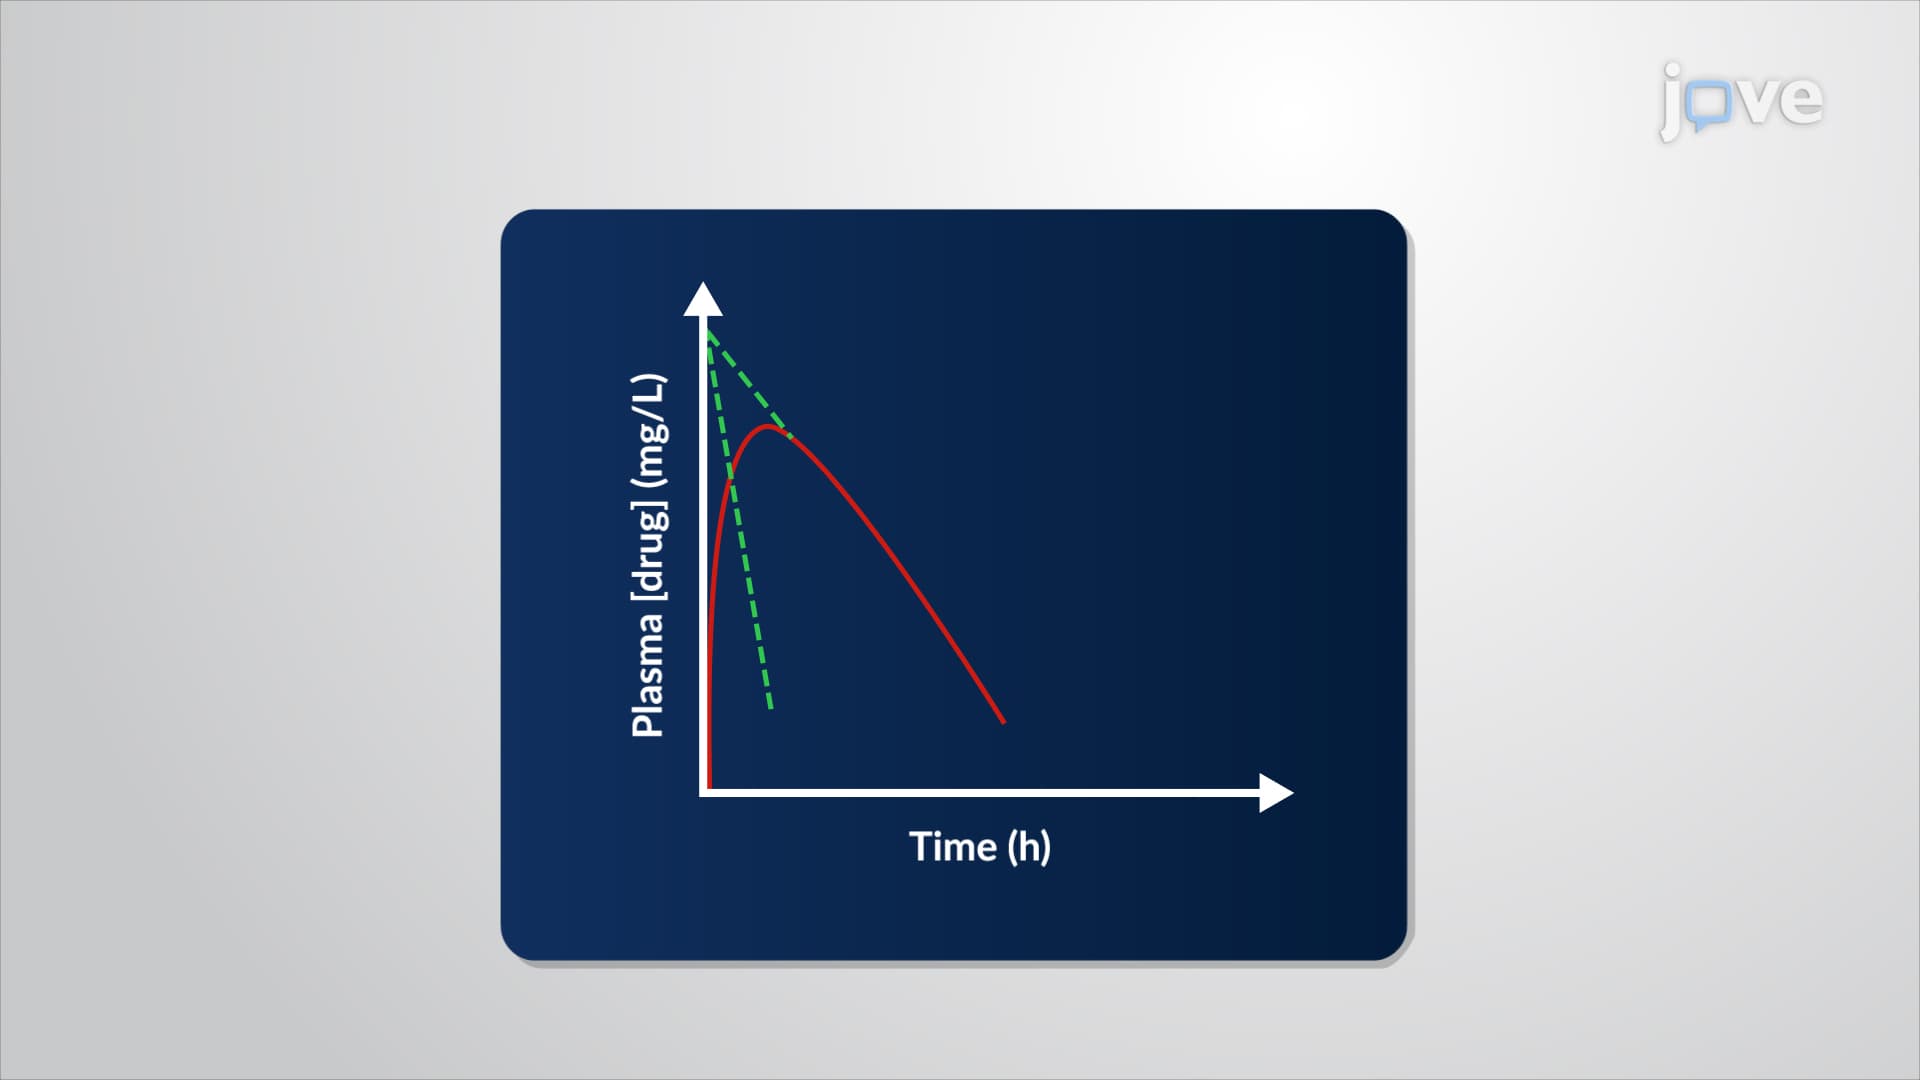
The absorption exponent (ka) indicates the speed at which the drug is...
 01:09
Methods for Studying Drug Absorption: In situ
The Doluisio method involves perfusing a prepared segment of a rat's small intestine with a solution of radiolabeled drug and a non-absorbable marker. This helps to differentiate between absorbed and non-absorbed drug concentrations. The intestinal segment is connected at both ends using tubing and syringes,...
 01:20
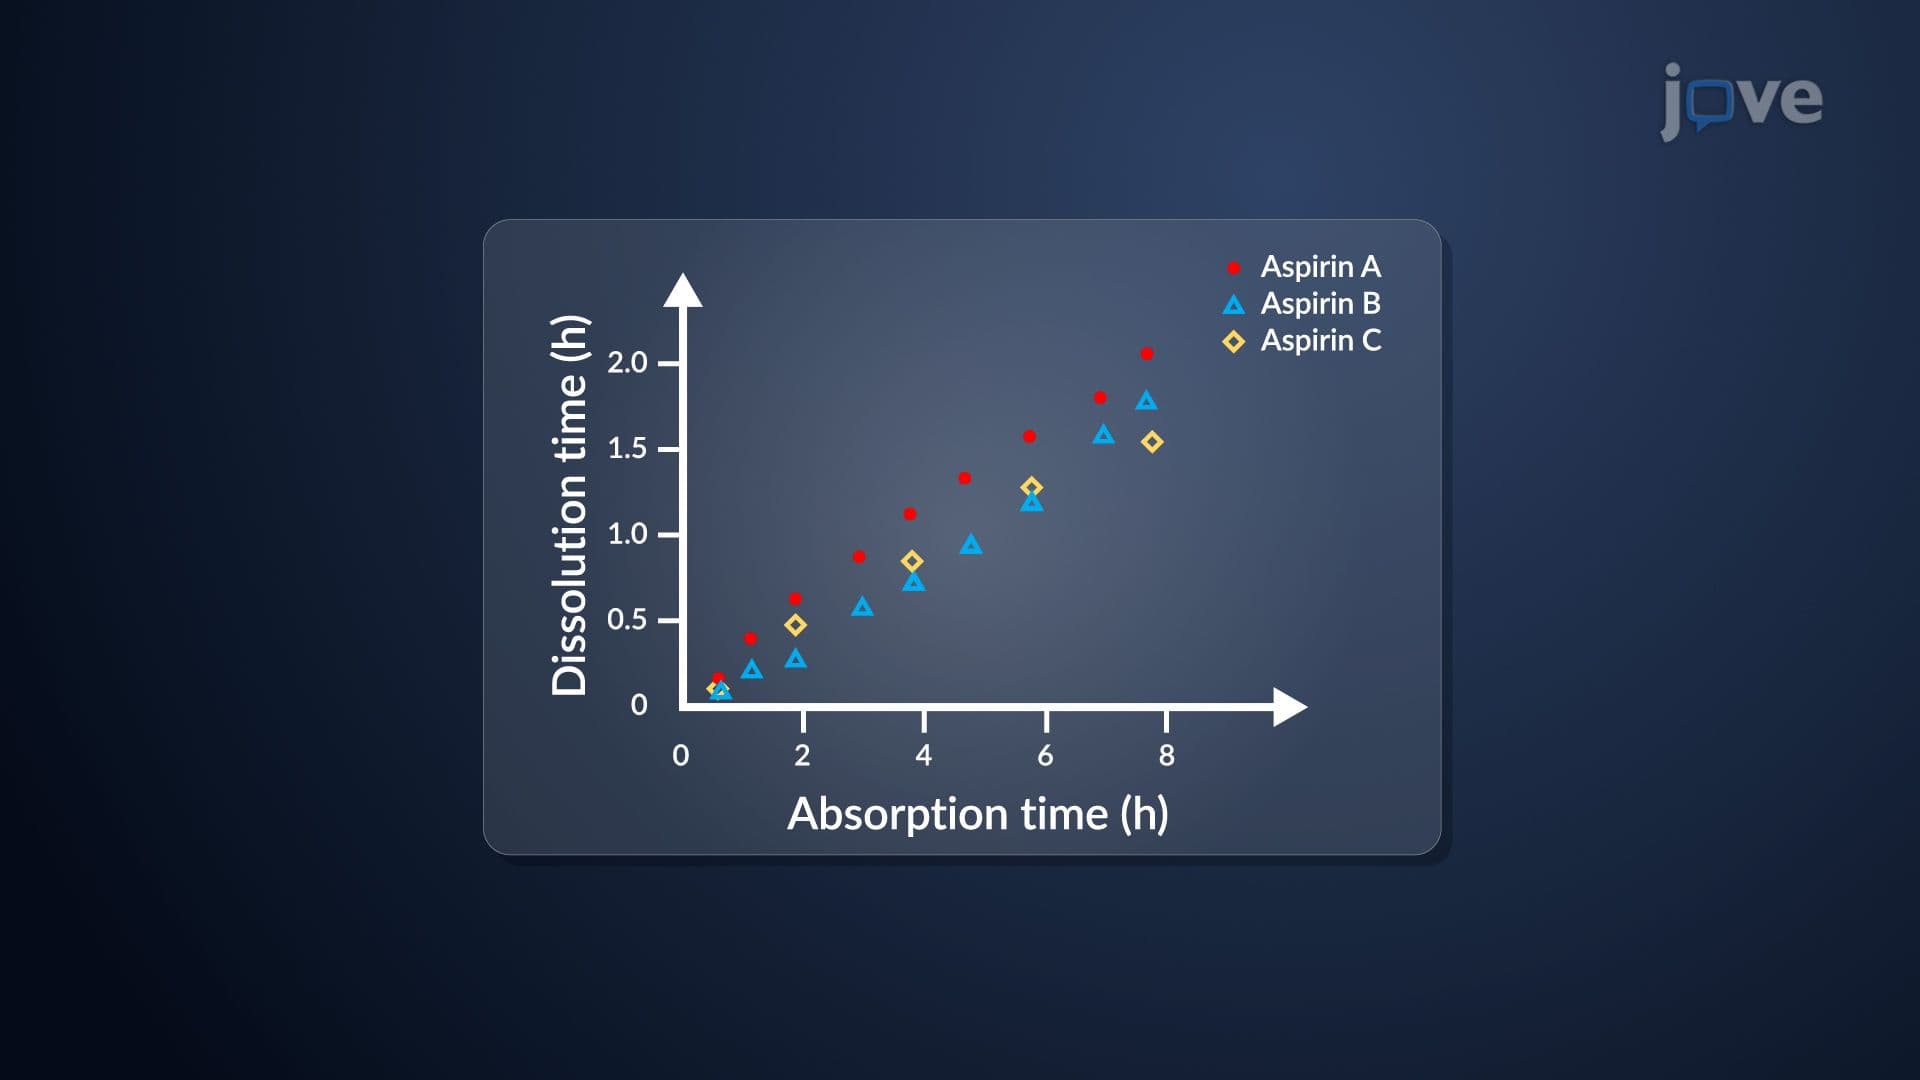
Drug Product Performance: In Vitro–In Vivo Correlation
 01:15
One-Compartment Open Model for Extravascular Administration: First-Order Absorption Model